Related Experiment Video
Updated: May 5, 2026

Refined Murine Model of Idiopathic Pulmonary Fibrosis
Published on: June 17, 2025
Natural history of pulmonary function in collagen VI-related myopathies
A Reghan Foley1, Susana Quijano-Roy, James Collins
11 Dubowitz Neuromuscular Centre, University College London Institute of Child Health and Great Ormond Street Hospital for Children, London, WC1N 1EH, UK.

Abstract:
The spectrum of clinical phenotypes associated with a deficiency or dysfunction of collagen VI in the extracellular matrix of muscle are collectively termed 'collagen VI-related myopathies' and include Ullrich congenital muscular dystrophy, Bethlem myopathy and intermediate phenotypes. To further define the clinical course of these variants, we studied the natural history of pulmonary function in correlation to motor abilities in the collagen VI-related myopathies by analysing longitudinal forced vital capacity data in a large international cohort. Retrospective chart reviews of genetically and/or pathologically confirmed collagen VI-related myopathy patients were performed at 10 neuromuscular centres: USA (n = 2), UK (n = 2), Australia (n = 2), Italy (n = 2), France (n = 1) and Belgium (n = 1). A total of 486 forced vital capacity measurements obtained in 145 patients were available for analysis. Patients at the severe end of the clinical spectrum, conforming to the original description of Ullrich congenital muscular dystrophy were easily identified by severe muscle weakness either preventing ambulation or resulting in an early loss of ambulation, and demonstrated a cumulative decline in forced vital capacity of 2.6% per year (P < 0.0001). Patients with better functional abilities, in whom walking with/without assistance was achieved, were initially combined, containing both intermediate and Bethlem myopathy phenotypes in one group. However, one subset of patients demonstrated a continuous decline in pulmonary function whereas the other had stable pulmonary function. None of the patients with declining pulmonary function attained the ability to hop or run; these patients were categorized as intermediate collagen VI-related myopathy and the remaining patients as Bethlem myopathy. Intermediate patients had a cumulative decline in forced vital capacity of 2.3% per year (P < 0.0001) whereas the relationship between age and forced vital capacity in patients with Bethlem myopathy was not significant (P = 0.1432). Nocturnal non-invasive ventilation was initiated in patients with Ullrich congenital muscular dystrophy by 11.3 years (±4.0) and in patients with intermediate collagen VI-related myopathy by 20.7 years (±1.5). The relationship between maximal motor ability and forced vital capacity was highly significant (P < 0.0001). This study demonstrates that pulmonary function profiles can be used in combination with motor function profiles to stratify collagen VI-related myopathy patients phenotypically. These findings improve our knowledge of the natural history of the collagen VI-related myopathies, enabling proactive optimization of care and preparing this patient population for clinical trials.
More Related Videos
 03:38
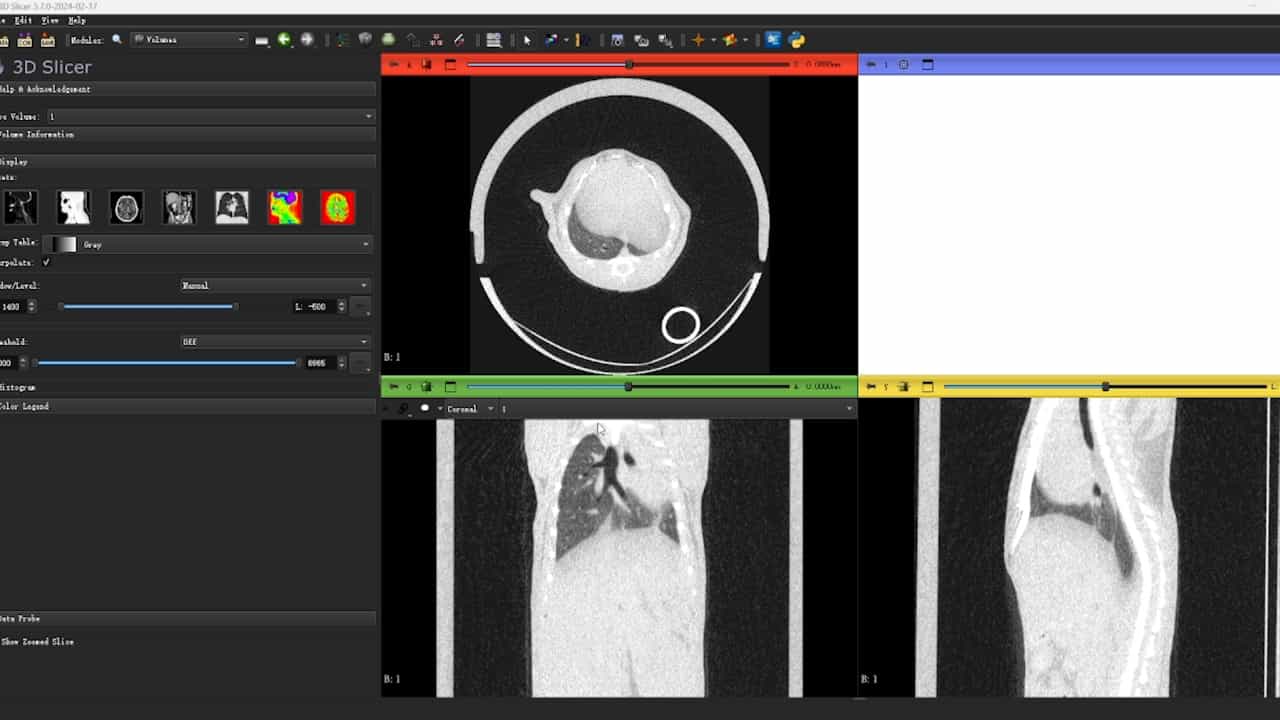
Unilateral Lung Volume Analysis Using Micro-CT for Enhanced Assessment of Pulmonary Fibrosis in Preclinical Models
Published on: June 20, 2025
 04:42
Author Spotlight: Analyzing Fibrosis Development in Chronic Lung Allograft Rejection Using Picrosirius Red Staining in Mouse Models
Published on: March 21, 2025
Related Concept Videos
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:30
Pulmonary Hypertension: Classification and Pathogenesis
There are various classifications for PH, each relating to different underlying causes and also...
 01:20
COPD: Pathogenesis and Clinical Features
The primary cause for the onset of COPD is cigarette smoking and exposure to air pollution. These hazardous factors initiate a chain reaction within the lungs, resulting in chronic inflammation, damage to the airways, and a...
 01:24
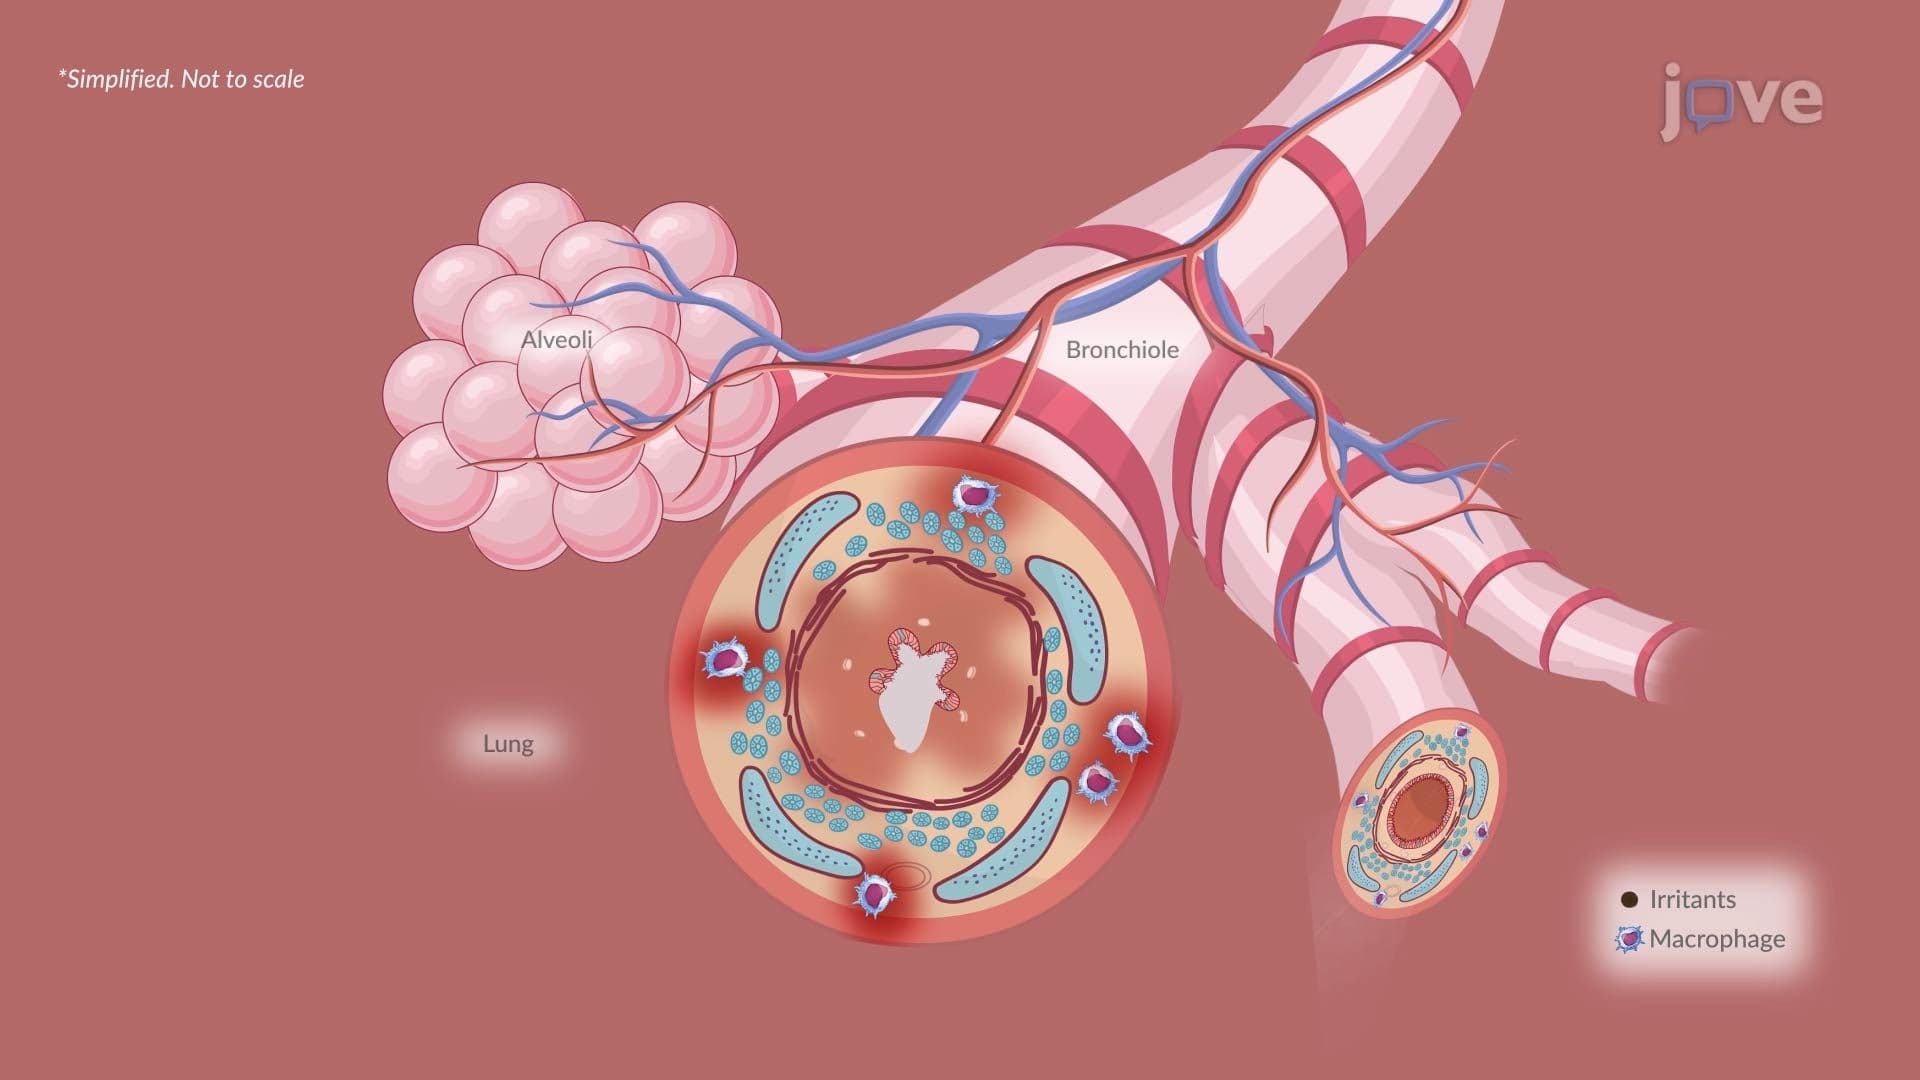
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:23
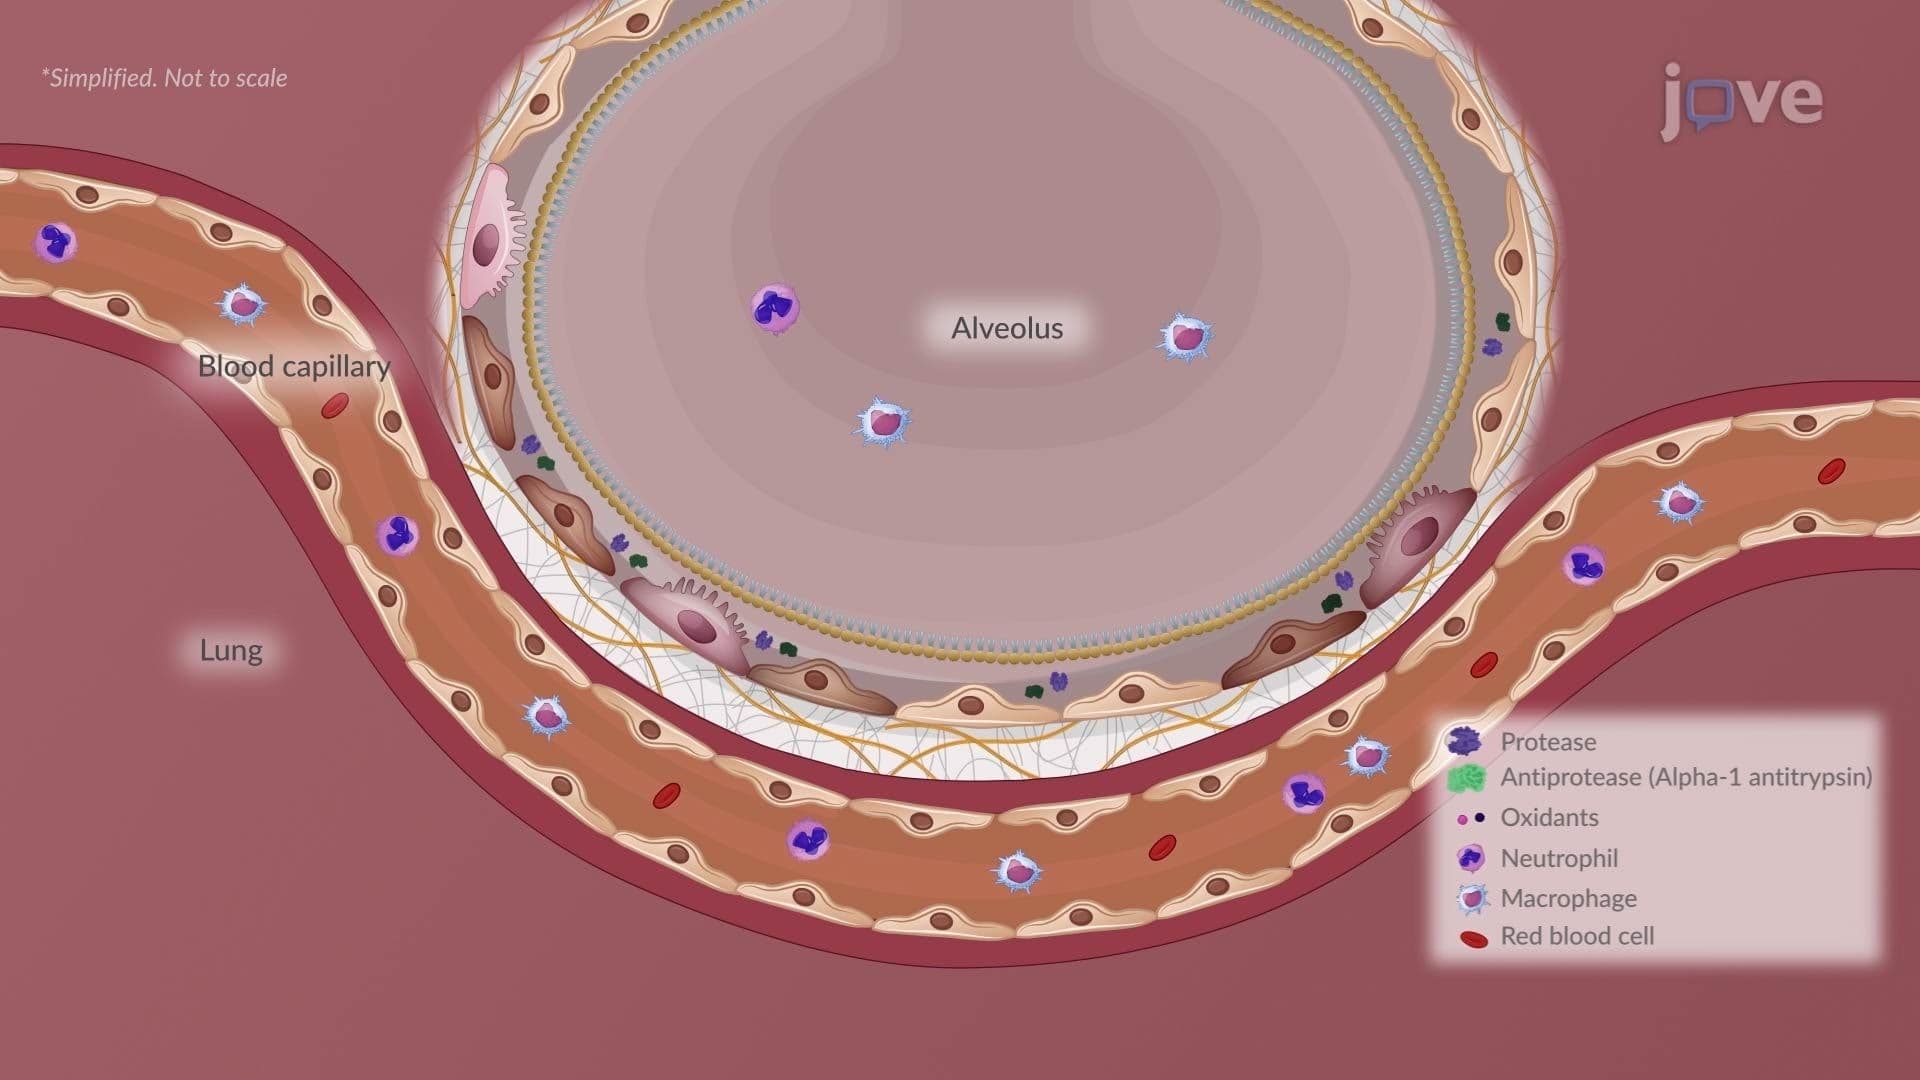
Chronic Obstructive Pulmonary Disease II: Emphysema