Characterising ChIP-seq binding patterns by model-based peak shape deconvolution
Related Experiment Video
Updated: May 5, 2026

A Novel Bayesian Change-point Algorithm for Genome-wide Analysis of Diverse ChIPseq Data Types
Published on: December 10, 2012
Marco-Antonio Mendoza-Parra1, Malgorzata Nowicka, Wouter Van Gool
1Equipe Labellisée Ligue Contre le Cancer, Department of Functional Genomics and Cancer, Institut de Génétique et de Biologie Moléculaire et Cellulaire (IGBMC)/CNRS/INSERM/Université de Strasbourg, BP 10142, Illkirch Cedex 67404, France. marco@igbmc.fr.

Background:
Chromatin immunoprecipitation combined with massive parallel sequencing (ChIP-seq) is widely used to study protein-chromatin interactions or chromatin modifications at genome-wide level. Sequence reads that accumulate locally at the genome (peaks) reveal loci of selectively modified chromatin or specific sites of chromatin-binding factors. Computational approaches (peak callers) have been developed to identify the global pattern of these sites, most of which assess the deviation from background by applying distribution statistics.
Results:
We have implemented MeDiChISeq, a regression-based approach, which--by following a learning process--defines a representative binding pattern from the investigated ChIP-seq dataset. Using this model MeDiChISeq identifies significant genome-wide patterns of chromatin-bound factors or chromatin modification. MeDiChISeq has been validated for various publicly available ChIP-seq datasets and extensively compared with other peak callers.
Conclusions:
MeDiChI-Seq has a high resolution when identifying binding events, a high degree of peak-assessment reproducibility in biological replicates, a low level of false calls and a high true discovery rate when evaluated in the context of gold-standard benchmark datasets. Importantly, this approach can be applied not only to 'sharp' binding patterns--like those retrieved for transcription factors (TFs)--but also to the broad binding patterns seen for several histone modifications. Notably, we show that at high sequencing depths, MeDiChISeq outperforms other algorithms due to its powerful peak shape recognition capacity which facilitates discerning significant binding events from spurious background enrichment patterns that are enhanced with increased sequencing depths.
Related Concept Videos
 01:20
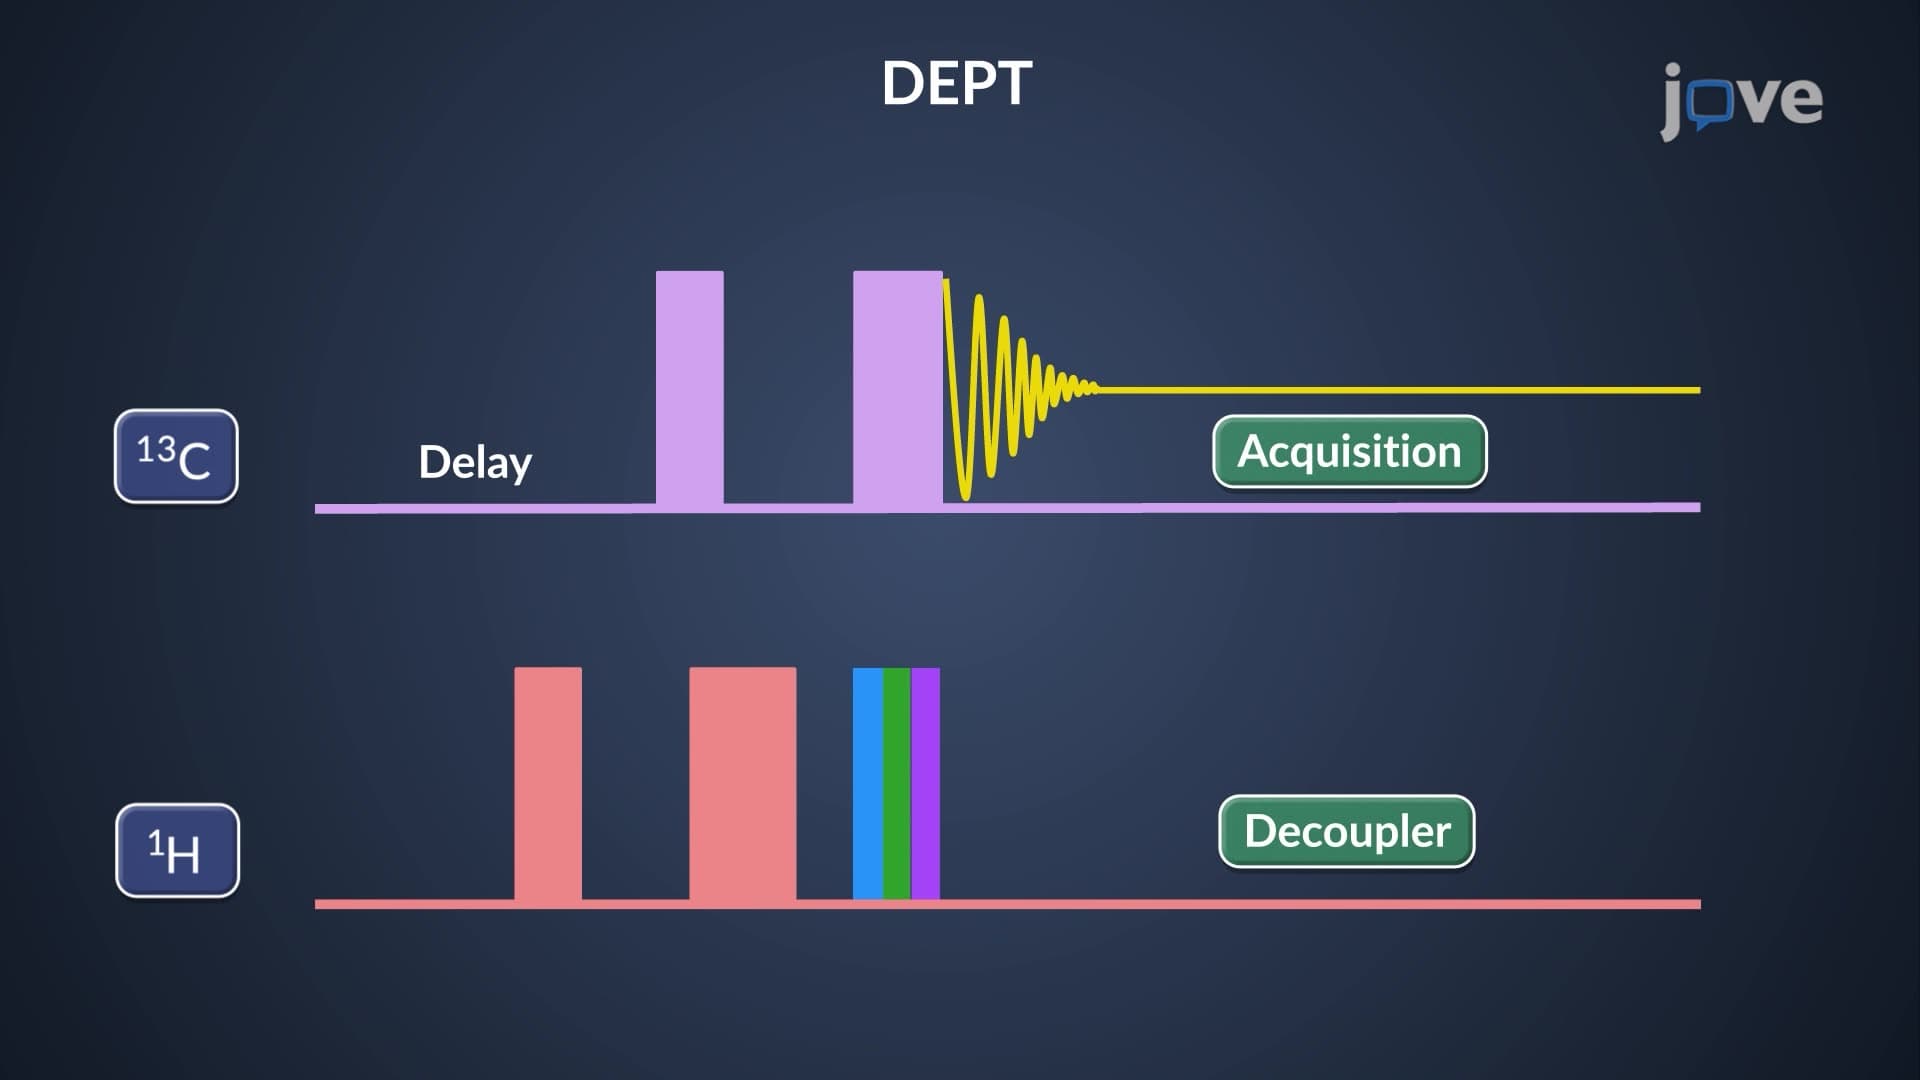
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:04
¹³C NMR: ¹H–¹³C Decoupling
A broadband decoupling technique is used to simplify these complex, sometimes overlapping, signals. Broadband decoupling relies on a...