Related Experiment Video
Updated: May 5, 2026
Ultrafast Time-resolved Near-IR Stimulated Raman Measurements of Functional π-conjugate Systems
Published on: February 10, 2020
Lowest triplet (n,π*) state of 2-cyclohexen-1-one: characterization by cavity ringdown spectroscopy and
Michael O McAnally1, Katherine L Zabronsky, Daniel J Stupca
1Department of Chemistry, University of Wisconsin-Eau Claire, Eau Claire, Wisconsin 54702-4004, USA.

Abstract:
The cavity ringdown (CRD) absorption spectrum of 2-cyclohexen-1-one (2CHO) was recorded over the range 401.5-410.5 nm in a room-temperature gas cell. The very weak band system (ε ≤ 0.1 M(-1) cm(-1)) in this spectral region is due to the T1(n, π*) ← S0 electronic transition. The 0(0)(0) origin band was assigned to the feature observed at 24,558.8 ± 0.3 cm(-1). We have assigned 46 vibronic transitions in a region extending from -200 to +350 cm(-1) relative to the origin band. For the majority of these transitions, we have made corresponding assignments in the spectrum of the deuterated derivative 2CHO-2,6,6-d3. From the assignments, we determined fundamental frequencies for several vibrational modes in the T1(n, π*) excited state of 2CHO, including the lowest ring-twisting (99.6 cm(-1)) and ring-bending (262.2 cm(-1)) modes. These values compare to fundamentals of 122.2 cm(-1) and 251.9 cm(-1), respectively, determined previously for the isoconfigurational S1(n, π*) excited state of 2CHO and 99 cm(-1) and 248 cm(-1), respectively, for the S0 ground state. With the aid of quantum-mechanical calculations, we have also ascertained descriptions for these two modes, thereby resolving ambiguities appearing in the previous literature. The ring-twisting mode (ν39) contains a significant contribution from O=C-C=C torsion, whereas the ring-bending mode (ν38 in the ground state) involves mainly the motion of C-5 with respect to the plane containing the other heavy atoms. The CRD spectroscopic data for the T1(n, π*) state have allowed us to benchmark several computational methods for treating excited states, including time-dependent density functional theory and an equation-of-motion coupled cluster method. In turn, the computational results provide an explanation for observed differences in the T1(n, π*) vs. S1(n, π*) ring frequencies.
Related Concept Videos
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:18
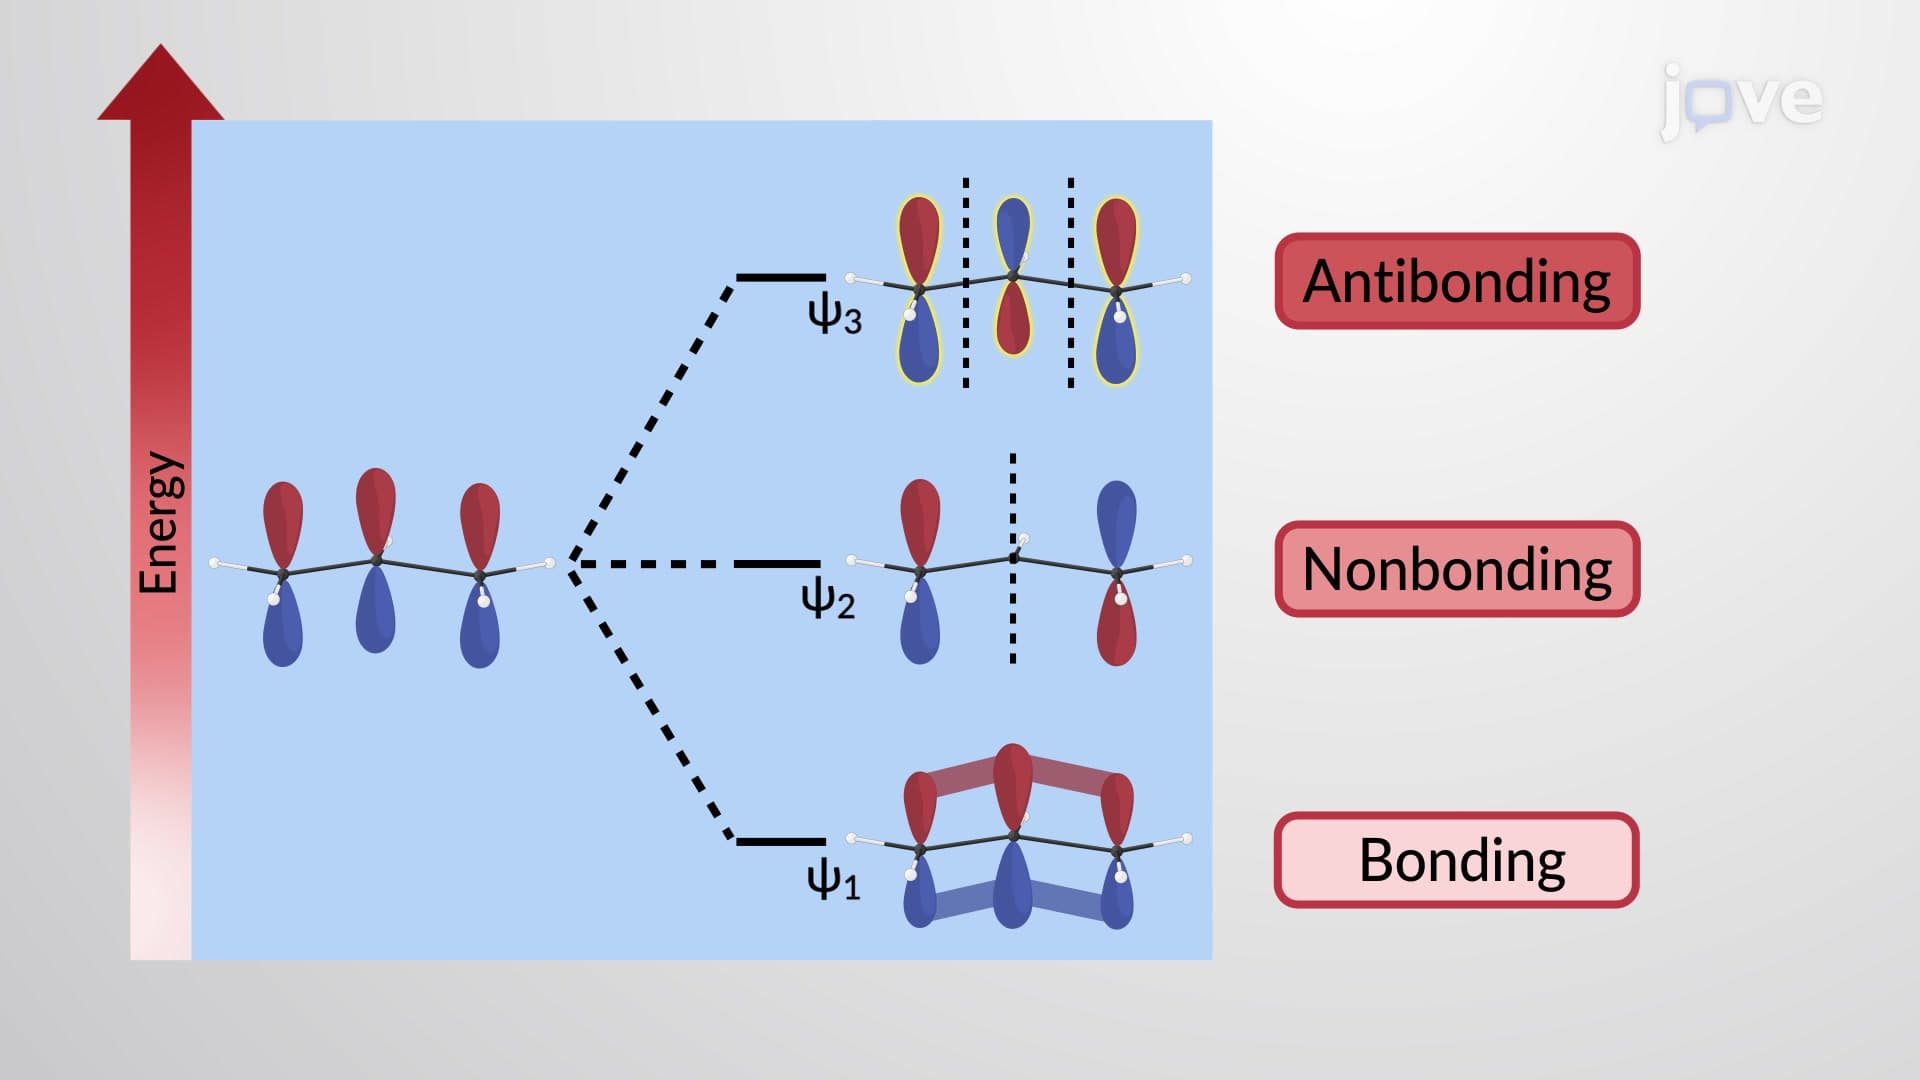
π Molecular Orbitals of the Allyl Cation and Anion
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:18
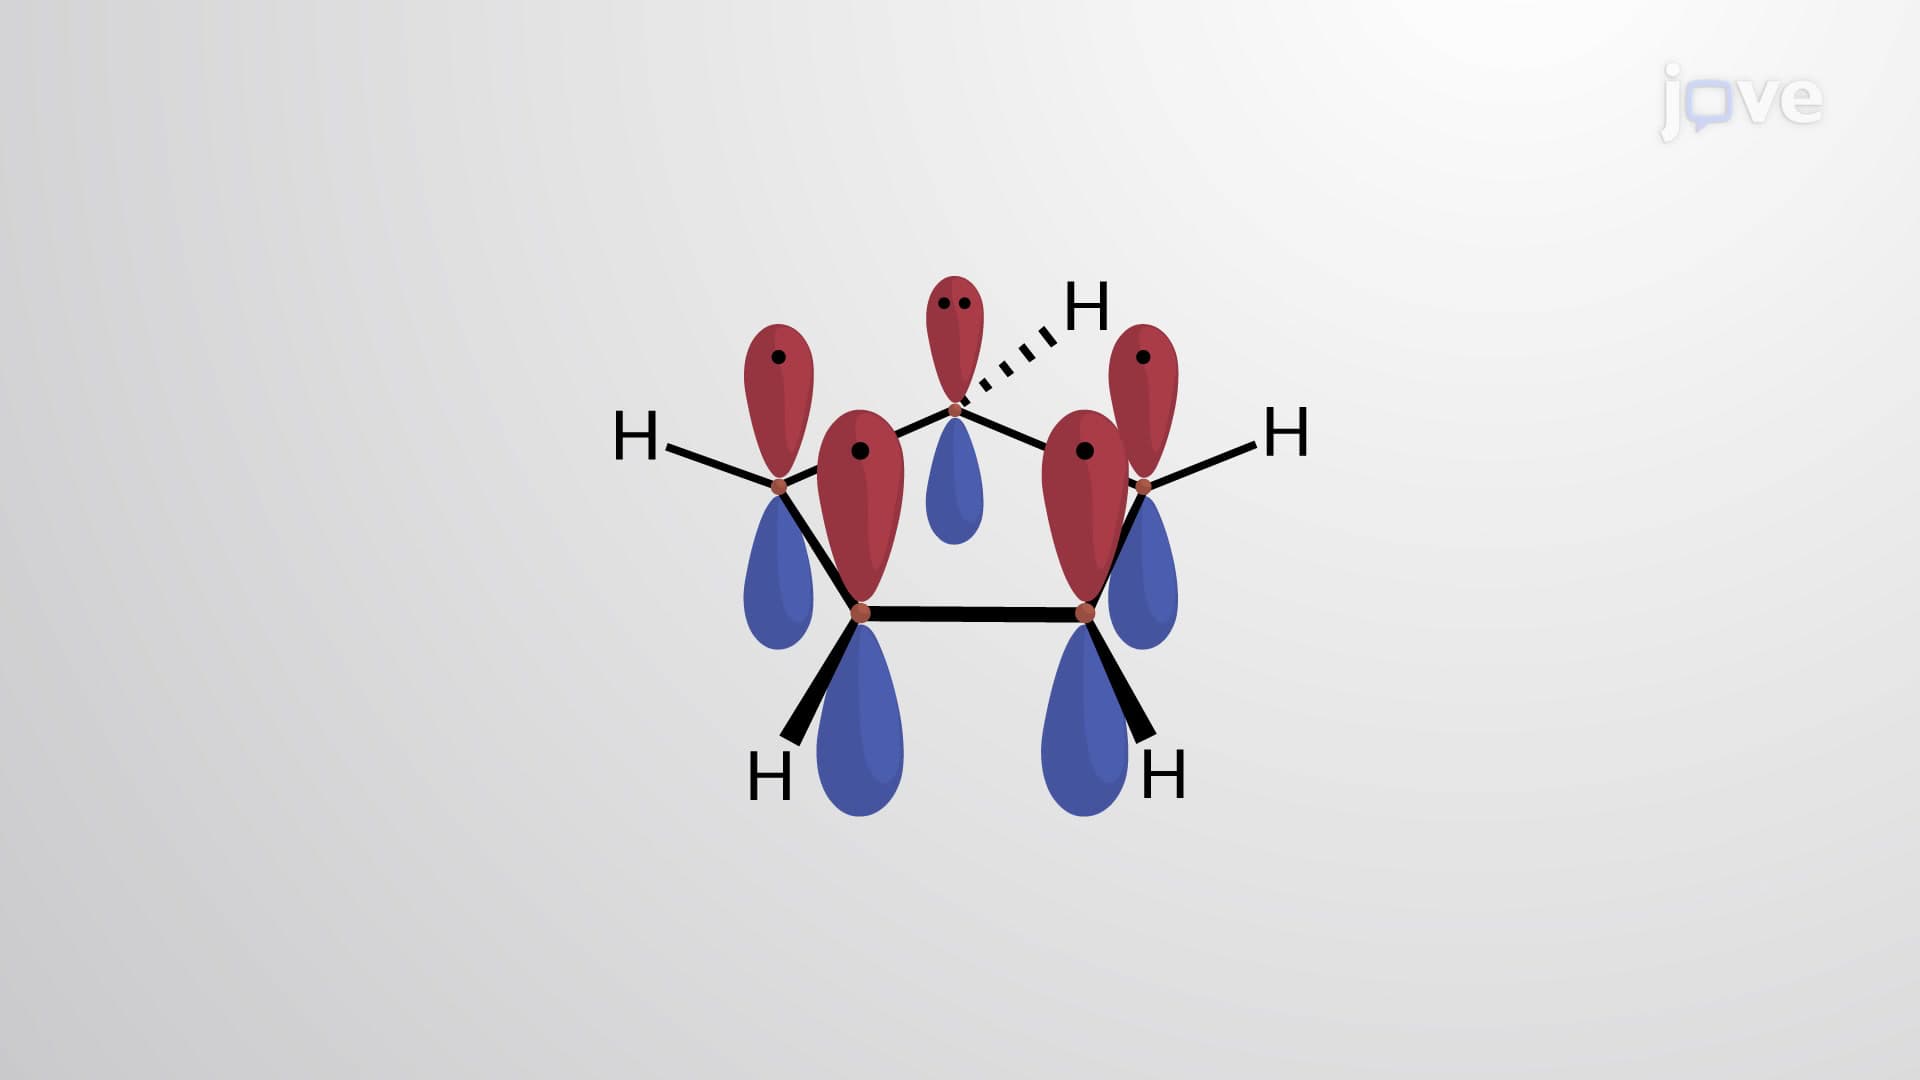
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...