Related Experiment Video
Updated: May 4, 2026


Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Predicting protein-ligand interactions based on chemical preference features with its application to new D-amino acid
Mingzhu Zhao, Hao-Teng Chang, Qiang Zhou
1State Key Laboratory of Microbial Metabolism and College of Life Sciences and Biotechnology, Shanghai Jiao Tong University, Shanghai 200240, China. zpliu@sibs.ac.cn.

Abstract:
In silico prediction of the new drug-target interactions from existing databases is of important value for the drug discovery process. Currently, the amount of protein targets that have been identified experimentally is still very small compared with the entire human proteins. In order to predict protein-ligand interactions in an accurate manner, we have developed a support vector machine (SVM) model based on the chemical-protein interactions from STITCH. New features from ligand chemical space and interaction networks have been selected and encoded as the feature vectors for SVM analysis. Both the 5-fold cross validation and independent test show high predictive accuracy that outperforms the state-of-the-art method based on ligand similarity. Moreover, 91 distinct pairs of features have been selected to rebuild a simplifier model, which still maintains the same performance as that based on all 332 features. Then, this refined model is used to search for the potential D-amino acid oxidase inhibitors from STITCH database and the predicted results are finally validated by our wet experiments. Out of 10 candidates obtained, seven D-amino acid oxidase inhibitors have been verified, in which four are newly found in the present study, and one may have a new application in therapy of psychiatric disorders other than being an antineoplastic agent. Clearly, our model is capable of predicting potential new drugs or targets on a large scale with high efficiency.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 00:49
Ligand Binding and Linkage
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:22
Protein-Drug Binding: Determination Methods
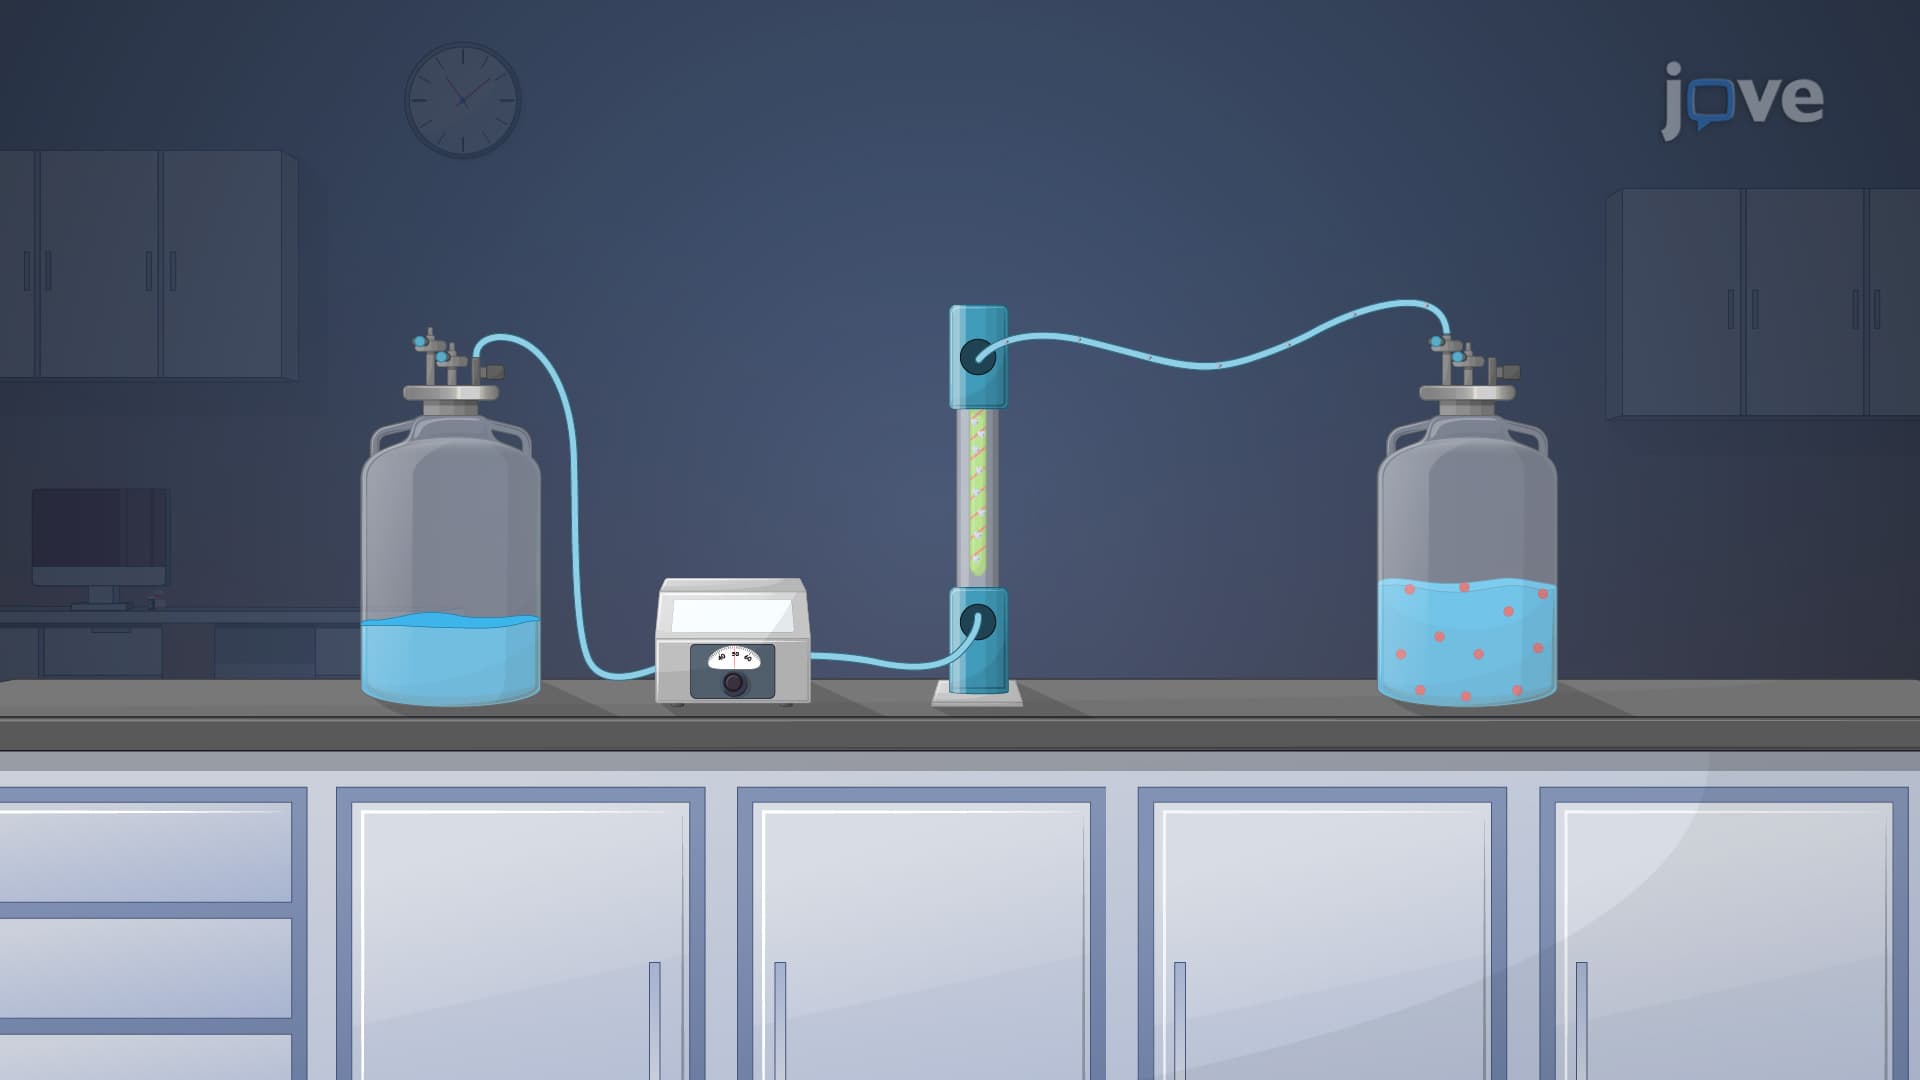
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...