Related Experiment Video
Updated: May 3, 2026

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Assessing hERG pore models as templates for drug docking using published experimental constraints: the inactivated
Christopher E Dempsey1, Dominic Wright, Charlotte K Colenso
1School of Biochemistry, Medical Sciences Building, University of Bristol , University Walk, Bristol BS8 1TD, U.K.

Many structurally and therapeutically diverse drugs interact with the human heart K+ channel hERG by binding within the K+ permeation pathway of the open channel, leading to drug-induced 'long QT syndrome'. Drug binding to hERG is often stabilized by inactivation gating. In the absence of a crystal structure, hERG pore homology models have been used to characterize drug interactions. Here we assess potentially inactivated states of the bacterial K+ channel, KcsA, as templates for inactivated state hERG pore models in the context of drug binding using computational docking. Although Flexidock and GOLD docking produced low energy score poses in the models tested, each method selected a MthK K+ channel-based model over models based on the putative inactivated state KcsA structures for each of the 9 drugs tested. The variety of docking poses found indicates that an optimal arrangement for drug binding of aromatic side chains in the hERG pore can be achieved in several different configurations. This plasticity of the drug "binding site" is likely to be a feature of the hERG inactivated state. The results demonstrate that experimental data on specific drug interactions can be used as structural constraints to assess and refine hERG homology models.
Many structurally and therapeutically diverse drugs interact with the human heart K+ channel hERG by binding within the K+ permeation pathway of the open channel, leading to drug-induced 'long QT syndrome'. Drug binding to hERG is often stabilized by inactivation gating. In the absence of a crystal structure, hERG pore homology models have been used to characterize drug interactions. Here we assess potentially inactivated states of the bacterial K+ channel, KcsA, as templates for inactivated state hERG pore models in the context of drug binding using computational docking. Although Flexidock and GOLD docking produced low energy score poses in the models tested, each method selected a MthK K+ channel-based model over models based on the putative inactivated state KcsA structures for each of the 9 drugs tested. The variety of docking poses found indicates that an optimal arrangement for drug binding of aromatic side chains in the hERG pore can be achieved in several different configurations. This plasticity of the drug "binding site" is likely to be a feature of the hERG inactivated state. The results demonstrate that experimental data on specific drug interactions can be used as structural constraints to assess and refine hERG homology models.
More Related Videos
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 01:18
Patch Clamp
In this method, a glass micropipette containing electrolyte solution is tightly sealed against a small portion of the cell membrane. As a result, a patch of the cell...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:29
Drug-Receptor Interactions
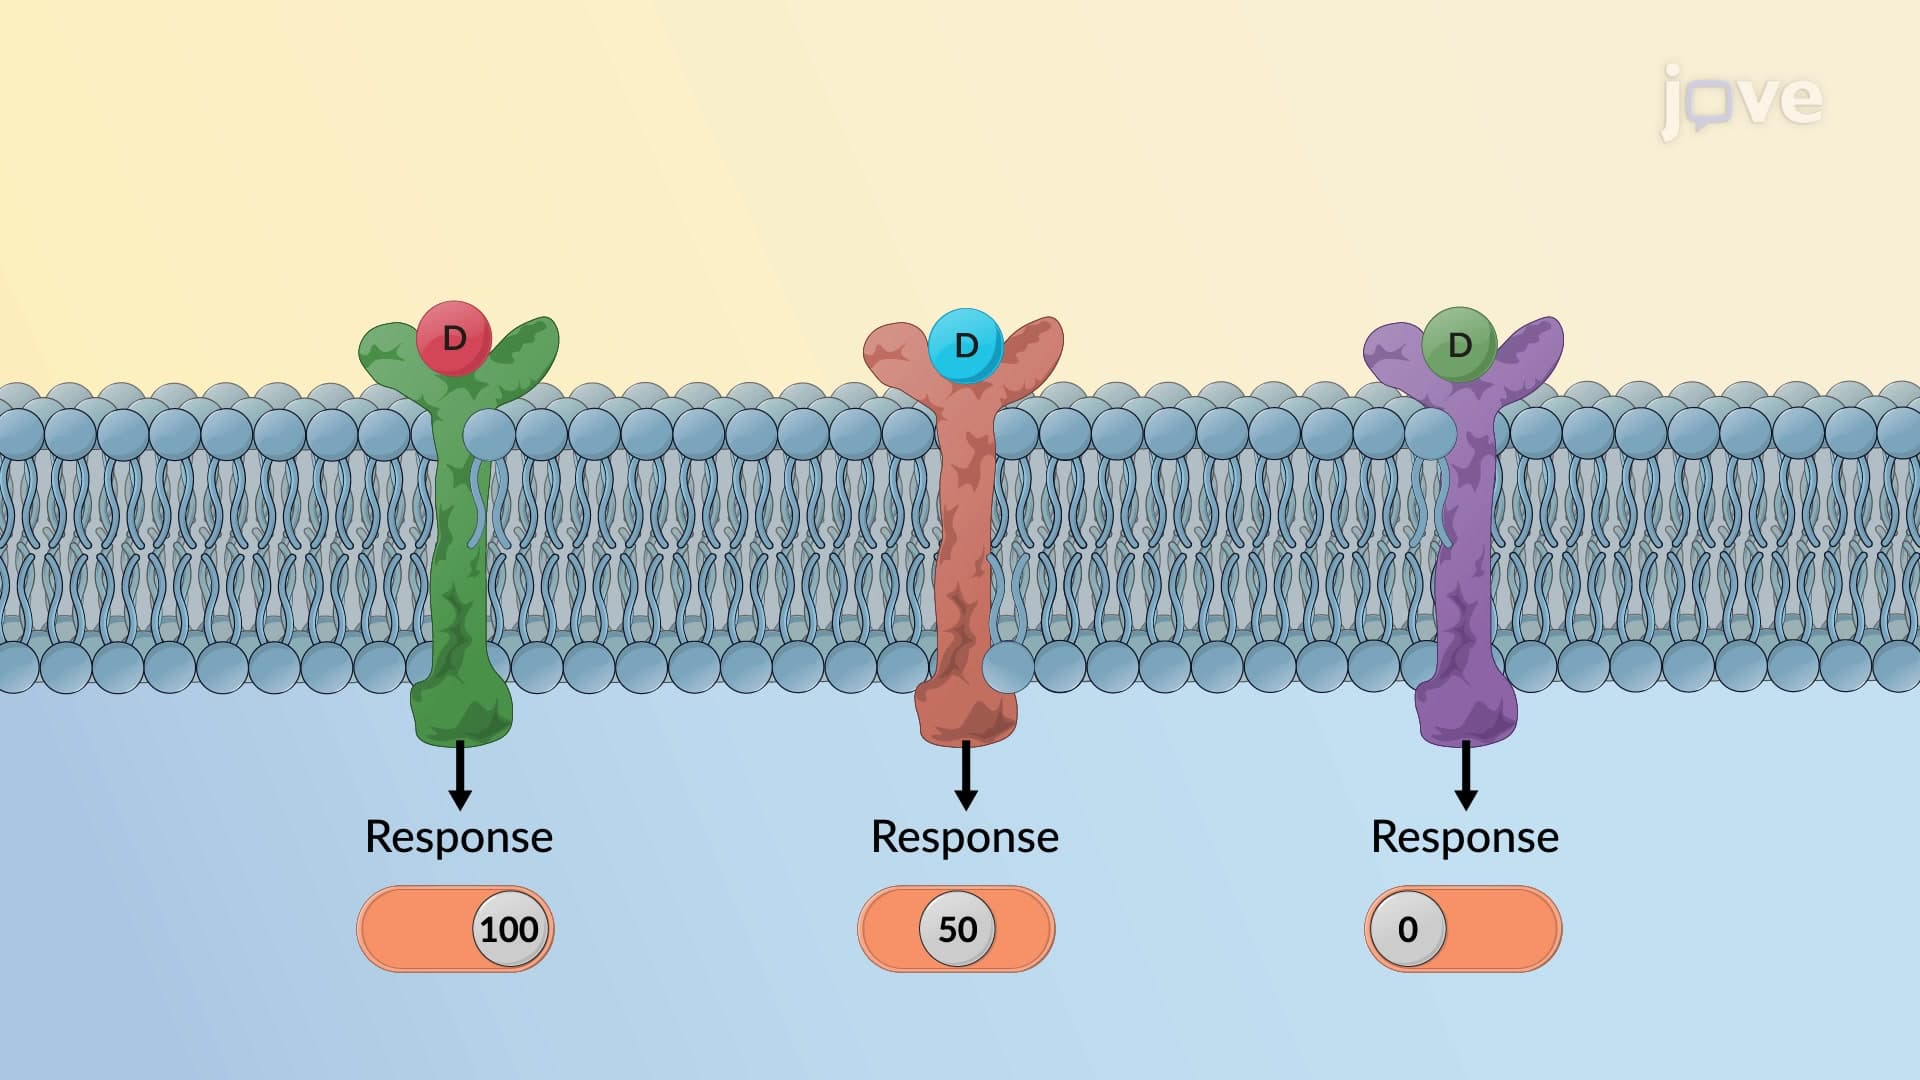
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....