Related Experiment Video
Updated: May 2, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
An interatomic potential for saturated hydrocarbons based on the modified embedded-atom method
S Nouranian1, M A Tschopp, S R Gwaltney
1Center for Advanced Vehicular Systems (CAVS), Mississippi State University, Mississippi State, MS 39762, USA.

Abstract:
In this work, we developed an interatomic potential for saturated hydrocarbons using the modified embedded-atom method (MEAM), a reactive semi-empirical many-body potential based on density functional theory and pair potentials. We parameterized the potential by fitting to a large experimental and first-principles (FP) database consisting of (1) bond distances, bond angles, and atomization energies at 0 K of a homologous series of alkanes and their select isomers from methane to n-octane, (2) the potential energy curves of H2, CH, and C2 diatomics, (3) the potential energy curves of hydrogen, methane, ethane, and propane dimers, i.e., (H2)2, (CH4)2, (C2H6)2, and (C3H8)2, respectively, and (4) pressure-volume-temperature (PVT) data of a dense high-pressure methane system with the density of 0.5534 g cc(-1). We compared the atomization energies and geometries of a range of linear alkanes, cycloalkanes, and free radicals calculated from the MEAM potential to those calculated by other commonly used reactive potentials for hydrocarbons, i.e., second-generation reactive empirical bond order (REBO) and reactive force field (ReaxFF). MEAM reproduced the experimental and/or FP data with accuracy comparable to or better than REBO or ReaxFF. The experimental PVT data for a relatively large series of methane, ethane, propane, and butane systems with different densities were predicted reasonably well by the MEAM potential. Although the MEAM formalism has been applied to atomic systems with predominantly metallic bonding in the past, the current work demonstrates the promising extension of the MEAM potential to covalently bonded molecular systems, specifically saturated hydrocarbons and saturated hydrocarbon-based polymers. The MEAM potential has already been parameterized for a large number of metallic unary, binary, ternary, carbide, nitride, and hydride systems, and extending it to saturated hydrocarbons provides a reliable and transferable potential for atomistic/molecular studies of complex material phenomena involving hydrocarbon-metal or polymer-metal interfaces, polymer-metal nanocomposites, fracture and failure in hydrocarbon-based polymers, etc. The latter is especially true since MEAM is a reactive potential that allows for dynamic bond formation and bond breaking during simulation. Our results show that MEAM predicts the energetics of two major chemical reactions for saturated hydrocarbons, i.e., breaking a C-C and a C-H bond, reasonably well. However, the current parameterization does not accurately reproduce the energetics and structures of unsaturated hydrocarbons and, therefore, should not be applied to such systems.
More Related Videos
Related Concept Videos
 03:13
Intermolecular Forces
03:13Intermolecular Forces
 02:05
Degree of Unsaturation
The degree of unsaturation for hydrocarbons is U = (2C + 2 − H) / 2, where C is the number of carbon atoms and H is the number of hydrogen atoms.
 02:56
Intermolecular Forces and Physical Properties
 02:23
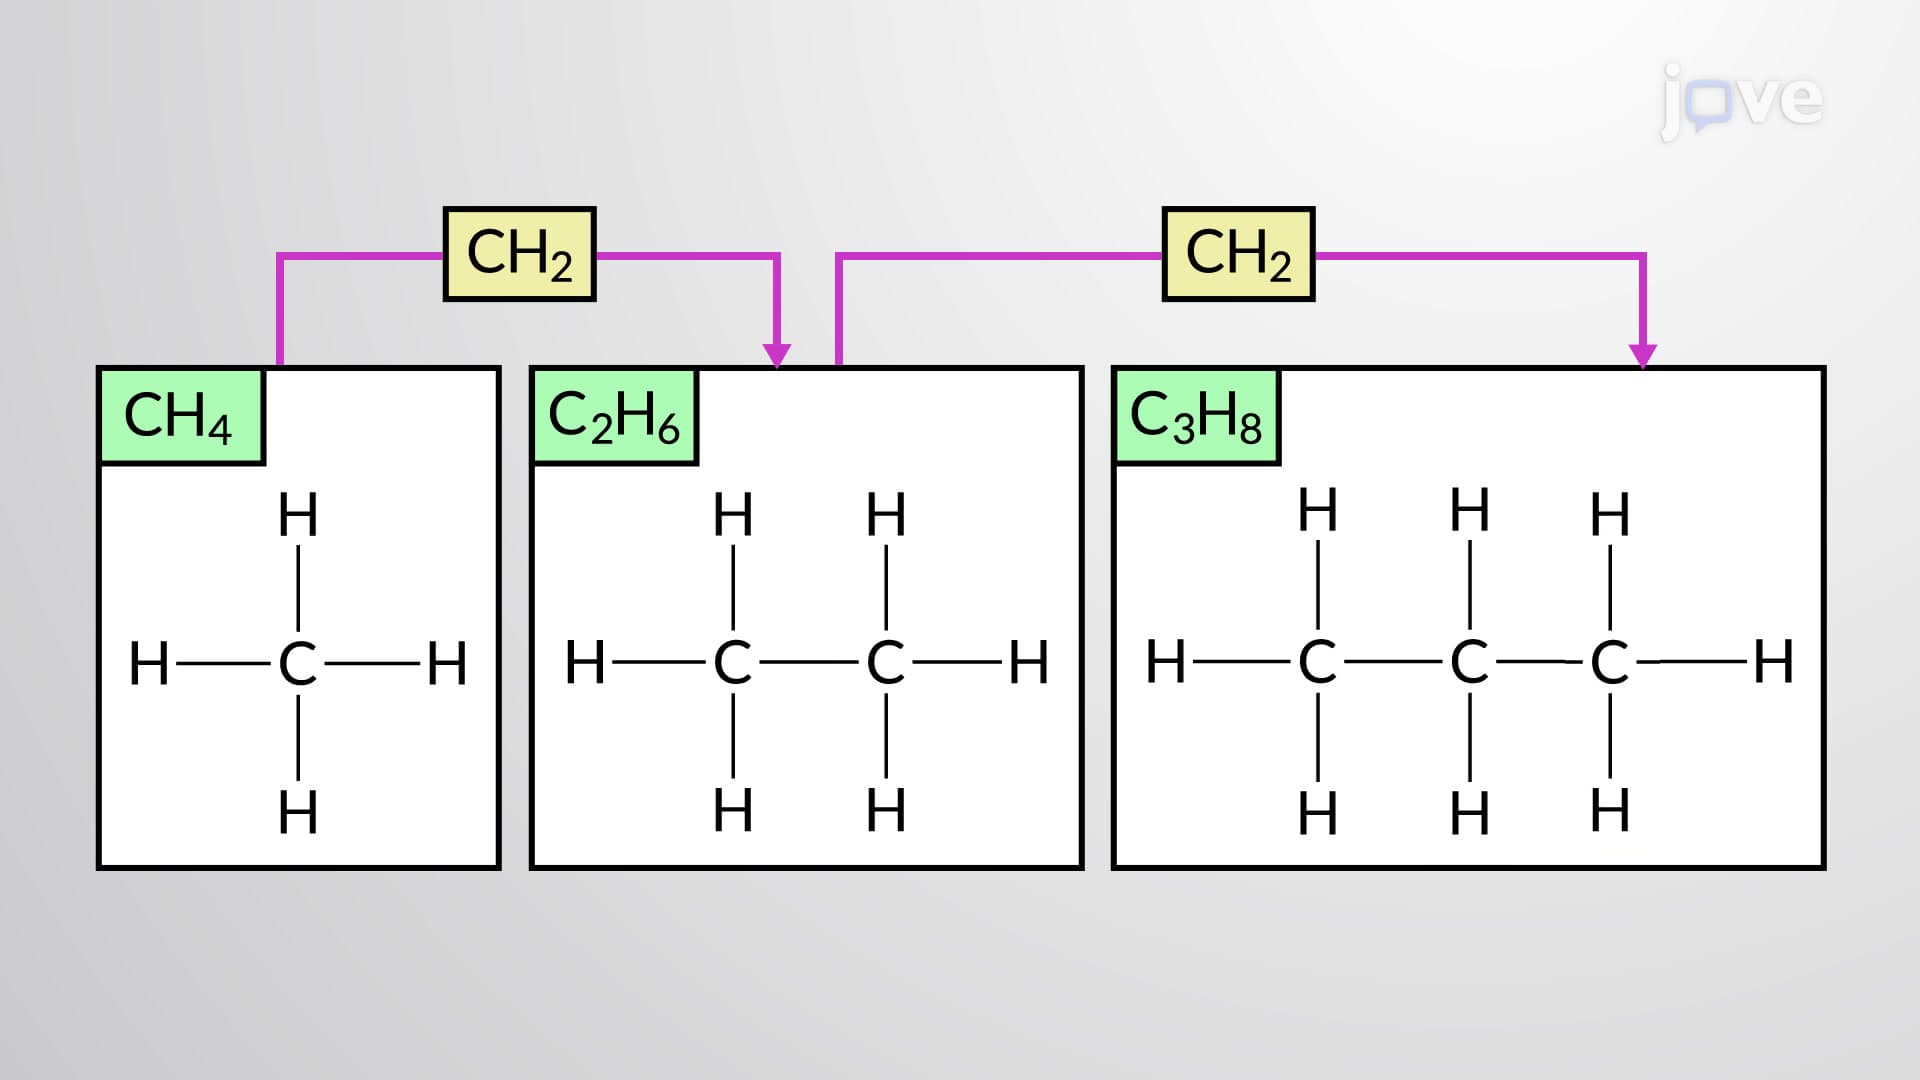
Structure of Alkanes
Hydrocarbons are the simplest organic compounds composed of carbons and hydrogens. Based on the bond order between carbons, the hydrocarbons are further classified into alkanes, alkenes, and alkynes.
Alkanes are the simplest hydrocarbons with sp3 hybrid carbon atoms....
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the...