
Resolving hot spots in the C-terminal dimerization domain that determine the stability of the molecular chaperone
Emanuele Ciglia1, Janina Vergin2, Sven Reimann3
1Institute for Pharmaceutical and Medicinal Chemistry, Heinrich-Heine-University, Düsseldorf, Germany.

Abstract:
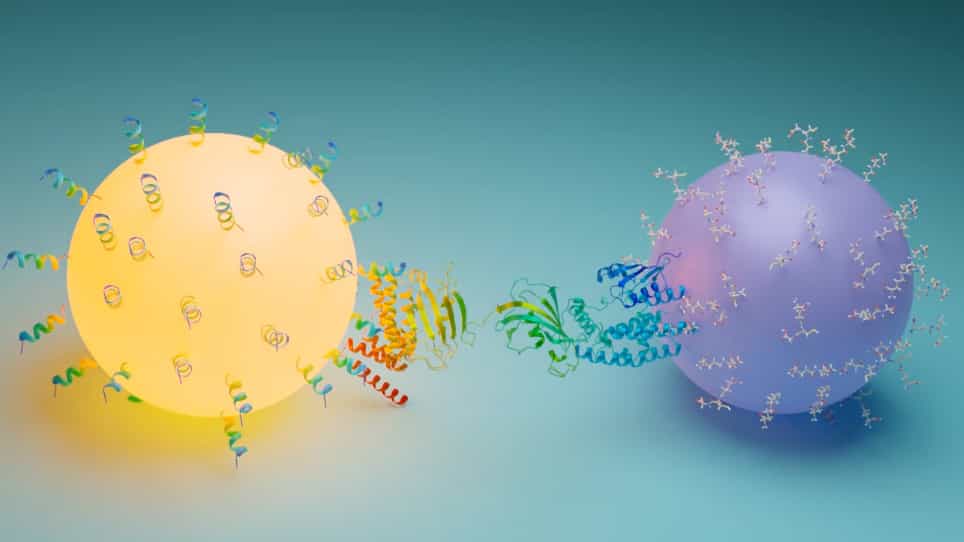
Human heat shock protein of 90 kDa (hHsp90) is a homodimer that has an essential role in facilitating malignant transformation at the molecular level. Inhibiting hHsp90 function is a validated approach for treating different types of tumors. Inhibiting the dimerization of hHsp90 via its C-terminal domain (CTD) should provide a novel way to therapeutically interfere with hHsp90 function. Here, we predicted hot spot residues that cluster in the CTD dimerization interface by a structural decomposition of the effective energy of binding computed by the MM-GBSA approach and confirmed these predictions using in silico alanine scanning with DrugScore(PPI). Mutation of these residues to alanine caused a significant decrease in the melting temperature according to differential scanning fluorimetry experiments, indicating a reduced stability of the mutant hHsp90 complexes. Size exclusion chromatography and multi-angle light scattering studies demonstrate that the reduced stability of the mutant hHsp90 correlates with a lower complex stoichiometry due to the disruption of the dimerization interface. These results suggest that the identified hot spot residues can be used as a pharmacophoric template for identifying and designing small-molecule inhibitors of hHsp90 dimerization.
Insights
Identifying key residues in human heat shock protein 90 kDa (hHsp90) C-terminal domain is crucial for developing novel cancer therapies. Disrupting hHsp90 dimerization offers a promising strategy to inhibit malignant transformation.
Area of Science:
- Biochemistry
- Molecular Biology
- Structural Biology
Background:
- Human heat shock protein 90 kDa (hHsp90) is a molecular chaperone essential for malignant transformation.
- Inhibiting hHsp90 function is a validated strategy for cancer treatment.
- Targeting hHsp90 dimerization presents a novel therapeutic approach.
Purpose of the Study:
- To identify critical residues in the hHsp90 C-terminal domain (CTD) involved in dimerization.
- To validate these residues as potential targets for small-molecule inhibitors.
Main Methods:
- Computational methods: MM-GBSA and in silico alanine scanning (DrugScorePPI) to predict hot spot residues.
- Experimental validation: Differential scanning fluorimetry, size exclusion chromatography, and multi-angle light scattering.
Main Results:
- Predicted hot spot residues in the hHsp90 CTD dimerization interface were identified.
- Mutating these residues significantly decreased complex stability and disrupted dimerization.
- Reduced stability correlated with lower complex stoichiometry, confirming the disruption of the dimerization interface.
Conclusions:
- The identified hot spot residues serve as a pharmacophoric template for designing hHsp90 dimerization inhibitors.
- Targeting hHsp90 dimerization offers a promising strategy for cancer therapy.
Related Concept Videos
 03:00
Molecular Chaperones and Protein Folding
The...
03:00Molecular Chaperones and Protein Folding
 01:26
Bacterial Protein Maturation
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...