[DFT/TDDFT study on the structure and absorption spectra of free base, N-/neo-confused porphyrin]
Related Experiment Video
Updated: Apr 30, 2026
A Technical Guide for Performing Spectroscopic Measurements on Metal-Organic Frameworks
Published on: April 28, 2023

Abstract:
Porphyrin is an important class of photochemical materials, which has been widely used in various fields. Computational investigations into the ground state structures and orbital energy levels of free base porphyrin (FBP), neo-confused porphyrin (NECP) and N-confused porphyrin (NCP) were performed with density functional theory(DFT). Absorption spectra were calculated at TD-B3LYP/6-31+G(d). Degeneracy of HOMO and HOMO-1 is lost, which would account for the shoulder peaks about the most intense transitions of FBP and NECP. Following FBP, NECP and NCP order, the orbital energy level (OEL) of LUMO decreases while the OEL of HOMO increases, which lead to the red shift of adsorption spectra. The energy difference between LUMO and LUMO+1 is almost the same as the difference between HOMO and HOMO-1, which would account for the only most intense transition of NCP. Solvent effect on ground state structures and absorption spectra was also investigated. The data shows that the character peak of Soret band and Q band changes in different solvent (benzene, chloroform, acetonitrile and water). So we further focus on discussing the N atom position and solvent effects on the energy level and Soret/Q bands of FBP, NCP and NECP, as well as clarifying its variation regularity and mechanism.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 10:03
Proton Transfer and Protein Conformation Dynamics in Photosensitive Proteins by Time-resolved Step-scan Fourier-transform Infrared Spectroscopy
Published on: June 27, 2014
Related Concept Videos
 01:29
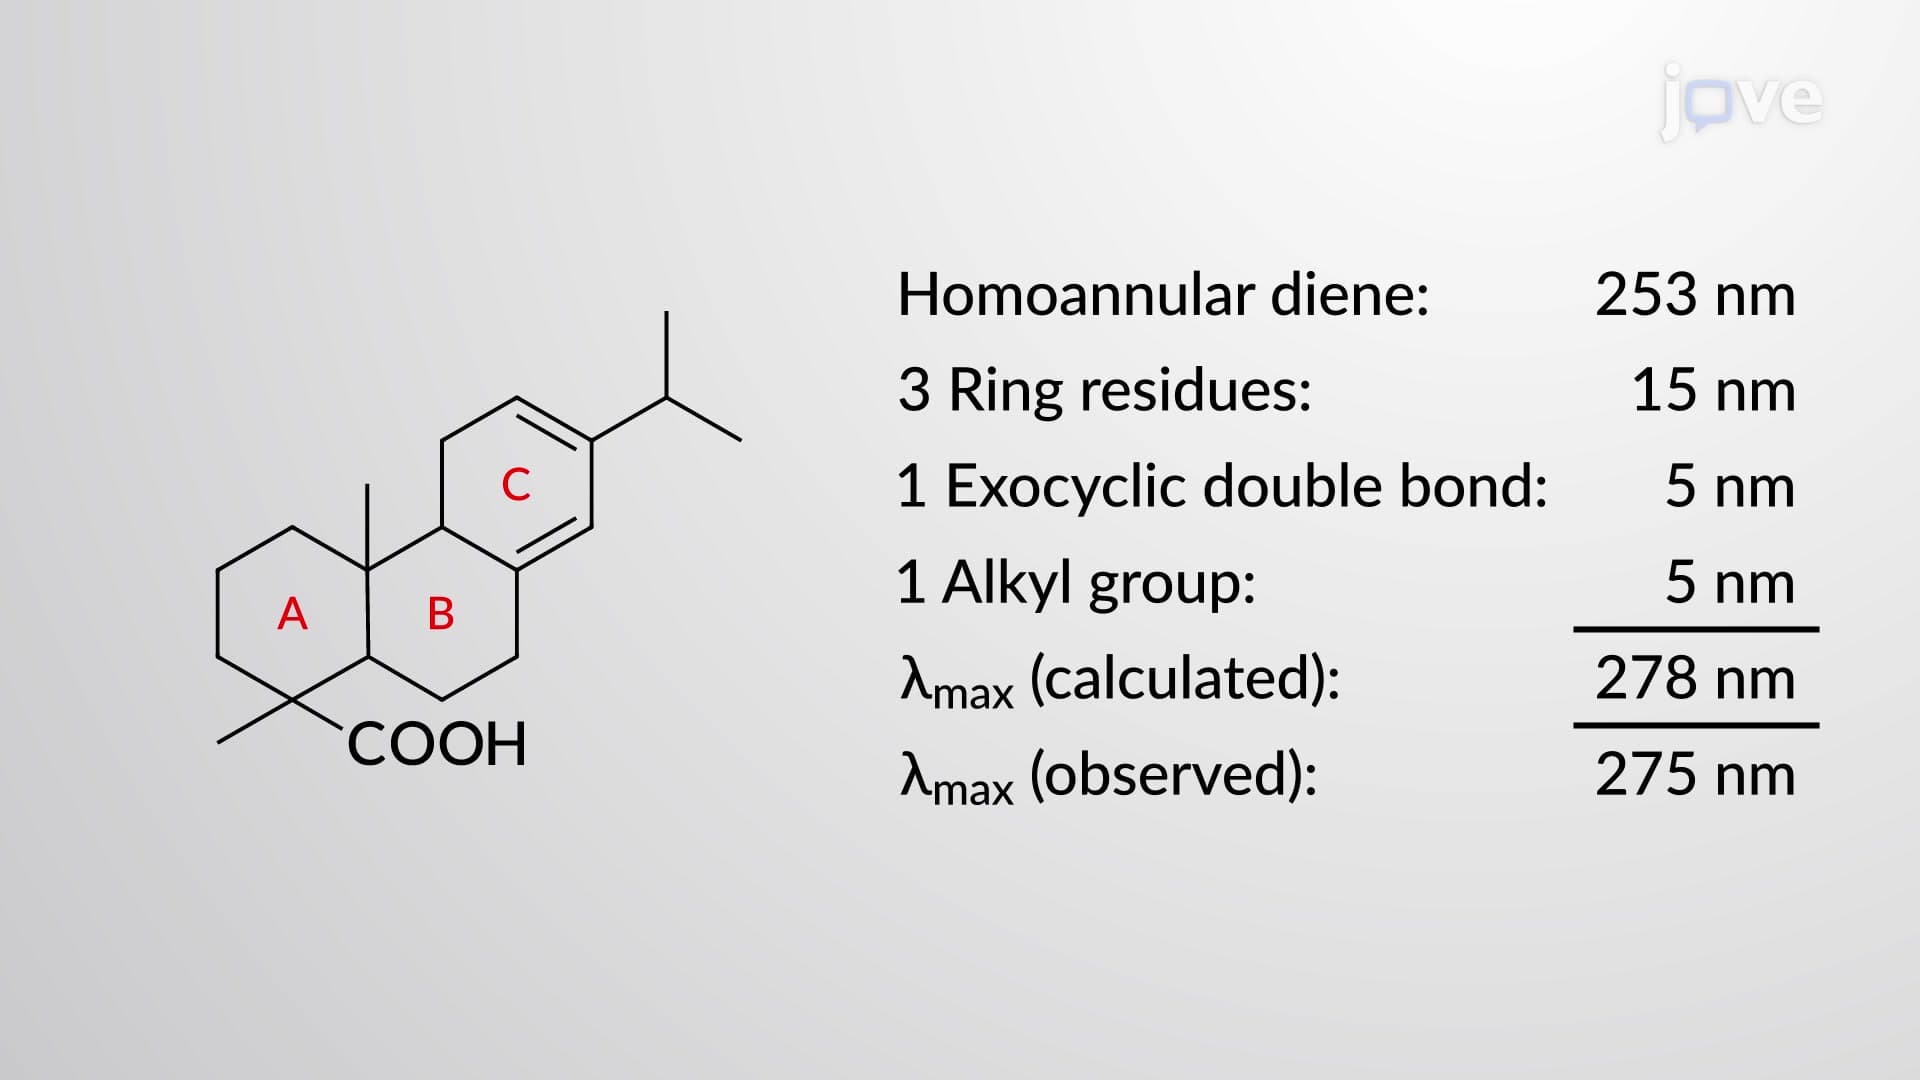
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)