Related Experiment Video
Updated: Apr 30, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Benchmarking quantum chemical methods for the calculation of molecular dipole moments and polarizabilities
A Leif Hickey1, Christopher N Rowley
1Department of Chemistry, Memorial University of Newfoundland , St. John's, NL A1B 3X7, Canada.

Abstract:
The calculation of molecular electric moments, polarizabilities, and electrostatic potentials is a widespread application of quantum chemistry. Although a range of wave function and density functional theory (DFT) methods have been applied in these calculations, combined with a variety of basis sets, there has not been a comprehensive evaluation of how accurate these methods are. To benchmark the accuracy of these methods, the dipole moments and polarizabilities of a set of 46 molecules were calculated using a broad set of quantum chemical methods and basis sets. Wave function methods Hartree-Fock (HF), second-order Møller-Plesset (MP2), and coupled cluster-singles and doubles (CCSD) were evaluated, along with the PBE, TPSS, TPSSh, PBE0, B3LYP, M06, and B2PLYP DFT functionals. The cc-pVDZ, cc-pVTZ, aug-cc-pVDZ, aug-cc-pVTZ, and Sadlej cc-pVTZ basis sets were tested. The aug-cc-pVDZ, Sadlej cc-pVTZ, and aug-cc-pVTZ basis sets all yield results with comparable accuracy, with the aug-cc-pVTZ calculations being the most accurate. CCSD, MP2, or hybrid DFT methods using the aug-cc-pVTZ basis set are all able to predict dipole moments with RMSD errors in the 0.12-0.13 D range and polarizabilities with RMSD errors in the 0.30-0.38 Å(3) range. Calculations using Hartree-Fock theory systematically overestimated dipole moments and underestimate polarizabilities. The pure DFT functionals included in this study (PBE and TPSS) slightly underestimate dipole moments and overestimate polarizability. Polarization anisotropy and implications for charge fitting are discussed.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:36
Molecular Geometry and Dipole Moments
 01:27
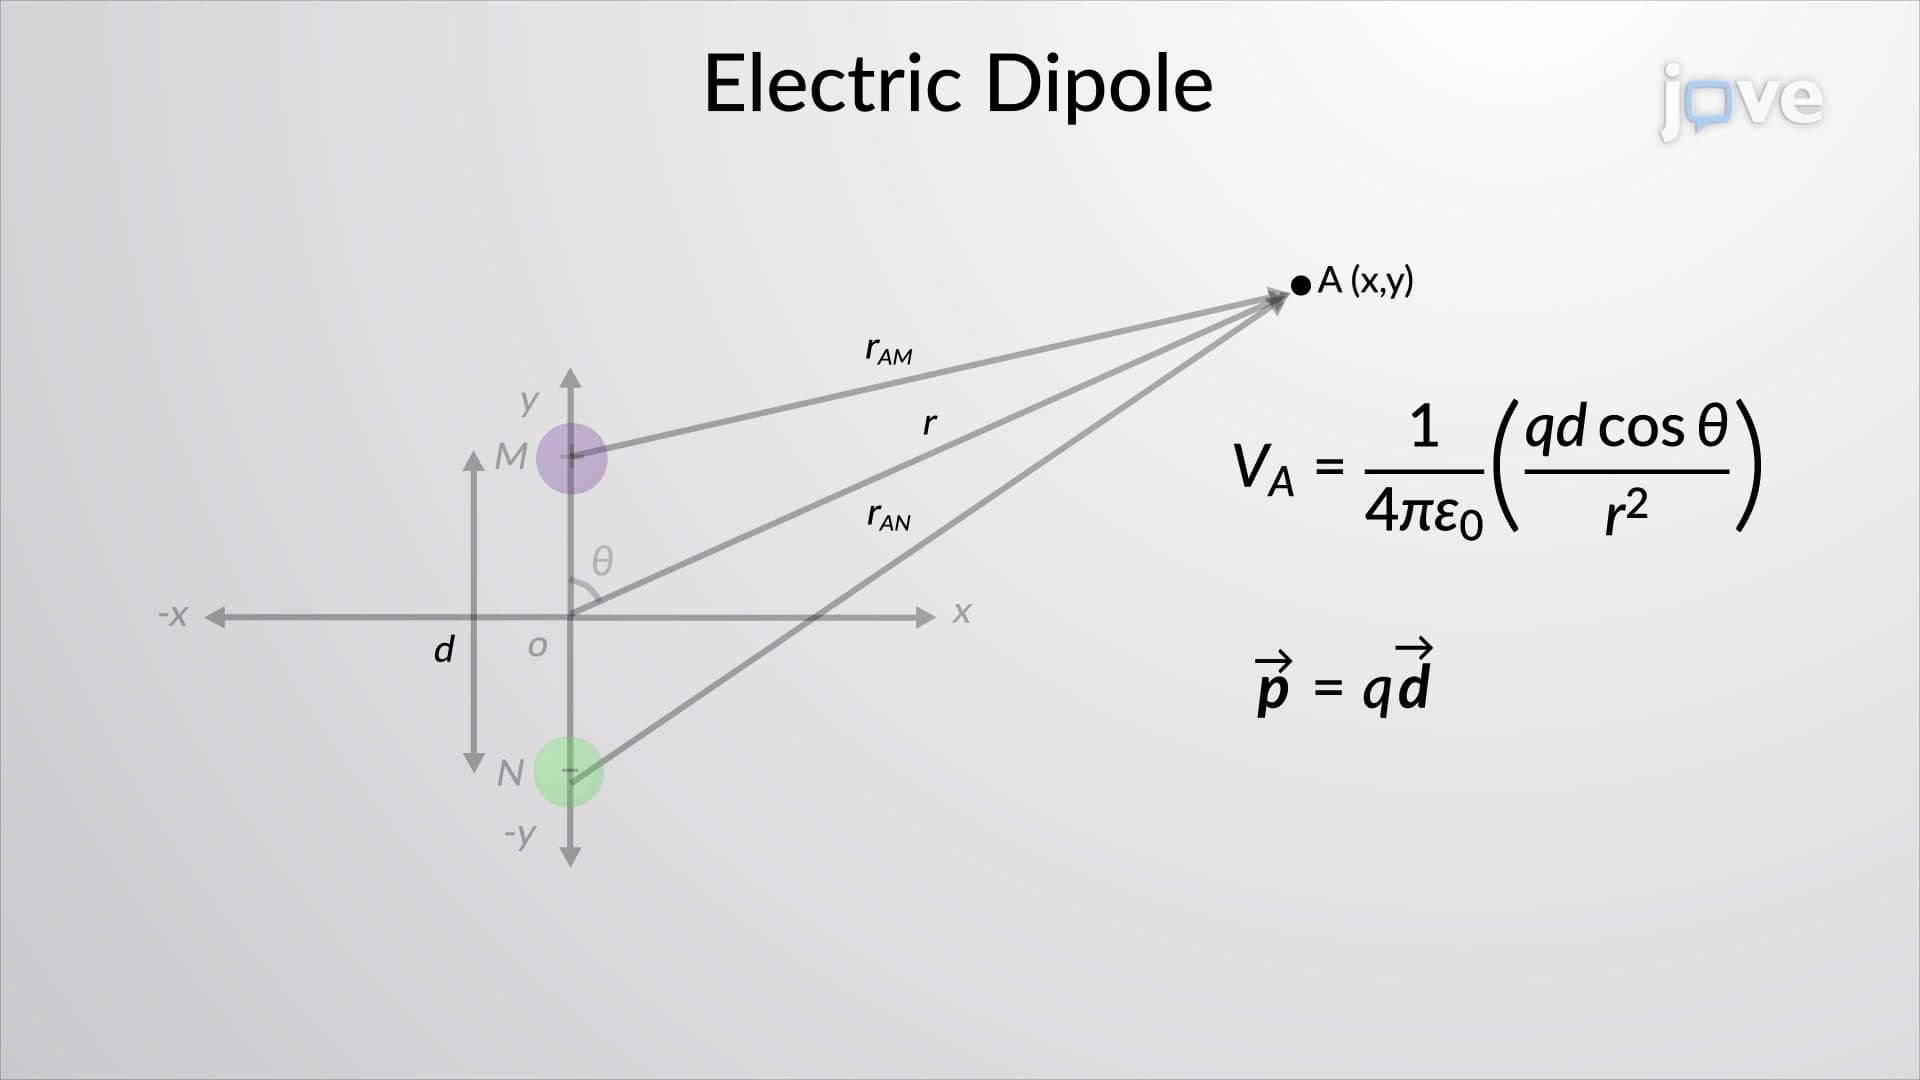
Calculations of Electric Potential II
Consider a...
 01:30
Electric Dipoles and Dipole Moment
Theoretically, studying electric dipoles leads to understanding why the resultant electric forces around us are weak. Since electric forces are strong, remnant net charges are rare. Hence, the interaction between dipoles helps us understand electrical interactions in...
 03:37
Molecular Shape and Polarity
 02:48
Bond Polarity, Dipole Moment, and Percent Ionic Character
 01:28
Induced Electric Dipoles
Since the absolute value of potential energy holds no physical meaning, its zero value can be chosen as per...