Conformation-dependent backbone geometry restraints set a new standard for protein crystallographic refinement
Related Experiment Video
Updated: Apr 28, 2026
Neutron Crystallography Data Collection and Processing for Modelling Hydrogen Atoms in Protein Structures
Published on: December 1, 2020
Nigel W Moriarty1, Dale E Tronrud, Paul D Adams
1Lawrence Berkeley National Laboratory, Berkeley, CA, USA.

Abstract:
Ideal values of bond angles and lengths used as external restraints are crucial for the successful refinement of protein crystal structures at all but the highest of resolutions. The restraints in common use today have been designed on the assumption that each type of bond or angle has a single ideal value that is independent of context. However, recent work has shown that the ideal values are, in fact, sensitive to local conformation, and, as a first step towards using such information to build more accurate models, ultra-high-resolution protein crystal structures have been used to derive a conformation-dependent library (CDL) of restraints for the protein backbone [Berkholz et al. (2009) Structure 17, 1316-1325]. Here, we report the introduction of this CDL into the phenix package and the results of test refinements of thousands of structures across a wide range of resolutions. These tests show that use of the CDL yields models that have substantially better agreement with ideal main-chain bond angles and lengths and, on average, a slightly enhanced fit to the X-ray data. No disadvantages of using the backbone CDL are apparent. In phenix, use of the CDL can be selected by simply specifying the cdl = True option. This successful implementation paves the way for further aspects of the context dependence of ideal geometry to be characterized and applied to improve experimental and predictive modeling accuracy.
More Related Videos
Related Concept Videos
 01:29
Determination of Crystal Structures
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:22
Protein Folding
 02:18
X-ray Crystallography
Diffraction
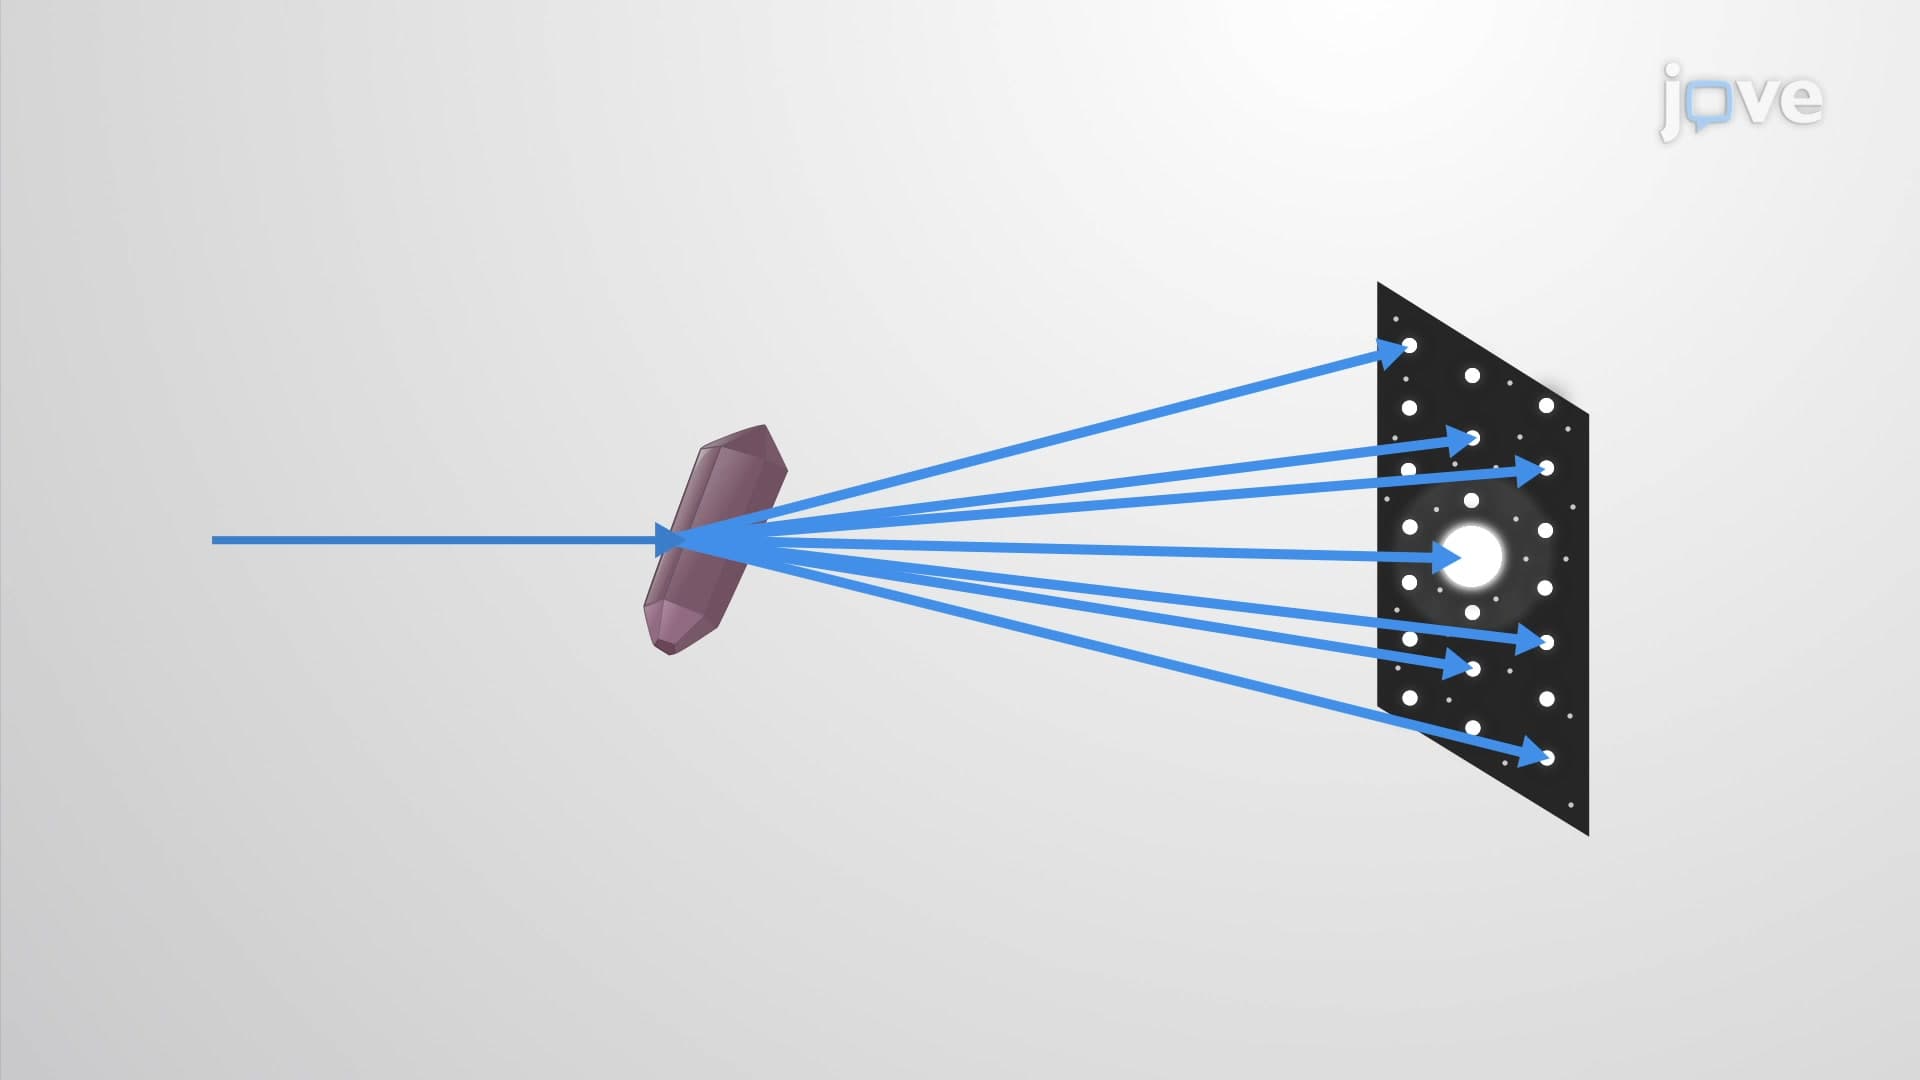
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...