Native structure-based modeling and simulation of biomolecular systems per mouse click
Related Experiment Video
Updated: Apr 24, 2026
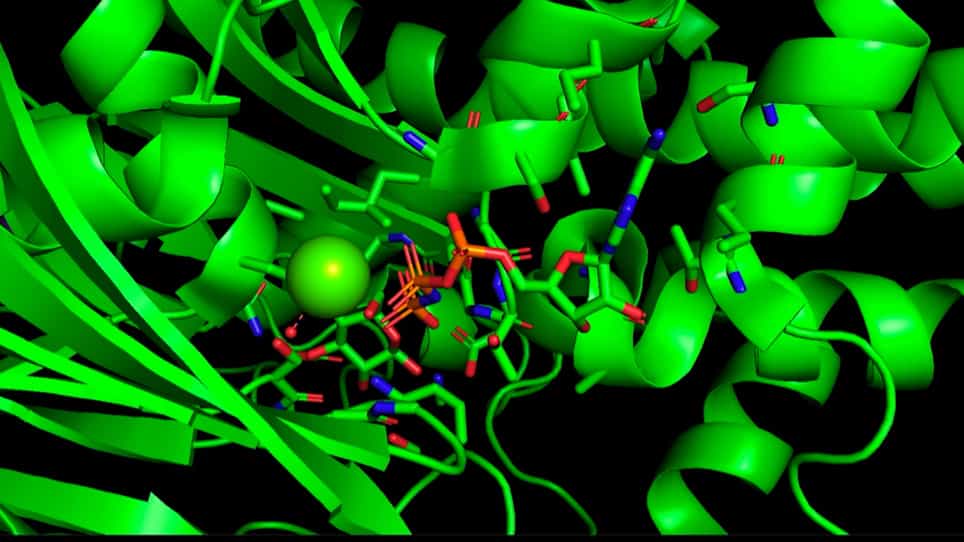
Modeling an Enzyme Active Site using Molecular Visualization Freeware
Published on: December 25, 2021
Benjamin Lutz, Claude Sinner, Stefan Bozic
1Steinbuch Centre for Computing, Karlsruhe Institute of Technology (KIT), Hermann-von-Helmholtz-Platz 1, 76344 Eggenstein-Leopoldshafen, Germany. ivan.kondov@kit.edu.

Background:
Molecular dynamics (MD) simulations provide valuable insight into biomolecular systems at the atomic level. Notwithstanding the ever-increasing power of high performance computers current MD simulations face several challenges: the fastest atomic movements require time steps of a few femtoseconds which are small compared to biomolecular relevant timescales of milliseconds or even seconds for large conformational motions. At the same time, scalability to a large number of cores is limited mostly due to long-range interactions. An appealing alternative to atomic-level simulations is coarse-graining the resolution of the system or reducing the complexity of the Hamiltonian to improve sampling while decreasing computational costs. Native structure-based models, also called Gō-type models, are based on energy landscape theory and the principle of minimal frustration. They have been tremendously successful in explaining fundamental questions of, e.g., protein folding, RNA folding or protein function. At the same time, they are computationally sufficiently inexpensive to run complex simulations on smaller computing systems or even commodity hardware. Still, their setup and evaluation is quite complex even though sophisticated software packages support their realization.
Results:
Here, we establish an efficient infrastructure for native structure-based models to support the community and enable high-throughput simulations on remote computing resources via GridBeans and UNICORE middleware. This infrastructure organizes the setup of such simulations resulting in increased comparability of simulation results. At the same time, complete workflows for advanced simulation protocols can be established and managed on remote resources by a graphical interface which increases reusability of protocols and additionally lowers the entry barrier into such simulations for, e.g., experimental scientists who want to compare their results against simulations. We demonstrate the power of this approach by illustrating it for protein folding simulations for a range of proteins.
Conclusions:
We present software enhancing the entire workflow for native structure-based simulations including exception-handling and evaluations. Extending the capability and improving the accessibility of existing simulation packages the software goes beyond the state of the art in the domain of biomolecular simulations. Thus we expect that it will stimulate more individuals from the community to employ more confidently modeling in their research.
More Related Videos
 07:31
Author Spotlight: Advancing Cell Membrane Biophysics - Exploring Interactions and Challenges Through Experimental and Computational Approaches
Published on: September 1, 2023
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:00
Molecular Models
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence....