Related Experiment Video
Updated: Apr 24, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Accuracy of dispersion interactions in semiempirical and molecular mechanics models: the benzene dimer case
Karol Strutyński1, José A N F Gomes, Manuel Melle-Franco
1REQUIMTE, Departamento de Química, Faculdade de Ciências, Universidade do Porto , Rua do Campo Alegre S/N, 4169-007 Porto, Portugal.

Abstract:
The benzene dimer is arguably the simplest molecular analogue of graphitic materials. We present the systematic study of minima and transition states of the benzene dimer with semiempirical and molecular mechanics (MM) methods. Full minimizations on all conformations were performed and the results, geometries, and binding energies were compared with CCSD(T) and DFT-D results. MM yields the best results with three force fields MM3, OPLS, and AMOEBA, which reproduced nine out of the ten stationary points of the benzene dimer. We obtained new parameters for MM3 and OPLS that successfully reproduce all structures of the benzene dimer and showed improved accuracy over DFT-D in most dimer geometries. Semiempirical models were, unexpectedly, less accurate than MM methods. The most accurate semiempirical method for the benzene dimer is PM6-DH2. DFT-D was the only Hamiltonian that reproduced the variations of energy with geometry from CCSD(T) calculations accurately and is the method of choice for energies of periodic and molecular calculations of graphitic systems. In contrast, MM represents an accurate alternative to calculate geometries.

Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:24
Van der Waals Interactions
 03:51
Molecular Orbital Theory II
 01:07
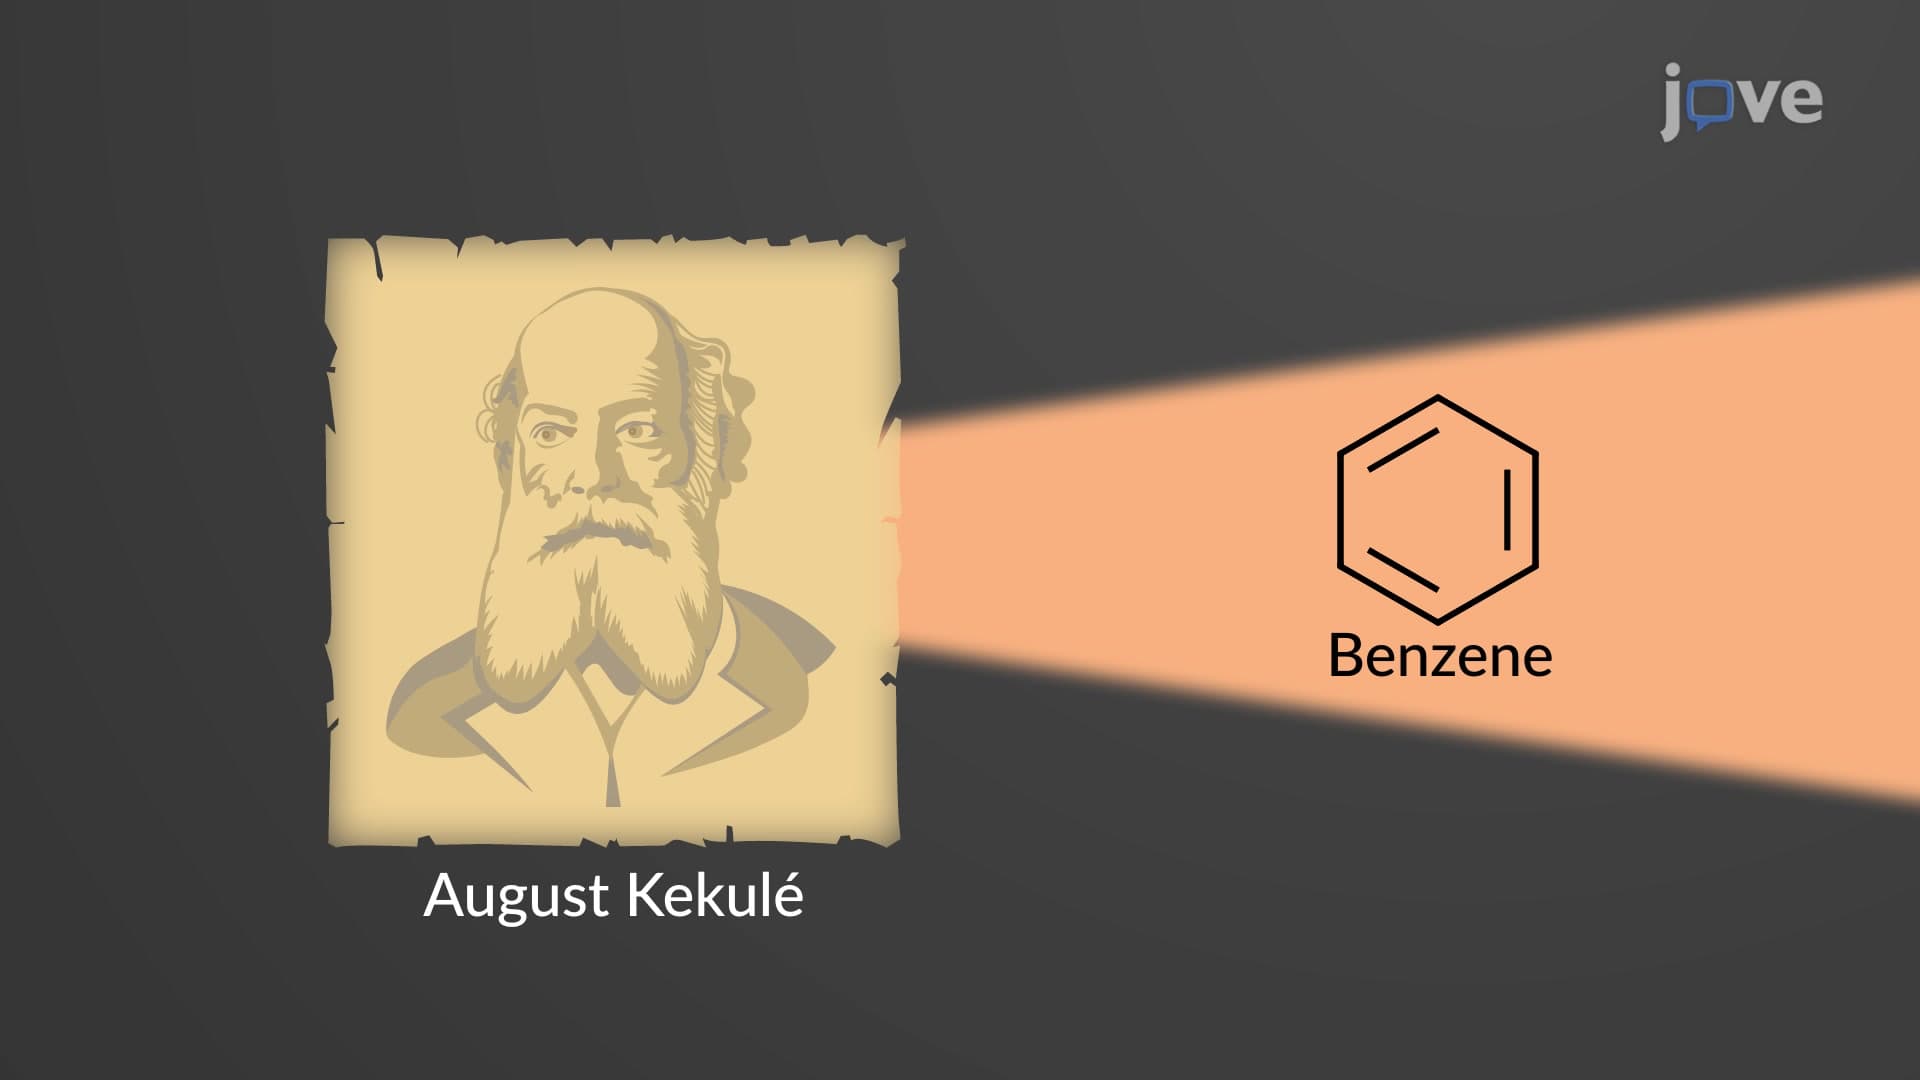
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 01:37
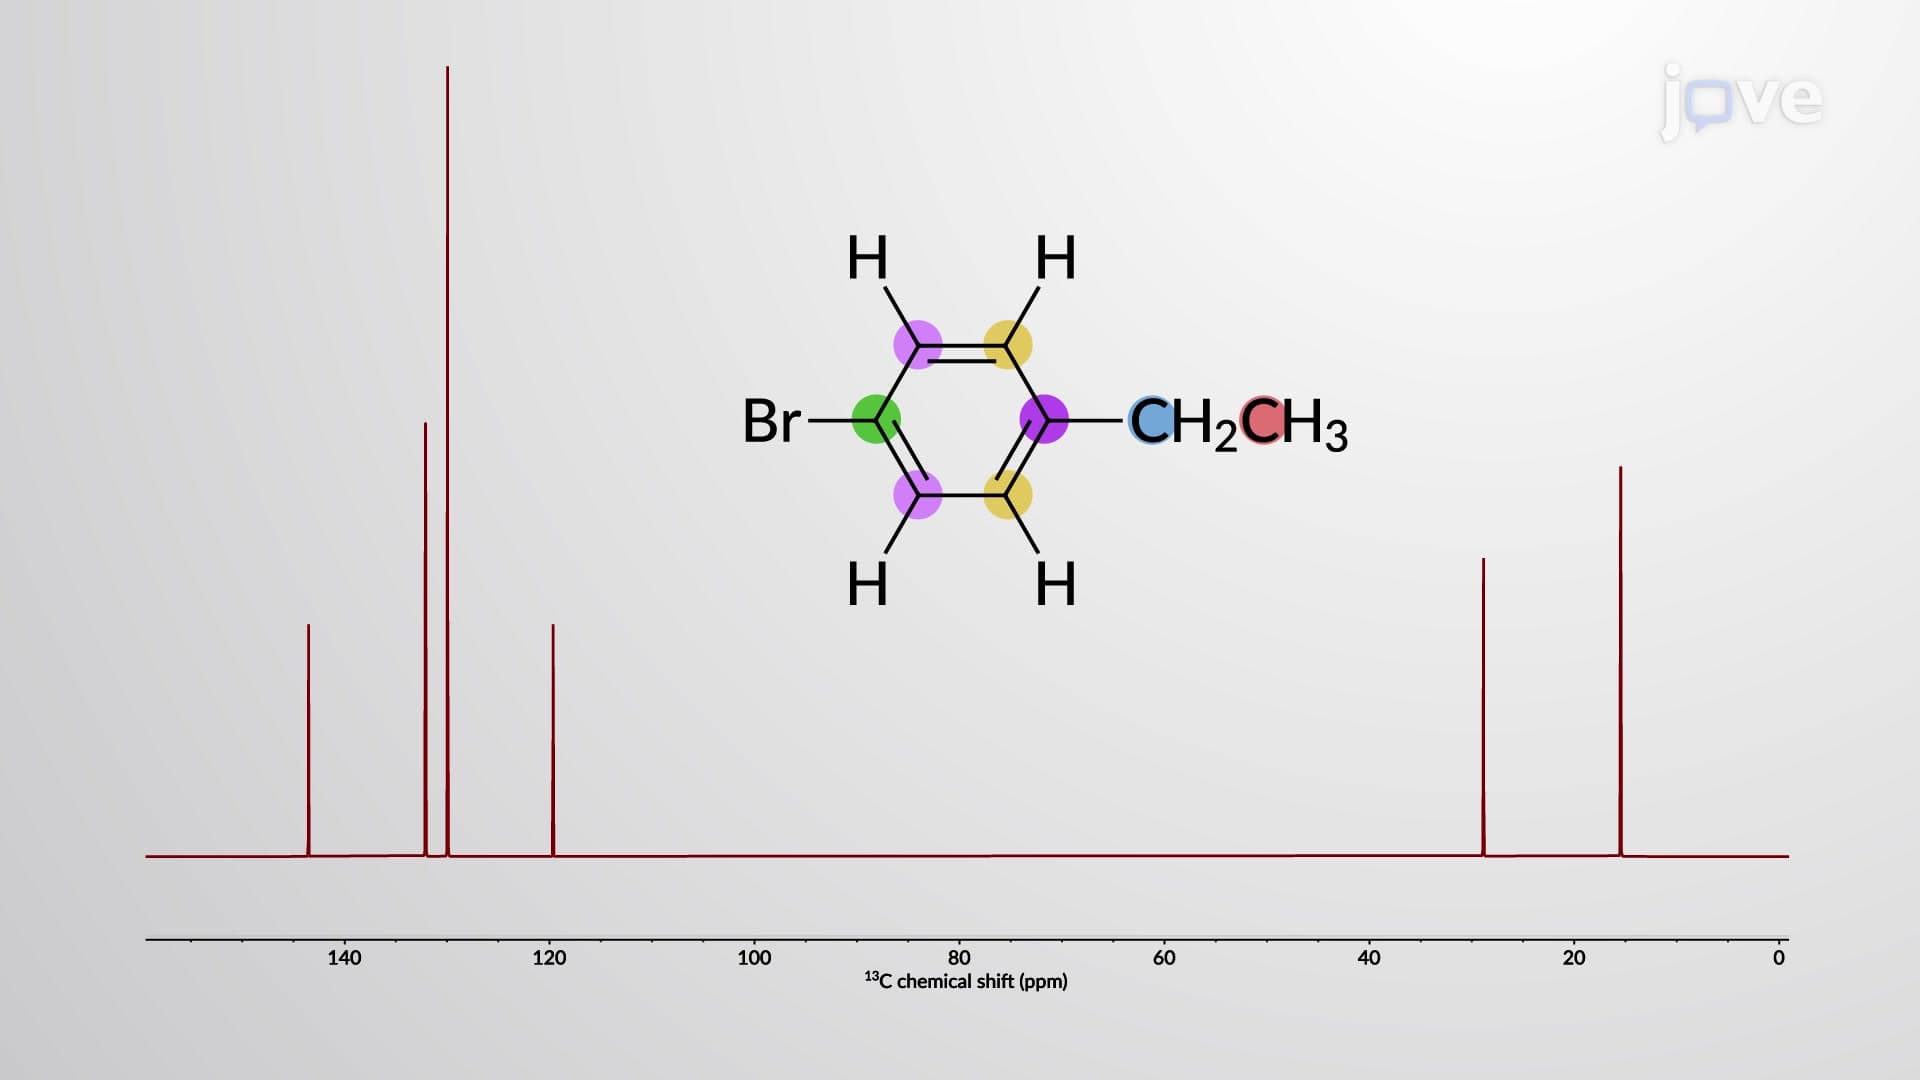
NMR Spectroscopy of Benzene Derivatives
 03:13
Intermolecular Forces