Related Experiment Video
Updated: Apr 24, 2026
Protein WISDOM: A Workbench for In silico De novo Design of BioMolecules
Published on: July 25, 2013
Density functional theory calculations on entire proteins for free energies of binding: application to a model polar
Stephen J Fox1, Jacek Dziedzic, Thomas Fox
1School of Chemistry, University of Southampton, Southampton, Hampshire, SO17 1BJ, United Kingdom.

Abstract:
In drug optimization calculations, the molecular mechanics Poisson-Boltzmann surface area (MM-PBSA) method can be used to compute free energies of binding of ligands to proteins. The method involves the evaluation of the energy of configurations in an implicit solvent model. One source of errors is the force field used, which can potentially lead to large errors due to the restrictions in accuracy imposed by its empirical nature. To assess the effect of the force field on the calculation of binding energies, in this article we use large-scale density functional theory (DFT) calculations as an alternative method to evaluate the energies of the configurations in a "QM-PBSA" approach. Our DFT calculations are performed with a near-complete basis set and a minimal parameter implicit solvent model, within the self-consistent calculation, using the ONETEP program on protein-ligand complexes containing more than 2600 atoms. We apply this approach to the T4-lysozyme double mutant L99A/M102Q protein, which is a well-studied model of a polar binding site, using a set of eight small aromatic ligands. We observe that there is very good correlation between the MM and QM binding energies in vacuum but less so in the solvent. The relative binding free energies from DFT are more accurate than the ones from the MM calculations, and give markedly better agreement with experiment for six of the eight ligands. Furthermore, in contrast to MM-PBSA, QM-PBSA is able to correctly predict a nonbinder.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 09:15
Measuring Biomolecular DSC Profiles with Thermolabile Ligands to Rapidly Characterize Folding and Binding Interactions
Published on: November 21, 2017
Related Concept Videos
 02:18
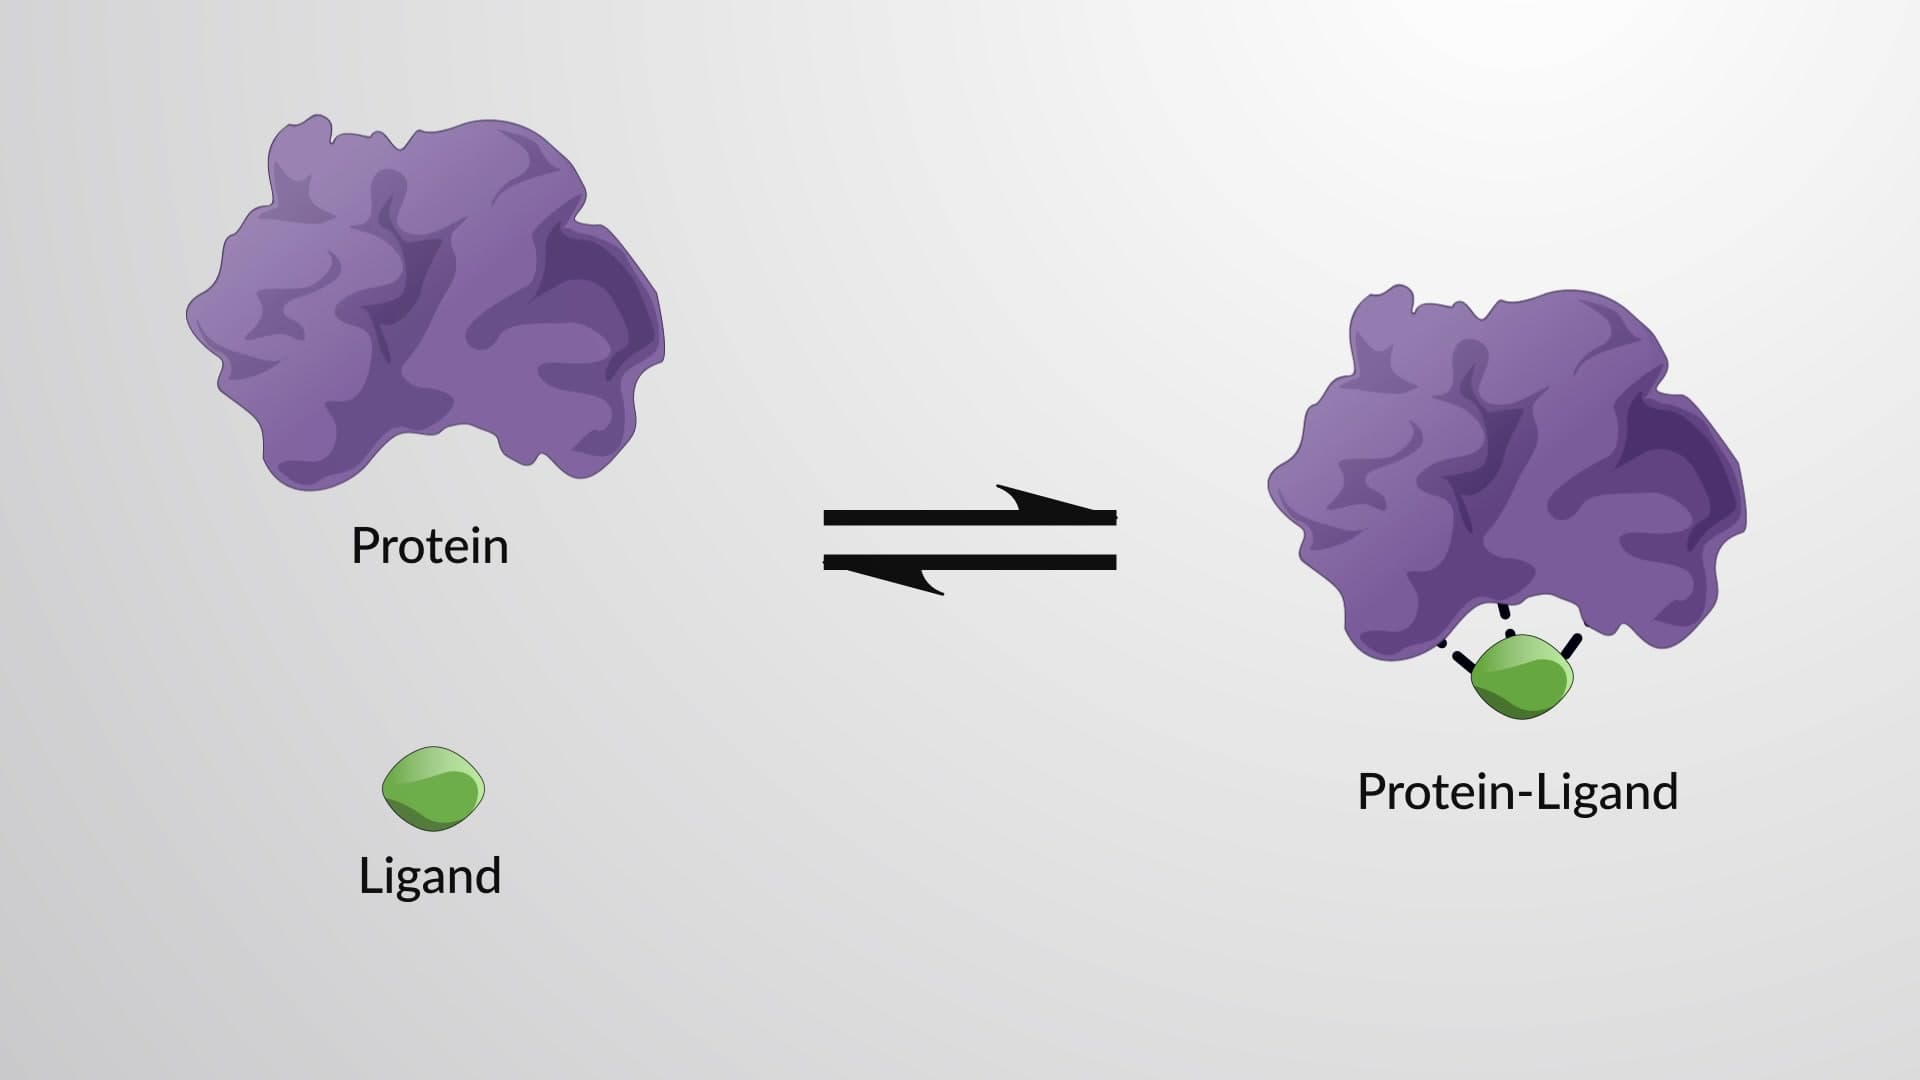
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:22
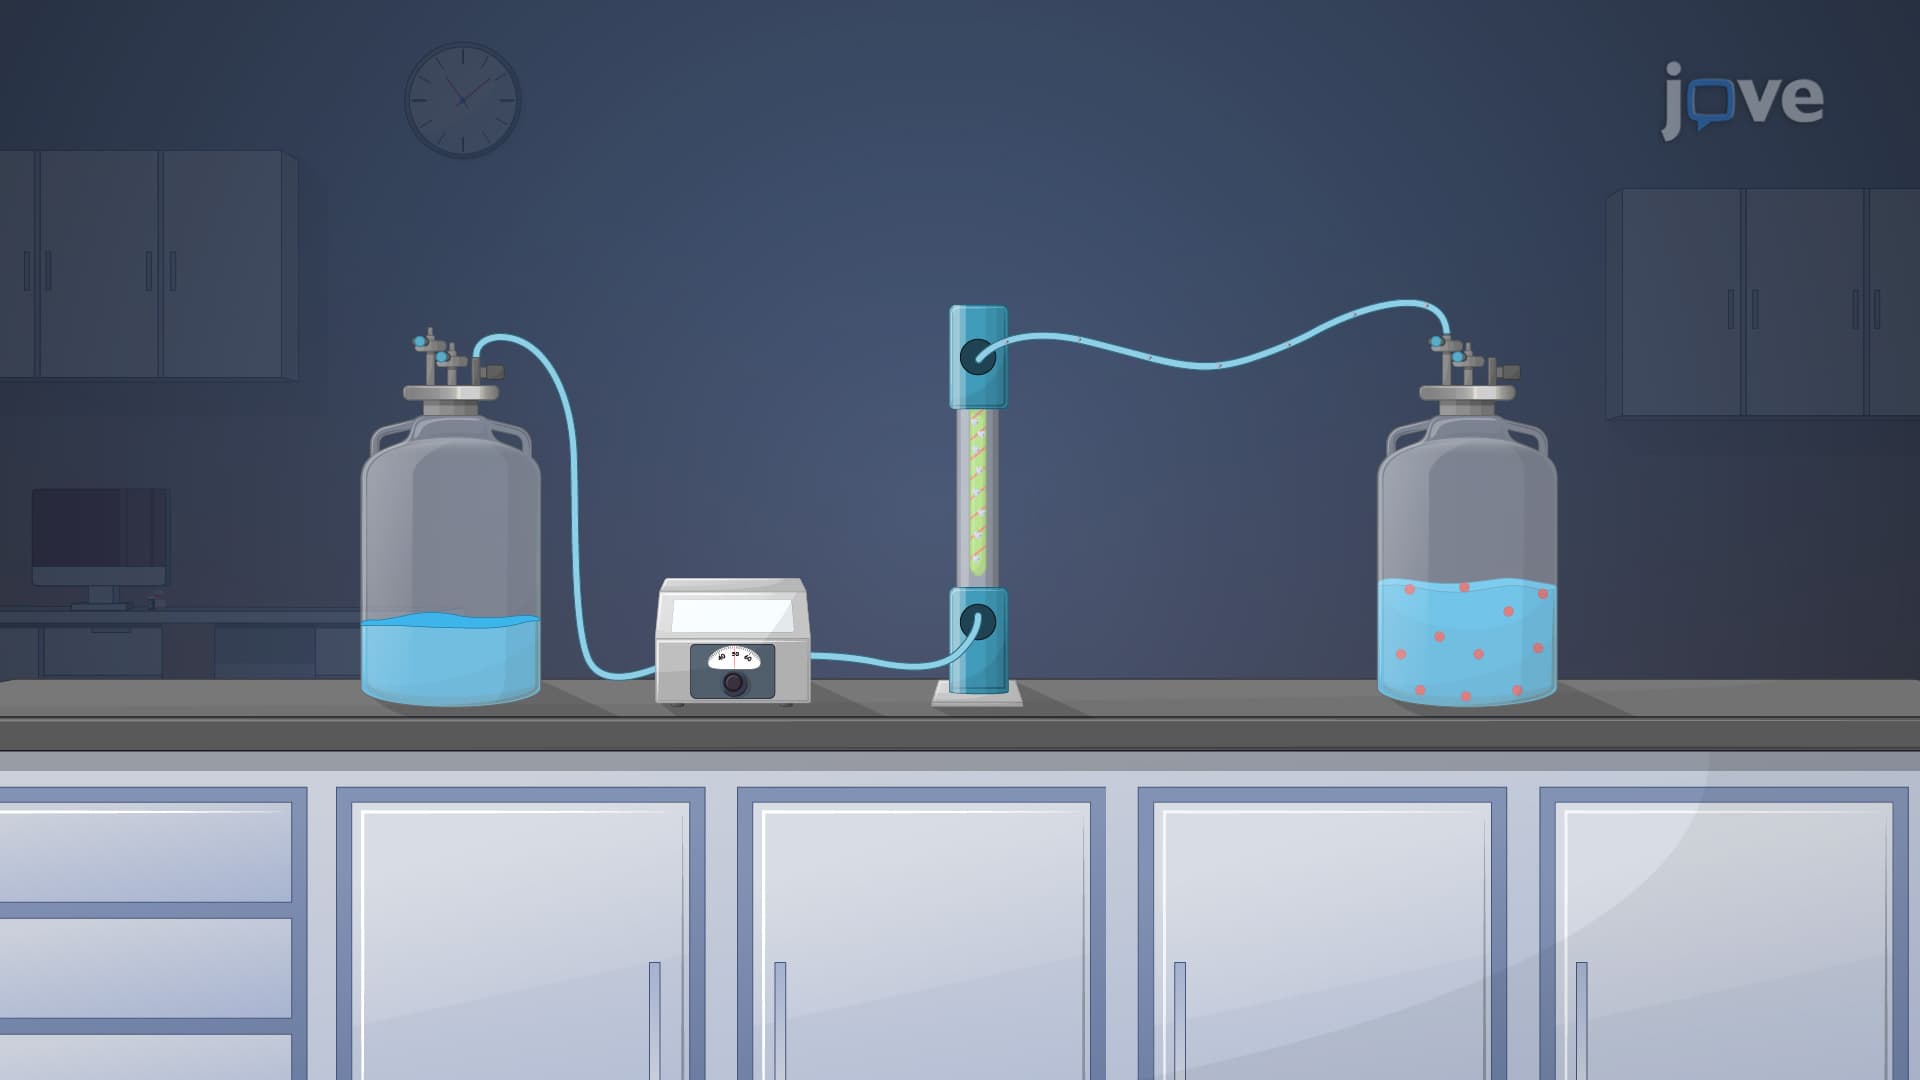
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...