Related Experiment Video
Updated: Apr 21, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
A comparison of two types of explicitly correlated Gaussian functions for non-Born-Oppenheimer molecular calculations
Martin Formanek1, Keeper L Sharkey2, Nikita Kirnosov1
1Department of Physics, University of Arizona, Tucson, Arizona 85721, USA.

Abstract:
A new functional form of the explicitly correlated Gaussian-type functions (later called Gaussians or ECGs) for performing non-Born-Oppenheimer (BO) calculations of molecular systems with an arbitrary number of nuclei is presented. In these functions, the exponential part explicitly depends on all interparticle distances and the preexponential part depends only on the distances between the nuclei. The new Gaussians are called sin/cos-Gaussians and their preexponential part is a product of sin and/or cos factors. The effectiveness of the new Gaussians in describing non-BO pure vibrational states is investigated by comparing them with r(m)-Gaussians containing preexponential multipliers in the form of non-negative powers of internuclear distances (the internuclear distance in the diatomic case). The testing is performed for a diatomic system with the nuclei interacting through a Morse potential. It shows that the new sin/cos-Gaussian basis set is capable of providing equally accurate results as obtained with the r(m)-Gaussians. However, especially for lower vibrational states, more sin/cos-Gaussians are needed to reach a similar accuracy level as obtained with the r(m)-Gaussians. Implementation of the sin/cos-Gaussians in non-BO calculations of diatomic and, in particular, of triatomic systems, which will follow, will provide further assessment of the efficiency of the new functions.


Related Concept Videos
 03:51
Molecular Orbital Theory II
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 02:35
Molecular Orbital Theory I
 01:10
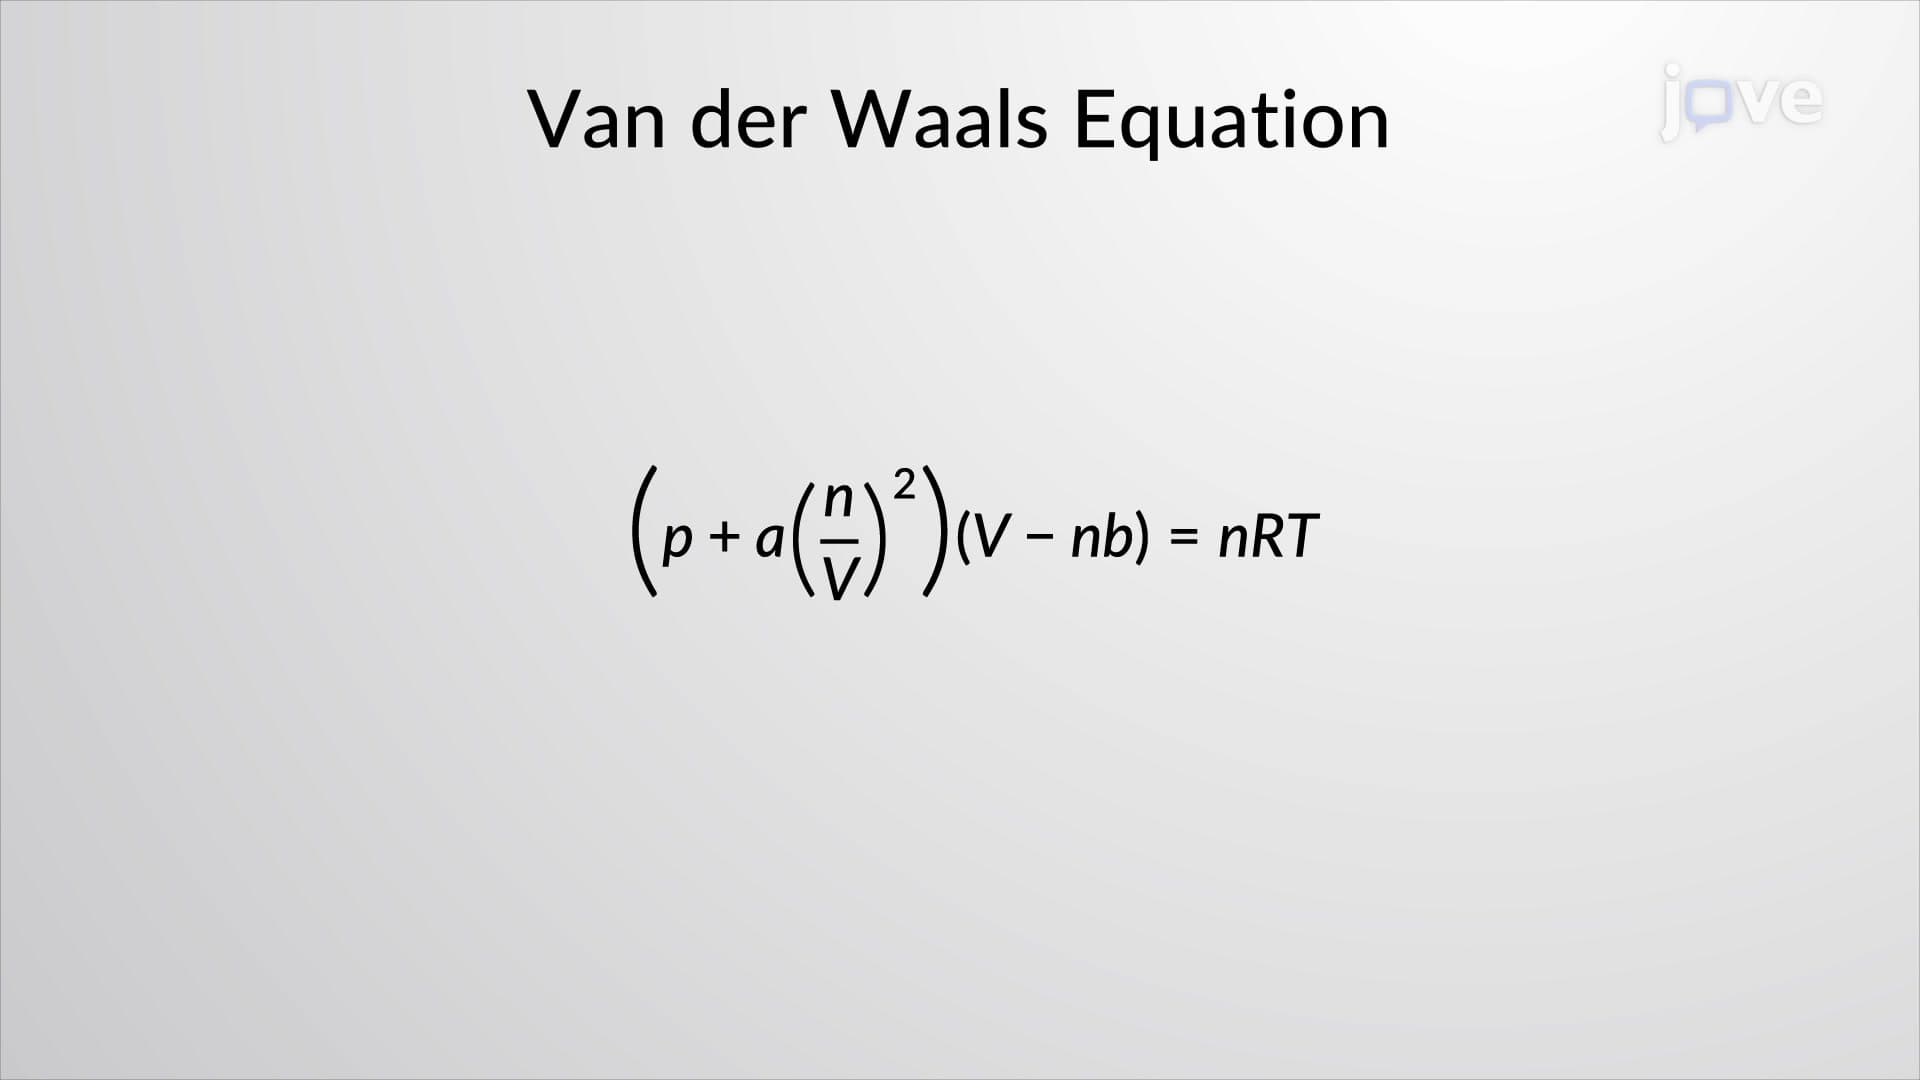
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the...
 02:40
MO Theory and Covalent Bonding
 02:00
Molecular Models