Related Experiment Video
Updated: Apr 14, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Probing the origins of 17β-hydroxysteroid dehydrogenase type 1 inhibitory activity via QSAR and molecular docking
Kakanand Srungboonmee1, Napat Songtawee1, Teerawat Monnor1
1Center of Data Mining and Biomedical Informatics, Faculty of Medical Technology, Mahidol University, Bangkok 10700, Thailand.

Abstract:
It is generally known that proliferation of human breast cancer cells is stimulated by excess estrogen namely 17β-estradiol. Therefore, reduction of 17β-estradiol production by inhibiting 17β-hydroxysteroid dehydrogenase type 1 (17β-HSD1) is an interesting route for breast cancer treatment particularly during adjuvant therapy. This study investigated the structure-activity relationship of 17β-HSD1 inhibitors as to gain insights and understanding on the origins of 17β-HSD1 inhibitory activities. To meet this goal, multiple linear regression model was constructed and correspondingly the results revealed good predictivity (N = 31, R(2) = 0.9438, Q(2) = 0.8530). The model suggested that low molecular weight and energy were preferred as 17β-HSD1 inhibitors. Additionally, high molecular flexibility and high number of hydrogen bond donors were also shown to be important that is in correspondence to previously reported pharmacophore model of 17β-HSD1 inhibitors. Furthermore, molecular docking of inhibitors to 17β-HSD1 followed by anchor analysis suggested that three different pockets comprising of hydrogen bonding sites 1 and 2 as well as van der Waals contacts contributed to protein-ligand interactions. Post-docking analysis of potent compound 9 with 17β-HSD1 suggested that the binding modality was similar to the binding of substrate (i.e. estradiol) and its analog (i.e. equilin). Such information is useful in guiding the further design of novel and robust 17β-HSD1 inhibitors.
Insights
Inhibiting 17β-hydroxysteroid dehydrogenase type 1 (17β-HSD1) can treat breast cancer. This study reveals key structural features of effective 17β-HSD1 inhibitors, guiding the development of new treatments.
Area of Science:
- Medicinal Chemistry
- Computational Chemistry
- Oncology
Background:
- Human breast cancer cell proliferation is stimulated by excess estrogen, specifically 17β-estradiol.
- Inhibiting 17β-hydroxysteroid dehydrogenase type 1 (17β-HSD1) offers a therapeutic strategy for breast cancer, particularly in adjuvant therapy.
- Understanding the structure-activity relationships of 17β-HSD1 inhibitors is crucial for designing effective drugs.
Purpose of the Study:
- To investigate the structure-activity relationship of 17β-HSD1 inhibitors.
- To gain insights into the origins of 17β-HSD1 inhibitory activities.
- To guide the design of novel and robust 17β-HSD1 inhibitors.
Main Methods:
- Construction of a multiple linear regression model (N = 31) to predict inhibitory activity.
- Analysis of molecular descriptors including molecular weight, energy, flexibility, and hydrogen bond donors.
- Molecular docking and anchor analysis of inhibitors with 17β-HSD1.
Main Results:
- The multiple linear regression model demonstrated good predictivity (R(2) = 0.9438, Q(2) = 0.8530).
- Preferred inhibitor characteristics include low molecular weight and energy, high molecular flexibility, and numerous hydrogen bond donors.
- Molecular docking revealed interactions within three distinct pockets of 17β-HSD1, involving hydrogen bonding and van der Waals forces.
- The binding mode of a potent inhibitor (compound 9) resembles that of the natural substrate (estradiol).
Conclusions:
- The study successfully elucidated key structural features essential for potent 17β-HSD1 inhibition.
- Findings provide valuable guidance for the rational design of new breast cancer therapeutic agents targeting 17β-HSD1.
- The identified structure-activity relationships and binding interactions can accelerate the development of novel inhibitors.
More Related Videos
 05:47
In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
 10:51
Reverse Yeast Two-hybrid System to Identify Mammalian Nuclear Receptor Residues that Interact with Ligands and/or Antagonists
Published on: November 15, 2013
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:30
Enzyme Inhibition
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:16
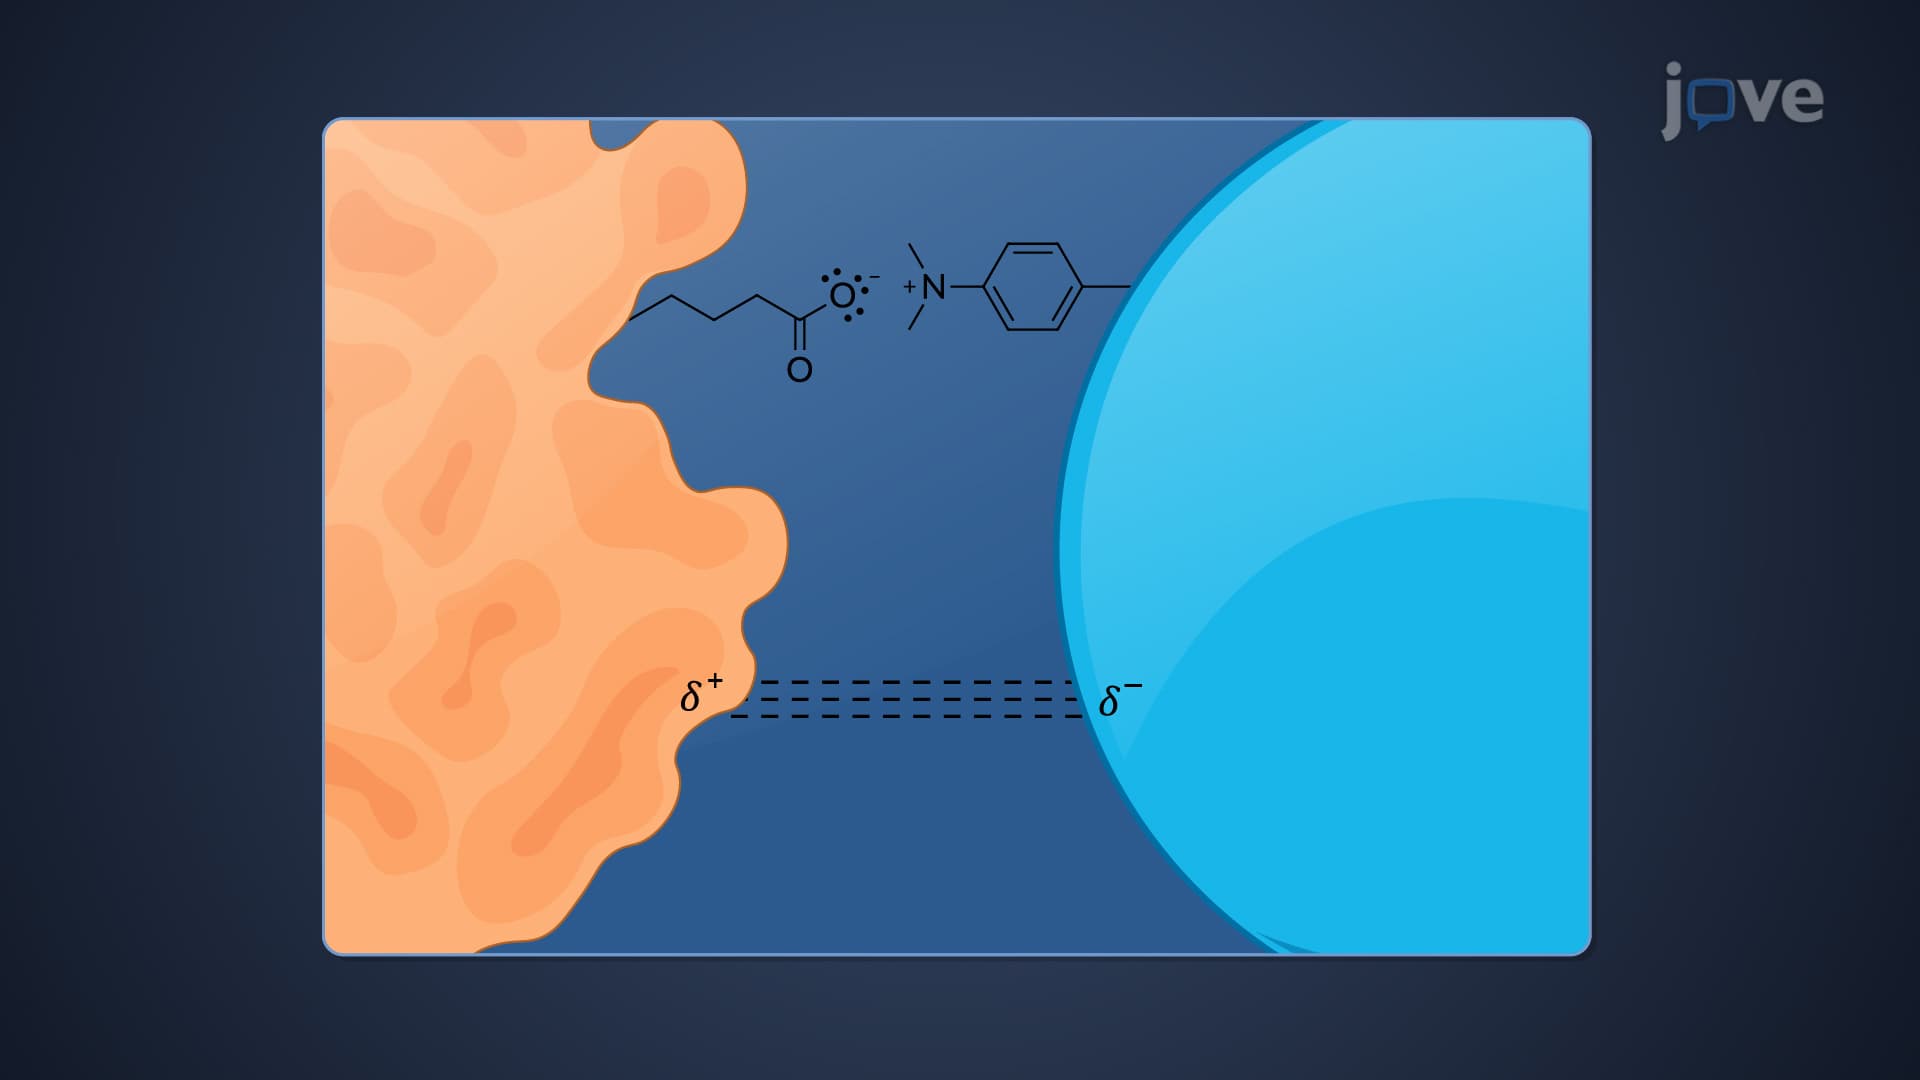
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:28
Drug-Receptor Interaction: Antagonist
Antagonists can be classified as competitive or noncompetitive based on their...
 01:22
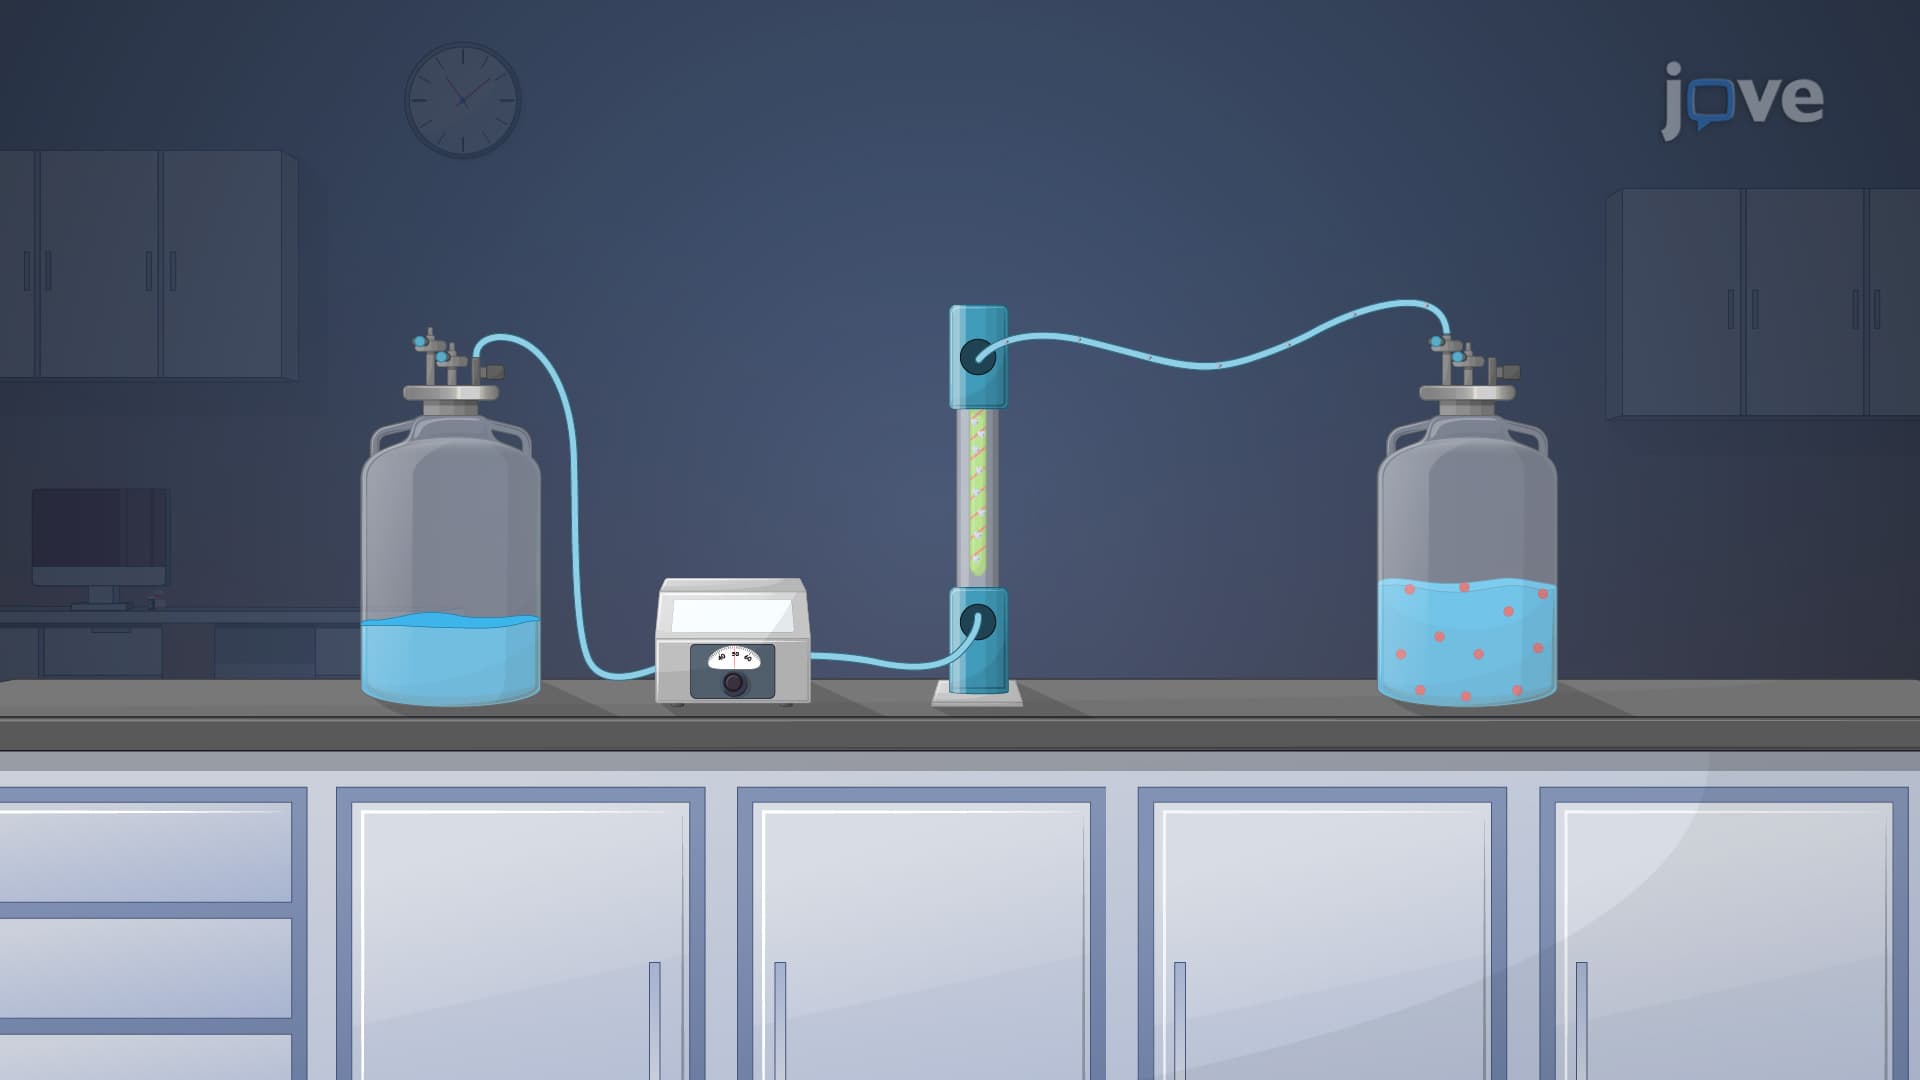
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...