Efficient parametrization of complex molecule-surface force fields
David Z Gao1, Filippo Federici Canova2, Matthew B Watkins1
1Department of Physics and Astronomy, University College London, Gower Street-London, United Kingdom.

Abstract:
We present an efficient scheme for parametrizing complex molecule-surface force fields from ab initio data. The cost of producing a sufficient fitting library is mitigated using a 2D periodic embedded slab model made possible by the quantum mechanics/molecular mechanics scheme in CP2K. These results were then used in conjunction with genetic algorithm (GA) methods to optimize the large parameter sets needed to describe such systems. The derived potentials are able to well reproduce adsorption geometries and adsorption energies calculated using density functional theory. Finally, we discuss the challenges in creating a sufficient fitting library, determining whether or not the GA optimization has completed, and the transferability of such force fields to similar molecules.
More Related Videos

Related Concept Videos
 02:36
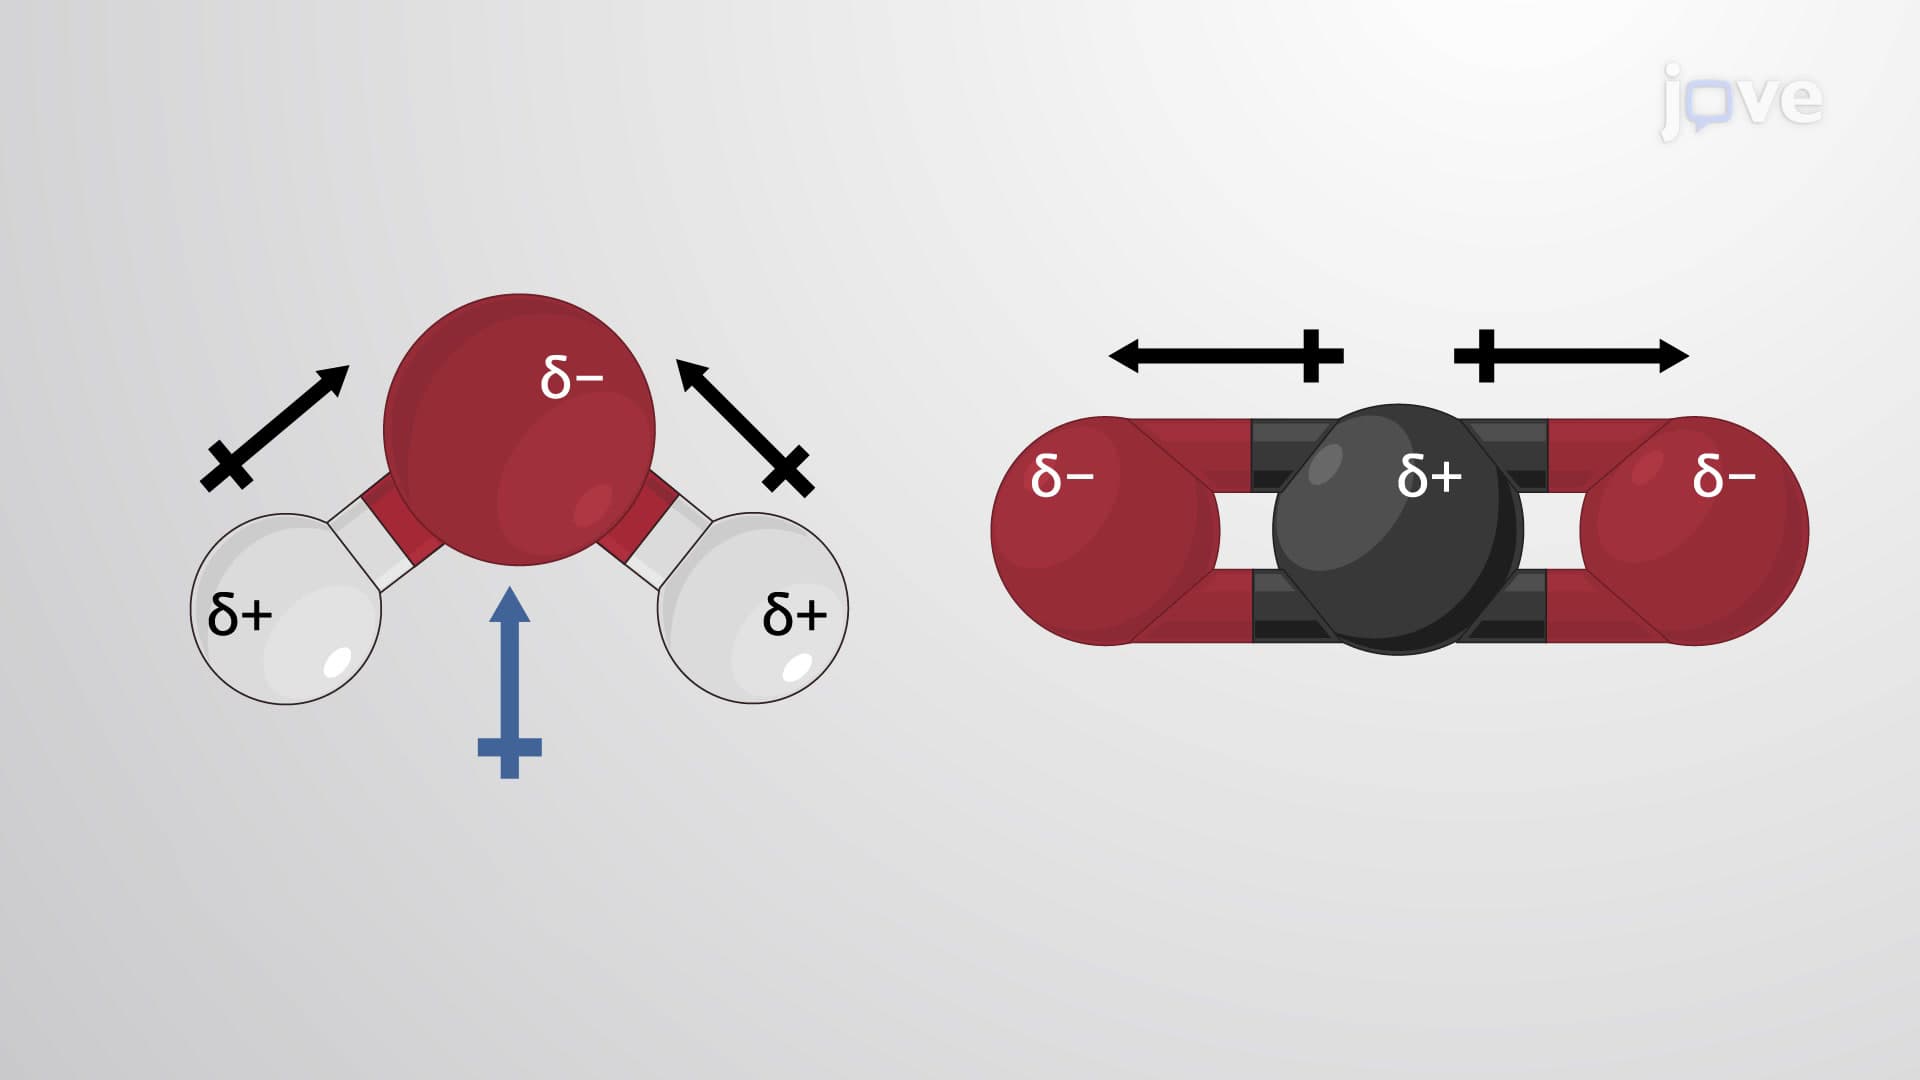
Molecular Geometry and Dipole Moments
 02:00
Molecular Models
 02:18
Fischer Projections
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as...
 01:04
Force and Potential Energy in Three Dimensions
 01:10
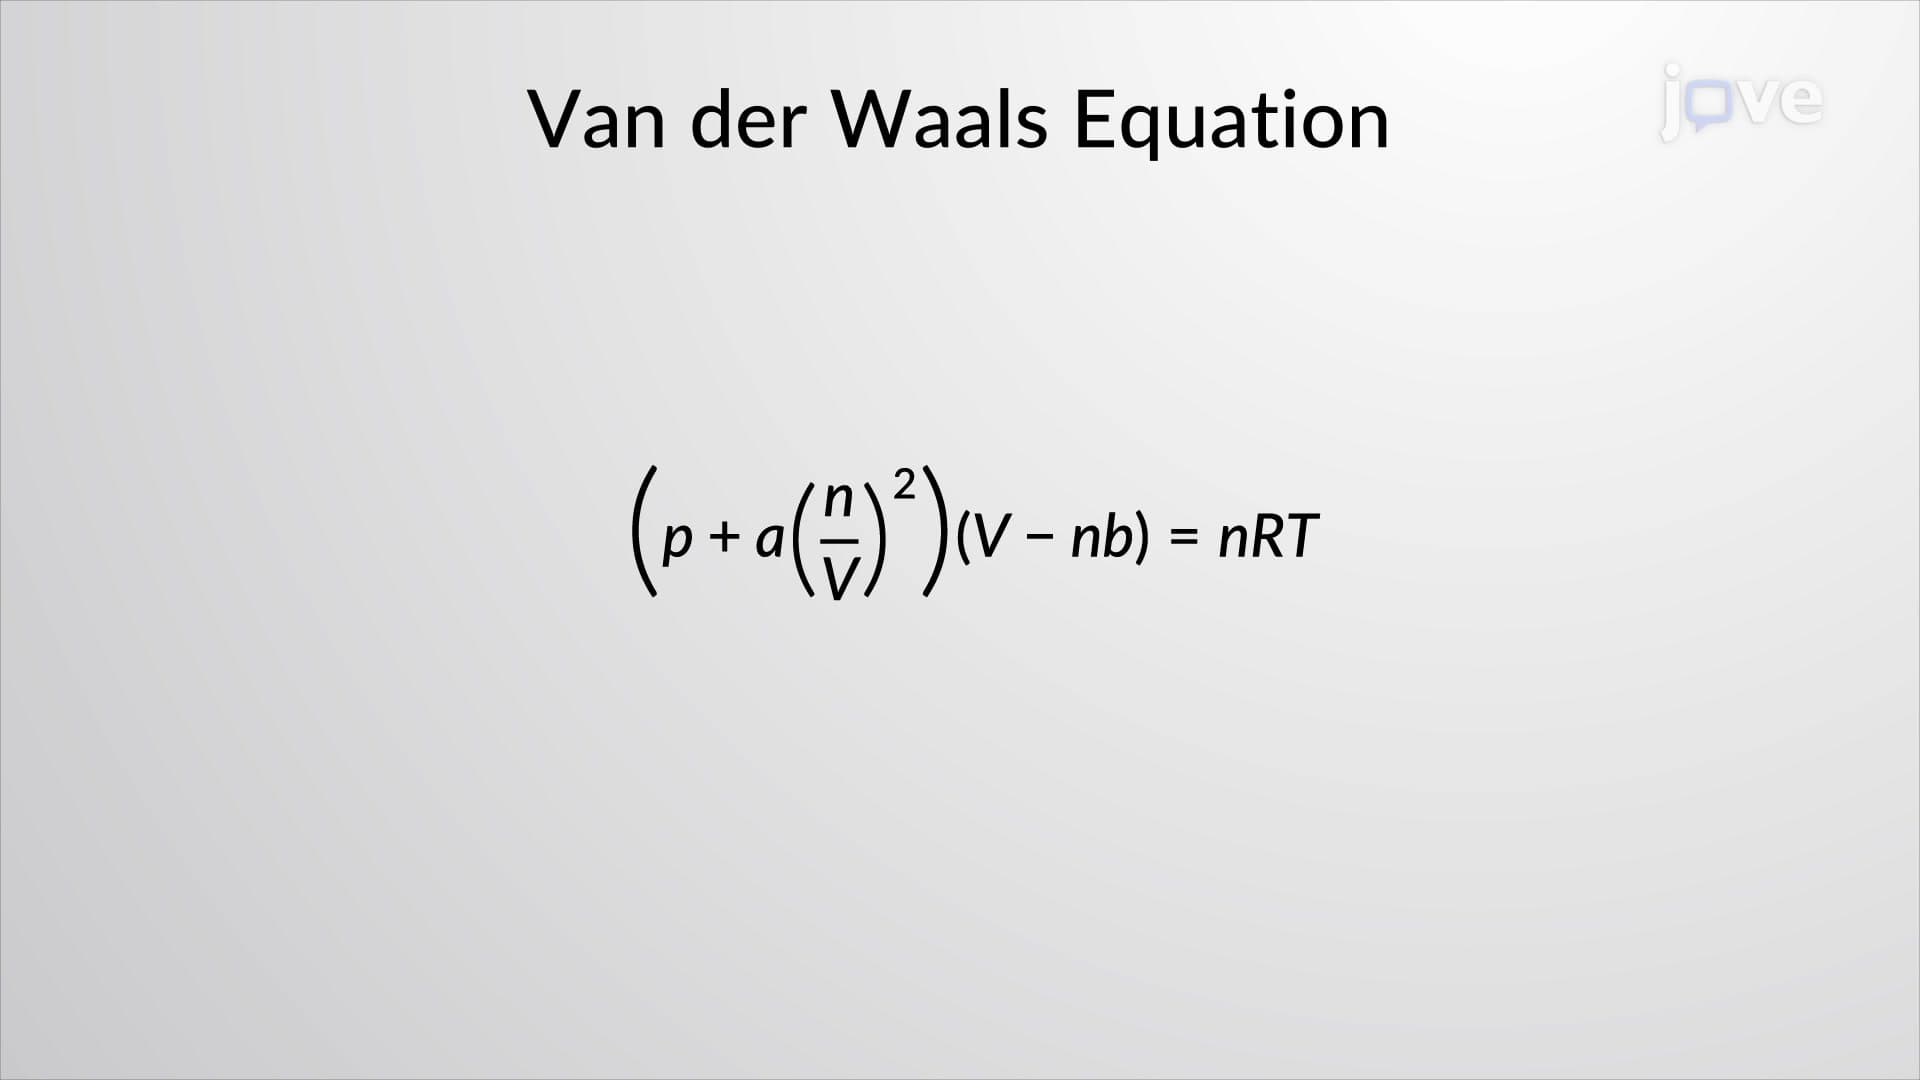
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the...