Teaching NeuroImages: MRI findings in carbamoyl phosphate synthetase 1 deficiency
Sunjay Nunley1, Debabrata Ghosh2
1From Nationwide Children's Hospital, Columbus, OH. sunjay.nunley@nationwidechildrens.org.
Neurology
|May 6, 2015

Abstract
No abstract available in PubMed .
More Related Videos
Related Concept Videos
 01:20
Inborn Errors of Metabolism
1.0K
Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
1.0K
 01:48
ATP Synthase: Mechanism
19.0K
In animals, the mitochondrial F1F0 ATP synthase is the key protein that synthesizes ATP molecules through a complex catalytic mechanism. While the nuclear genome encodes the majority of ATP synthase subunits, the mitochondrial genome encodes some of the enzyme's most critical components. The formation of this multi-subunit enzyme is a complex multi-step process regulated at the level of transcription, translation, and assembly. Defects in one or more of these steps can result in decreased...
19.0K
 01:18
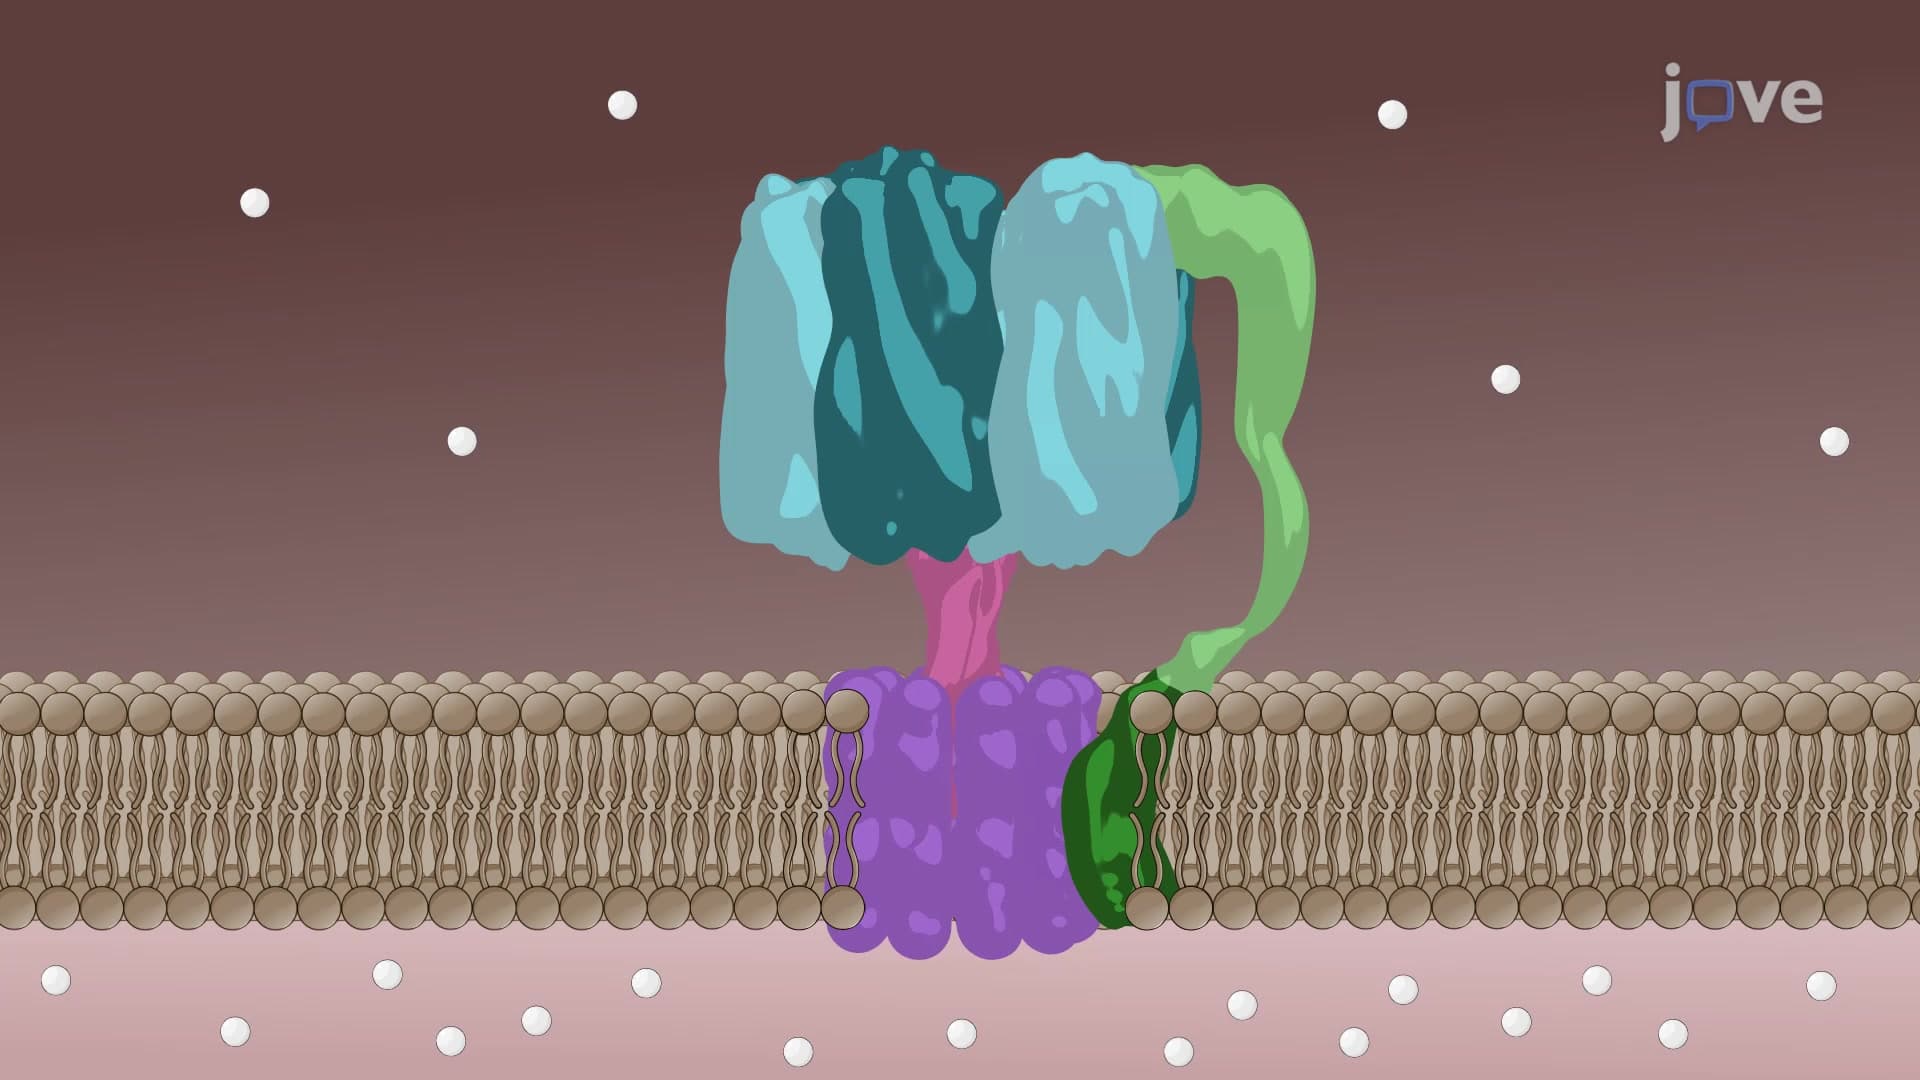
ATP Synthase: Structure
18.0K
ATP synthase or ATPase is among the most conserved proteins found in bacteria, mammals, and plants. This enzyme can catalyze a forward reaction in response to the electrochemical gradient, producing ATP from ADP and inorganic phosphate. ATP synthase can also work in a reverse direction by hydrolyzing ATP and generating an electrochemical gradient. Different forms of ATP synthases have evolved special features to meet the specific demands of the cell. Based on their specific feature, ATP...
18.0K
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
767
Hypertrophic cardiomyopathy, or HCM, is an autosomal dominant genetic disorder characterized by asymmetric left ventricular hypertrophy without ventricular dilation. It is more common in men and is typically diagnosed in young, athletic adults.EtiologyHCM is primarily genetic and is caused by mutations in genes encoding sarcomeric proteins. Researchers have identified over 1400 mutations across at least 11 different genes. Among these, the most frequently occurring mutations are found in the...
767
 01:29
Cardiomyopathy IV: Restrictive Cardiomyopathy
884
Restrictive cardiomyopathy (RCM) is a rare heart muscle disease characterized by impaired ventricular filling due to stiffened ventricular walls, leading to significant diastolic dysfunction.EtiologyRestrictive cardiomyopathy can arise from both inherited and acquired diseases, many of which are systemic. It is categorized into four main types: infiltrative, storage, non-infiltrative, and endomyocardial diseases.Infiltrative diseases, such as amyloidosis, lead to RCM by depositing amyloid...
884
 01:19
Allosteric Proteins-ATCase
7.0K
Binding sites linkages can regulate a protein's function. For example, enzyme activity is often regulated through a feedback mechanism where the end product of the biochemical process serves as an inhibitor.
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...
7.0K