Nucleic acid reactivity: challenges for next-generation semiempirical quantum models
Related Experiment Video
Updated: Apr 13, 2026
Author Spotlight: In Silico Creation and Impact of Carbonylated Amino Acids on Protein Structure and Function
Published on: April 26, 2024
Ming Huang1,2, Timothy J Giese2, Darrin M York2
1Department of Chemistry, Scientific Computation, University of Minnesota, 207 Pleasant St. SE, Minneapolis, Minnesota, 55455-0431.

Abstract:
Semiempirical quantum models are routinely used to study mechanisms of RNA catalysis and phosphoryl transfer reactions using combined quantum mechanical (QM)/molecular mechanical methods. Herein, we provide a broad assessment of the performance of existing semiempirical quantum models to describe nucleic acid structure and reactivity to quantify their limitations and guide the development of next-generation quantum models with improved accuracy. Neglect of diatomic differential overlap and self-consistent density-functional tight-binding semiempirical models are evaluated against high-level QM benchmark calculations for seven biologically important datasets. The datasets include: proton affinities, polarizabilities, nucleobase dimer interactions, dimethyl phosphate anion, nucleoside sugar and glycosidic torsion conformations, and RNA phosphoryl transfer model reactions. As an additional baseline, comparisons are made with several commonly used density-functional models, including M062X and B3LYP (in some cases with dispersion corrections). The results show that, among the semiempirical models examined, the AM1/d-PhoT model is the most robust at predicting proton affinities. AM1/d-PhoT and DFTB3-3ob/OPhyd reproduce the MP2 potential energy surfaces of 6 associative RNA phosphoryl transfer model reactions reasonably well. Further, a recently developed linear-scaling "modified divide-and-conquer" model exhibits the most accurate results for binding energies of both hydrogen bonded and stacked nucleobase dimers. The semiempirical models considered here are shown to underestimate the isotropic polarizabilities of neutral molecules by approximately 30%. The semiempirical models also fail to adequately describe torsion profiles for the dimethyl phosphate anion, the nucleoside sugar ring puckers, and the rotations about the nucleoside glycosidic bond. The modeling of pentavalent phosphorus, particularly with thio substitutions often used experimentally as mechanistic probes, was problematic for all of the models considered. Analysis of the strengths and weakness of the models suggests that the creation of robust next-generation models should emphasize the improvement of relative conformational energies and barriers, and nonbonded interactions.
More Related Videos
 10:34
Probing RNA Structure with Dimethyl Sulfate Mutational Profiling with Sequencing In Vitro and in Cells
Published on: December 9, 2022
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 02:00
Molecular Models
 01:13
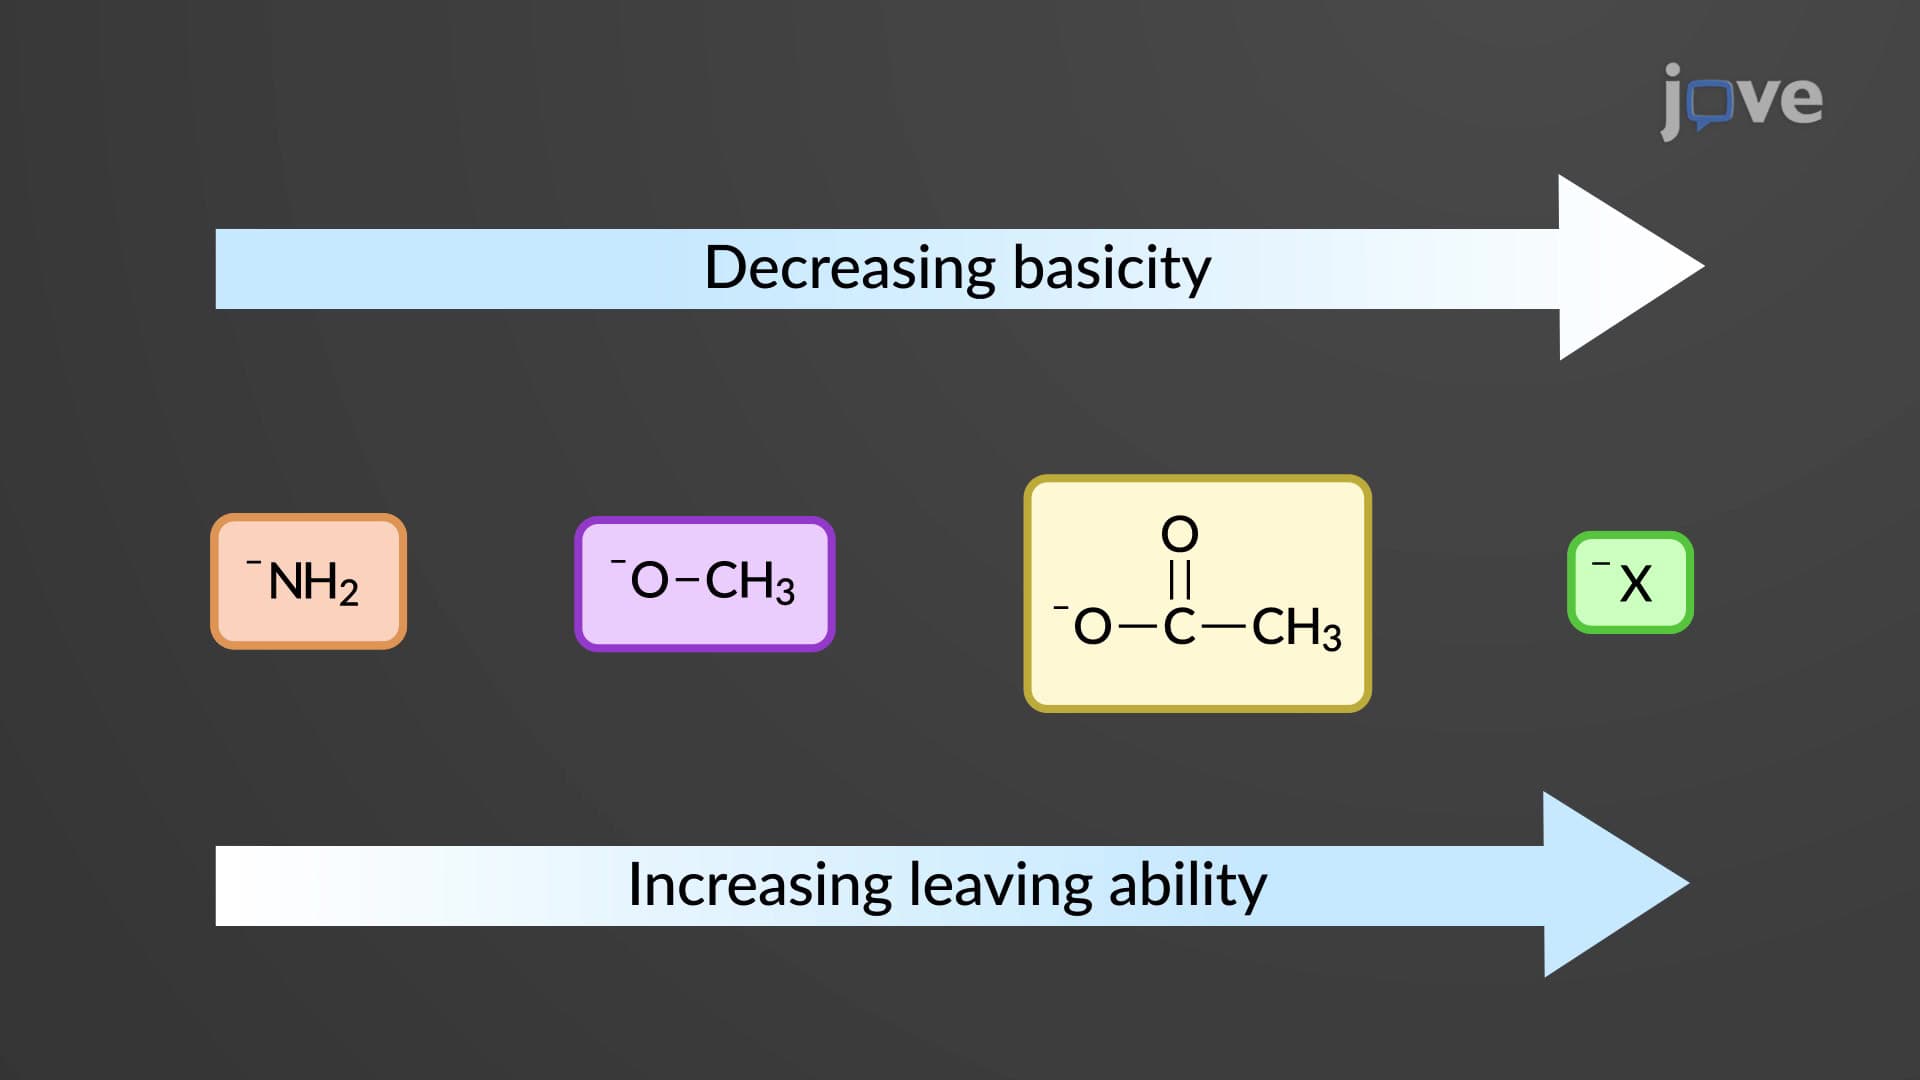
Relative Reactivity of Carboxylic Acid Derivatives
A key factor in assessing the reactivity of the acid derivatives is the basicity of the substituent or the leaving group. The lower the basicity of the leaving group, the higher the reactivity of the derivative. The basicity of the leaving group follows this order:
Halide ions < Acyloxy ions < Alkoxy ions < Amine ions
 01:25
Nucleic Acid Structure
DNA Structure
DNA...
 02:45
The Quantum-Mechanical Model of an Atom
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:16
Radical Reactivity: Nucleophilic Radicals