Related Experiment Video
Updated: Apr 12, 2026


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Grand Canonical Monte Carlo Simulations Guided by an Analytic Equation of State-Transferable Anisotropic Mie
Andrea Hemmen1, Athanassios Z Panagiotopoulos2, Joachim Gross1
1†Institute of Thermodynamics and Thermal Process Engineering, University of Stuttgart, Pfaffenwaldring 9, 70569 Stuttgart, Germany.

Abstract:
In this study, we propose using an analytical equation of state for guiding molecular simulations in the grand canonical ensemble. Molecular simulations in the grand canonical ensemble deliver phase equilibrium properties with low statistical uncertainty. The entire phase envelope can be obtained when histograms of several simulations along the phase envelope are combined. In this study, we explore the use of an analytical equation of state for defining chemical potentials, temperatures, and intervals of molecule numbers for simulations in the grand canonical ensemble, such that the phase envelope is traced. We limit particle numbers to intervals and ensure even sampling of molecule numbers in each interval by applying a bias potential determined from transition-matrix sampling. The methodology is described for pure components and binary mixtures. We apply the simulation method to develop parameters of the transferable anisotropic Mie (TAMie) force field for ethers. We find that the partial charges optimized individually for diethyl ether and for dipropyl ether differ substantially from the partial charges optimized simultaneously to both substances. The concept of transferable partial charges is thus a significant assumption. For developing the (TAMie) force field, we constrained the partial charge to a range, where individually optimized partial charges were found.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:17
Physical Properties of Ethers
An ether molecule has a net dipole moment due to the polarity of C–O bonds. Subsequently, boiling points of ethers are lower than those of alcohols of comparable molecular weight and slightly higher than those of hydrocarbons of comparable molecular weight (Table 1).
Ethers can act as hydrogen bond acceptors, making them more water-soluble than hydrocarbons, but since ethers cannot act as hydrogen bond donors, they are much less soluble in water than alcohols. Ethers are considered...
 01:07
Equation of State
 01:27
Variables and Equations of State
 01:28
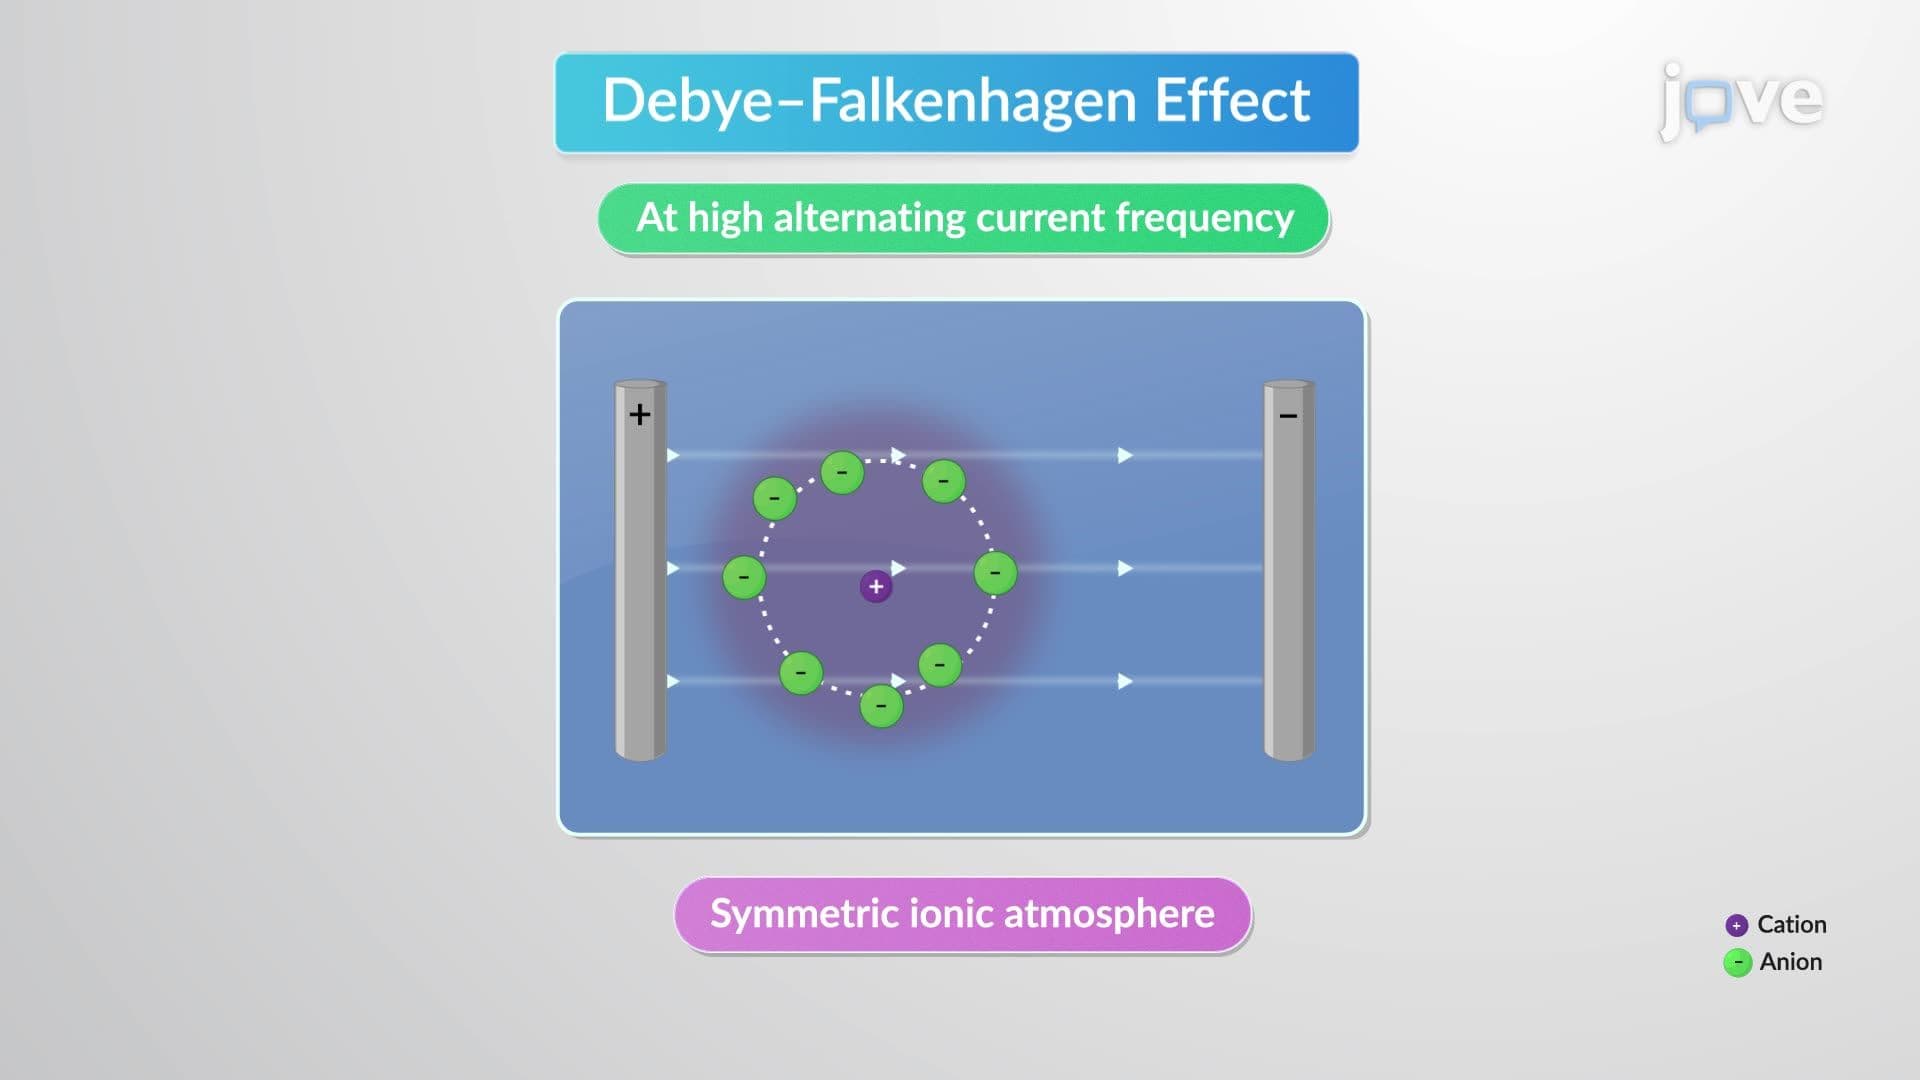
Debye–Huckel–Onsager Conductance Equation
 02:35
Ethers from Alkenes: Alcohol Addition and Alkoxymercuration-Demercuration
Ethers can also be prepared from alkenes through acid-catalyzed addition of alcohols and alkoxymercuration–demercuration.
Preparation of Ethers by Acid-Catalyzed Addition of Alcohol to Alkenes
The acid-catalyzed addition of alcohol to an alkene involves treating the alkene with an excess of alcohol in the presence of an acid catalyst to form an ether under suitable conditions. The hydrogen will add to the less substituted carbon so that the nucleophile can attack the more substituted...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...