Related Experiment Video
Updated: Apr 7, 2026
Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
Conformational preferences of monohydrated clusters of imidazole derivatives revisited
Aditi Bhattacherjee1, Sanjay Wategaonkar
1Department of Chemical Sciences, Tata Institute of Fundamental Research, Homi Bhabha Road, Mumbai 400 005, India. abhattacherjee@lbl.gov sanwat@tifr.res.in.

Abstract:
We present the IR spectroscopic investigations of benzimidazole (BIM), N-methylbenzimidazole (MBIM), and their monohydrates (W1) carried out in a supersonic jet in the region of N-H, C-H, and O-H stretching fundamentals. The C-H stretching modes in the monomers were studied with the aim of identifying the C(2)H mode (the C atom between the two N atoms in the imidazole moiety) which is known to play an important role as a H-bond donor in enzymes and ionic liquids. The assignment was aided by quantum chemical calculations as well as the literature data for FTIR measurements in the matrix. The monohydrated clusters were investigated for a global comparison with previously reported conformations of hydrated imidazole and related derivatives in the gas phase, matrices, and He nanodroplets. The BIM-W1 complex showed the presence of three conformers; an N-H∙∙∙O bound conformer (A') and two O-H∙∙∙N bound conformers, tilted towards the phenyl side (A) and the imidazole side (B), respectively. Although both A' and B type conformers have been reported in the literature, our experiments identify a new conformer (conformer A) in the gas phase for the first time which has also been reported in crystal structures of histidine containing proteins. The binding energies of the three conformers were found to be of comparable magnitude, with the N-HO bound structure lying in between (∼0.02-0.04 kcal mol(-1)) the O-H∙∙∙N bound ones at the counterpoise corrected (cp) MP2/aug-cc-pVDZ level of theory. The formation of two distinct but closely related O-H∙∙∙N bound conformers (A and B) was additionally confirmed by studying the MBIM-W1 complex. Binding energies of the MBIM-W1 conformers were found to be higher than those of the analogous BIM-W1 conformers by 0.2 kcal mol(-1) at the cp-MP2/aug-cc-pVDZ level. The C(2)-H∙∙∙O or π bound water structures were not observed in the beam for monohydrated clusters of either monomer. While QTAIM calculations predicted secondary stabilization in the A type conformer by a C(4)-HO hydrogen bond, such an effect due to a possible C(2)-HO interaction was not found for conformer B. Experimentally, however, no spectral evidence was found for either the C(4)-H∙∙∙O or the C(2)-H∙∙∙O interaction.
Related Concept Videos
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 02:37
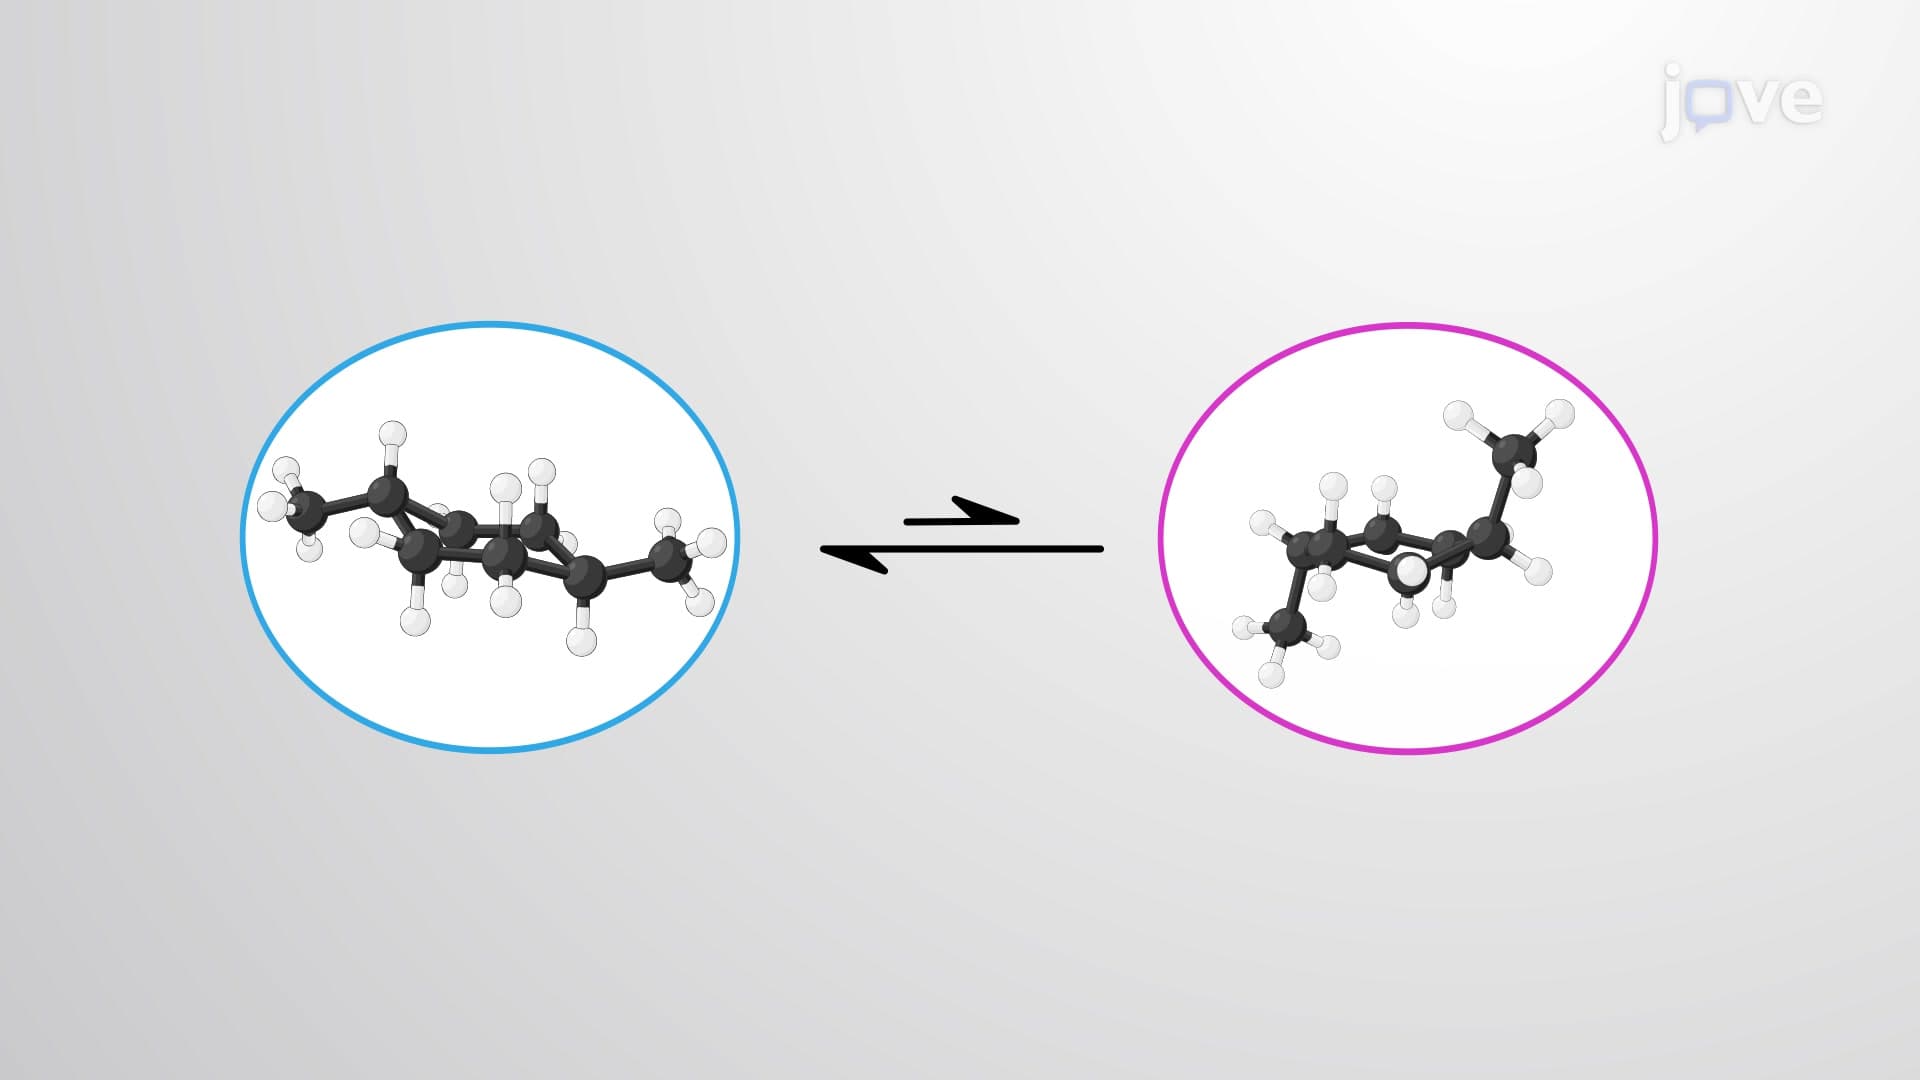
Disubstituted Cyclohexanes: cis-trans Isomerism
In cyclohexane, the substituents can occupy different positions generating distinct isomers....
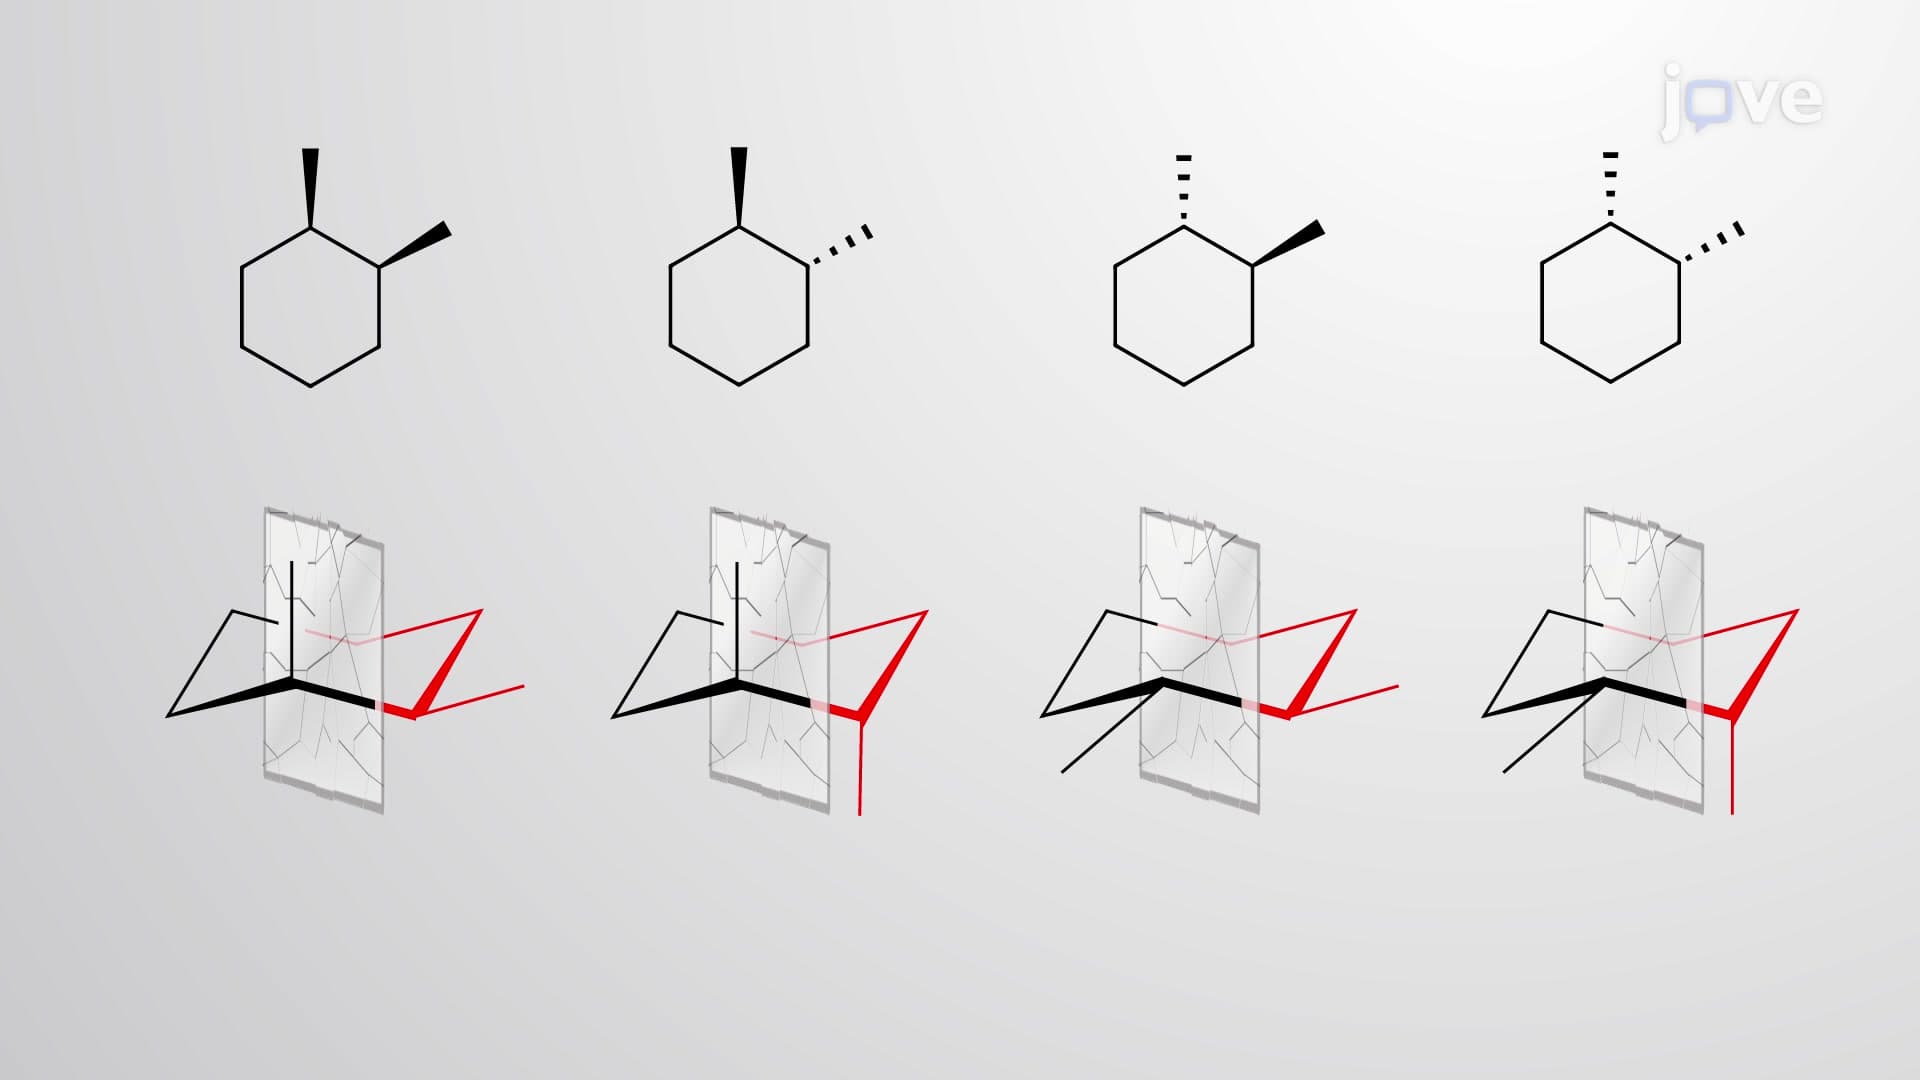 02:33
Stereoisomerism of Cyclic Compounds
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this...