Related Experiment Video
Updated: Apr 6, 2026

In Vitro Analysis of PDZ-dependent CFTR Macromolecular Signaling Complexes
Published on: August 13, 2012
Cystic Fibrosis Transmembrane Conductance Regulator (CFTR): CLOSED AND OPEN STATE CHANNEL MODELS
Valentina Corradi1, Paola Vergani2, D Peter Tieleman3
1From the Department of Biological Sciences and Centre for Molecular Simulation, University of Calgary, Calgary, Alberta T2N 1N4, Canada and.

Abstract:
The cystic fibrosis transmembrane conductance regulator (CFTR) is a member of the ATP-binding cassette (ABC) transporter superfamily. CFTR controls the flow of anions through the apical membrane of epithelia. Dysfunctional CFTR causes the common lethal genetic disease cystic fibrosis. Transitions between open and closed states of CFTR are regulated by ATP binding and hydrolysis on the cytosolic nucleotide binding domains, which are coupled with the transmembrane (TM) domains forming the pathway for anion permeation. Lack of structural data hampers a global understanding of CFTR and thus the development of "rational" approaches directly targeting defective CFTR. In this work, we explored possible conformational states of the CFTR gating cycle by means of homology modeling. As templates, we used structures of homologous ABC transporters, namely TM(287-288), ABC-B10, McjD, and Sav1866. In the light of published experimental results, structural analysis of the transmembrane cavity suggests that the TM(287-288)-based CFTR model could correspond to a commonly occupied closed state, whereas the McjD-based model could represent an open state. The models capture the important role played by Phe-337 as a filter/gating residue and provide structural information on the conformational transition from closed to open channel.
More Related Videos
 14:56
Expression and Purification of the Cystic Fibrosis Transmembrane Conductance Regulator Protein in Saccharomyces cerevisiae
Published on: March 10, 2012
 09:59
Functional Reconstitution and Channel Activity Measurements of Purified Wildtype and Mutant CFTR Protein
Published on: March 9, 2015
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:05
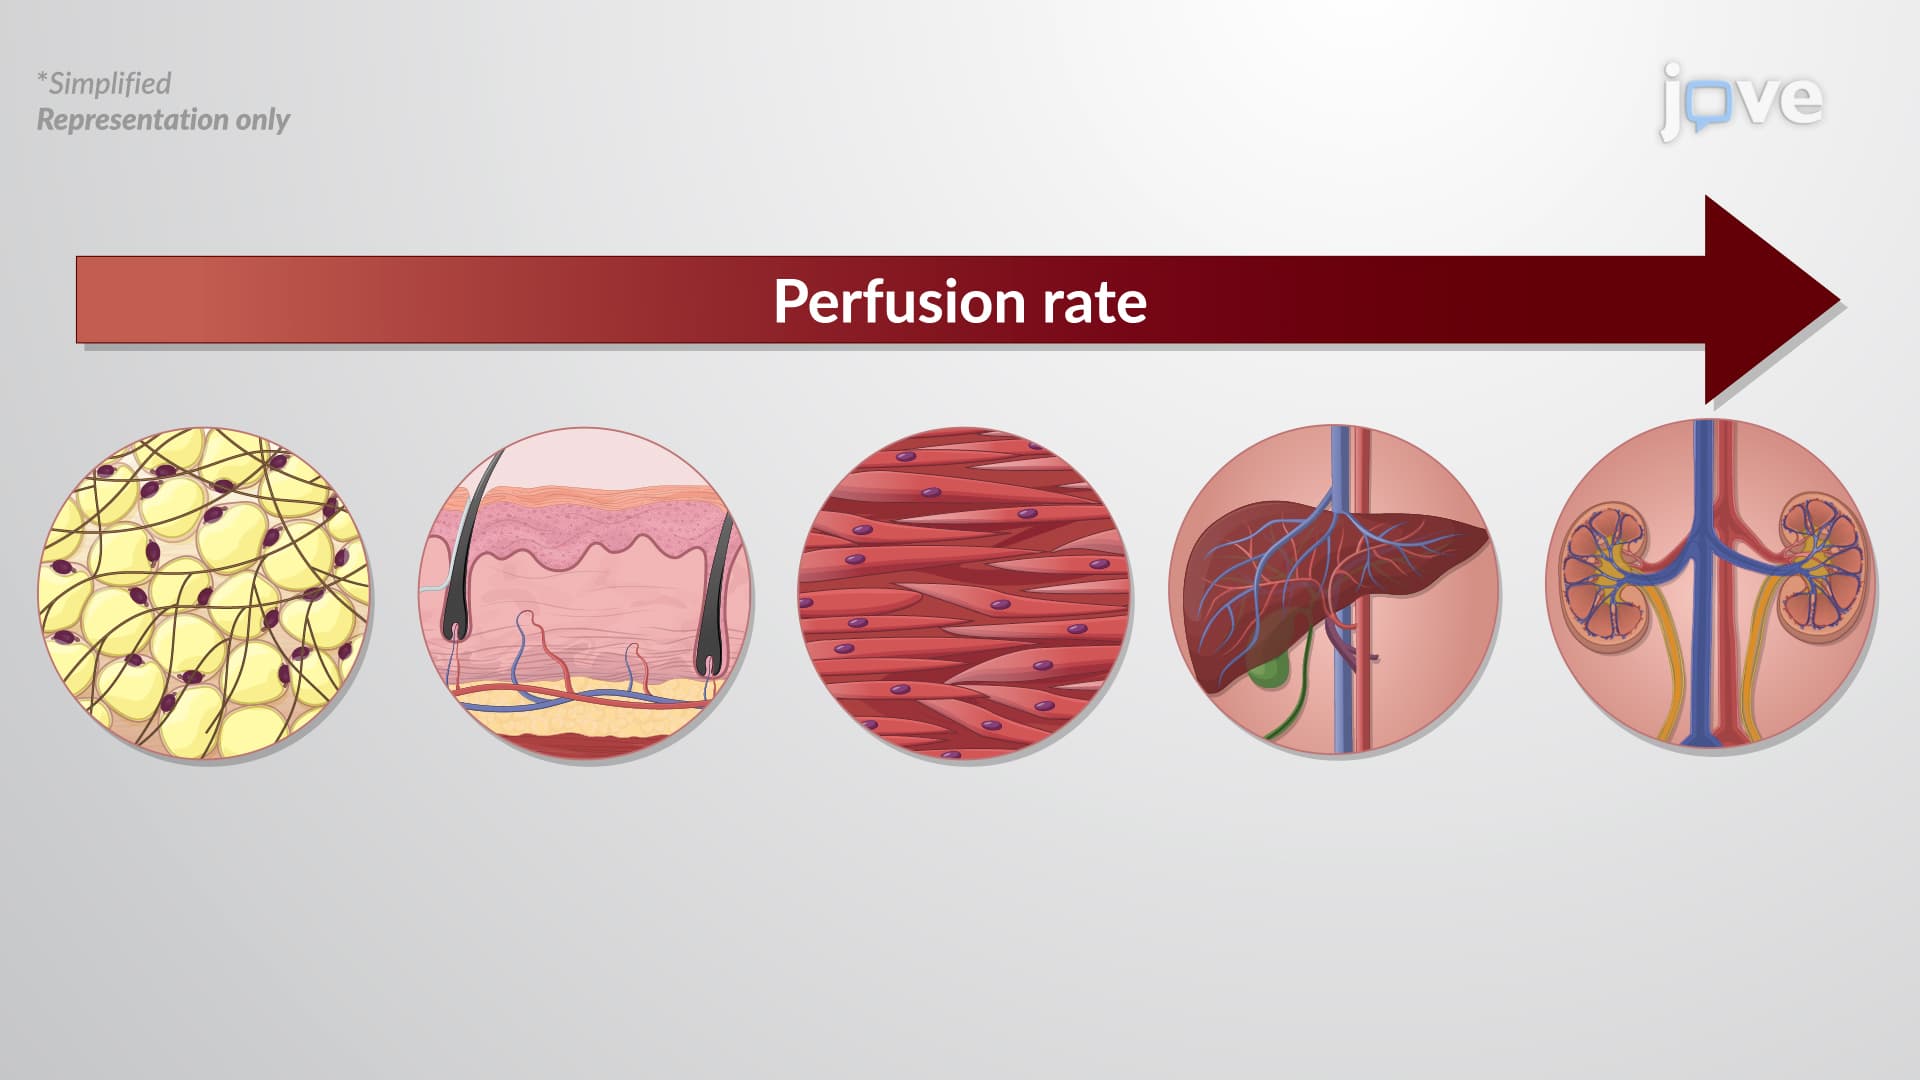
Two-Compartment Open Model: Overview
The...
 01:26
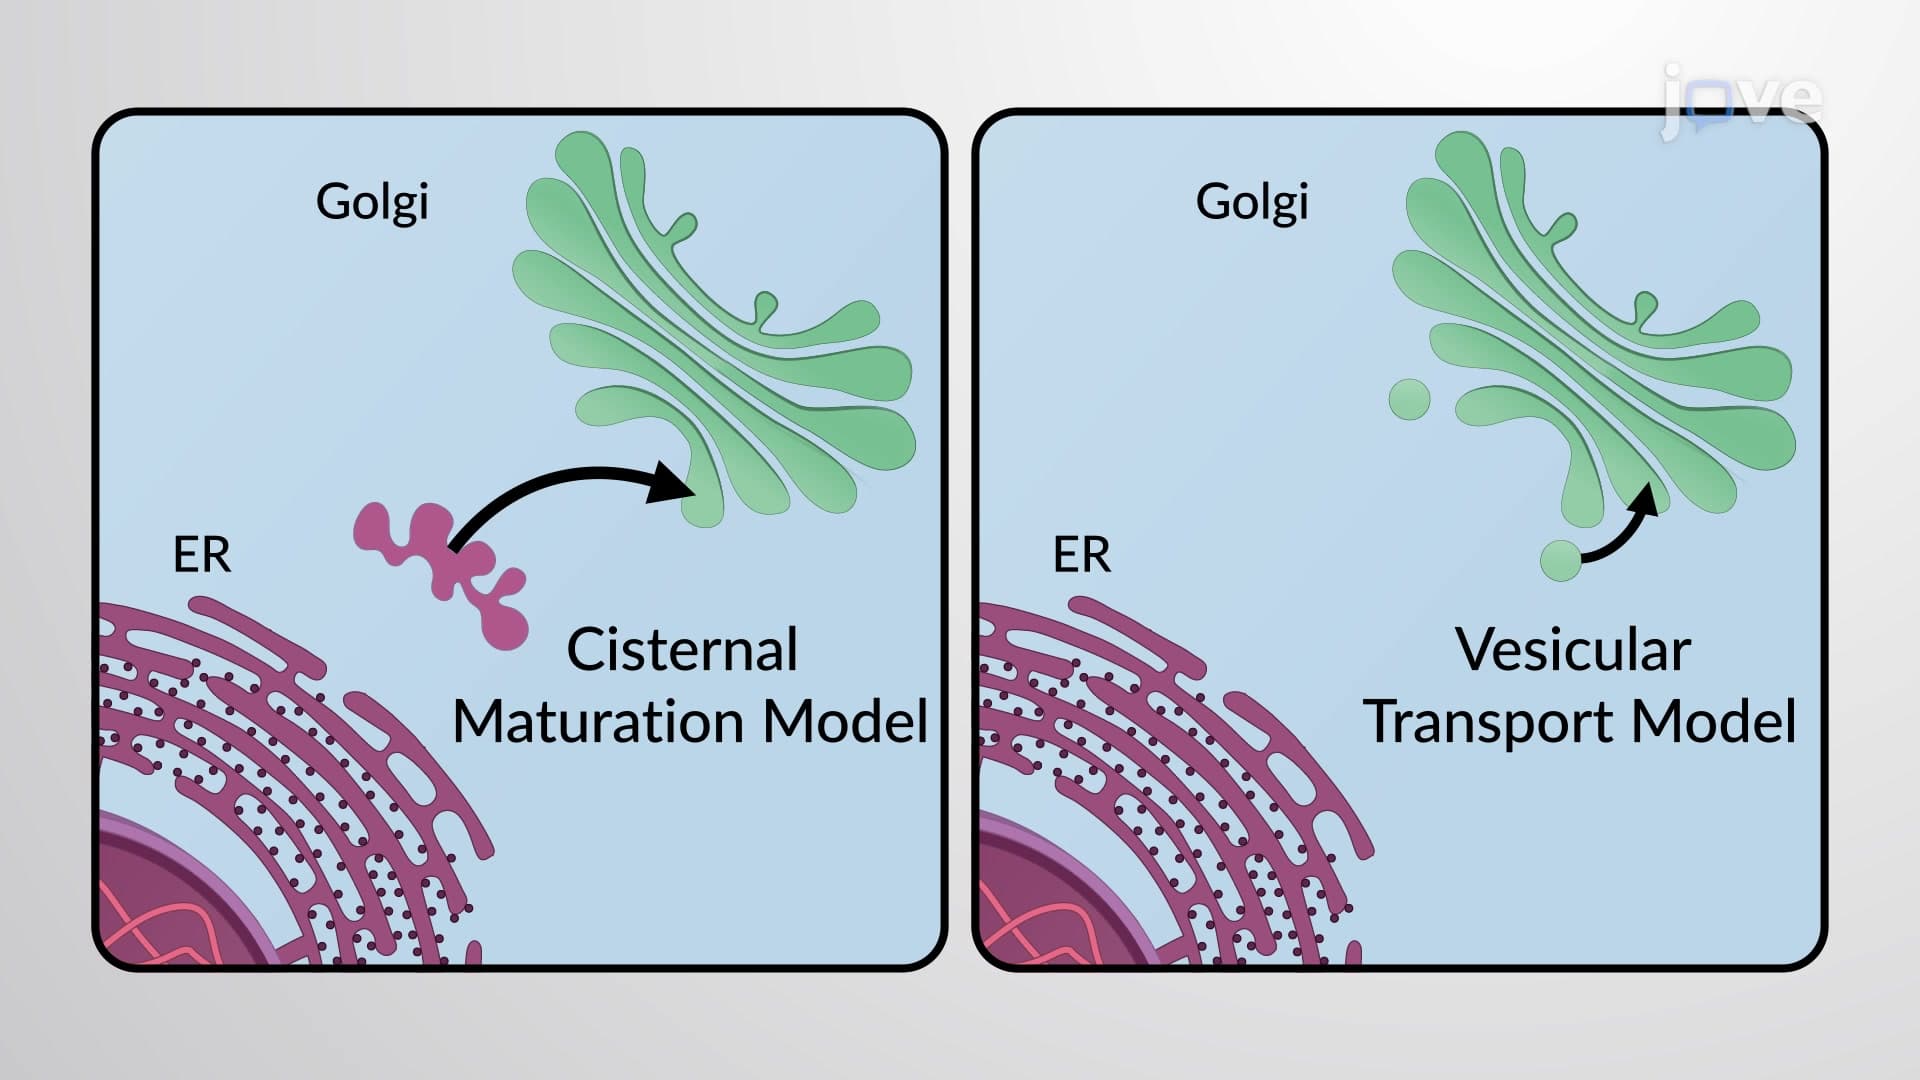
Transport Across the Golgi
 01:09
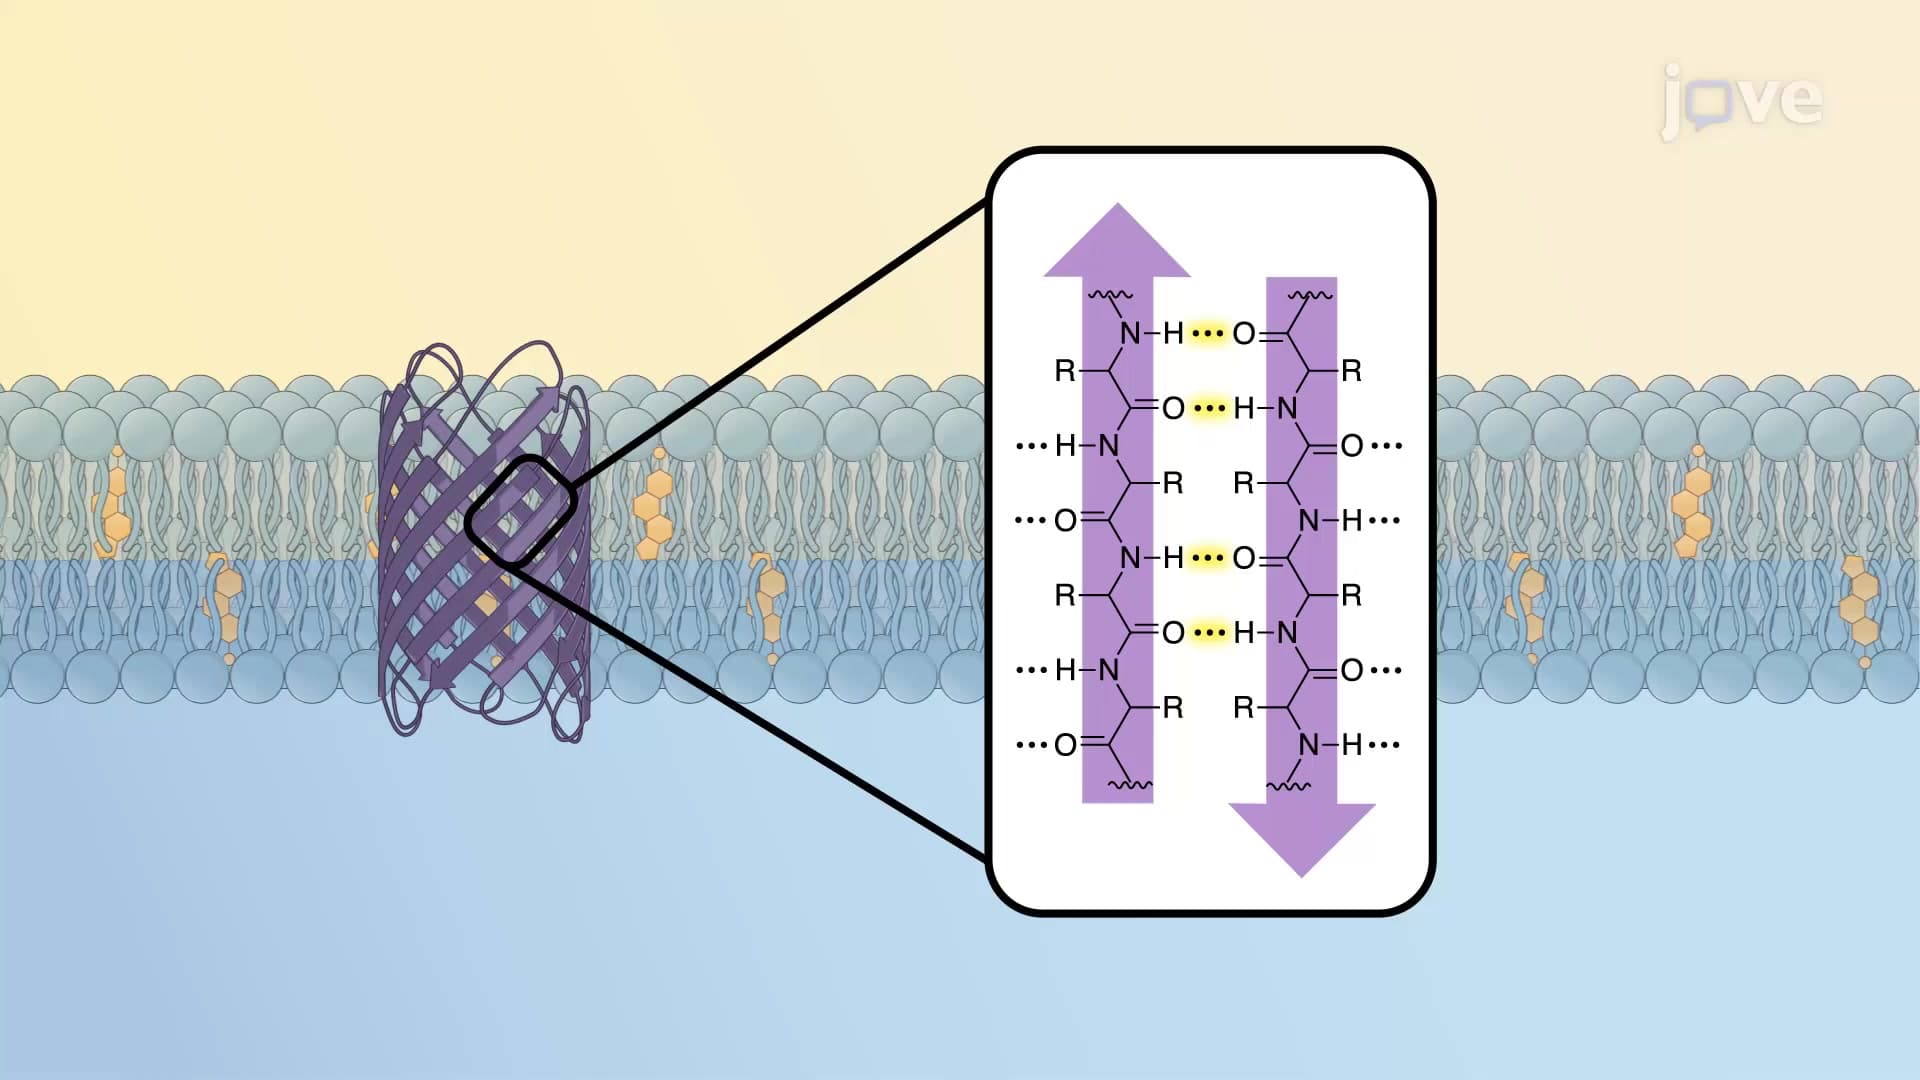
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as...
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism