Related Experiment Video
Updated: Apr 4, 2026


A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Template-based protein structure prediction in CASP11 and retrospect of I-TASSER in the last decade
Jianyi Yang1,2, Wenxuan Zhang1,2, Baoji He1,2
1Department of Computational Medicine and Bioinformatics, University of Michigan, Ann Arbor, Michigan, 48109.

Abstract:
We report the structure prediction results of a new composite pipeline for template-based modeling (TBM) in the 11th CASP experiment. Starting from multiple structure templates identified by LOMETS based meta-threading programs, the QUARK ab initio folding program is extended to generate initial full-length models under strong constraints from template alignments. The final atomic models are then constructed by I-TASSER based fragment reassembly simulations, followed by the fragment-guided molecular dynamic simulation and the MQAP-based model selection. It was found that the inclusion of QUARK-TBM simulations as an intermediate modeling step could help improve the quality of the I-TASSER models for both Easy and Hard TBM targets. Overall, the average TM-score of the first I-TASSER model is 12% higher than that of the best LOMETS templates, with the RMSD in the same threading-aligned regions reduced from 5.8 to 4.7 Å. Nevertheless, there are nearly 18% of TBM domains with the templates deteriorated by the structure assembly pipeline, which may be attributed to the errors of secondary structure and domain orientation predictions that propagate through and degrade the procedures of template identification and final model selections. To examine the record of progress, we made a retrospective report of the I-TASSER pipeline in the last five CASP experiments (CASP7-11). The data show no clear progress of the LOMETS threading programs over PSI-BLAST; but obvious progress on structural improvement relative to threading templates was witnessed in recent CASP experiments, which is probably attributed to the integration of the extended ab initio folding simulation with the threading assembly pipeline and the introduction of atomic-level structure refinements following the reduced modeling simulations. Proteins 2016; 84(Suppl 1):233-246. © 2015 Wiley Periodicals, Inc.


Related Concept Videos
 01:24
Protein Organization
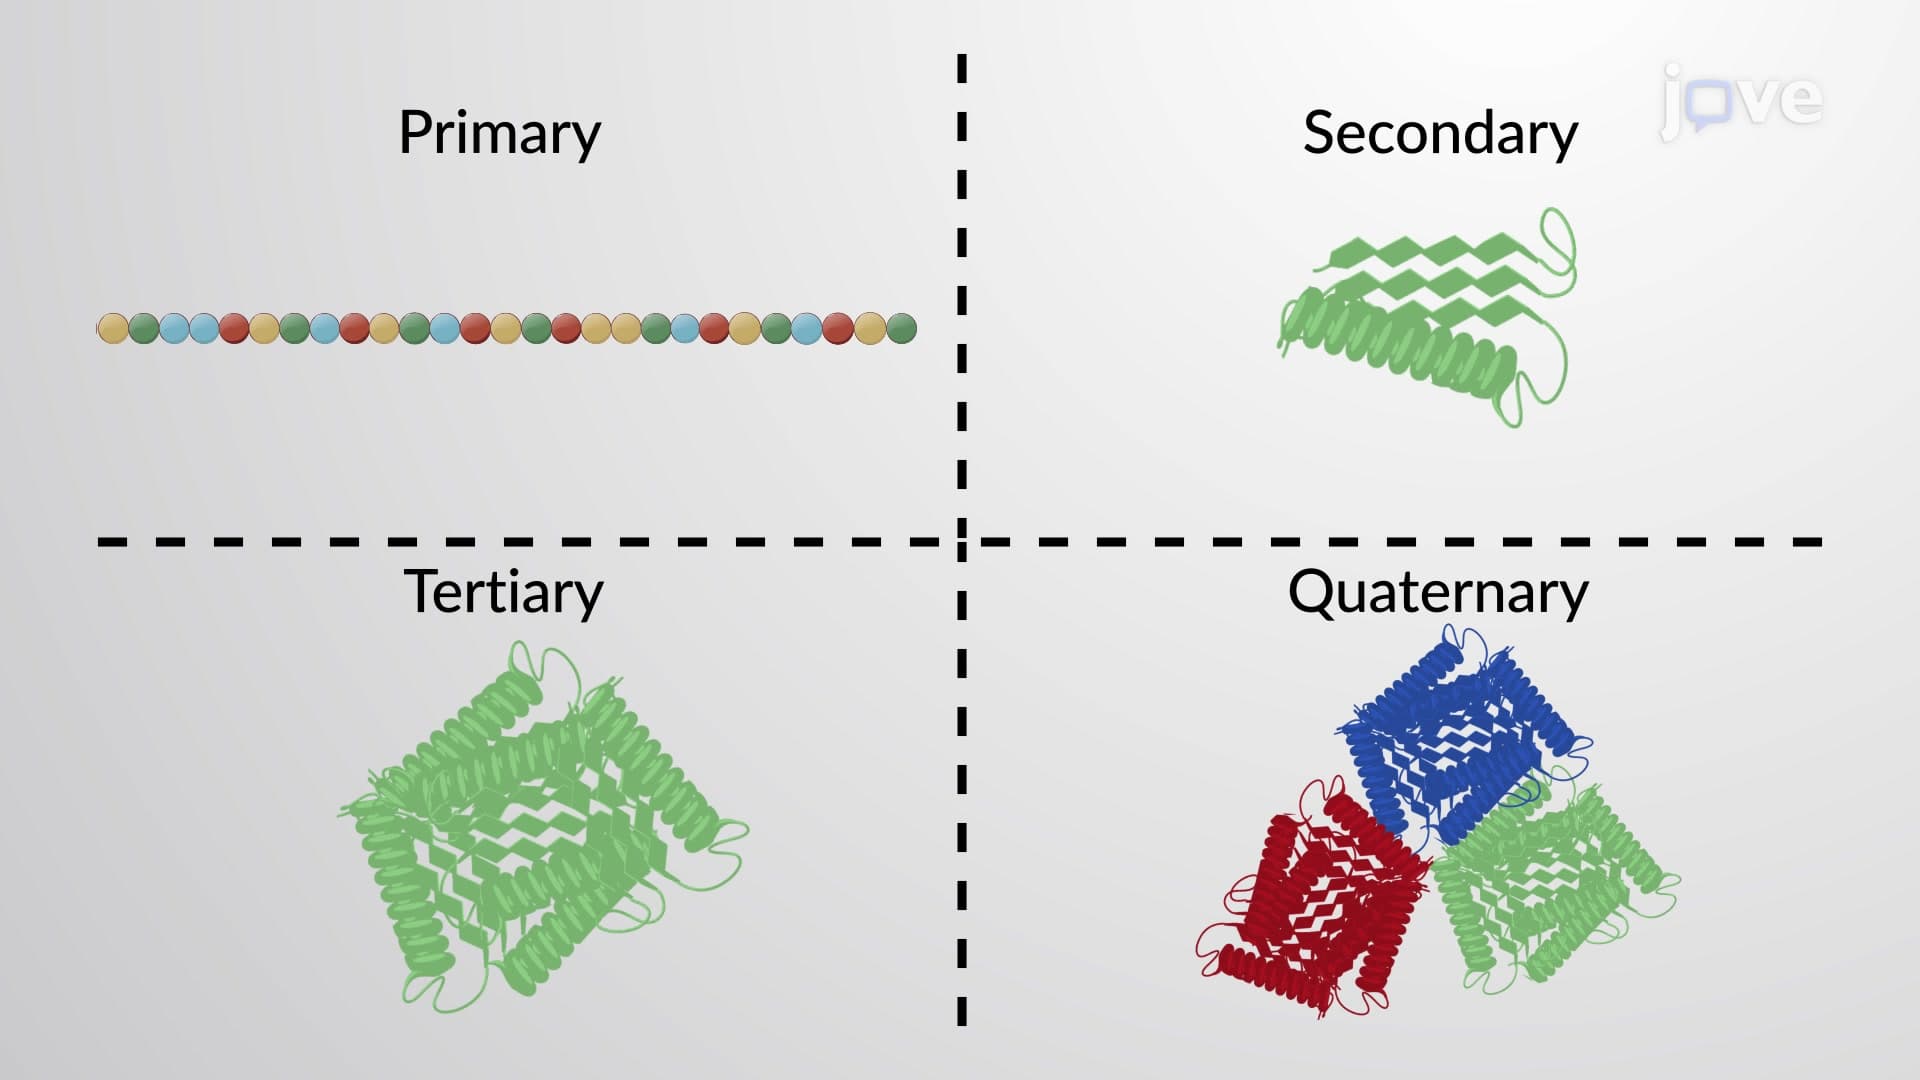
The primary structure of a protein is its amino acid sequence....
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:15
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
02:15Protein and Protein Structures
 02:04
Protein-protein Interfaces
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...