Related Experiment Video
Updated: Apr 3, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Quantum Monte Carlo calculation of the binding energy of bilayer graphene
E Mostaani1, N D Drummond1, V I Fal'ko2
1Department of Physics, Lancaster University, Lancaster LA1 4YB, United Kingdom.

Abstract:
We report diffusion quantum Monte Carlo calculations of the interlayer binding energy of bilayer graphene. We find the binding energies of the AA-and AB-stacked structures at the equilibrium separation to be 11.5(9) and 17.7(9) meV/atom, respectively. The out-of-plane zone-center optical phonon frequency predicted by our binding-energy curve is consistent with available experimental results. As well as assisting the modeling of interactions between graphene layers, our results will facilitate the development of van der Waals exchange-correlation functionals for density functional theory calculations.
Related Concept Videos
 02:13
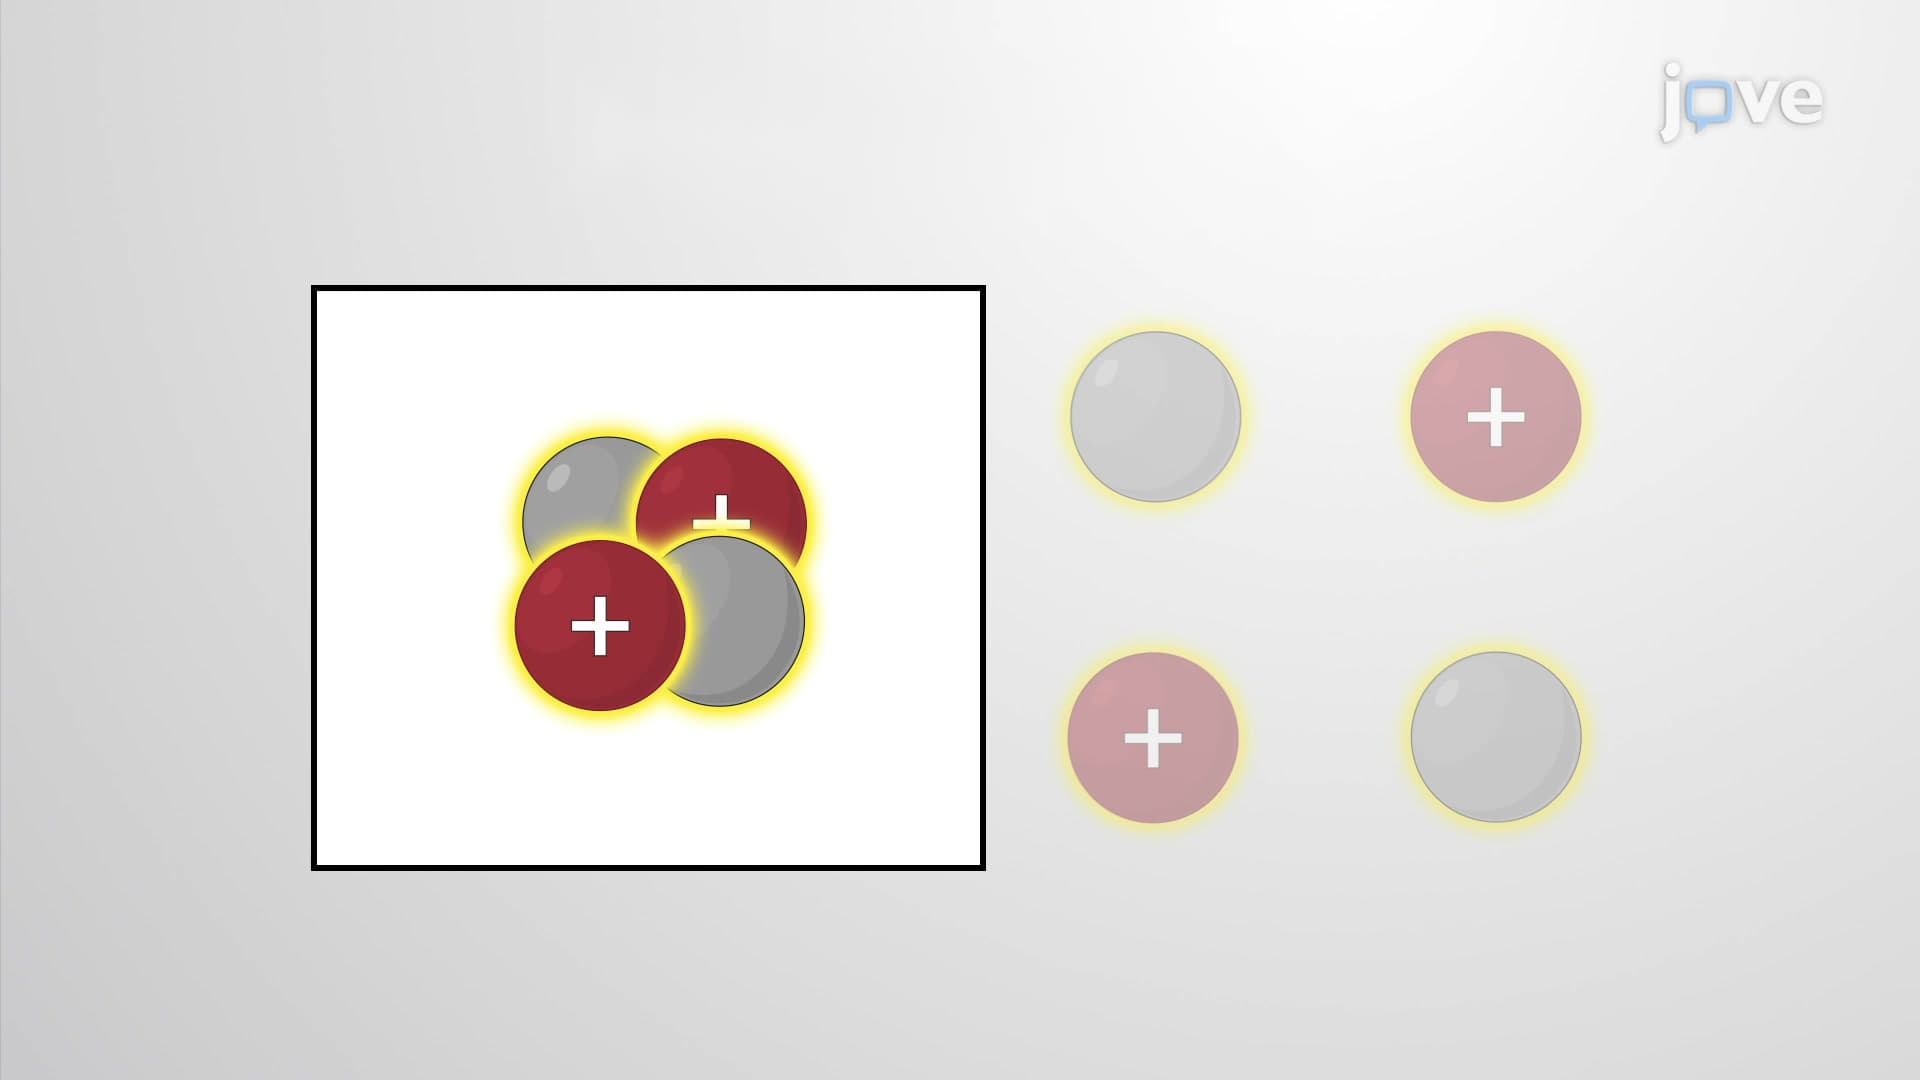
Nuclear Binding Energy
 02:45
Valence Bond Theory
 02:42
Valence Bond Theory
 02:40
MO Theory and Covalent Bonding
 03:51
Molecular Orbital Theory II
 02:13
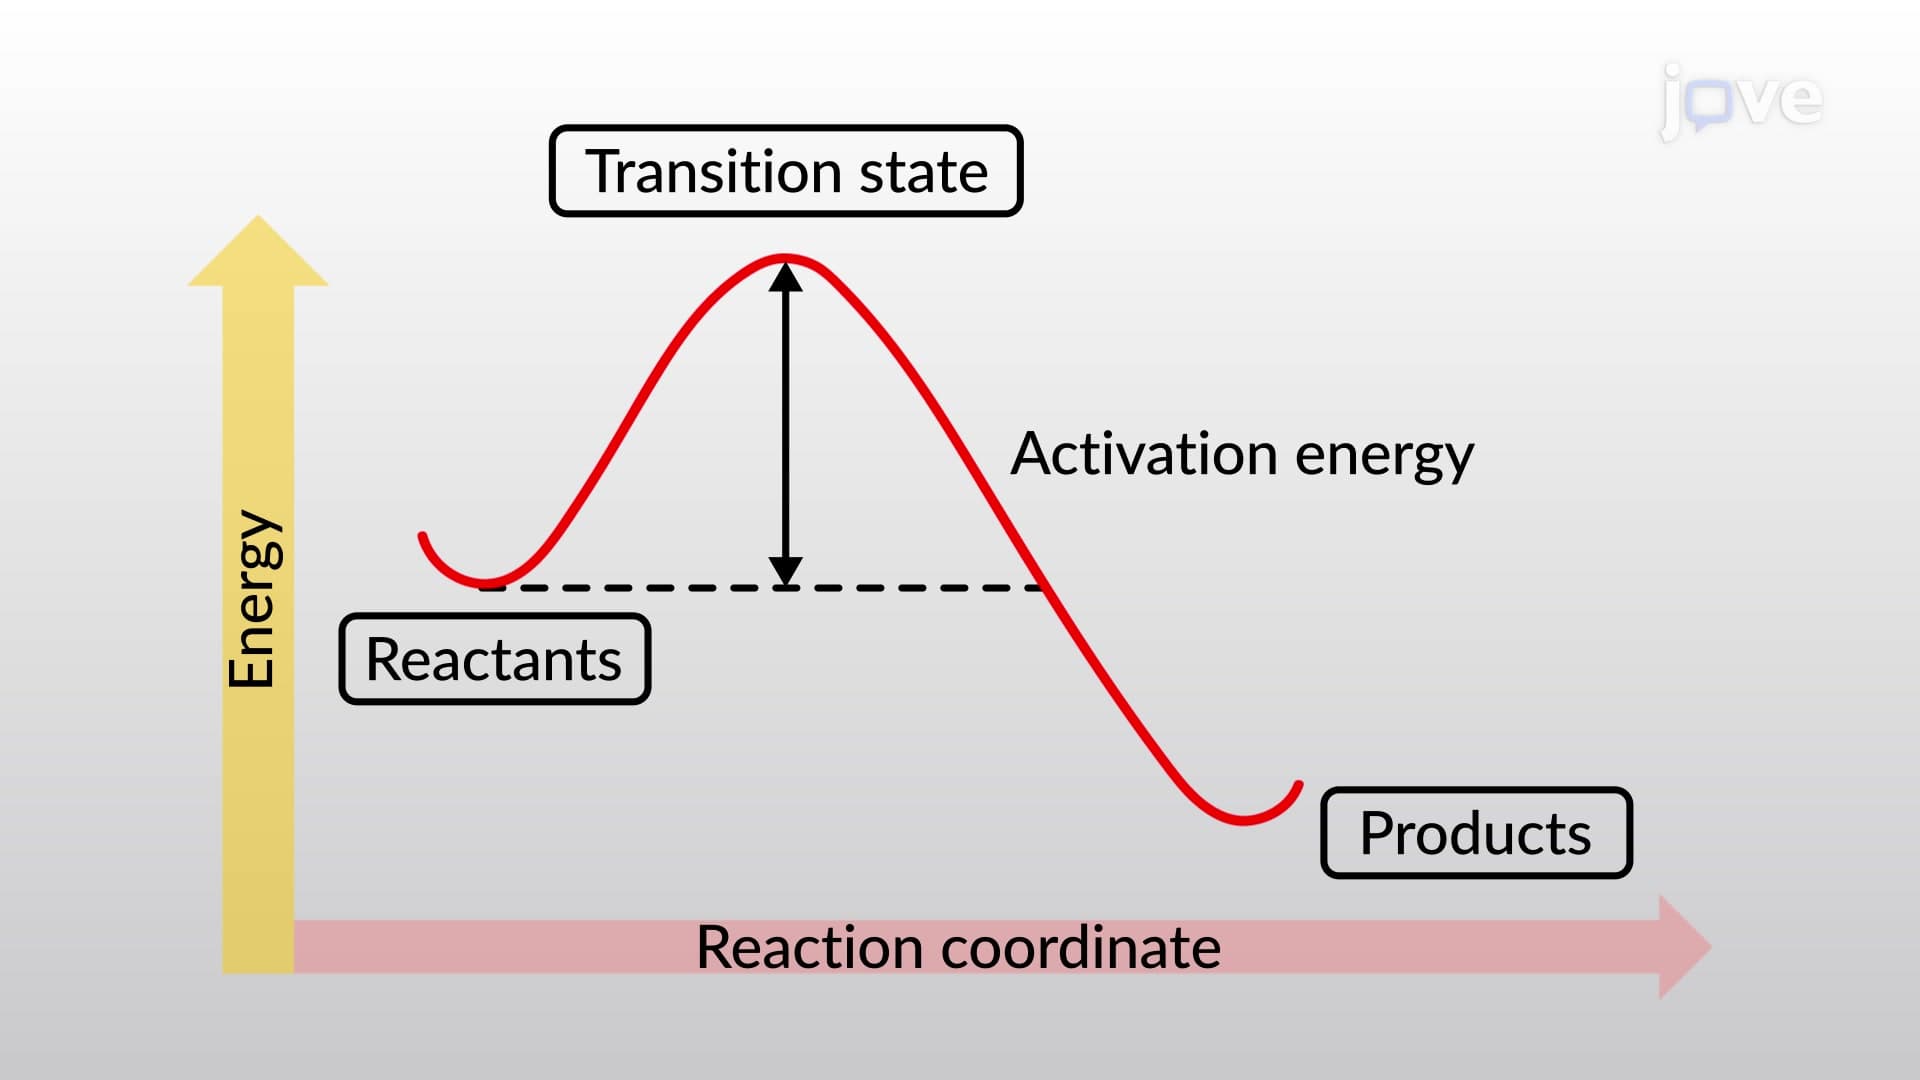
Bond Dissociation Energy and Activation Energy