
Properties of noncovalent tetraphenylporphine···C60 dyads as studied by different long-range and dispersion-corrected
Oscar Amelines-Sarria1, Vladimir A Basiuk, Víctor Duarte-Alaniz
1Instituto de Física, Universidad Nacional Autónoma de México, Sendero Bicipuma, Cd. Universitaria, C.P. 04510, Del. Coyoacán, Ciudad de México, Mexico. oscar.amelines@fisica.unam.mx.

Abstract:
The noncovalent dyad of tetraphenylporphine and C60 fullerene (H2TPP···C60) and the tetraphenylporphine dimer (H2TPP···H2TPP) were studied by density functional theory (DFT), using functionals that incorporate empirical dispersion correction (DFT-D), functionals that use a long-range correction (LC) scheme, a hybrid functional (B3LYP) and a highly parametrized empirical exchange-correlation functional (M05-2X). The results were compared to X-ray structures and interaction energies reported in previous experimental and theoretical studies. It was found that B3LYP and CAM-B3LYP functionals fail to reproduce the X-ray structures and binding energies of the TPP···C60 system. DFT-D functionals overestimated the π···π energy interactions for both systems, however, the optimized structures agree well with those observed experimentally. The LC-BLYP functional predicts geometries similar to X-ray structures; nevertheless, due to the lack of correction in the dispersion energy, the predicted energies for both model systems are low. On the other hand, the M05-2X functional exhibited the best performance. Both the structures and binding energies calculated with M05-2X are consistent with experimental and theoretical evidence reported by other authors, as well as with our experimental results obtained by means of atomic force microscopy on H2TPP thin films grown on the HOPG/C60 substrate by physical vapor deposition.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 04:01
VSEPR Theory and the Effect of Lone Pairs
 02:42
Valence Bond Theory
 01:18
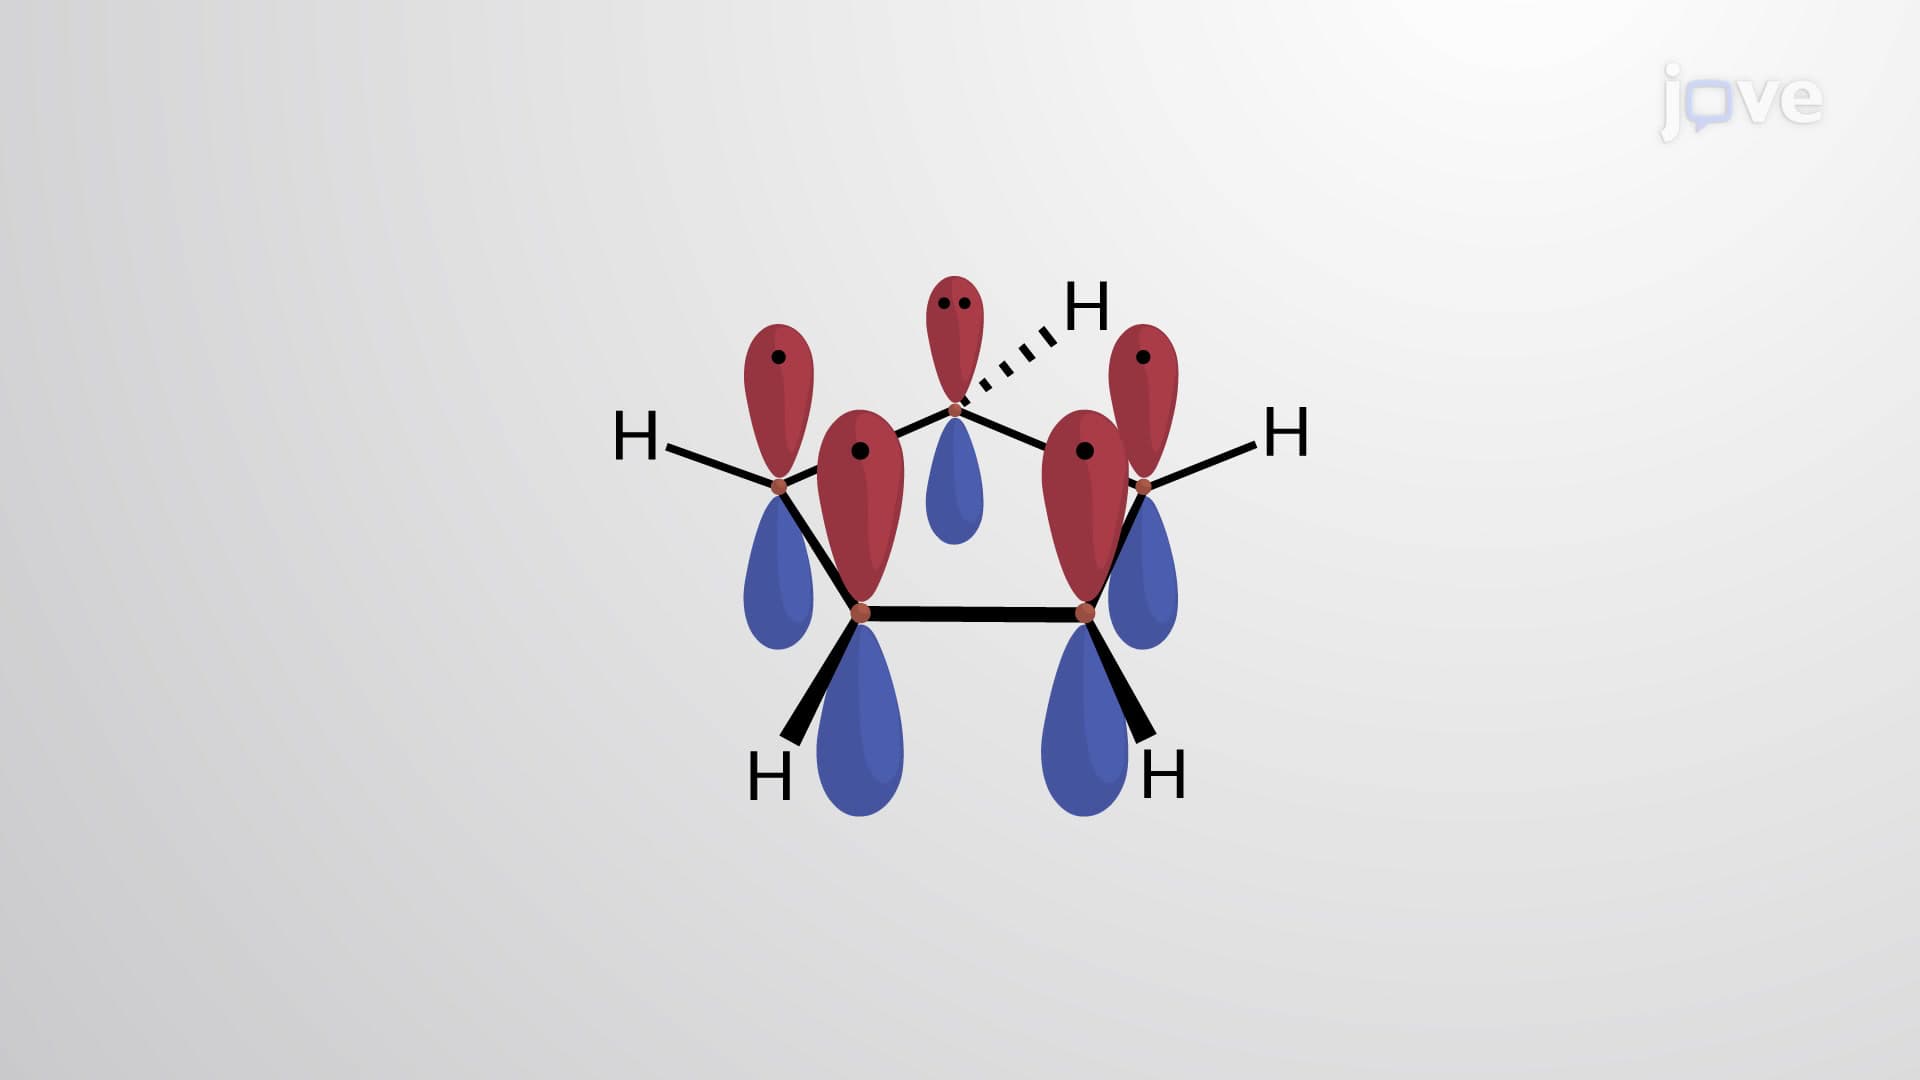
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...