Related Experiment Video
Updated: Mar 30, 2026

Hyperspectral Imaging as a Tool to Study Optical Anisotropy in Lanthanide-Based Molecular Single Crystals
Published on: April 14, 2020
Computational Investigation on the Spectroscopic Properties of Thiophene Based Europium β-Diketonate Complexes
Claudio Greco1, Giorgio Moro2, Luca Bertini2
1Dipartimento di Scienze dell'Ambiente e del Territorio e di Scienze della Terra, Università degli Studi di Milano-Bicocca ; Piazza della Scienza 1, 20126 Milano, Italy.

Abstract:
The adiabatic transition energies from the lowest triplet states of four Europium tris β-diketonate/phenantroline complexes have been determined in vacuo and in dicholomethane solution by the ΔSCF approach at the density functional theory level, using the PBE1PBE and the CAM-B3LYP hybrid functionals. The calculated adiabatic transition energies have been compared with the experimental 0-0 transitions of each complex determined from phosphorescence spectra of the corresponding Gd(3+) complexes and followed by direct comparison between simulated and experimental spectra line shapes. For compound 1, the Eu(TTA)3Phen system, triplet states other than the lowest one and conformational isomers other than the one present in the crystallographic structure have been considered. In the crystallographic structure, this compound presents three quasi-degenerate low energy triplet states, differing for the TTA ligand where the two unpaired electrons are localized and showing close adiabatic transition energies. For compound 1, the lowest triplet states of the four investigated conformational isomers show similar characteristics and close adiabatic transition energies. On the basis of these results, an investigation of compounds 2-4 (Eu(Br-TTA)3Phen, Eu(DTDK)3Phen, and Eu(MeT-TTA)3) has been performed by considering only the isomer present in the crystallographic structure and only the lowest triplet state of each compound. For compounds 1-3, the energies of the lowest triplet states calculated by both functionals in solution including zero-point energy corrections well reproduce the experimental trends as well as the values of the adiabatic transition energies: CAM-B3LYP, the best performing functional, provides energies of the lowest triplet state with deviations from experiments lower than 1200 cm(-1). Also, the calculated vibrationally resolved phosphorescence spectra and UV-vis absorptions well reproduce the main features of their experimental counterparts. Significant differences between calculated and experimental results are observed for compound 4, for which difficulties in the experimental determination of the triplet state energy were encountered: our results show that the negligible photoluminescence quantum yield of this compound is due to the fact that the energy of the most stable triplet state is significantly lower than that of the resonance level of the Europium ion, and thus the energy transfer process is prevented. These results confirm the reliability of the adopted computational approach in calculating the energy of the lowest triplet state energy of these systems, a key parameter in the design of new ligands for lanthanide complexes presenting large photoluminescence quantum yields.
More Related Videos
Related Concept Videos
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is the extent of conjugation in...
 01:29
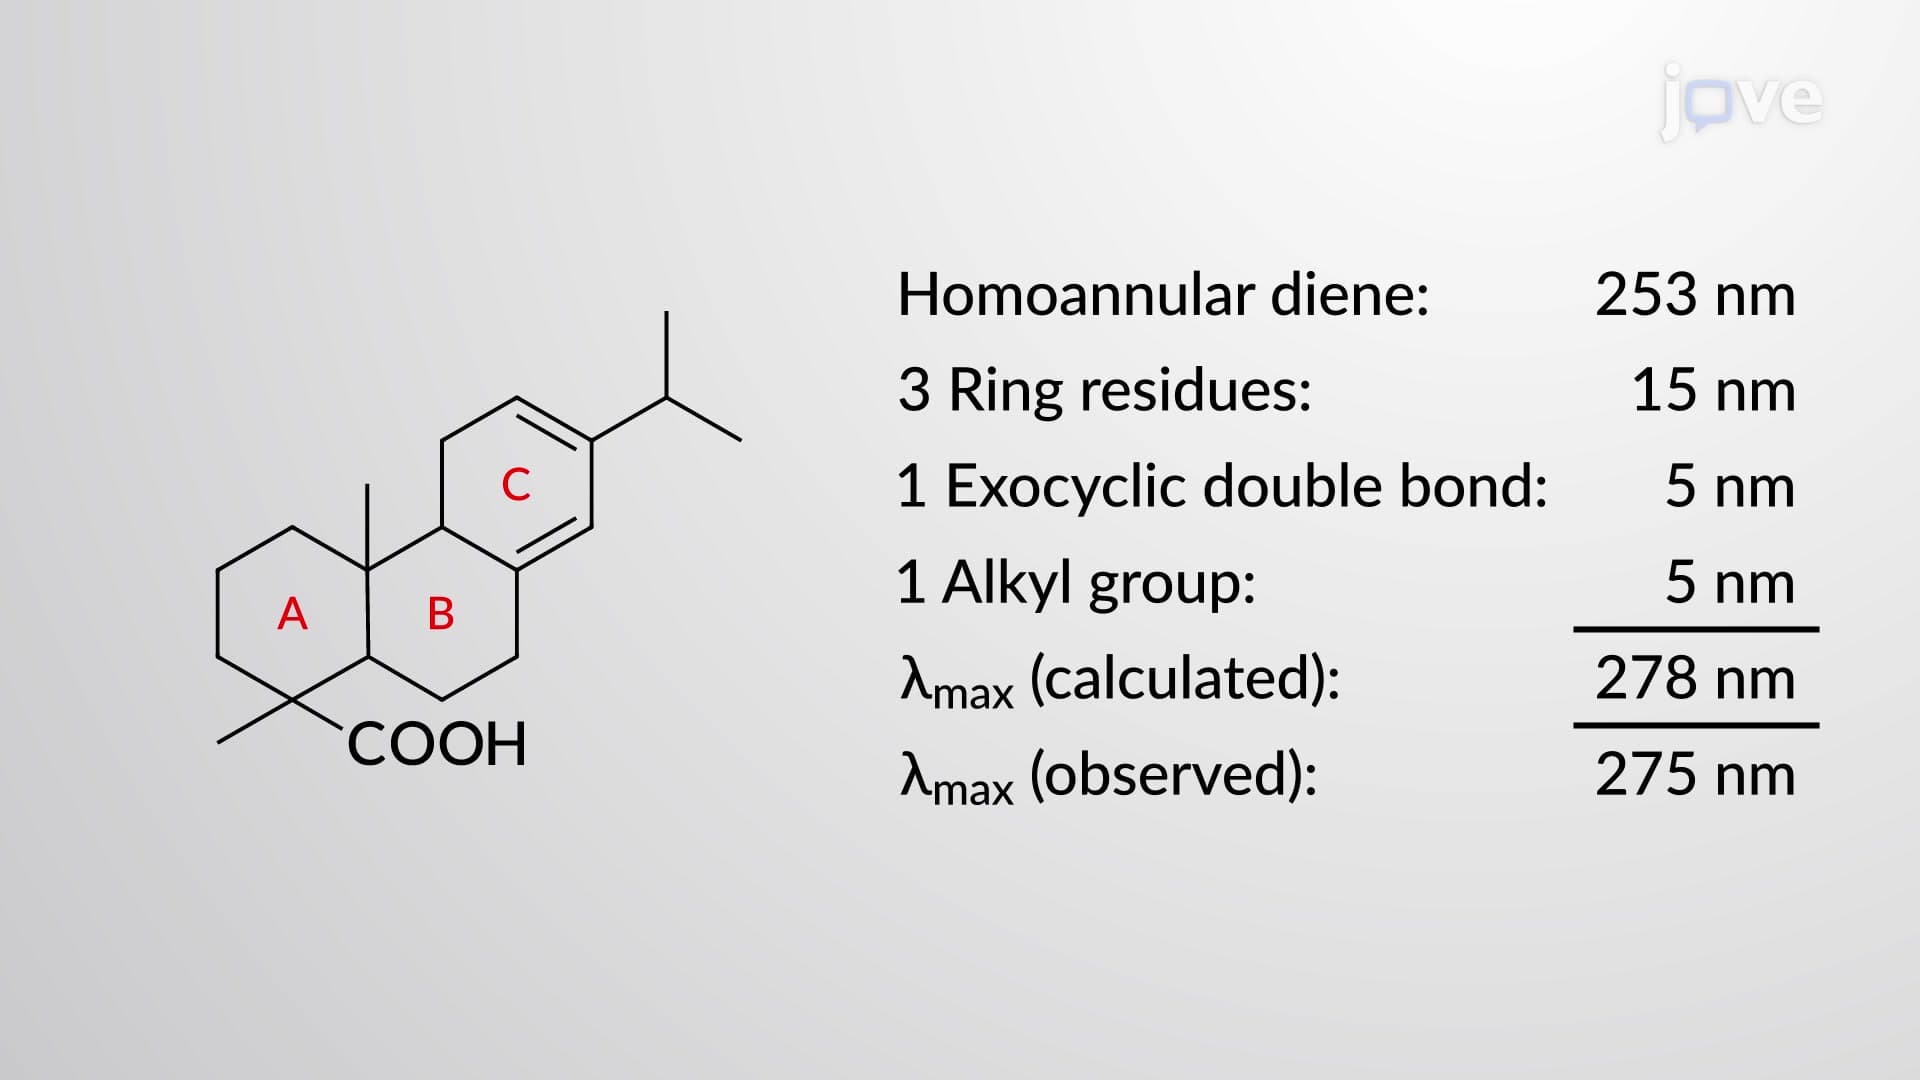
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:17
Electron Paramagnetic Resonance (EPR) Spectroscopy: Organic Radicals
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...