fireball/amber: An Efficient Local-Orbital DFT QM/MM Method for Biomolecular Systems
Related Experiment Video
Updated: Mar 30, 2026

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Jesús I Mendieta-Moreno1,2,3, Ross C Walker4, James P Lewis5
1Departamento de Física Teórica de la Materia Condensada and Condensed Matter Physics Center (IFIMAC), Universidad Autónoma de Madrid , ES-28049 Madrid, Spain.

Abstract:
In recent years, quantum mechanics/molecular mechanics (QM/MM) methods have become an important computational tool for the study of chemical reactions and other processes in biomolecular systems. In the QM/MM technique, the active region is described by means of QM calculations, while the remainder of the system is described using a MM approach. Because of the complexity of biomolecules and the desire to achieve converged sampling, it is important that the QM method presents a good balance between accuracy and computational efficiency. Here, we report on the implementation of a QM/MM technique that combines a DFT approach specially designed for the study of complex systems using first-principles molecular dynamics simulations (fireball) with the amber force fields and simulation programs. We also present examples of the application of this QM/MM approach to three representative biomolecular systems: the analysis of the effect of electrostatic embedding in the behavior of a salt bridge between an aspartic acid and a lysine residue, a study of the intermediate states for the triosephosphate isomerase catalyzed conversion of dihydroxyacetone phosphate into glyceraldehyde 3-phosphate, and the detailed description, using DFT QM/MM molecular dynamics, of the cleavage of a phosphodiester bond in RNA catalyzed by the enzyme RNase A.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I
 02:40
MO Theory and Covalent Bonding
 02:45
The Quantum-Mechanical Model of an Atom
 01:18
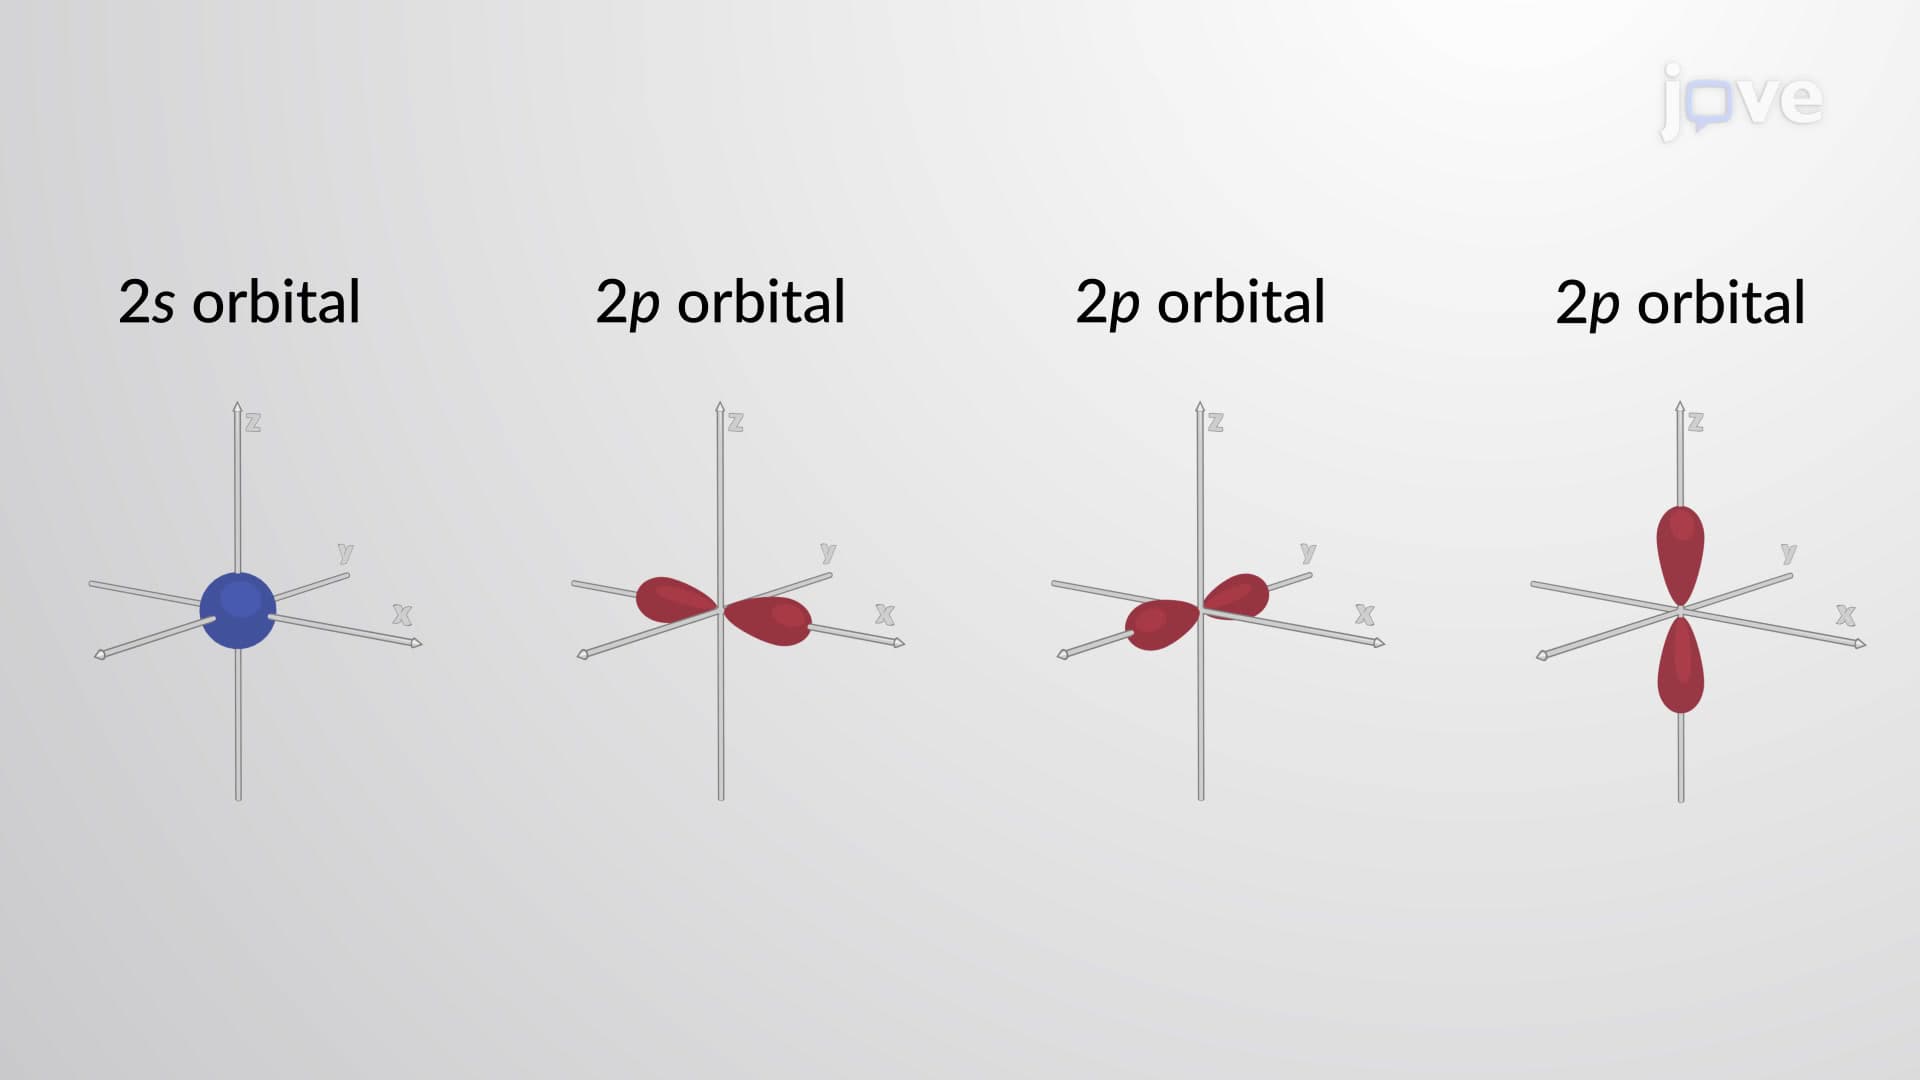
Electron Orbital Model
The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...
 02:44
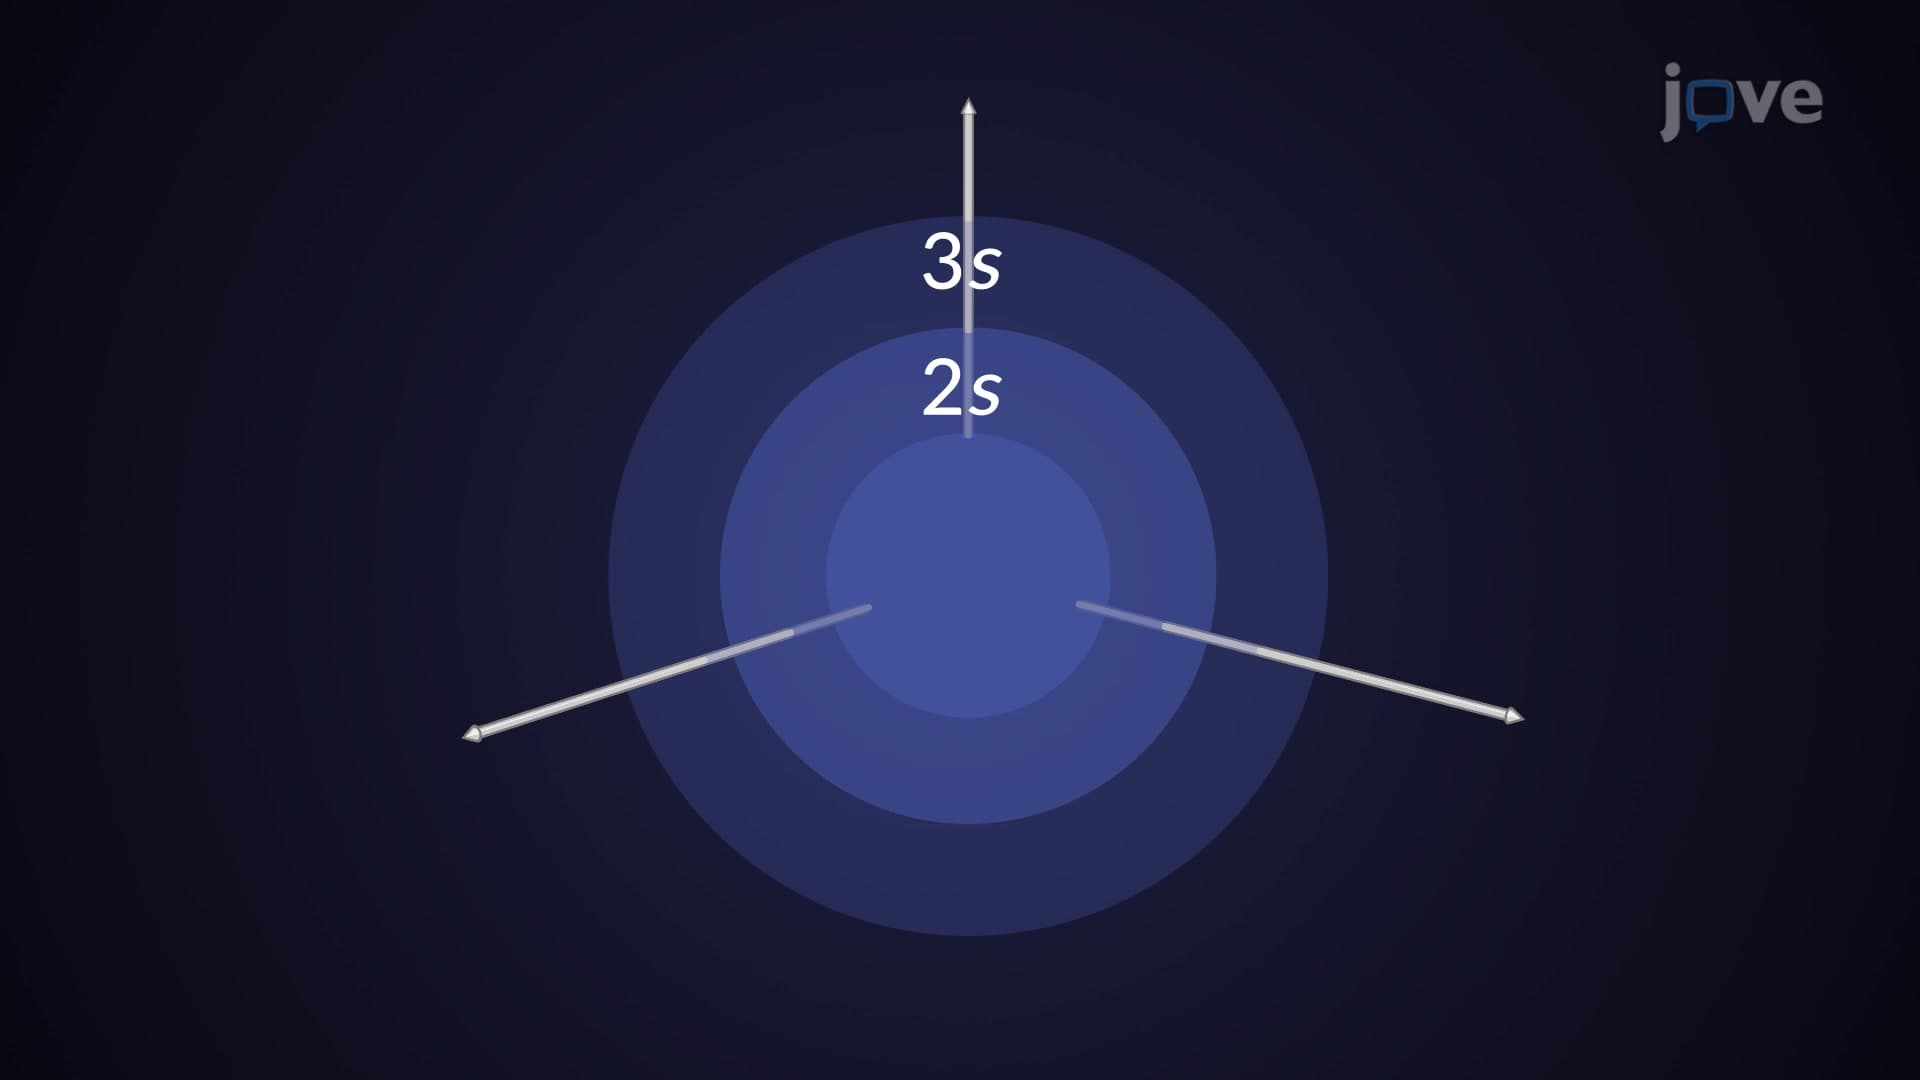
Atomic Orbitals