Related Experiment Video
Updated: Mar 30, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Accuracy and Efficiency of Coupled-Cluster Theory Using Density Fitting/Cholesky Decomposition, Frozen Natural
A Eugene DePrince1, C David Sherrill1
1Center for Computational Molecular Science and Technology, School of Chemistry and Biochemistry, and School of Computational Science and Engineering, Georgia Institute of Technology, Atlanta, Georgia 30332-0400, United States.

Abstract:
We present an algorithm for coupled-cluster through perturbative triples [CCSD(T)] based on a t1-dressed Hamiltonian and the use of density fitting (DF) or Cholesky decomposition (CD) approximations for the construction and contraction of all electron repulsion integrals (ERIs). An efficient implementation of this algorithm is then used to explore whether preoptimized density fitting basis sets [specifically, the (aug-)cc-pVXZ-RI series designed for DF-MP2 computations] are suitable for DF-CCSD(T) computations and how they compare to the CD representation of the integrals. The code is also used to systematically explore the accuracy and efficiency of DF/CD combined with frozen natural orbitals (FNOs) to reduce computational costs. The mean absolute errors due to DF/CD in the CCSD(T)/aug-cc-pVDZ interaction energies of 11 van der Waals dimers are only 0.001 kcal mol(-1) for the preoptimized RI basis set and only 0.002 and 0.001 kcal mol(-1) for CD with cutoffs of 10(-4) and 10(-5), respectively. The very similar performance of the aug-cc-pVDZ-RI auxiliary set is a bit surprising considering that the numbers of CD vectors using these thresholds are, on average, 28% and 73% larger than the dimension of the RI set. When FNOs are coupled with DF/CD, the DF/CD error is roughly an order of magnitude less than the FNO truncation error (at a conservative FNO occupation cutoff of 10(-5)). Utilizing t1-dressed three-index integrals, which remove the explicit dependence of the doubles residual equations on the t1-amplitudes, results in a moderate performance acceleration for the CCSD portion of the algorithm. Moreover, the t1-dressing results in a simpler code which will be more amenable to parallelization. Utilizing both CD and FNO techniques, we observe a speedup of four times for the evaluation of the three-body contribution to the interaction energy for the benzene trimer described by an aug-cc-pVDZ basis set; the error incurred by the CD and FNO approximations in the three-body contribution is only 0.002 kcal mol(-1).
More Related Videos

Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 02:38
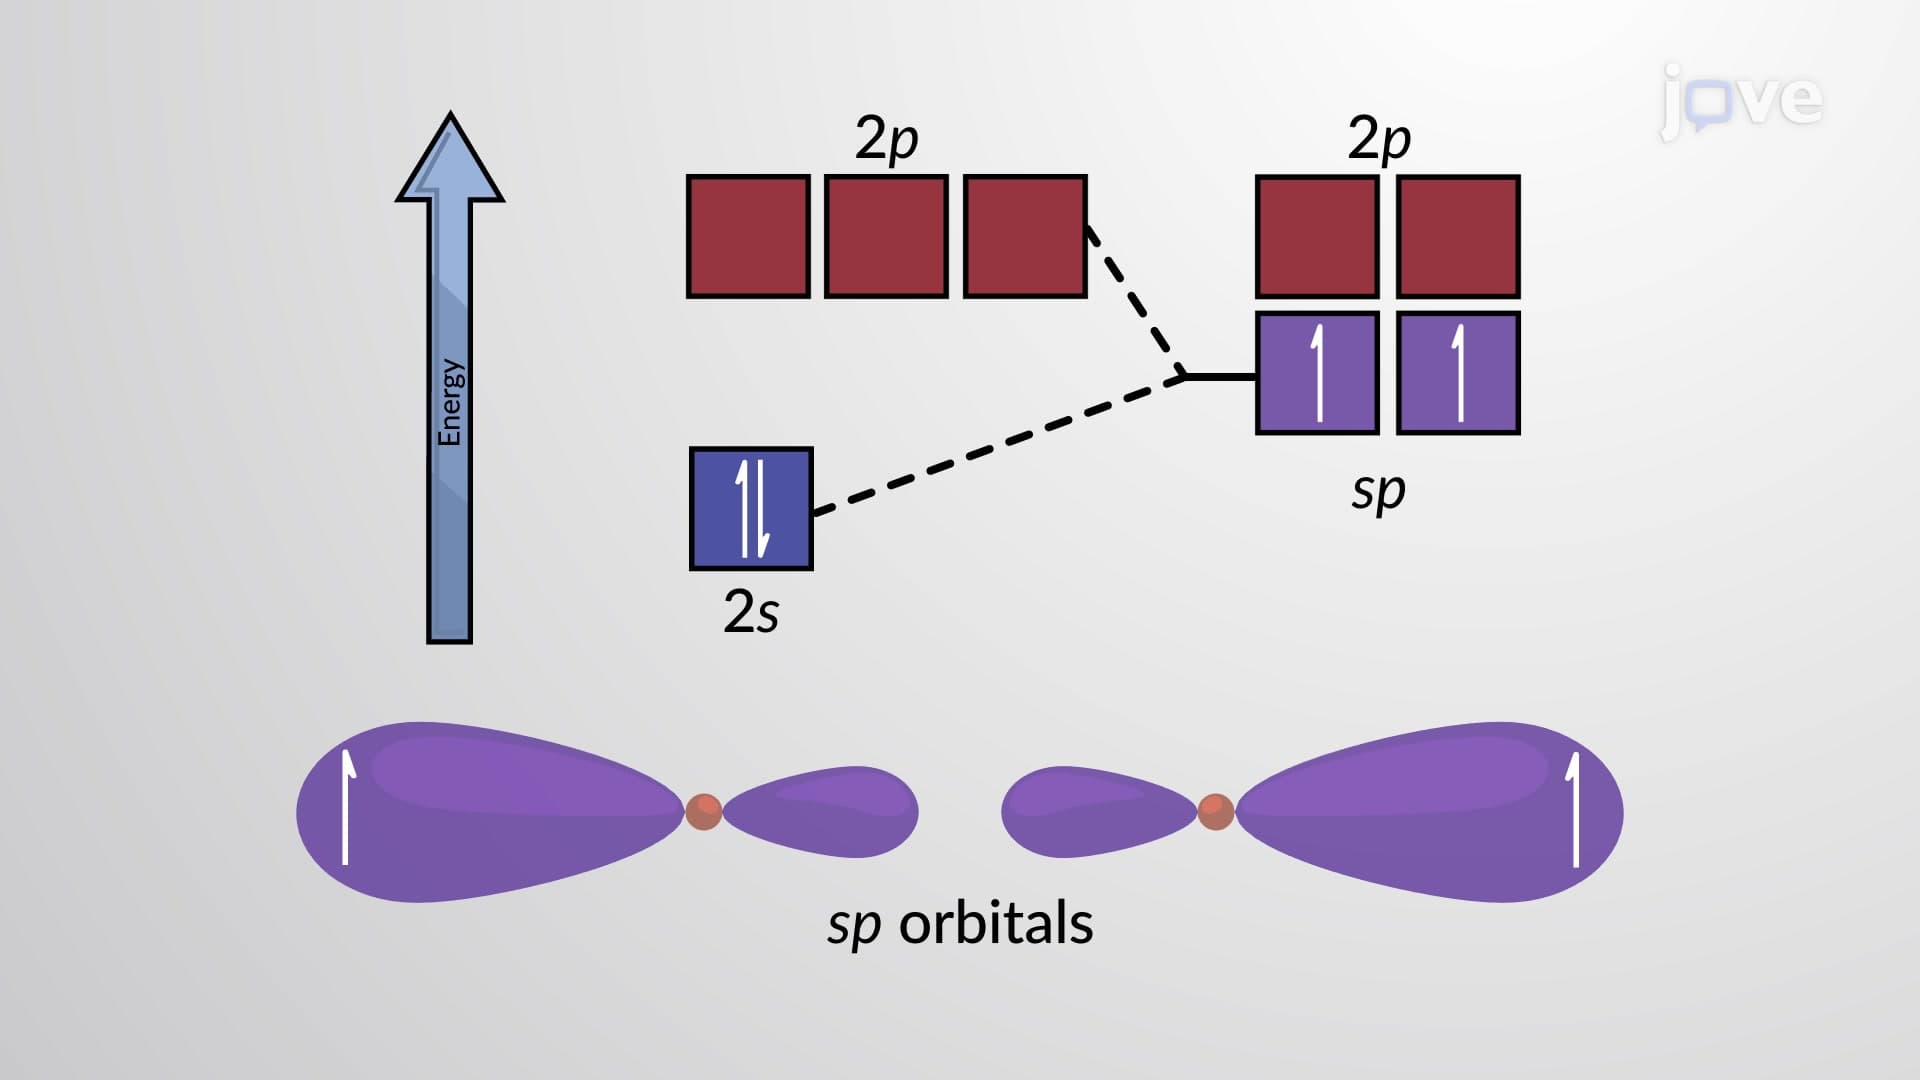
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...