Related Experiment Video
Updated: Mar 30, 2026

Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Establishing the Accuracy of Broadly Used Density Functionals in Describing Bulk Properties of Transition Metals
Patanachai Janthon1,2,3,4, Sergey M Kozlov1, Francesc Viñes1
1Departament de Química Física and Institut de Química Teòrica i Computacional (IQTCUB), Universitat de Barcelona, c/Martí i Franquès 1, 08028, Barcelona, Spain.

Abstract:
The performance of various commonly used density functionals is established by comparing calculated values of atomic structure data, cohesive energies, and bulk moduli of all transition metals to available experimental data. The functionals explored are the Ceperley-Alder (CA), Vosko-Wilk-Nussair (VWN) implementation of the Local Density Approximation (LDA); the Perdew-Wang (PW91) and Perdew-Burke-Ernzerhof (PBE) forms of the Generalized Gradient Approximation (GGA), and the RPBE and PBEsol modifications of PBE, aimed at better describing adsorption energies and bulk solid lattice properties, respectively. The present systematic study shows that PW91 and PBE consistently provide the smallest differences between the calculated and experimental values. Additional calculations of the (111) surface energy of several face centered cubic (fcc) transition metals reveal that LDA produces the most accurate results, while all other functionals significantly underestimate the experimental values. RPBE severely underestimates surface energy, which may be the origin for the reduced surface chemical activity and the better performance of RPBE describing adsorption energies.
More Related Videos
 07:24
Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
 08:55
Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
Related Concept Videos
 02:58
Properties of Transition Metals
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 04:00
Periodic Classification of the Elements
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:17
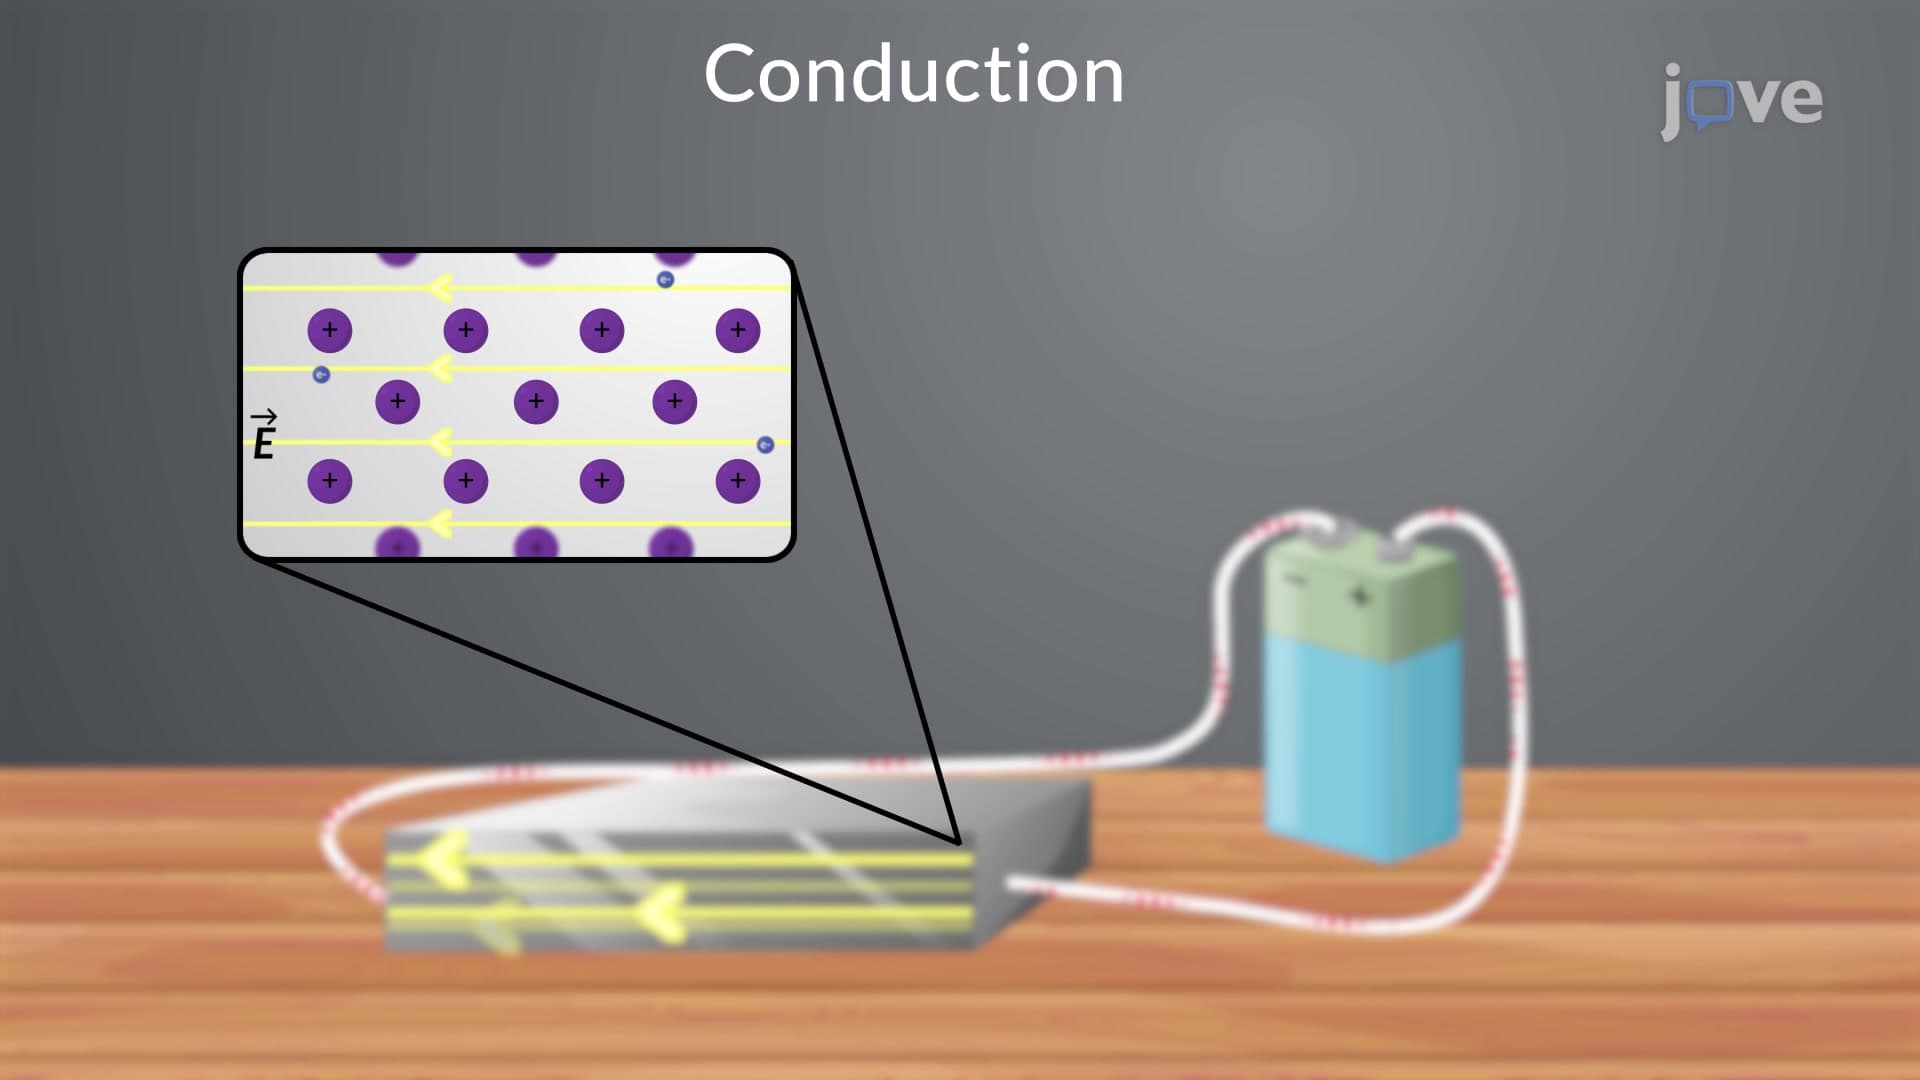
Theory of Metallic Conduction
In this theory, Newton's second law of motion is used to determine the acceleration of an electron in the presence of an applied electric field. Then, its velocity is expressed via this acceleration.
An electron moves through the crystal, containing positive ions,...