Related Experiment Video
Updated: Mar 29, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
A Parametrized Coupled-Pair Functional for Molecular Interactions: PCPF-MI
Daniel R Nascimento1, A Eugene DePrince1
1Department of Chemistry and Biochemistry, Florida State University , Tallahassee, Florida 32306-4390, United States.

Abstract:
We present a parametrized coupled-pair functional, optimized to describe molecular interactions (PCPF-MI). The method has the same computational cost as singles and doubles configuration interaction, is size extensive, and yields energies that are stationary with respect to variations in all of its excitation coefficients. This last property facilitates the construction of density matrices and the evaluation of one- and two-electron properties. For the S22, HSG, S66, and A24 databases of van der Waals dimers, PCPF-MI computations yield interaction energies with mean unsigned errors of 0.326, 0.149, 0.214, and 0.044 kcal mol(-1), respectively, relative to benchmark computations. PCPF-MI interaction energies consistently improve upon those obtained from several other coupled-pair methods, including the averaged coupled-pair functional (ACPF), the coupled electron-pair approximation [CEPA(n), n = 0,1,3], and averaged quadratic coupled cluster (AQCC). Optimal parameters for spin-component-scaled (SCS) variants of each of these methods are also presented. SCS-CEPA(0), SCS-CEPA(1), SCS-ACPF, and SCS-PCPF-MI all perform similarly, with average errors for the four databases of roughly 0.1 kcal mol(-1). The PCPF-MI method, without additional SCS parametrization, has an average error of 0.192 kcal mol(-1) over the four databases and, when compared to the SCS-parametrized coupled-pair methods, has the added benefit that the energy is stationary with respect to variations in all its excitation coefficients.
More Related Videos
 05:57
Author Spotlight: In Silico Creation and Impact of Carbonylated Amino Acids on Protein Structure and Function
Published on: April 26, 2024
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...
 02:00
Molecular Models
 01:17
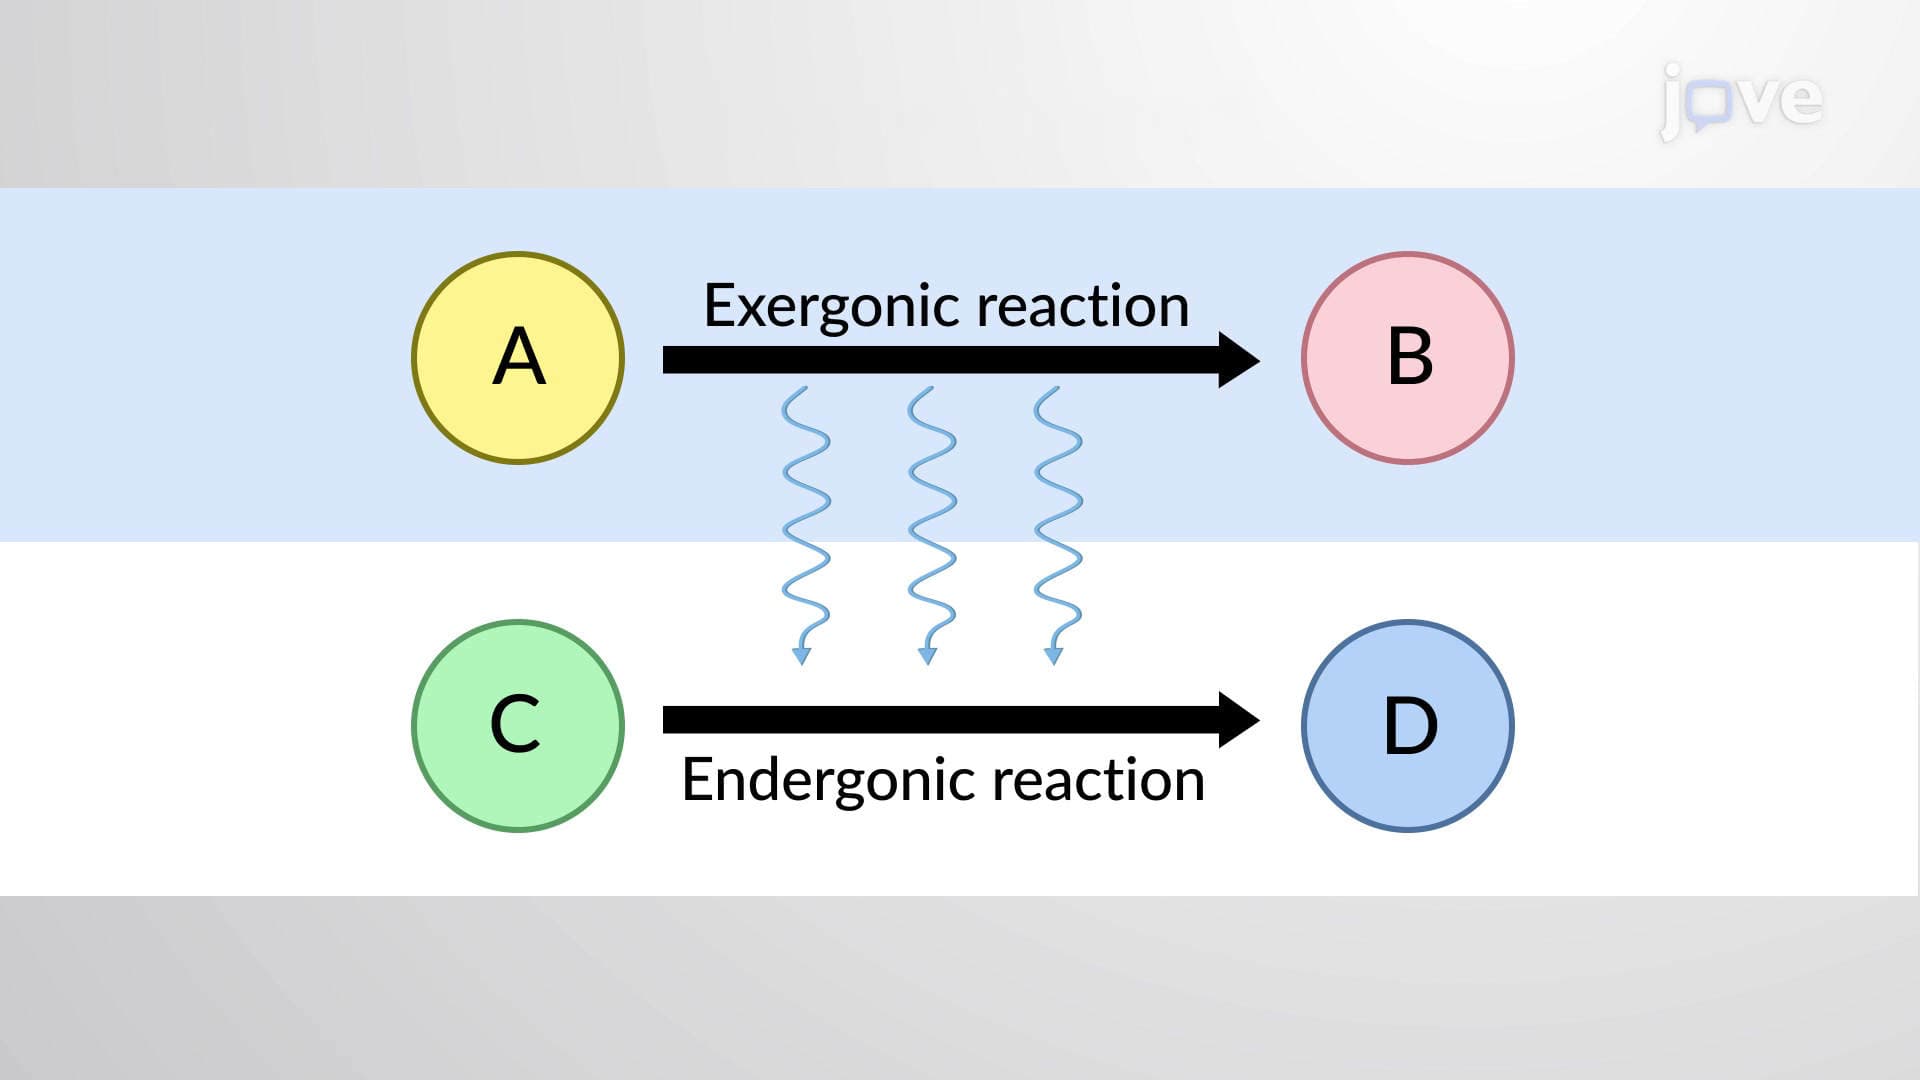
Coupled Reactions
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions....
 02:27
Predicting Molecular Geometry
 02:36
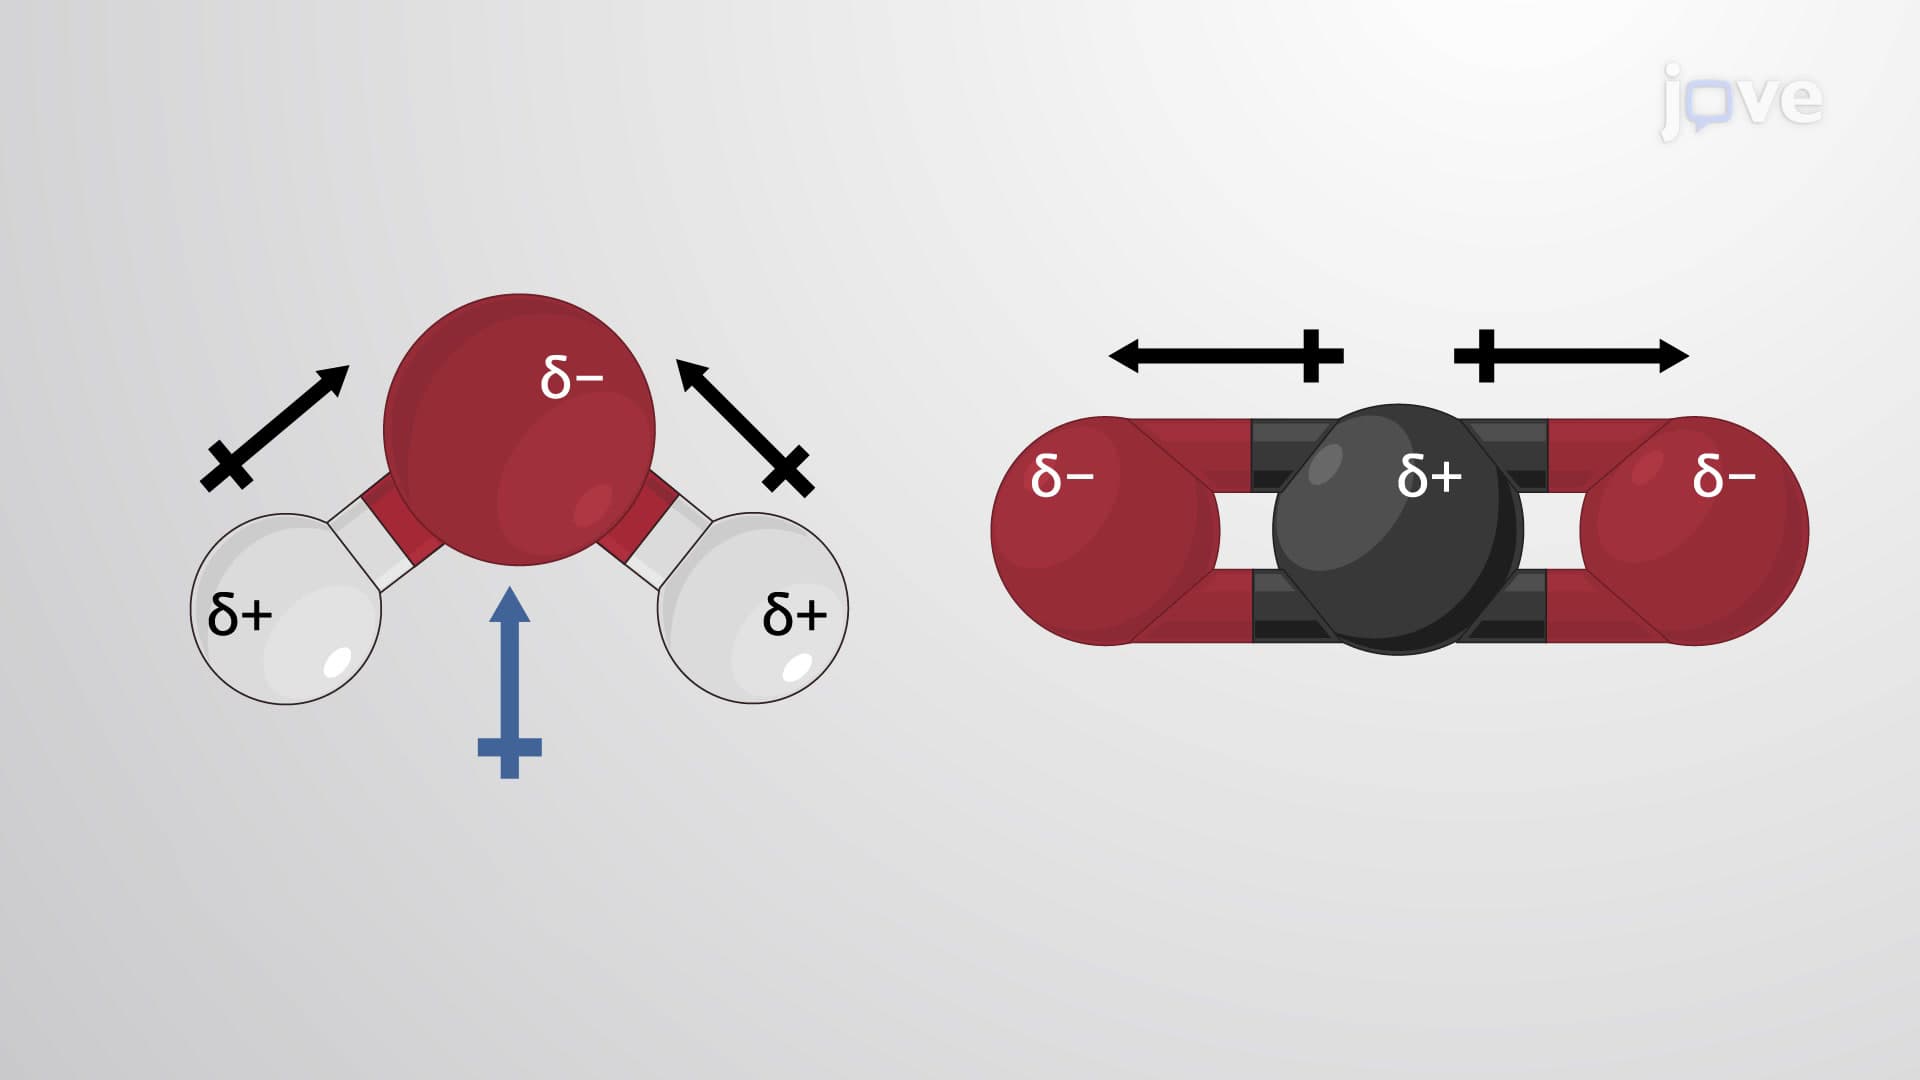
Molecular Geometry and Dipole Moments
 01:24
Van der Waals Interactions