Related Experiment Video
Updated: Mar 29, 2026


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Reaction Path Force Matching: A New Strategy of Fitting Specific Reaction Parameters for Semiempirical Methods in
1Department of Chemistry and Chemical Biology, Indiana University-Purdue University Indianapolis , 402 N. Blackford St., LD326, Indianapolis, Indiana 46202, United States.

Abstract:
We present a general strategy of reparametrizing semiempirical (SE) methods against ab initio (AI) methods for combined quantum mechanical and molecular mechanical (QM/MM) simulations of specific chemical reactions in condensed phases. The resulting approach, designated Reaction Path Force Matching (RP-FM), features cycles of sampling configurations along a reaction path on an efficient SE/MM potential energy surface (PES) and adjusting specific reaction parameters (SRPs) in the SE method such that the atomic forces computed at the target AI/MM level are reproduced. Iterative applications of the RP-FM cycle make possible achieving the accuracy of AI/MM simulations without explicitly sampling the computationally expensive AI/MM PES. The bypassed sampling, nevertheless, is implicitly accomplished through the aid of the efficient SE-SRP/MM PES, on which the target-level reaction path is expected to be obtained upon convergence. We demonstrate the effectiveness of the RP-FM procedure for a symmetric proton transfer reaction in the gas phase and in solution. The remarkable agreements between the RP-FM optimized SE-SRP methods and the target AI method on various properties, including energy profiles, potential of mean force free energy profiles, atomic forces, charge populations, and solvation effects, suggest that RP-FM can be used as an efficient and reliable strategy for simulating condensed-phase chemical reactions.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
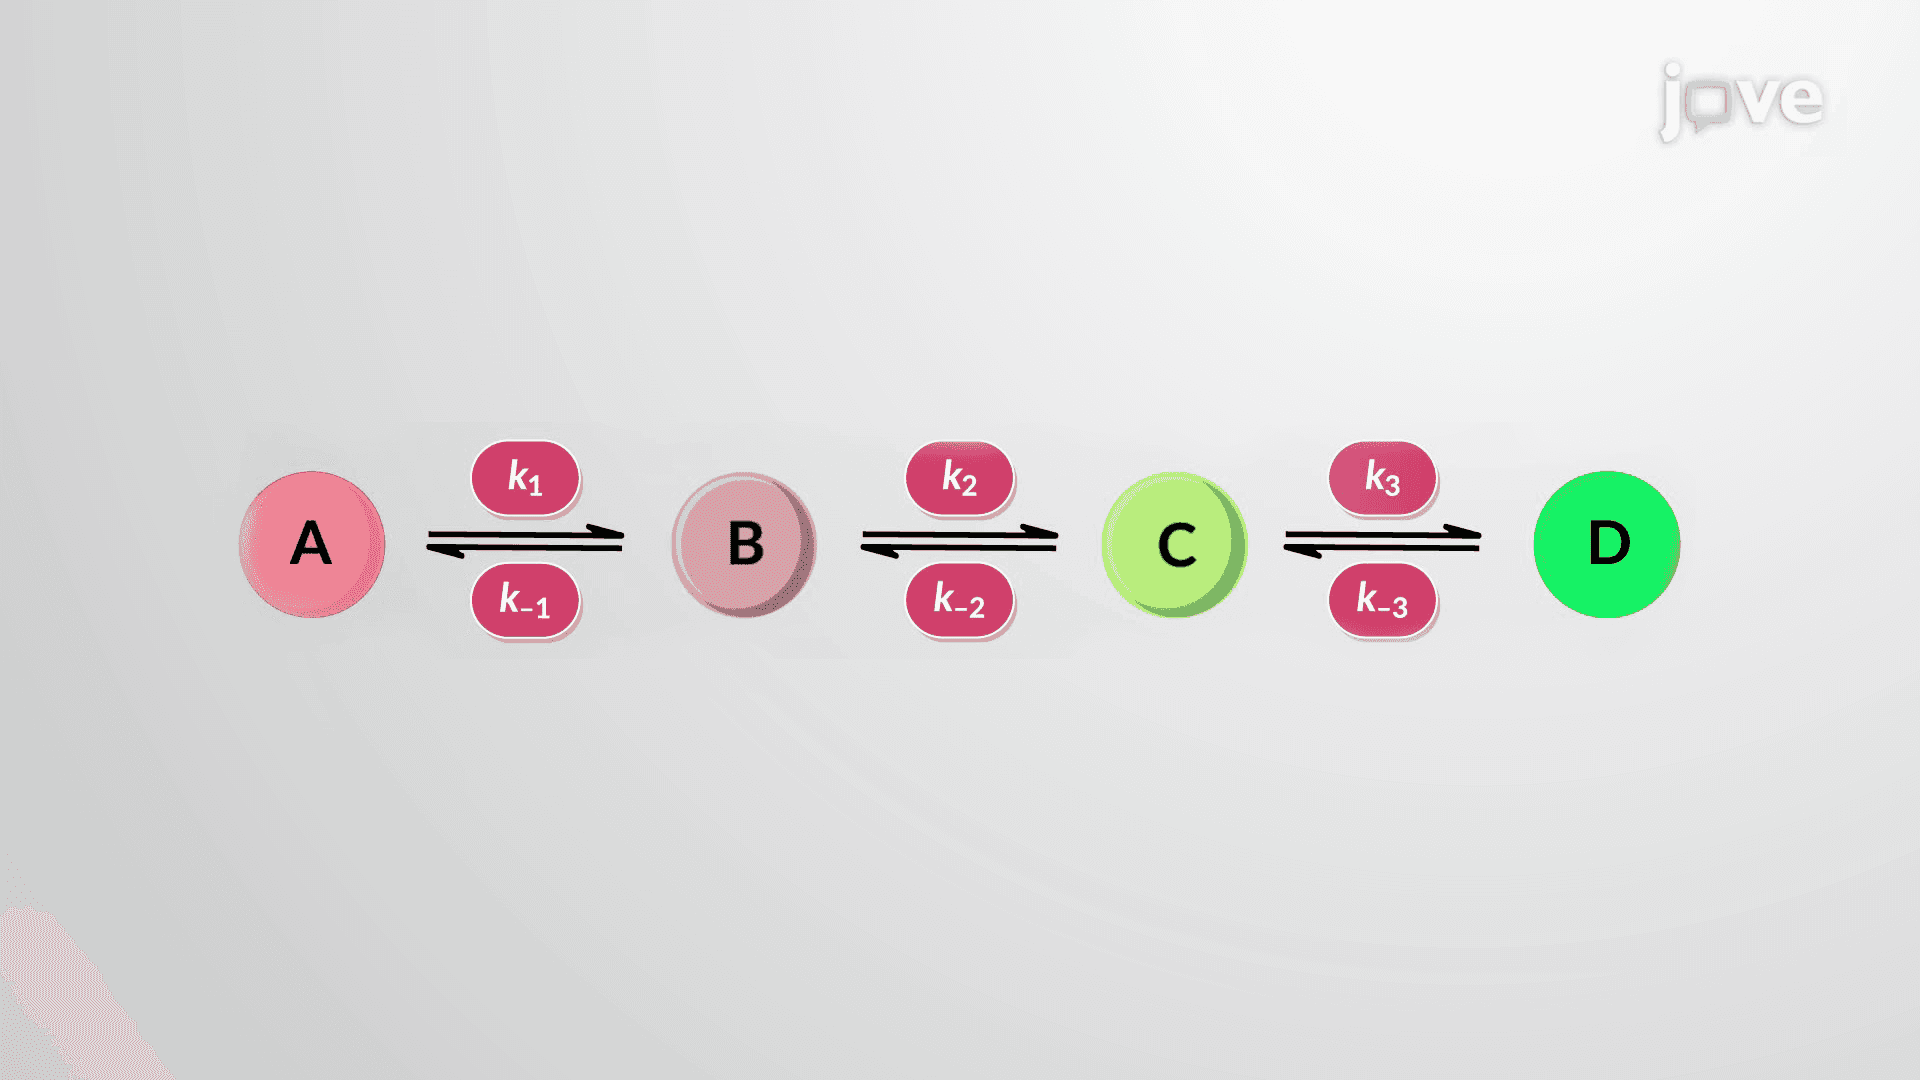 01:29
Reaction Mechanisms: Rate-limiting Step Approximation
 03:06
Reaction Mechanisms
For instance, the decomposition of ozone appears to follow a mechanism with two steps:
 02:24
Predicting Reaction Outcomes
 01:17
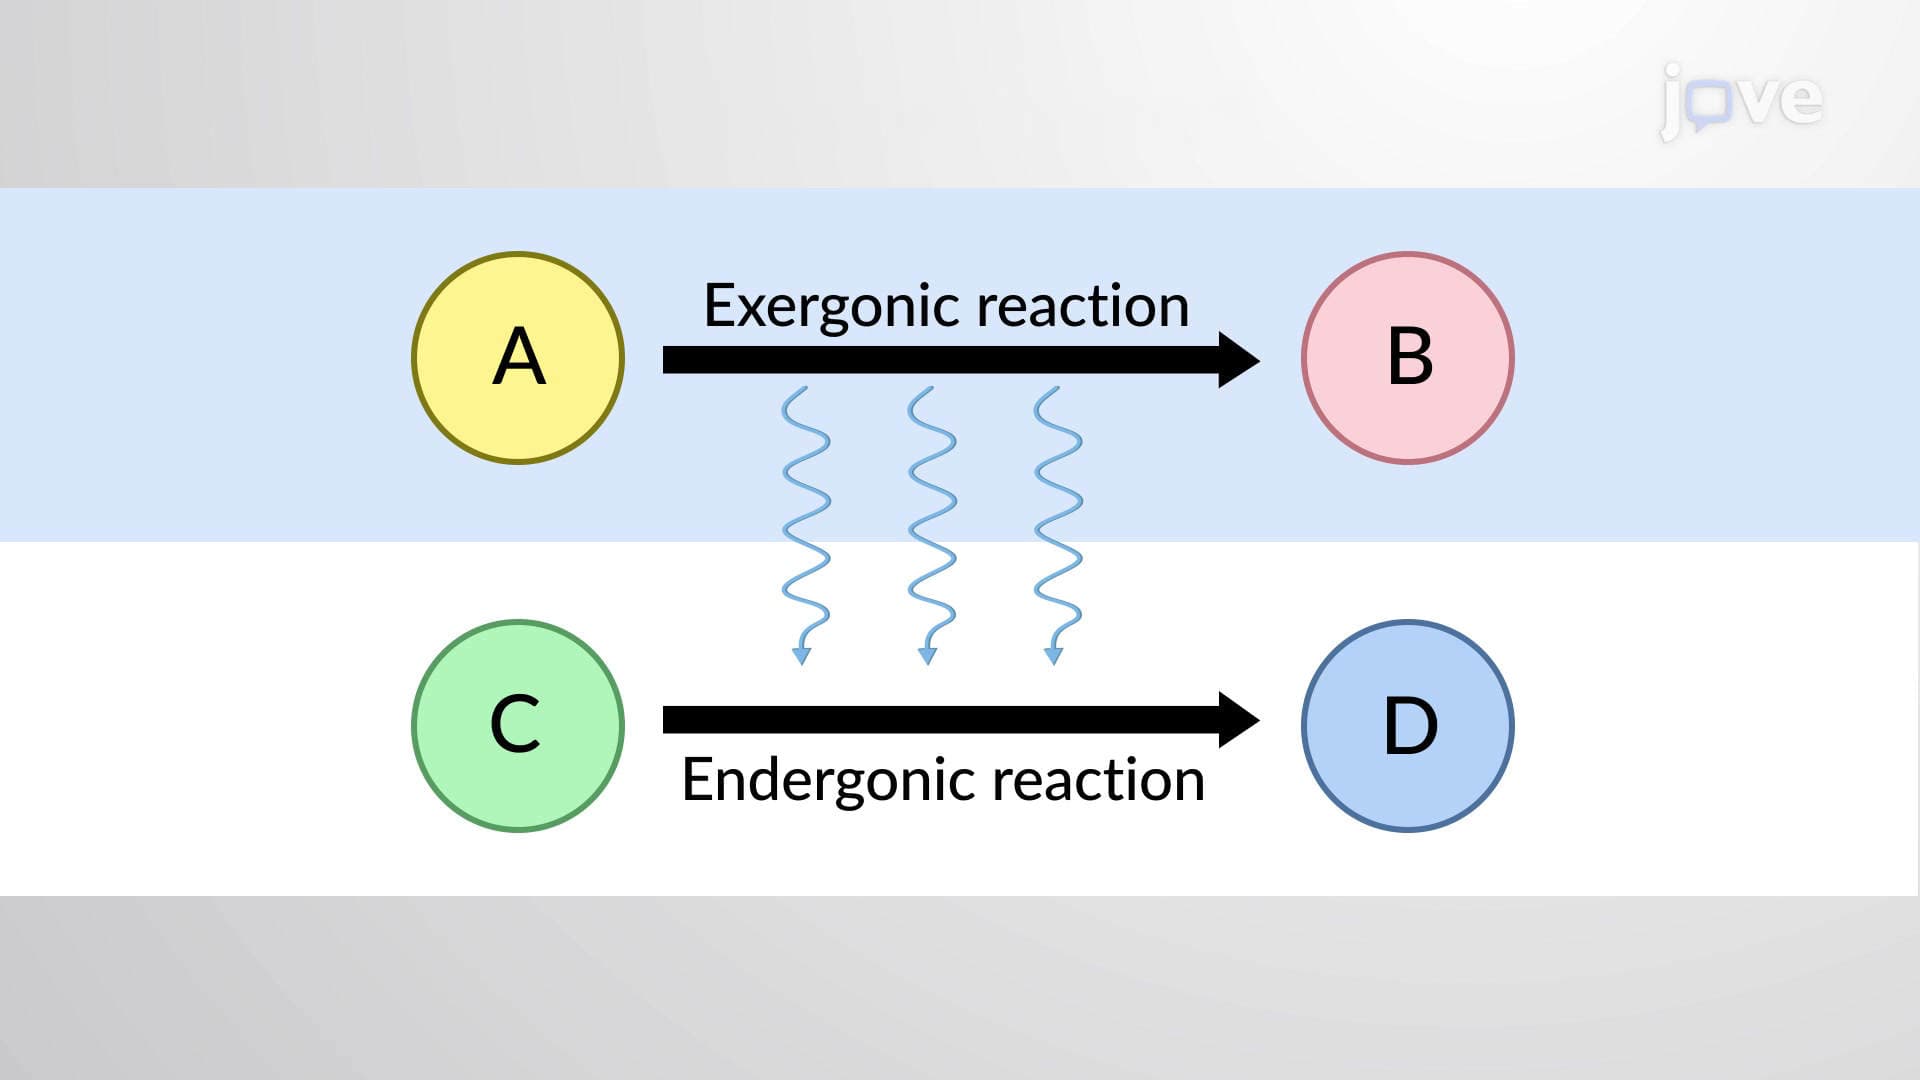
Coupled Reactions
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions....
 02:55
Temperature Dependence on Reaction Rate
Atoms, molecules, or ions must collide before they can react with each other. Atoms must be close together to form chemical bonds. This premise is the basis for a theory that explains many observations regarding chemical kinetics, including factors affecting reaction rates.
The collision theory is based on the postulates that (i) the reaction rate is proportional to the rate of reactant collisions, (ii) the reacting species collide in an orientation allowing contact between...
 01:11
Radical Reactivity: Overview