Related Experiment Video
Updated: Mar 29, 2026
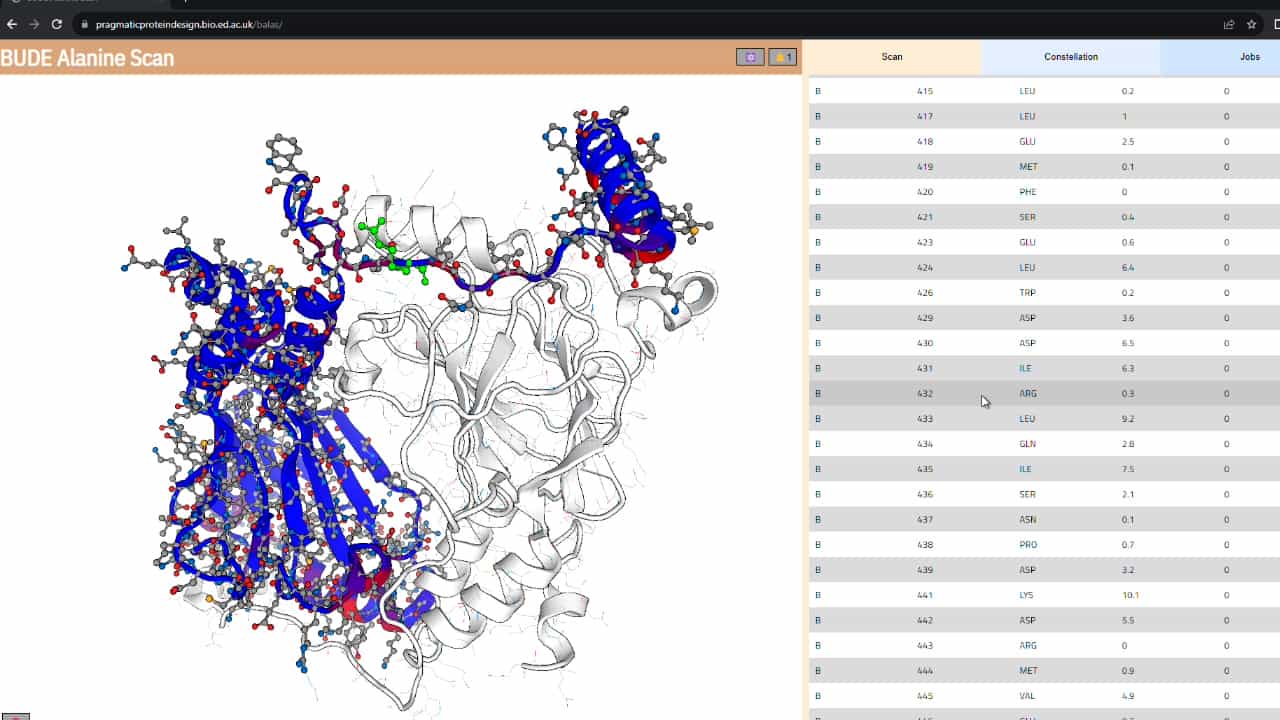
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Representative Amino Acid Side Chain Interactions in Proteins. A Comparison of Highly Accurate Correlated ab Initio
Karel Berka1, Roman Laskowski1, Kevin E Riley1
1Institute of Organic Chemistry and Biochemistry, Academy of Sciences of the Czech Republic and Center for Complex Molecular Systems and Biomolecules, Flemingovo náměstí 2, Prague 6, 166 10 Czech Republic, Department of Physical and Macromolecular Chemistry, Faculty of Natural Sciences, Charles University in Prague, Hlavova 8, Prague 2, 128 43 Czech Republic, EMBL Outstation - Hinxton, European Bioinformatics Institute, Wellcome Trust Genome Campus, Hinxton, Cambridge CB10 1SD, U.K., and Department of Chemistry, P.O. Box 23346, University of Puerto Rico, Rio Piedras, Puerto Rico 00931.

Abstract:
Interactions between amino acid side chains play a crucial role both within a folded protein and between the interacting protein molecules. Here we have selected a representative set of 24 of the 400 (20 × 20) possible interacting side chain pairs based on data from Atlas of Protein Side-Chain Interactions. For each pair, we obtained its most favorable interaction geometry from the structural data and computed the interaction energy in the gas phase using several different, commonly used, ab initio and force field methods, namely Møller-Plesset perturbation theory (MP2), density functional theory combined with symmetry-adapted perturbation theory (DFT-SAPT), density functional theory empirically augmented with an empirical dispersion term (DFT-D), and empirical potentials using the OPLS-AA/L and Amber03 force fields. All the methods were compared against a reference method taken to be the CCSD(T) level of theory extrapolated to the complete basis set limit. We found a high degree of agreement between the different methods, even though the range of binding energies obtained was extremely large. The most computationally intensive methods yielded the best results. Among the less computationally time-consuming methods, the DFT-D method as well as parm03 force field provided consistently good results when compared to the reference values. We also tested how representative the chosen geometries of the side chains were and investigated the effect on the binding energies of the dielectric constant of the surrounding medium.
More Related Videos
Related Concept Videos
 03:42
Amino acids
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:22
Protein Folding
 01:55
What are Proteins?
 01:28
What are Proteins?
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...