Related Experiment Video
Updated: Mar 29, 2026


Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Monoligand Zn(II) Complexes: Ab Initio Benchmark Calculations and Comparison with Density Functional Theory
Víctor M Rayón1, Haydee Valdés1, Natalia Díaz1
1Departamento de Química Física y Química Inorgánica, Facultad de Ciencias, Universidad de Valladolid, 47005 Valladolid, Spain, Center for Biomolecules and Complex Molecular Systems, Institute of Organic Chemistry and Biochemistry, Academy of Sciences of the Czech Republic, 16610 Prague 6, Czech Republic, and Departamento de Química Física y Analítica, Facultad de Química, Universidad de Oviedo, 33007 Oviedo, Spain.

Abstract:
A systematic theoretical study on several models of Zn(II) complexes has been carried out employing both ab initio correlated wave function and density functional methods. The performance of five different functionals namely PW91, PBE, B3LYP, MPWLYP1M, and TPSS in the prediction of metal-ligand bond distances, binding energies, and proton affinities has been assessed comparing the results to those obtained with the MP2 and CCSD(T) wave function methodologies. Several basis sets ranging from double-ζ up to quintuple-ζ quality have been used, including the recently developed all-electron correlation consistent basis sets for zinc. It is shown that all the tested functionals overestimate both the metal-ligand bond distances and the binding energies, being that the B3LYP and TPSS functionals are the ones that perform the best. An analysis of the metal-ligand interaction energy shows that induction and charge-transfer effects play a prominent role in the bonding of these systems, even for those complexes with the less polarizable ligands. This finding highlights the importance of a correct description of the polarization of the monomers' charge densities by any theoretical method which aims to be applied to the study of Zn(II) complexes.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:07
Ladder Diagrams: Complexation Equilibria
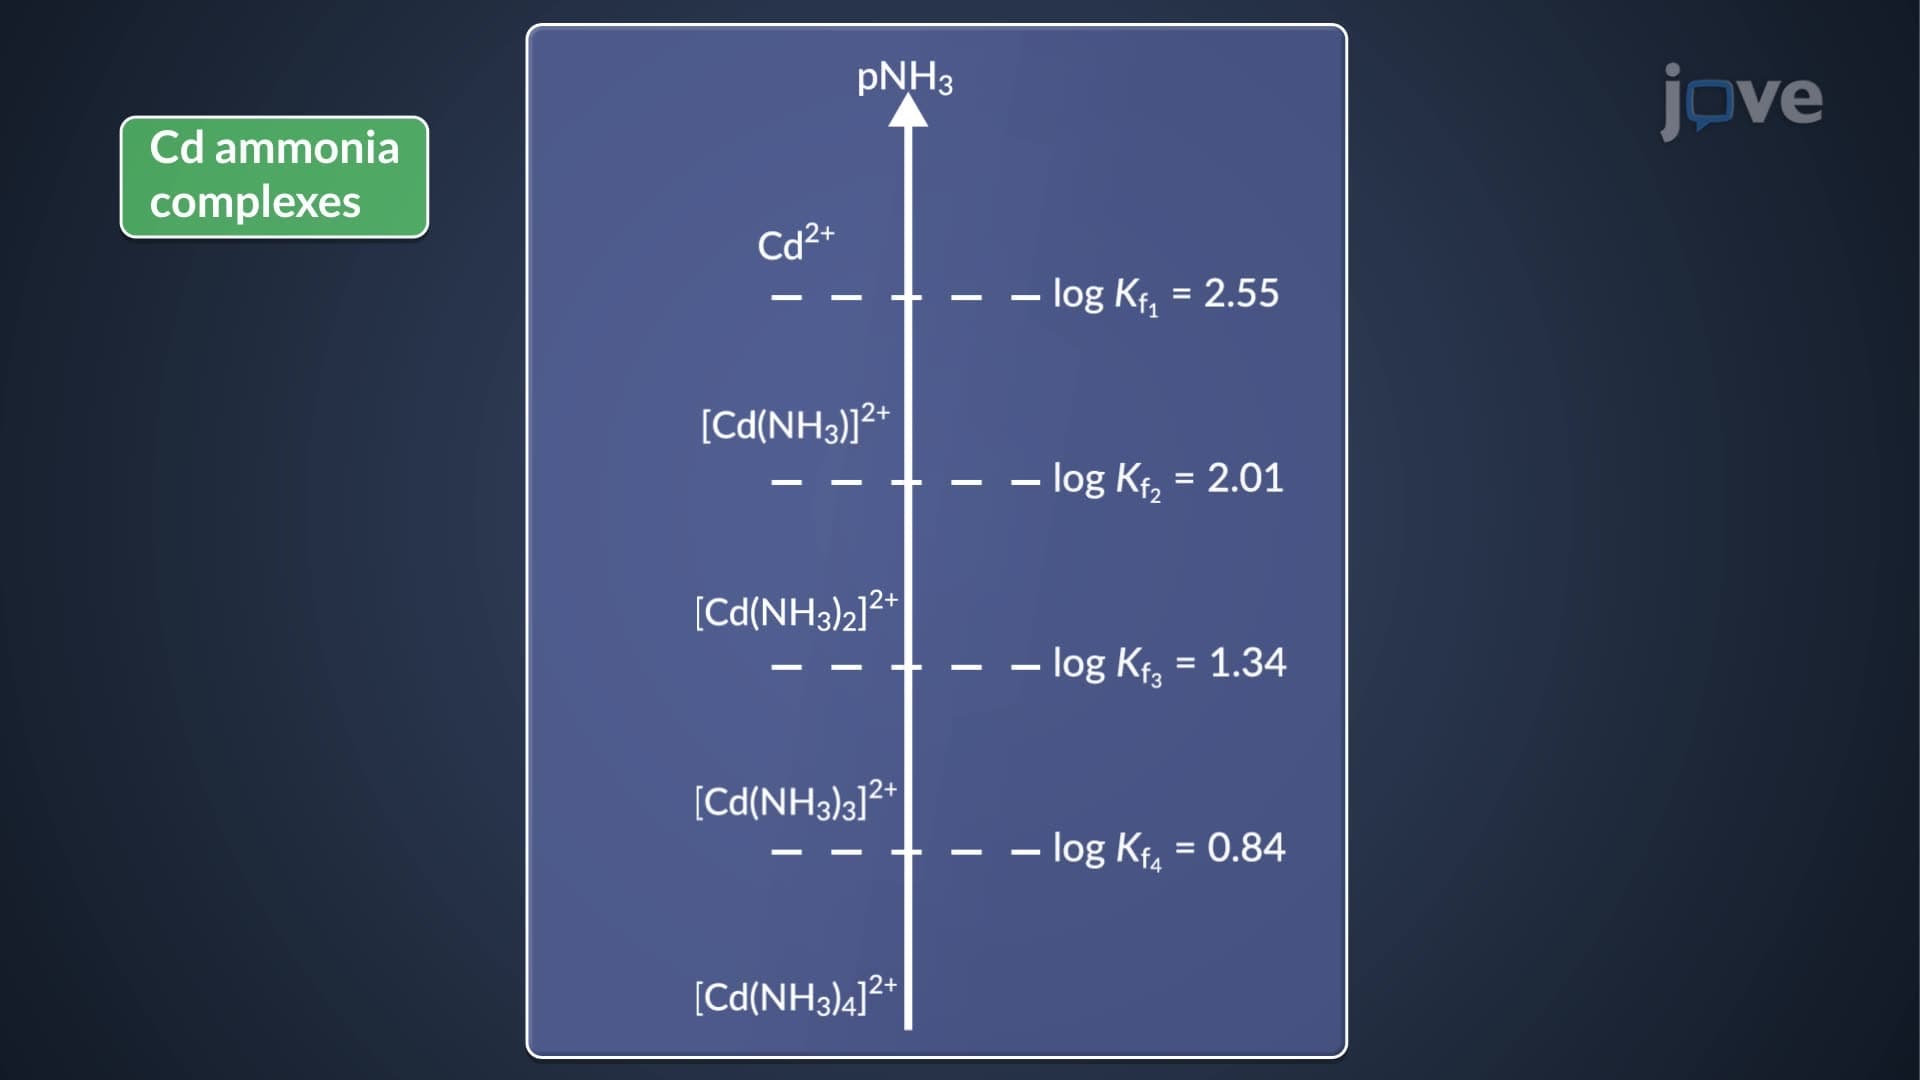
The formation constant, K1, for the formation of Cd(NH3)2+ complex from cadmium and ammonia is 3.55 × 10². Log K1 (i.e. pNH3) is 2.55, and...
 02:42
Valence Bond Theory
 00:43
Complexometric Titration: Ligands
 02:18
The Equilibrium Binding Constant and Binding Strength