Related Experiment Video
Updated: Mar 29, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Density Functional Static Dipole Polarizability and First-Hyperpolarizability Calculations of Nan (n = 2, 4, 6, 8)
K B Sophy1, Patrizia Calaminici1, Sourav Pal1
1Theory Group, Physical Chemistry Division, National Chemical Laboratory, Pune 411 008, India, and Departmento de Quimica, CINVESTAV, Av. Instituto Politecnico Nacional 2508, A. P. 14-740 Mexico, D. F. 07000, Mexico.

Abstract:
We report the static dipole polarizability and first-hyperpolarizability of the sodium atom clusters, Nan, n = 2, 4, 6 and 8, using our recent implementation of a numerical-analytical approach to the coupled-perturbed Kohn-Sham (CPKS) equations in deMon2k. The calculations are reported for VWN and BP86 exchange-correlation functionals using Sadlej and TZVP-FIP1 basis sets which have been previously optimized for polarizability calculations. All-electron calculations were performed for the optimizations at the VWN/DZVP/A2 and PW86/DZVP/A2 levels. Comparisons are made with Hartree-Fock (HF) and MP2 benchmark calculations.

Related Concept Videos
 01:29
Potential Due to a Polarized Object
 01:27
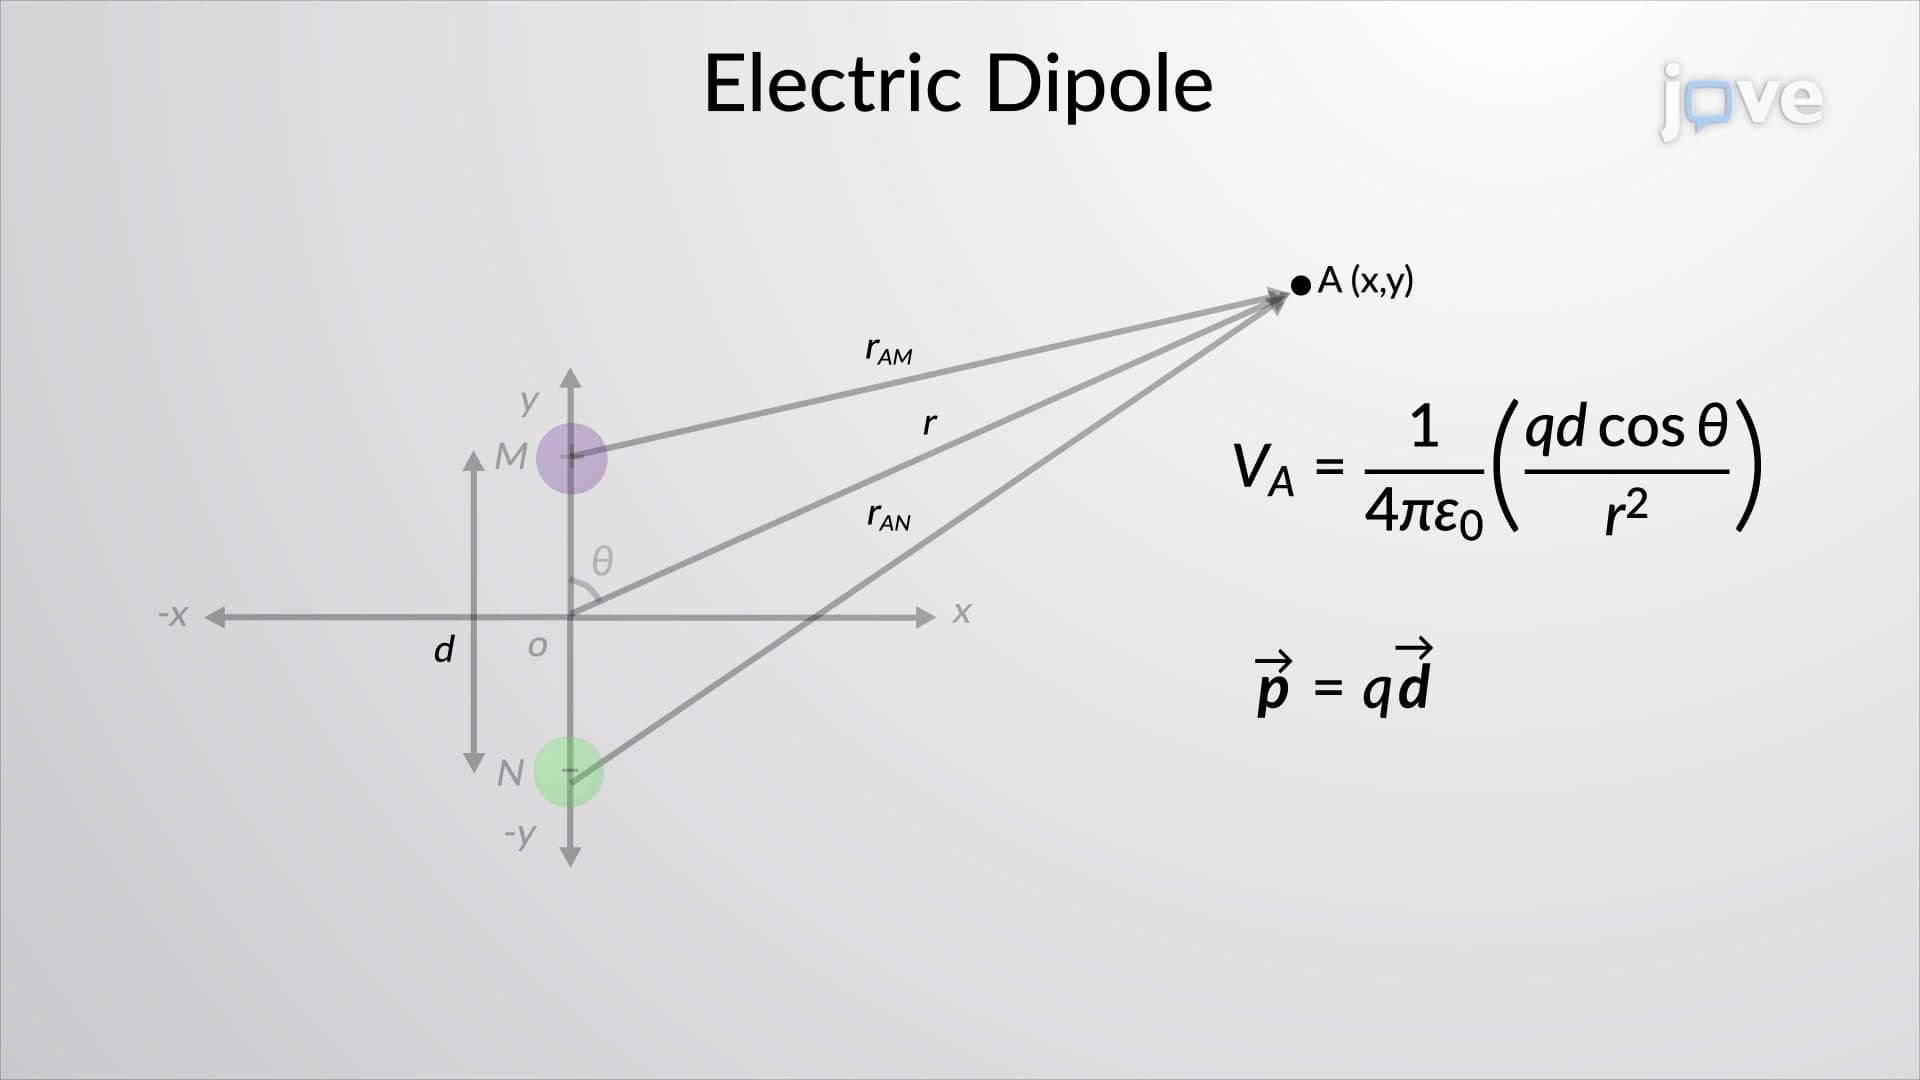
Calculations of Electric Potential II
Consider a...
 02:36
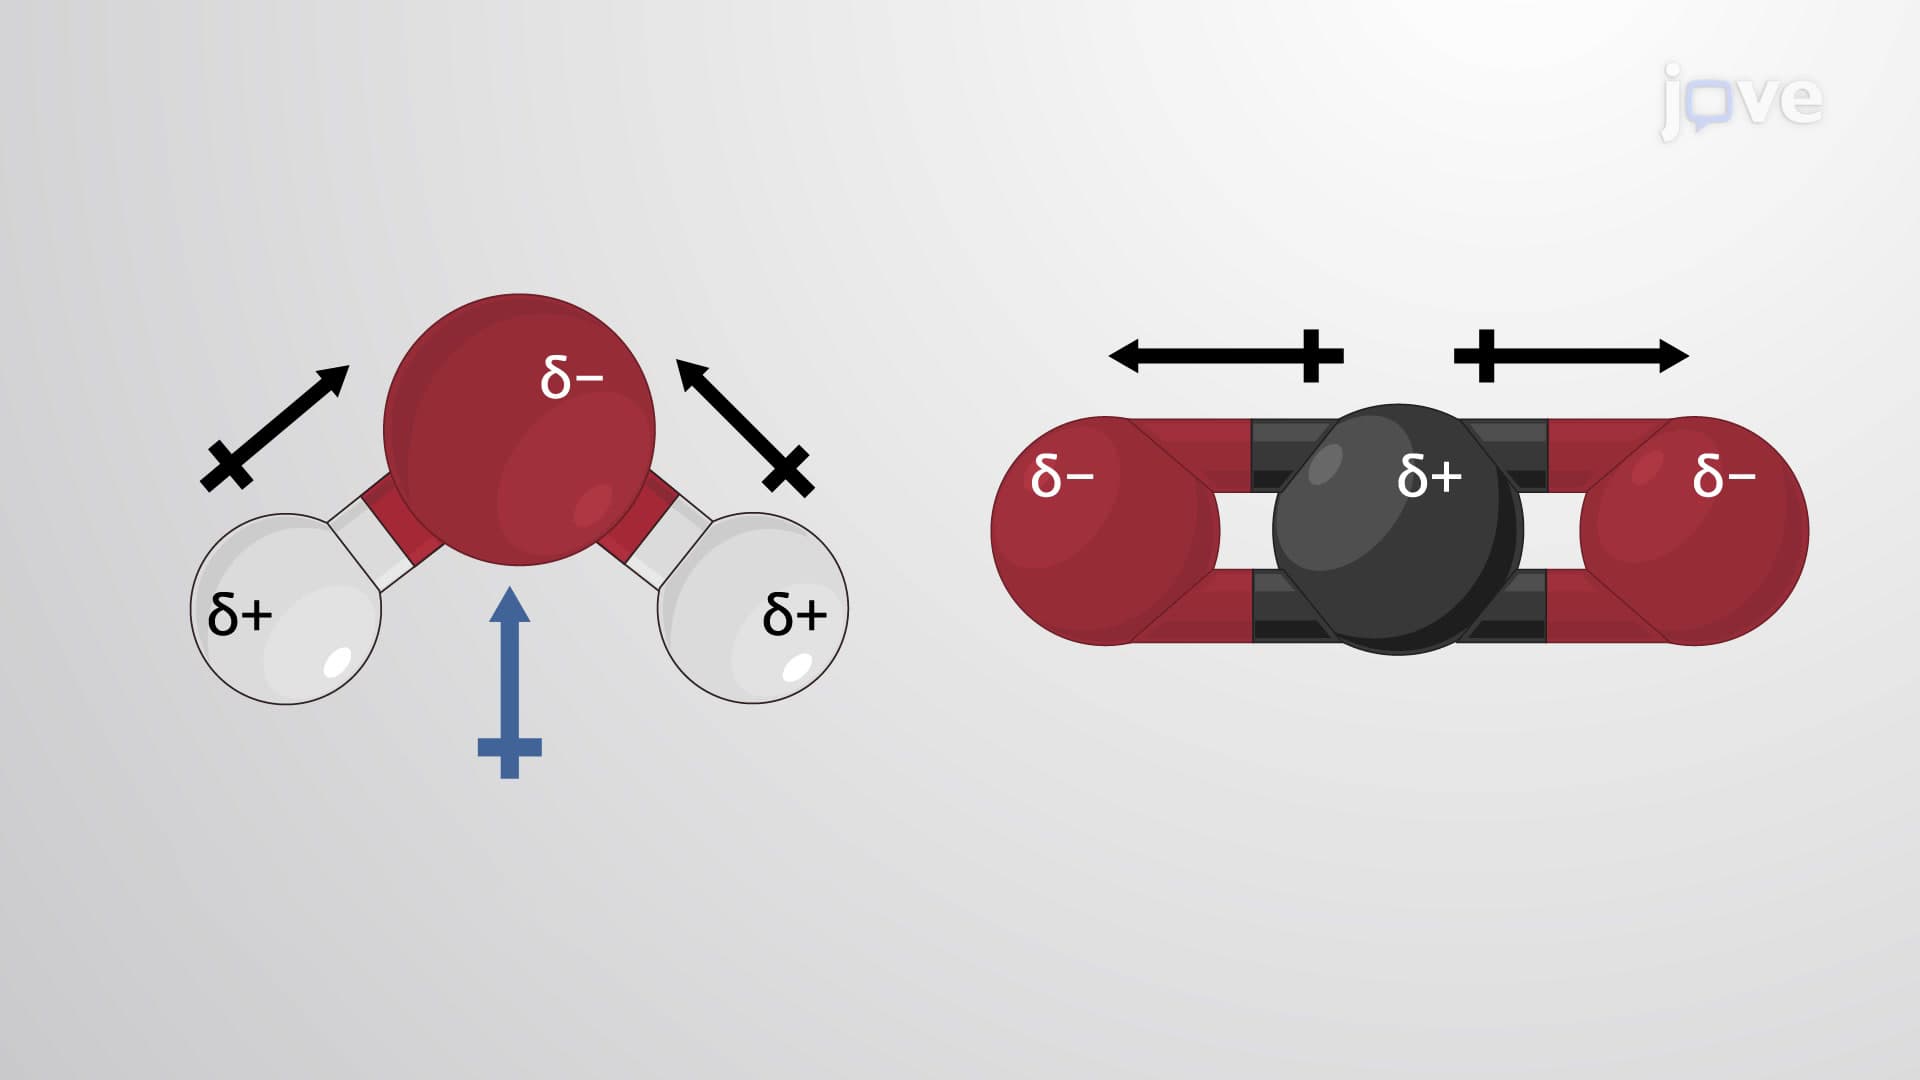
Molecular Geometry and Dipole Moments
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:48
Bond Polarity, Dipole Moment, and Percent Ionic Character
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...