
Atomistic Simulation of Adiabatic Reactive Processes Based on Multi-State Potential Energy Surfaces
Jonas Danielsson1, Markus Meuwly1
1Chemistry Department, University of Basel, Klingelbergstrasse 80, CH-4056 Basel, Switzerland.

Abstract:
The adiabatic reactive molecular dynamics (ARMD) method provides a framework to study chemical reactions using molecular dynamics simulations with minimal computational overhead. Here, ARMD is generalized to an arbitrary reactive process between two states in which reactants and products can be treated by an atomistic force field. The implementation is described, and the method is applied to two systems: the kinetics of NO rebinding to myoglobin (Mb) as a validation system and the conformational transition in neuroglobin (Ngb) which explores the full functionality of ARMD. For MbNO, the nonexponential kinetics observed both in experiment and earlier ARMD studies is reproduced. Furthermore, the sensitivity of the results with respect to the asymptotic separation between the two potential energy surfaces (NO bound and unbound) is studied.
More Related Videos
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:18
Adiabatic Processes for an Ideal Gas
 01:27
Pressure and Volume in an Adiabatic Process
 01:16
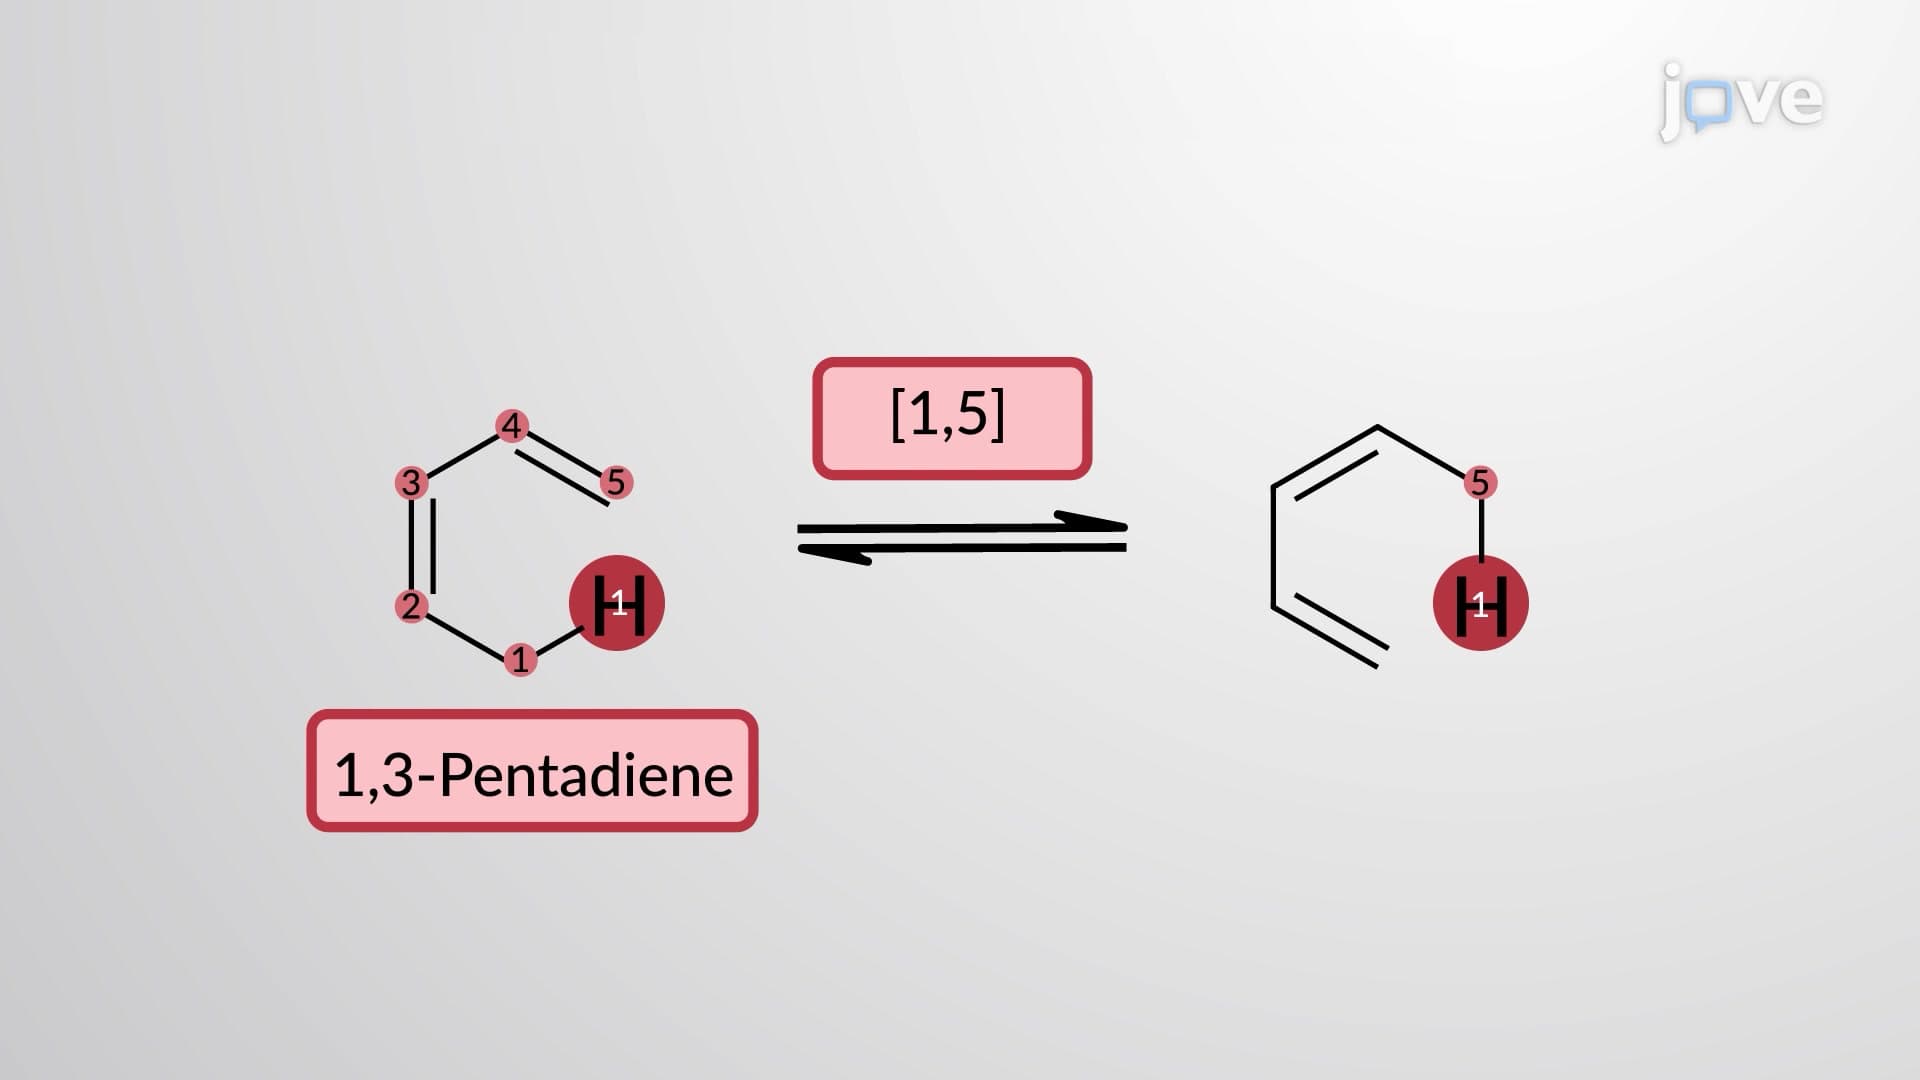
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred...
 02:13
Energy Diagrams, Transition States, and Intermediates
 01:20
Work Done in an Adiabatic Process
 01:26
Thermodynamic Potentials