Direct Free Energy Calculation in the Continuous Fractional Component Gibbs Ensemble
Related Experiment Video
Updated: Mar 24, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Ali Poursaeidesfahani1, Ariana Torres-Knoop2, David Dubbeldam2
1Engineering Thermodynamics, Process & Energy Department, Faculty of Mechanical, Maritime and Materials Engineering, Delft University of Technology , Leeghwaterstraat 39, 2628CB Delft, The Netherlands.

Abstract:
A new formulation of the Gibbs ensemble (GE) combined with the continuous fractional component Monte Carlo method is presented. In the proposed formulation, only a single fractional molecule per component is used instead of two in the original formulation by Shi and Maginn ( J. Comput. Chem. 2008 , 29 , 2520 - 2530 ). This has the following advantages: (1) one directly obtains chemical potentials, without using test particles. We show analytically that the expressions for the chemical potential are identical to those in the conventional Gibbs ensemble; (2) biasing is applied to each simulation box independently; (3) maximum allowed changes in the scaling parameter of intermolecular interactions can be chosen differently in each simulation box. Obtaining chemical potentials directly facilitates thermodynamic modeling using equations of state, and it can be used as an independent check to ensure that chemical equilibrium is achieved. As a proof of principle, our method is tested for Lennard-Jones (LJ) particles and the TIP3P-Ew water model. Results are compared with the conventional GE. Excellent agreement was found both for average densities and chemical potentials. In our new approach, the acceptance probability for molecule exchanges between the boxes is much higher (typically larger than 40% for LJ particles) than for the conventional GE (typically lower than 2% for LJ particles). It is also shown that the contribution of the fractional molecule should be disregarded when computing ensemble averages such as the average energy per molecule and the average densities. The algorithm can be easily extended to mixtures and molecules with intramolecular interactions.
More Related Videos
Related Concept Videos
 02:39
Gibbs Free Energy
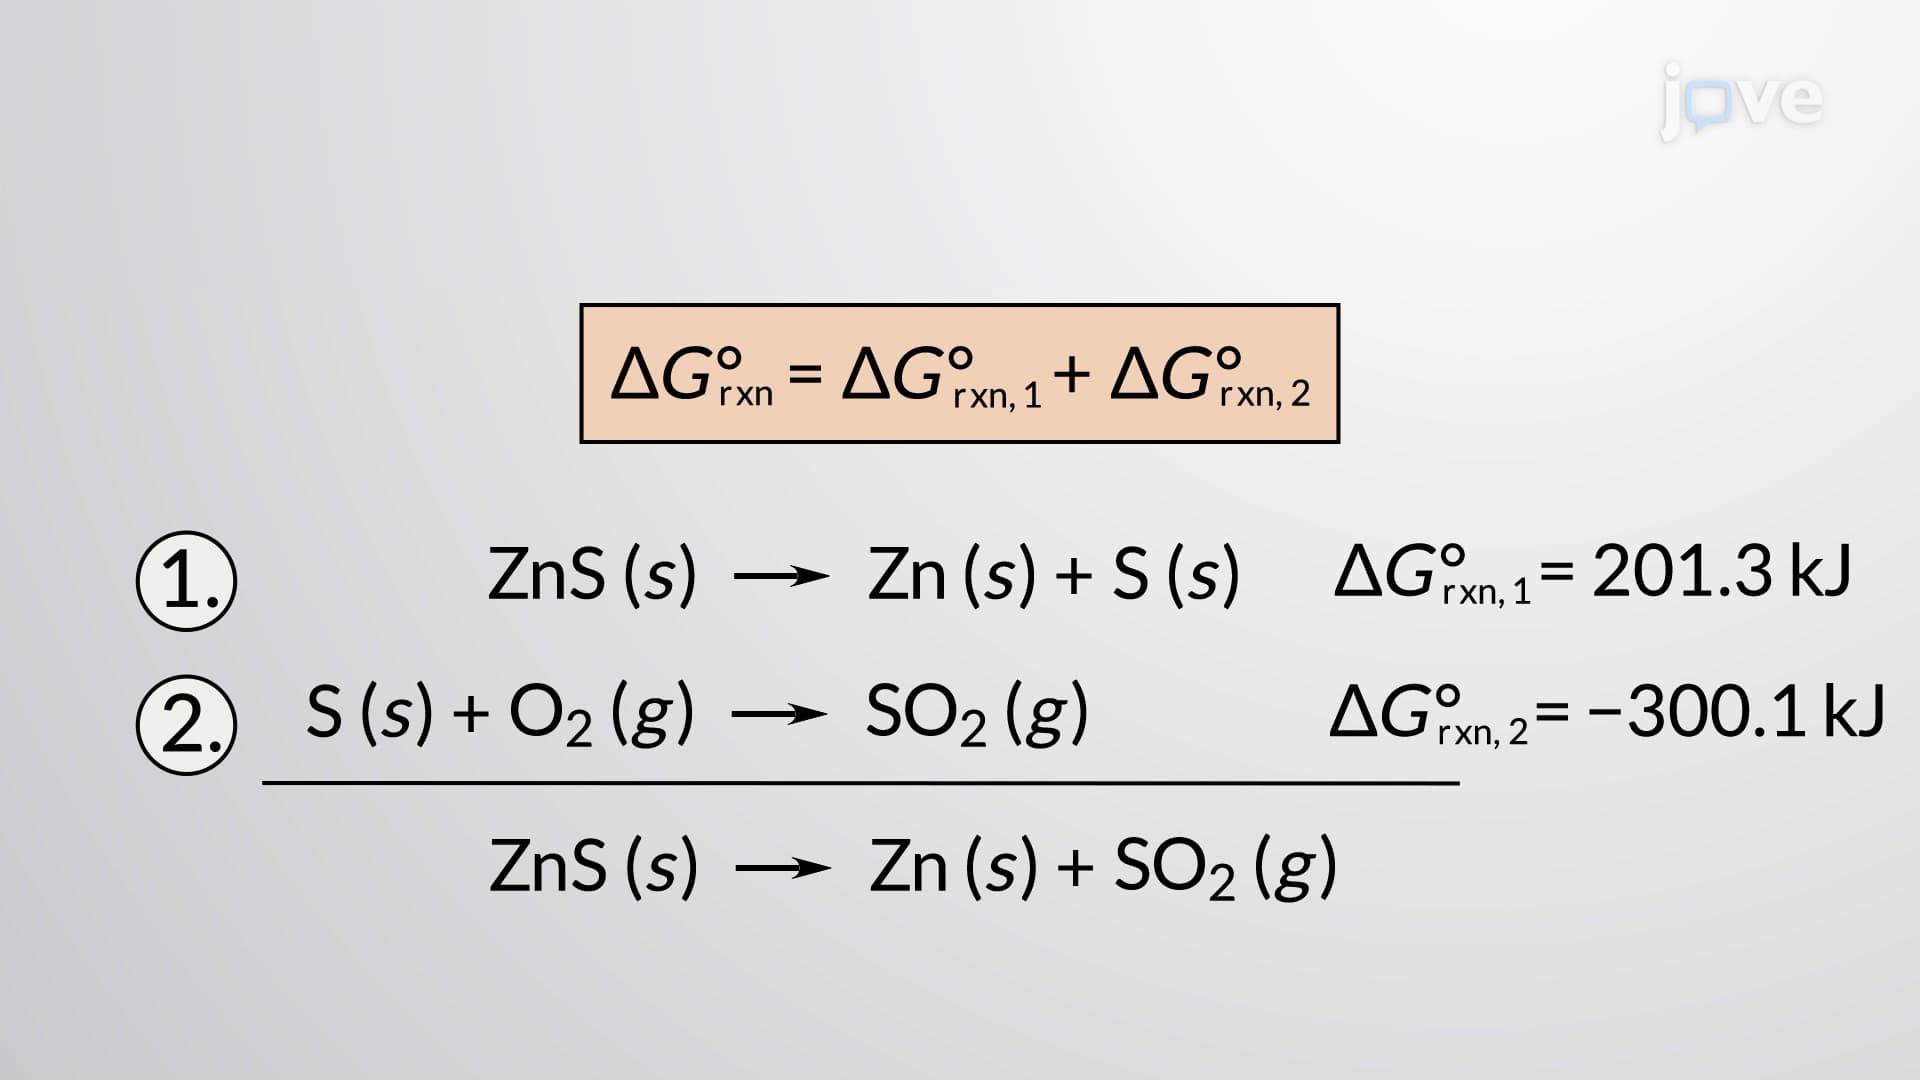 02:49
Calculating Standard Free Energy Changes
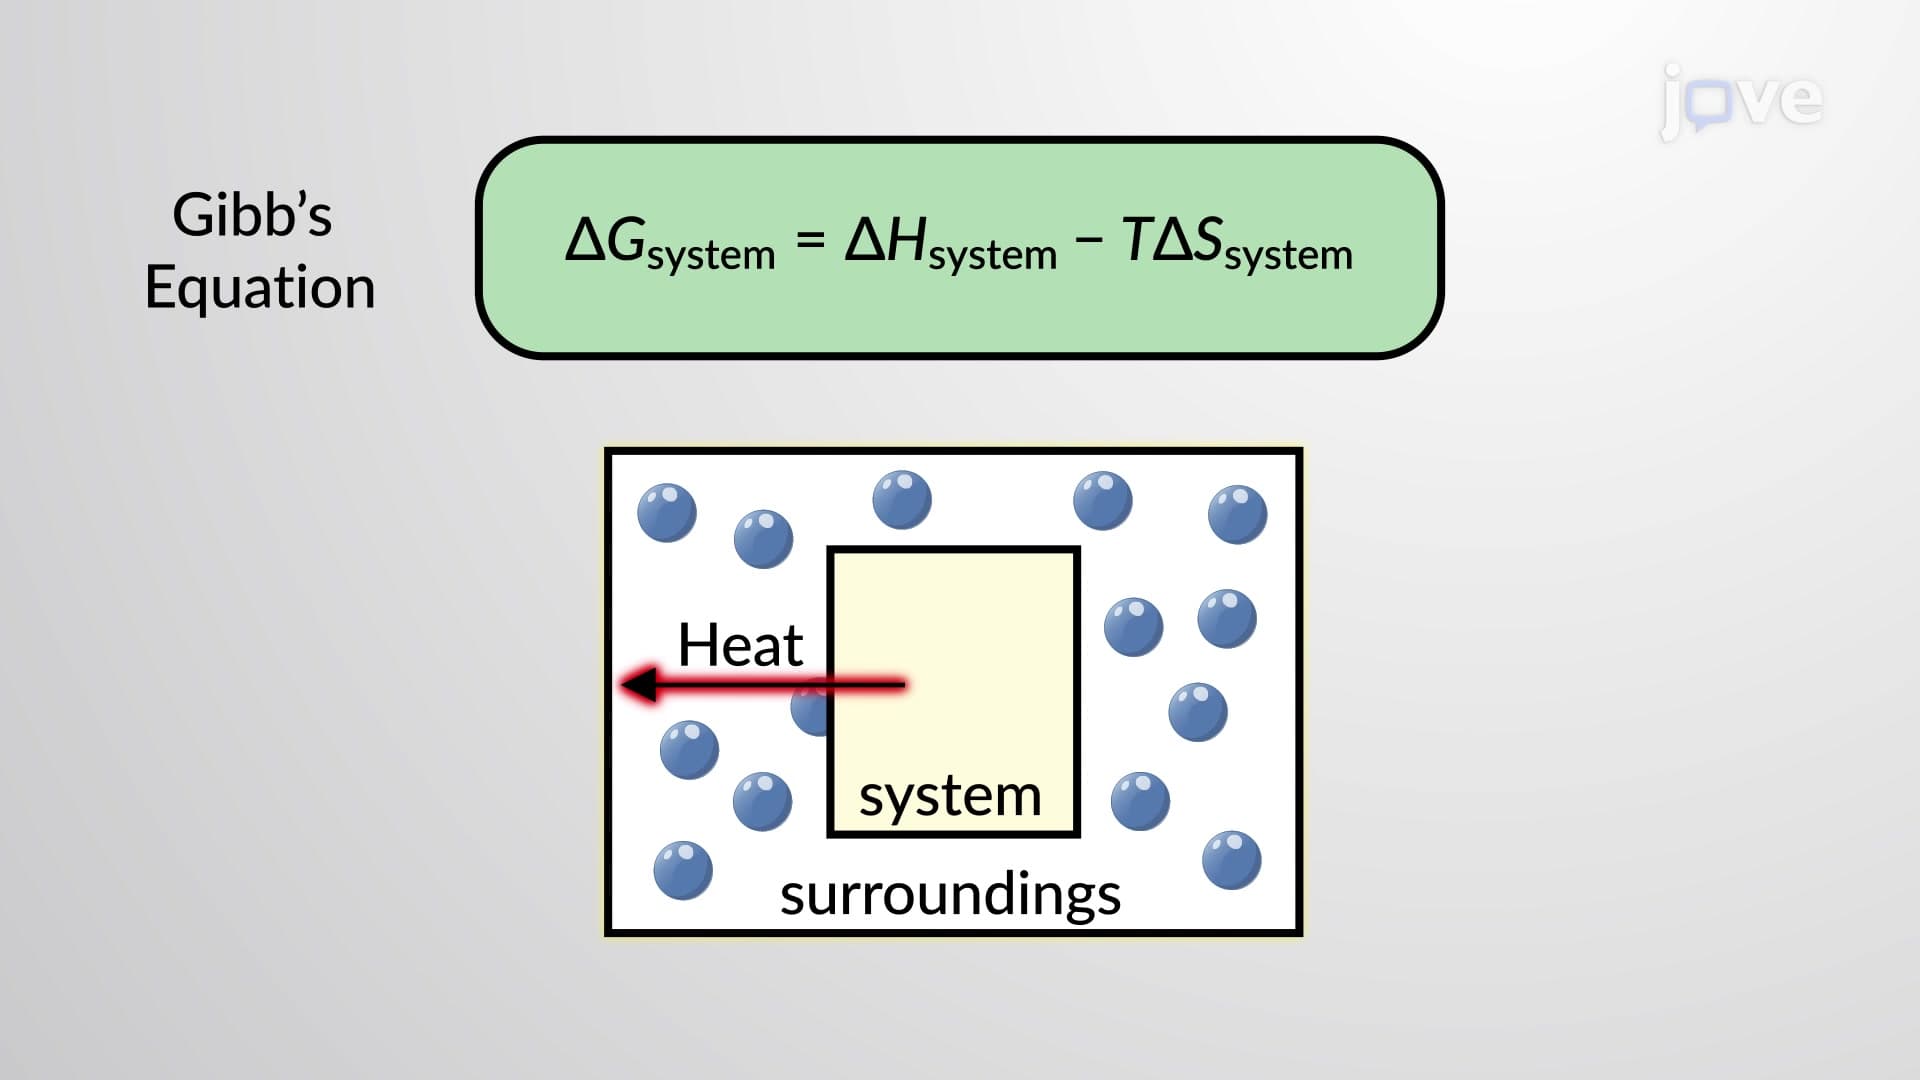 01:05
An Introduction to Free Energy
 03:25
Free Energy Changes for Nonstandard States
 02:23
Gibbs Free Energy and Thermodynamic Favorability
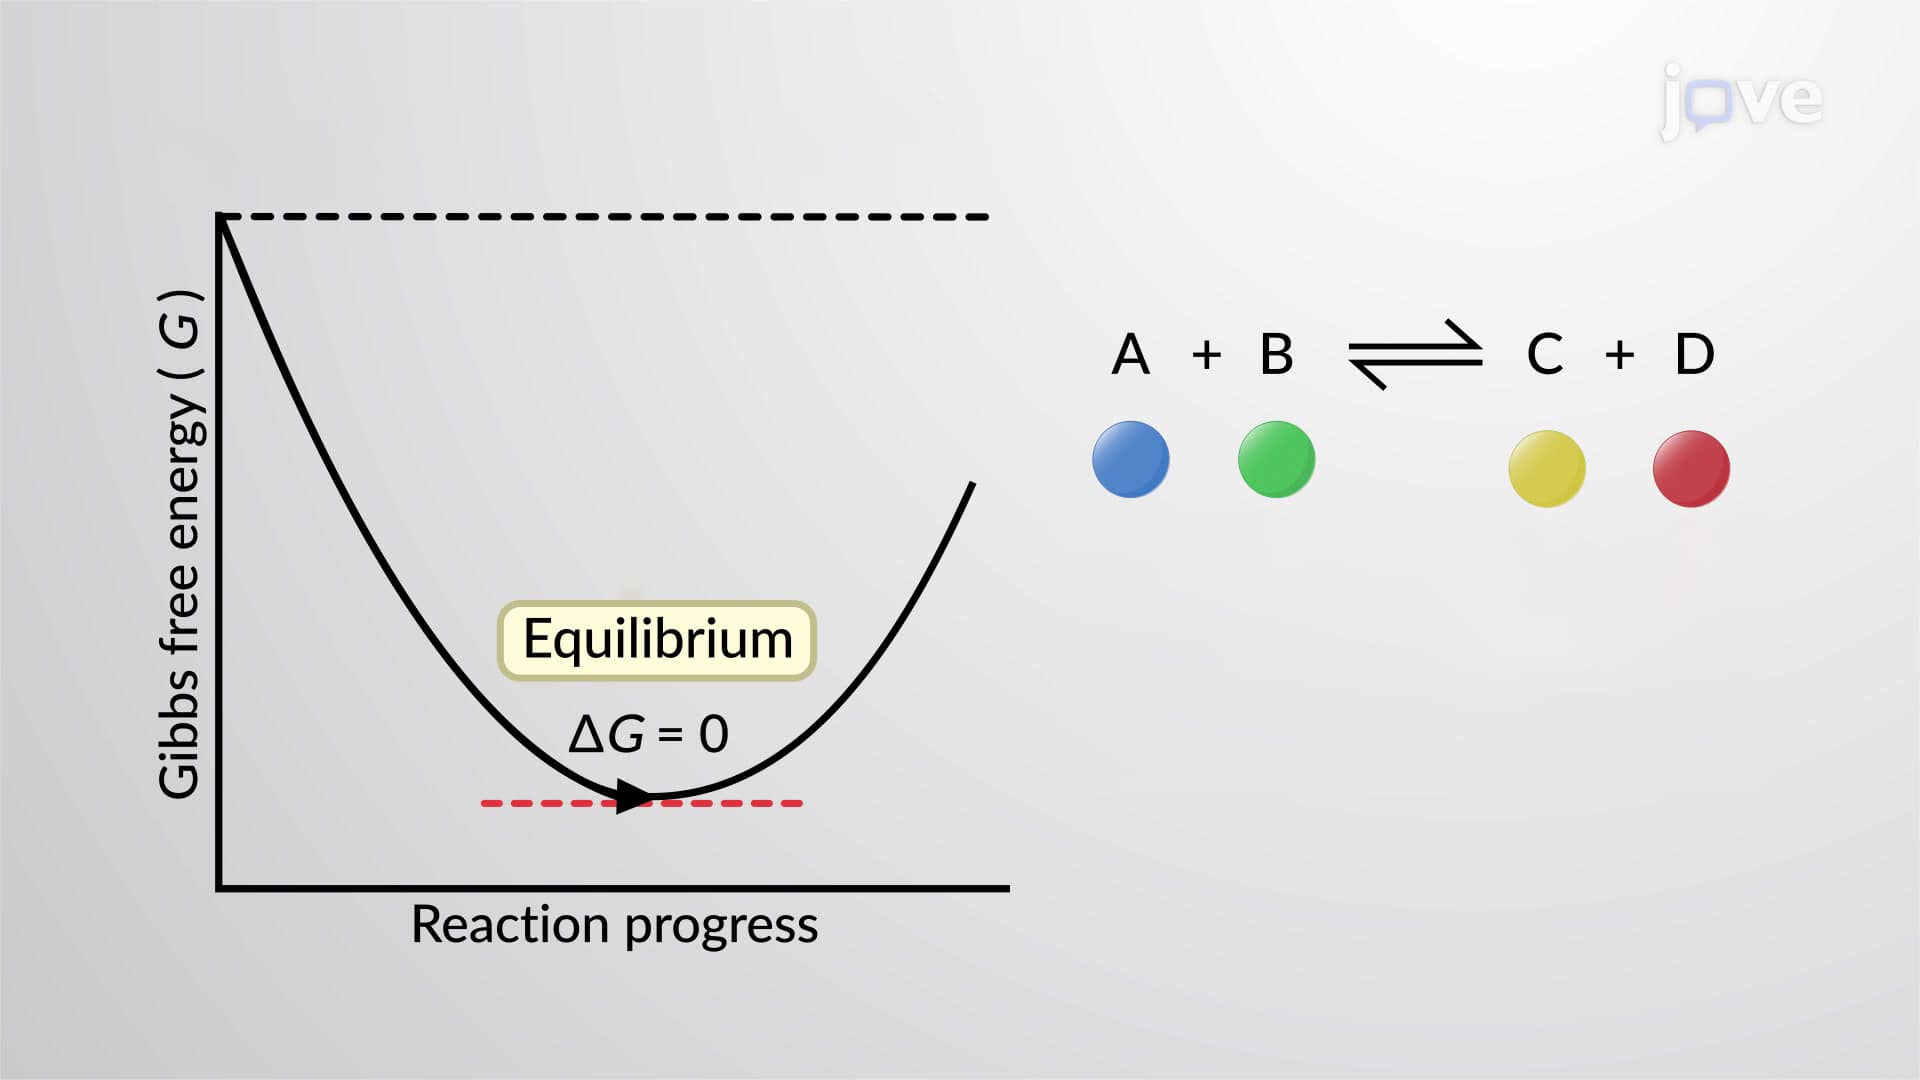 00:55
Free Energy and Equilibrium
The reaction quotient, Q, is a convenient measure of the...