Modeling a halogen dance reaction mechanism: A density functional theory study
Related Experiment Video
Updated: Mar 22, 2026
From Molecules to Materials: Engineering New Ionic Liquid Crystals Through Halogen Bonding
Published on: March 24, 2018
Leighton Jones1, Benjamin J Whitaker1
1School of Chemistry, University of Leeds, LS2 9JT, United Kingdom.

Abstract:
Since the discovery of the halogen dance (HD) reaction more than 60 years ago, numerous insights into the mechanism have been unveiled. To date however, the reaction has not been investigated from a theoretical perspective. Density functional theory (DFT) was used to model the potential energy surface linking the starting reagents to the lithiated products for each step in the mechanism using a thiophene substrate. It was found that the lithium-halogen exchange mechanism is critical to understand the HD mechanism in detail and yielded the knowledge that SN 2 transition states (TS) are favored over the four-center type for the lithium-bromine exchange steps. The overall driving force for the HD is thermodynamics, while the kinetic factors tightly control the reaction path through temperature. The SN 2 lithium-bromide TS are barrierless, except the second, which is the limiting step. Finally, the model for the HD is discovered to be a pseudo-clock type, due to a highly favorable bromide catalysis step and the reformation of 2-bromothiophene. © 2016 Wiley Periodicals, Inc.
Related Concept Videos
 03:01
Halogens
 02:41
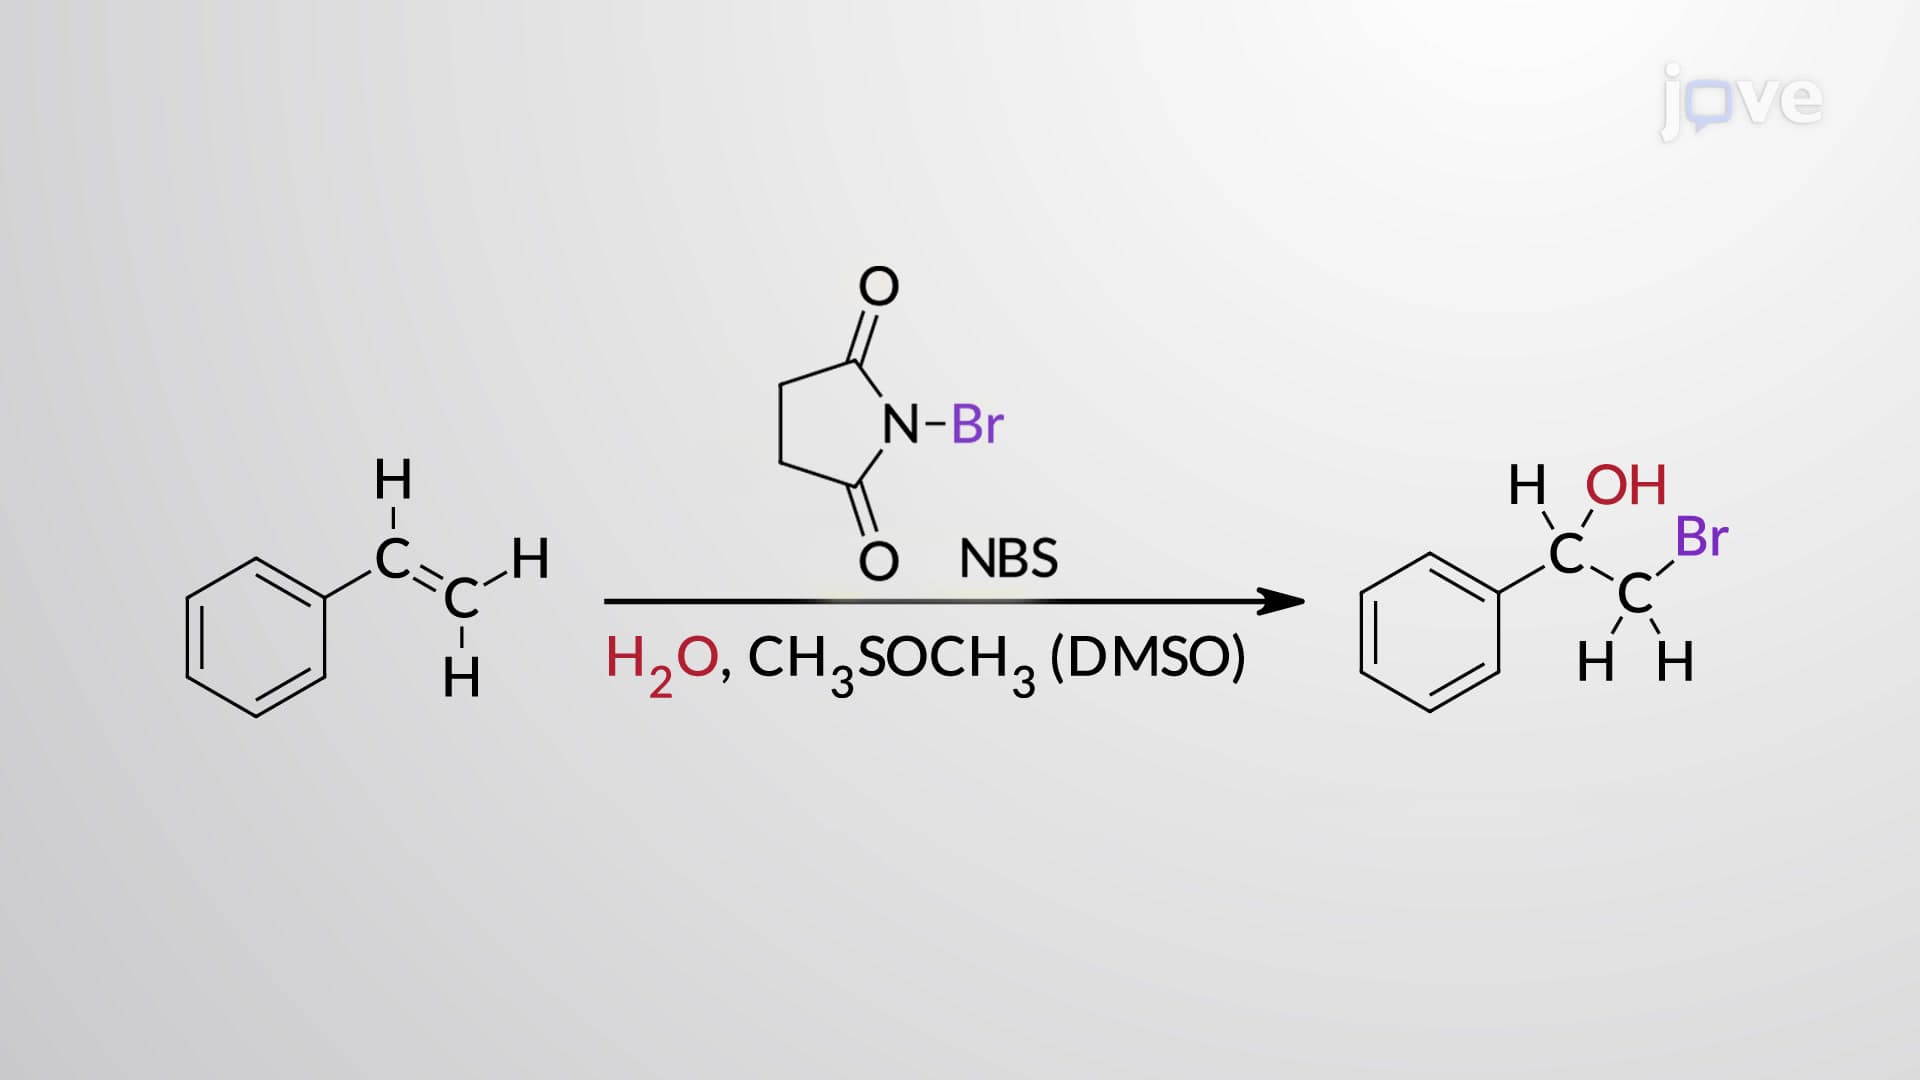
Formation of Halohydrin from Alkenes
 01:34
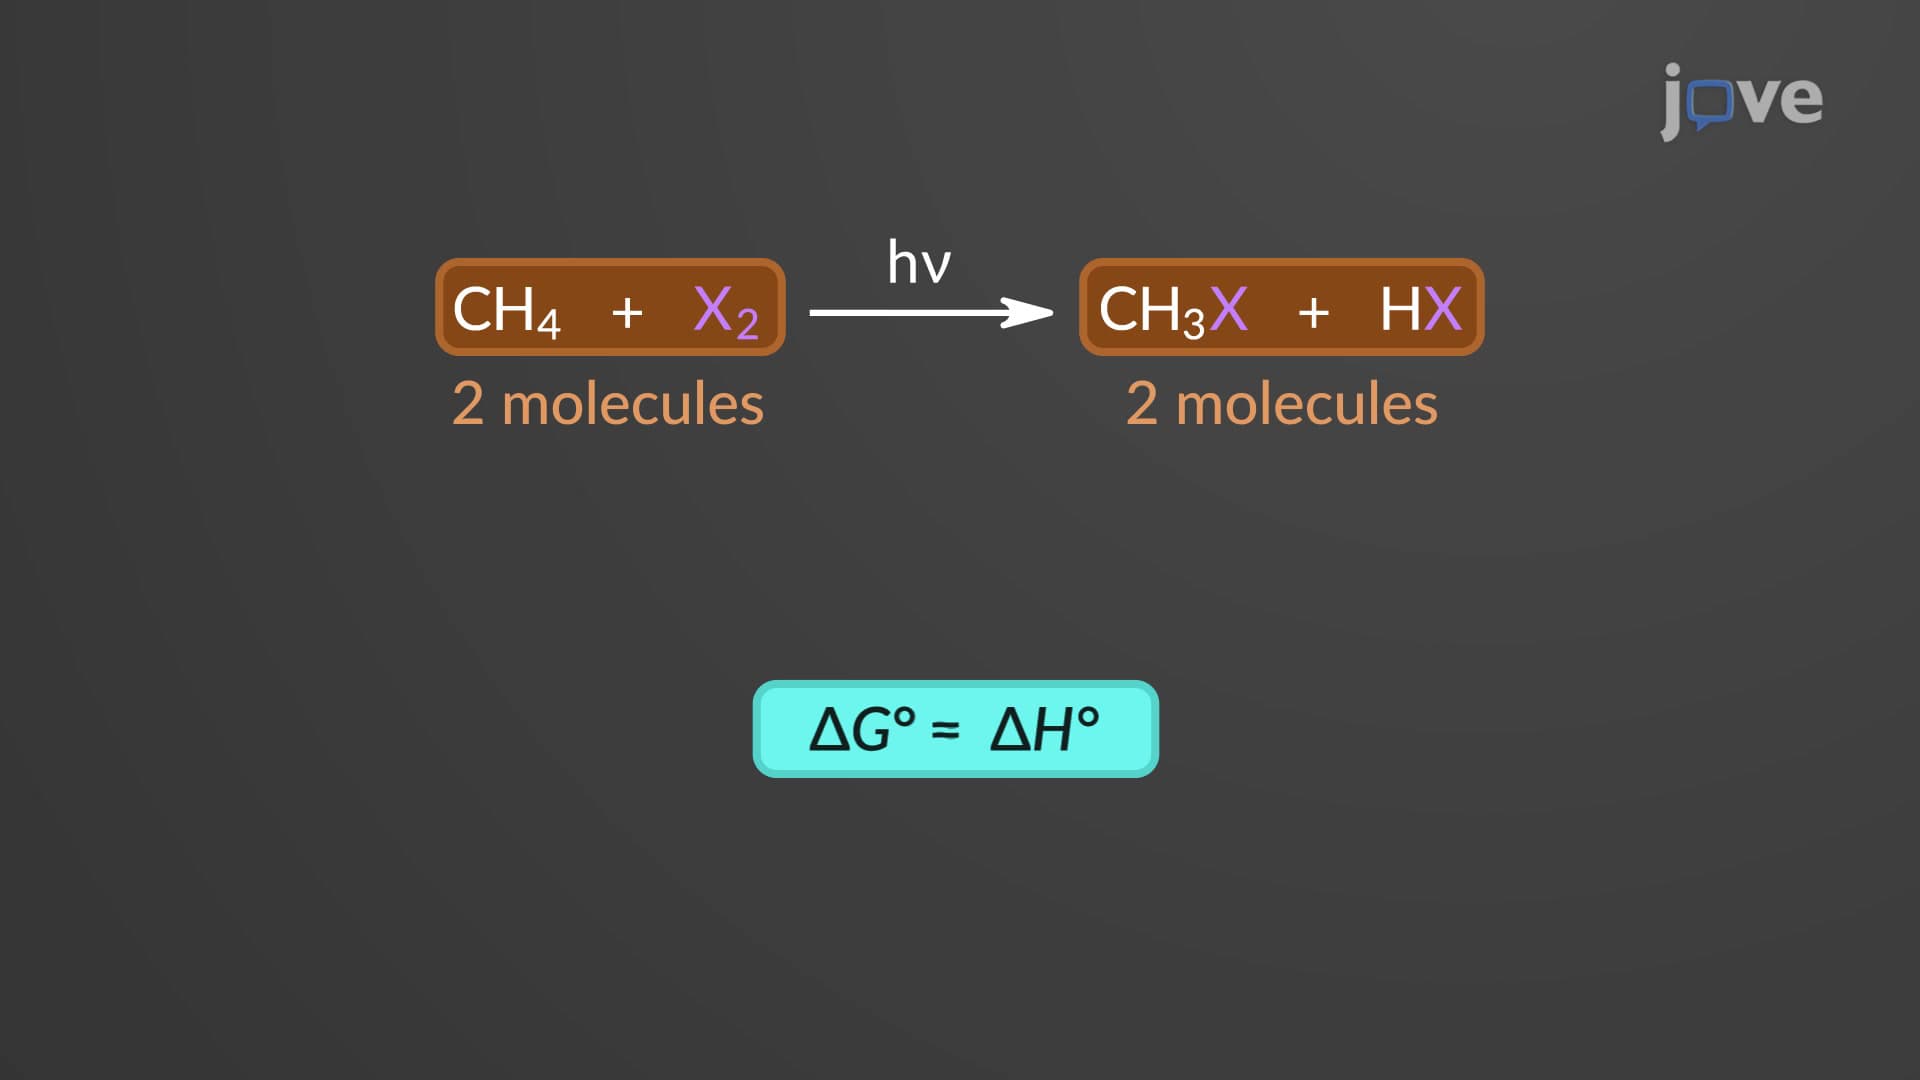
Radical Halogenation: Thermodynamics
 01:24
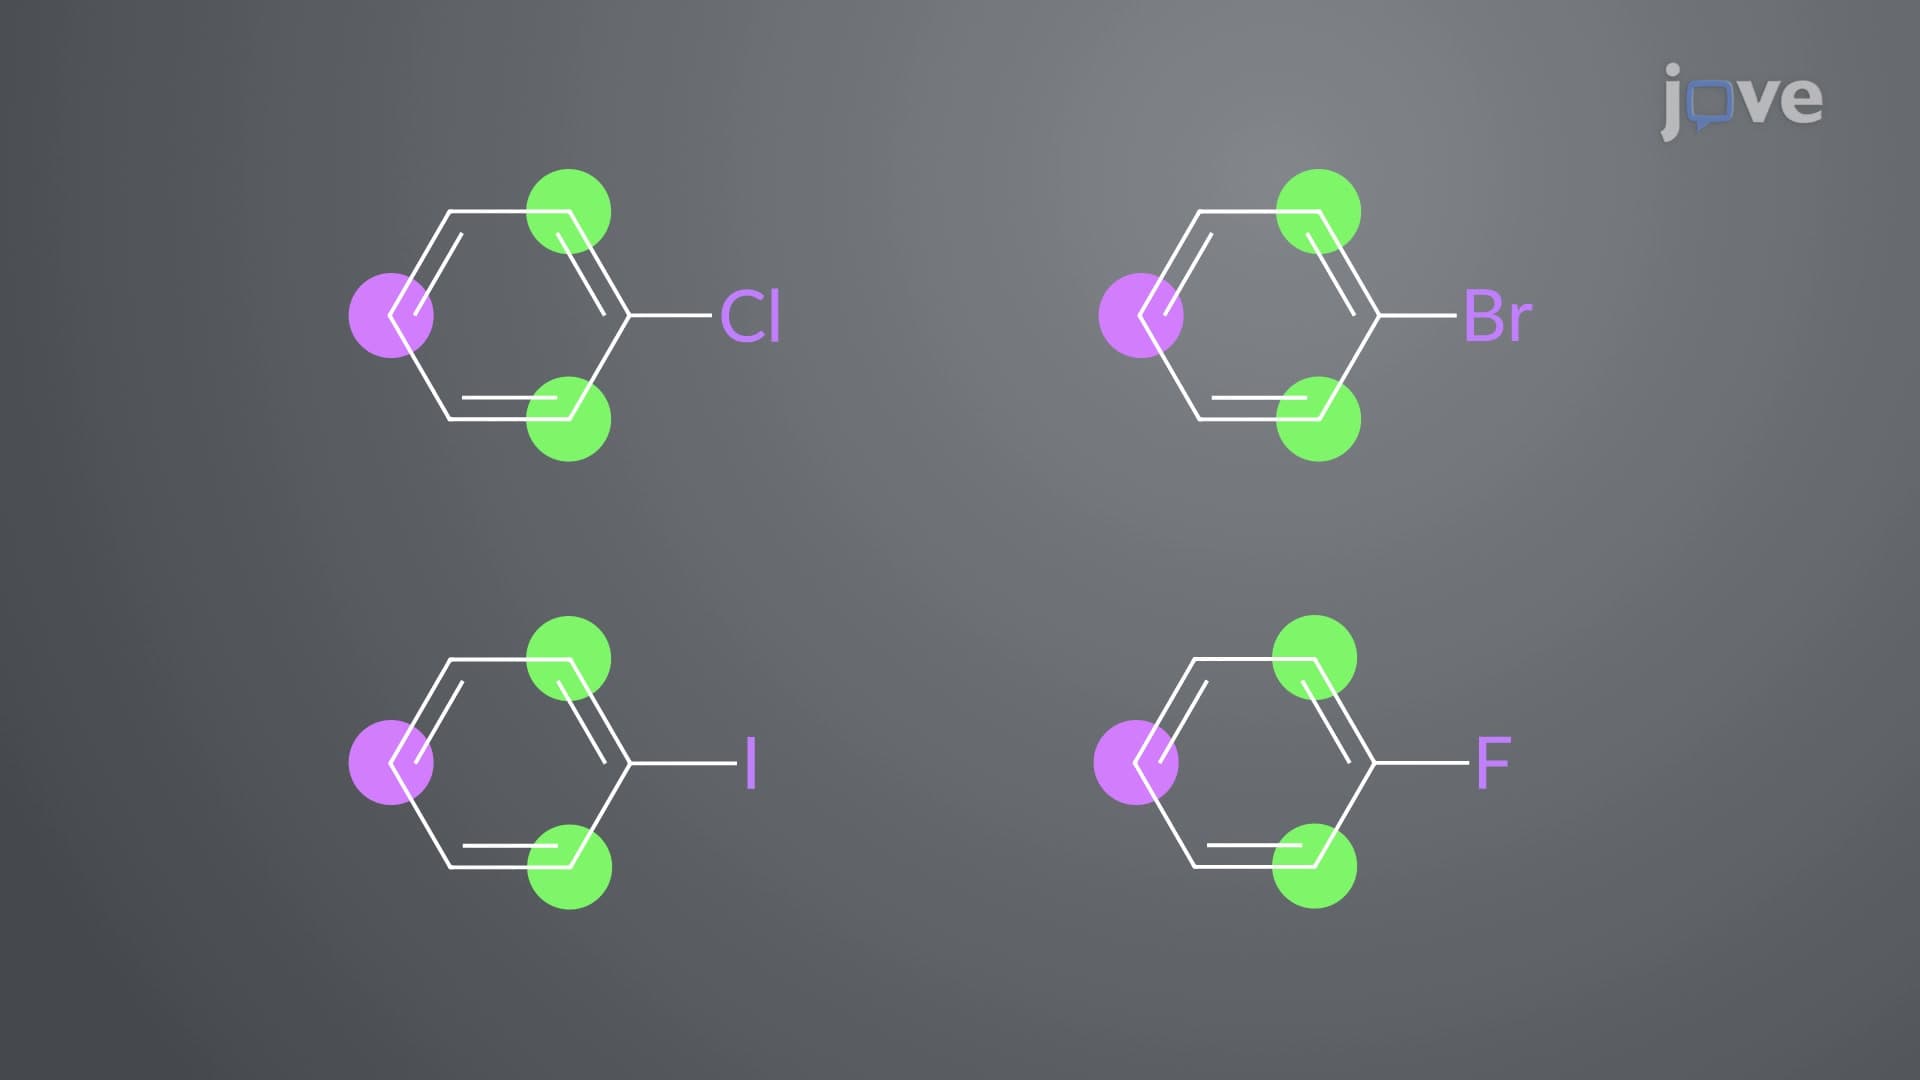
ortho–para-Directing Deactivators: Halogens
 02:46
Halogenation of Alkenes
Consider the bromination of cyclopentene. Molecular bromine is polarized in the proximity of the π electrons of cyclopentene. An electrophilic bromine atom adds across the double bond, forming a cyclic bromonium ion intermediate.
 01:14
Electrophilic 1,2- and 1,4-Addition of X2 to 1,3-Butadiene