Related Experiment Video
Updated: Mar 22, 2026
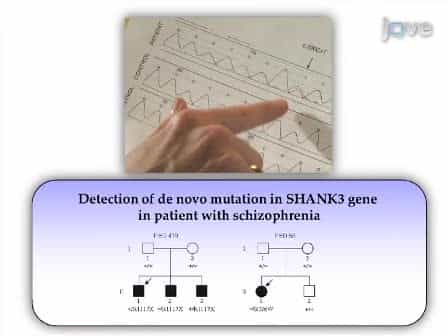

A Strategy to Identify de Novo Mutations in Common Disorders such as Autism and Schizophrenia
Published on: June 15, 2011
A Restricted Repertoire of De Novo Mutations in ITPR1 Cause Gillespie Syndrome with Evidence for Dominant-Negative
Meriel McEntagart1, Kathleen A Williamson2, Jacqueline K Rainger2
1Medical Genetics, St George's University Hospitals NHS Foundation Trust, Cranmer Terrace, London SW17 0RE, UK.

Abstract:
Gillespie syndrome (GS) is characterized by bilateral iris hypoplasia, congenital hypotonia, non-progressive ataxia, and progressive cerebellar atrophy. Trio-based exome sequencing identified de novo mutations in ITPR1 in three unrelated individuals with GS recruited to the Deciphering Developmental Disorders study. Whole-exome or targeted sequence analysis identified plausible disease-causing ITPR1 mutations in 10/10 additional GS-affected individuals. These ultra-rare protein-altering variants affected only three residues in ITPR1: Glu2094 missense (one de novo, one co-segregating), Gly2539 missense (five de novo, one inheritance uncertain), and Lys2596 in-frame deletion (four de novo). No clinical or radiological differences were evident between individuals with different mutations. ITPR1 encodes an inositol 1,4,5-triphosphate-responsive calcium channel. The homo-tetrameric structure has been solved by cryoelectron microscopy. Using estimations of the degree of structural change induced by known recessive- and dominant-negative mutations in other disease-associated multimeric channels, we developed a generalizable computational approach to indicate the likely mutational mechanism. This analysis supports a dominant-negative mechanism for GS variants in ITPR1. In GS-derived lymphoblastoid cell lines (LCLs), the proportion of ITPR1-positive cells using immunofluorescence was significantly higher in mutant than control LCLs, consistent with an abnormality of nuclear calcium signaling feedback control. Super-resolution imaging supports the existence of an ITPR1-lined nucleoplasmic reticulum. Mice with Itpr1 heterozygous null mutations showed no major iris defects. Purkinje cells of the cerebellum appear to be the most sensitive to impaired ITPR1 function in humans. Iris hypoplasia is likely to result from either complete loss of ITPR1 activity or structure-specific disruption of multimeric interactions.
More Related Videos
 09:34
Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
 09:37
Defining Gene Functions in Tumorigenesis by Ex vivo Ablation of Floxed Alleles in Malignant Peripheral Nerve Sheath Tumor Cells
Published on: August 25, 2021
Related Concept Videos
 02:57
piRNA - Piwi-interacting RNAs
 01:20
The Retinoblastoma Gene
The first-ever tumor suppressor gene called Rb was identified in retinoblastoma - a rare eye tumor in children. In inherited forms of the disease, a child inherits one defective copy of the Rb gene, which predisposes them to retinoblastoma. However,...
 01:33
Pleiotropy
 02:18
Alternative RNA Splicing
There are five types of alternative RNA splicing that vary in the ways the pre-mRNA segments are removed or retained in the mature mRNA. The first...
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...
 03:18
Non-LTR Retrotransposons