Related Experiment Video
Updated: Mar 19, 2026
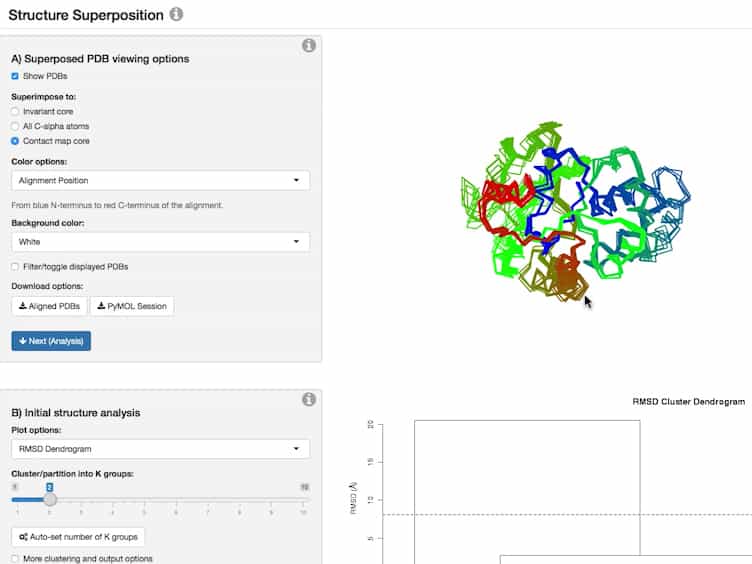

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Cluster analysis of molecular simulation trajectories for systems where both conformation and orientation of the
Tigran M Abramyan1, James A Snyder1, Aby A Thyparambil1
1Department of Bioengineering, 501 Rhodes Engineering Research Center, Clemson University, Clemson, South Carolina, 29634.

Abstract:
Clustering methods have been widely used to group together similar conformational states from molecular simulations of biomolecules in solution. For applications such as the interaction of a protein with a surface, the orientation of the protein relative to the surface is also an important clustering parameter because of its potential effect on adsorbed-state bioactivity. This study presents cluster analysis methods that are specifically designed for systems where both molecular orientation and conformation are important, and the methods are demonstrated using test cases of adsorbed proteins for validation. Additionally, because cluster analysis can be a very subjective process, an objective procedure for identifying both the optimal number of clusters and the best clustering algorithm to be applied to analyze a given dataset is presented. The method is demonstrated for several agglomerative hierarchical clustering algorithms used in conjunction with three cluster validation techniques. © 2016 Wiley Periodicals, Inc.
More Related Videos
 09:17
Structure-Based Simulation and Sampling of Transcription Factor Protein Movements along DNA from Atomic-Scale Stepping to Coarse-Grained Diffusion
Published on: March 1, 2022
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 03:37
Molecular Shape and Polarity
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as...
 02:27
Predicting Molecular Geometry
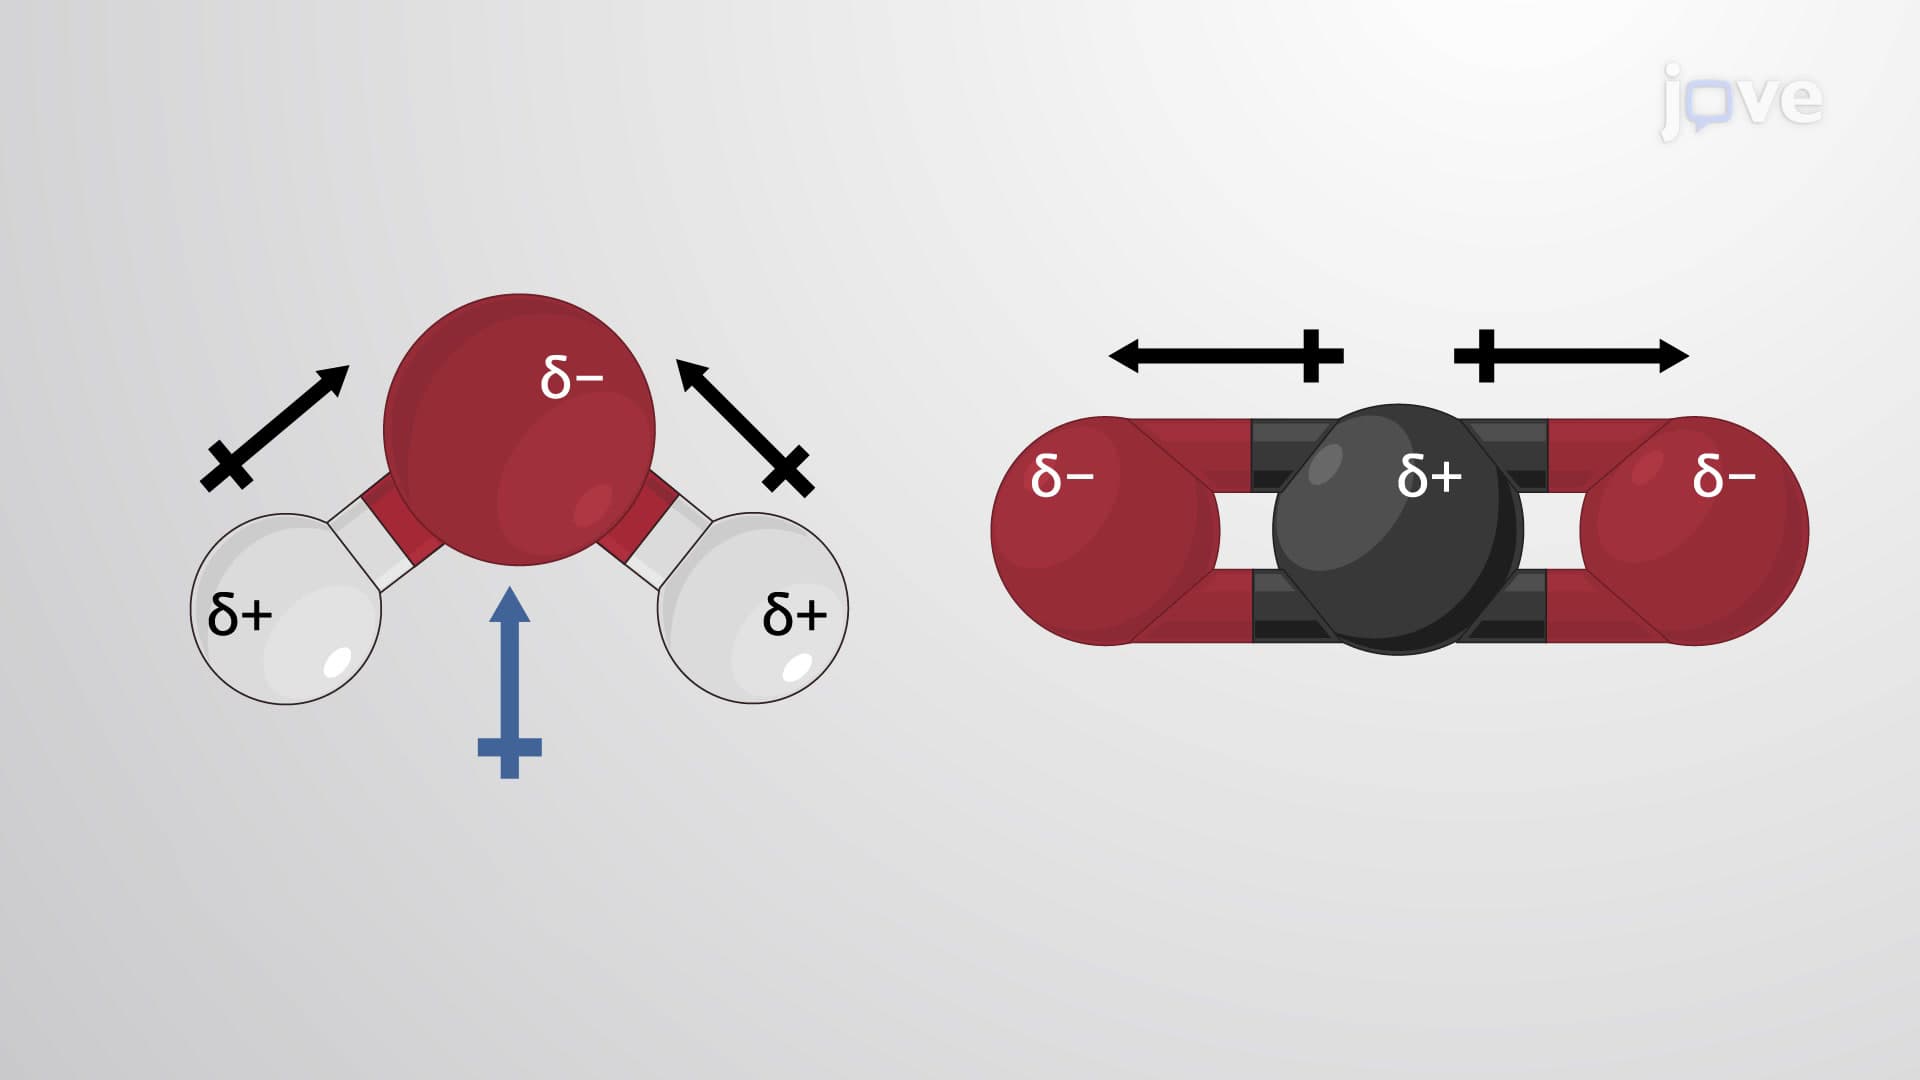 02:36
Molecular Geometry and Dipole Moments