Related Experiment Video
Updated: Mar 17, 2026


Targeted Next-generation Sequencing and Bioinformatics Pipeline to Evaluate Genetic Determinants of Constitutional Disease
Published on: April 4, 2018
Unravelling 5-oxoprolinuria (pyroglutamic aciduria) due to bi-allelic OPLAH mutations: 20 new mutations in 14
Jörn Oliver Sass1, Corinne Gemperle-Britschgi2, Maja Tarailo-Graovac3
1Bioanalytics & Biochemistry, Department of Natural Sciences, Bonn-Rhein-Sieg University of Applied Sciences, Rheinbach, Germany; Clinical Chemistry & Biochemistry, Children's Research Center, University Children's Hospital, Zürich, Switzerland; Laboratory of Clinical Biochemistry and Metabolism, University Children's Hospital, Freiburg, Germany.

Abstract:
Primary 5-oxoprolinuria (pyroglutamic aciduria) is caused by a genetic defect in the γ-glutamyl cycle, affecting either glutathione synthetase or 5-oxoprolinase. While several dozens of patients with glutathione synthetase deficiency have been reported, with hemolytic anemia representing the clinical key feature, 5-oxoprolinase deficiency due to OPLAH mutations is less frequent and so far has not attracted much attention. This has prompted us to investigate the clinical phenotype as well as the underlying genotype in patients from 14 families of various ethnic backgrounds who underwent diagnostic mutation analysis following the detection of 5-oxoprolinuria. In all patients with 5-oxoprolinuria studied, bi-allelic mutations in OPLAH were indicated. An autosomal recessive mode of inheritance for 5-oxoprolinase deficiency is further supported by the identification of a single mutation in all 9/14 parent sample sets investigated (except for the father of one patient whose result suggests homozygosity), and the absence of 5-oxoprolinuria in all tested heterozygotes. It is remarkable, that all 20 mutations identified were novel and private to the respective families. Clinical features were highly variable and in several sib pairs, did not segregate with 5-oxoprolinuria. Although a pathogenic role of 5-oxoprolinase deficiency remains possible, this is not supported by our findings. Additional patient ascertainment and long-term follow-up is needed to establish the benign nature of this inborn error of metabolism. It is important that all symptomatic patients with persistently elevated levels of 5-oxoproline and no obvious explanation are investigated for the genetic etiology.

Related Concept Videos
 01:16
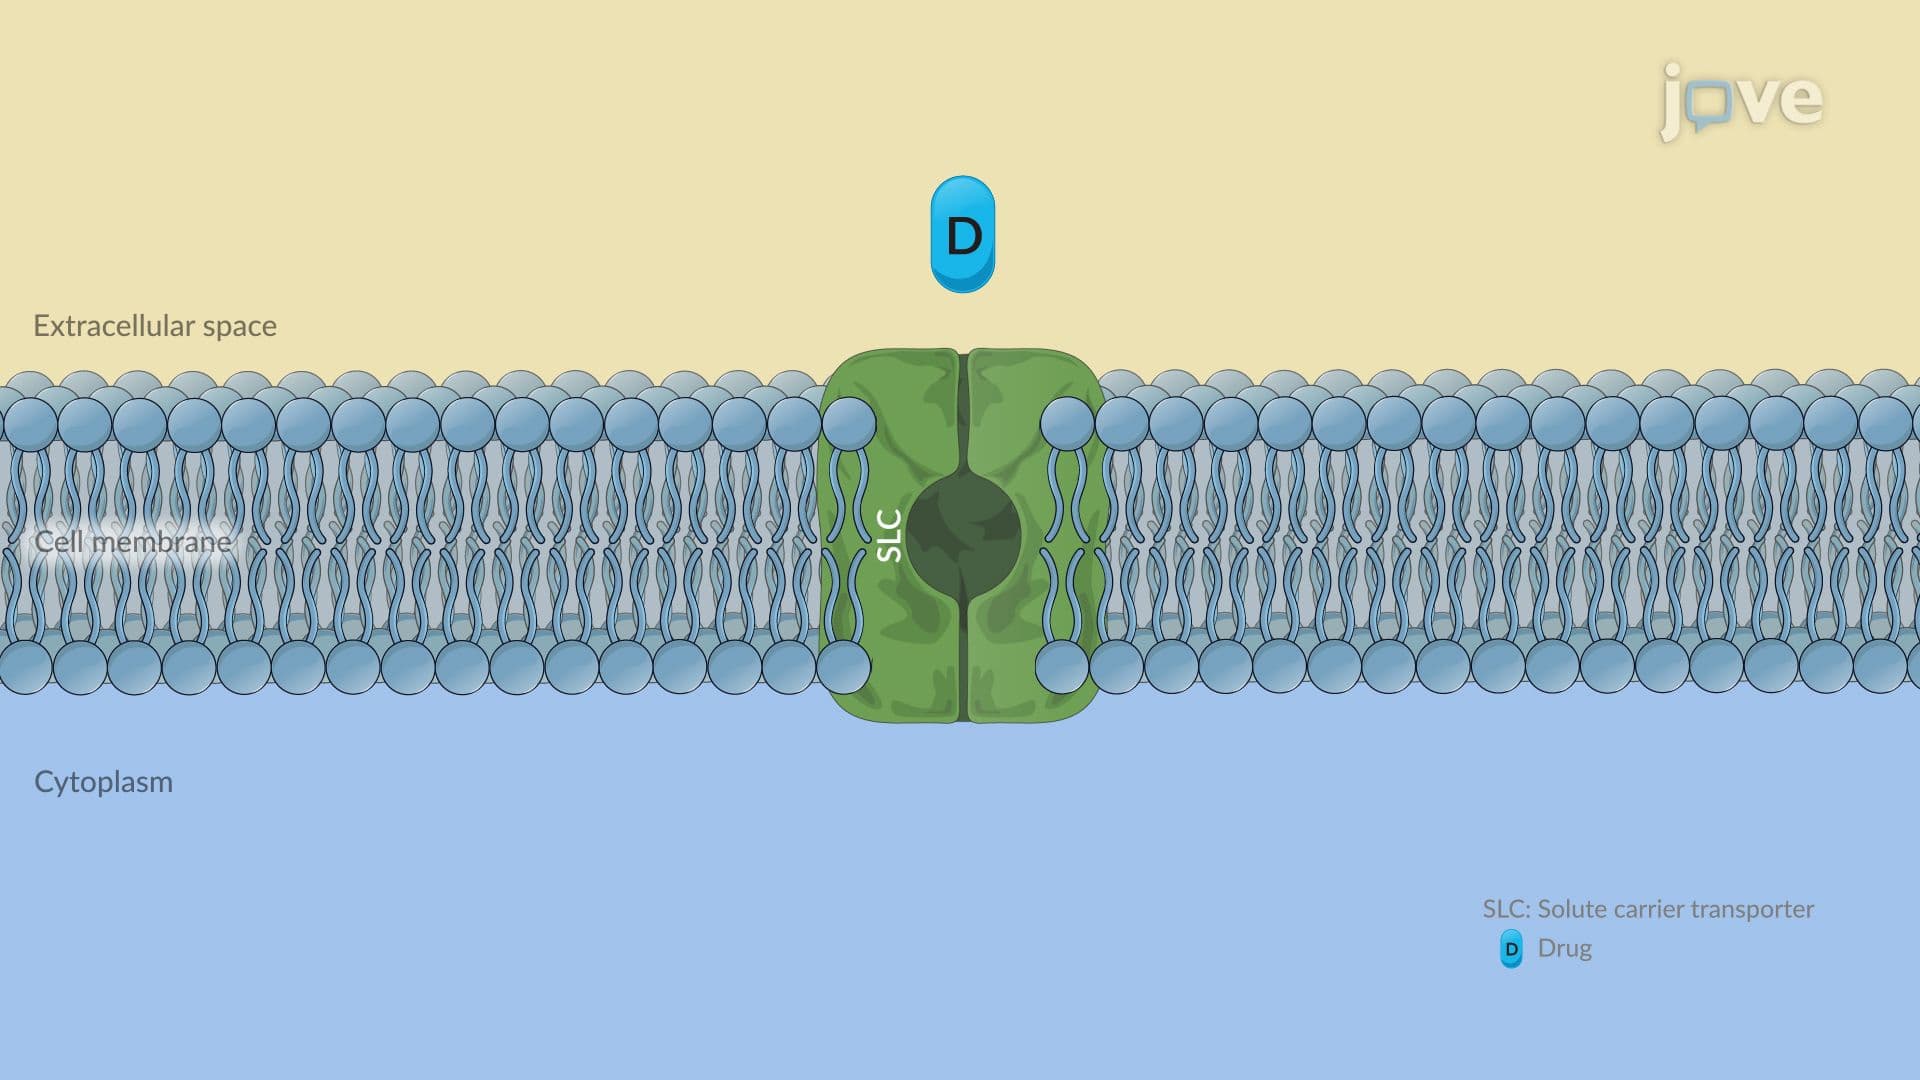
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...
01:31Translation
Translation Produces the Building Blocks of Life
Proteins are...