
Theoretical investigation of loratadine reactivity in order to understand its degradation properties: DFT and MD
Stevan Armaković1, Sanja J Armaković2, Biljana F Abramović2
1University of Novi Sad, Faculty of Sciences, Department of Physics, Trg D. Obradovića 4, 21000, Novi Sad, Serbia. stevan.armakovic@df.uns.ac.rs.

Abstract:
Antihistamines are frequently used pharmaceuticals that treat the symptoms of allergic reactions. Loratadine (LOR) is an active component of the second generation of selective antihistaminic pharmaceutical usually known as Claritin. Frequent usage of this type of pharmaceuticals imposes the need for understanding their fundamental reactive properties. In this study we have theoretically investigated reactive properties of LOR using both density functional theory (DFT) calculations and molecular dynamics (MD) simulations. DFT study is used for collecting information related to the molecule stability, structure, frontier molecular orbitals, quantum molecular descriptors, charge distribution, molecular electrostatic potential surfaces, charge polarization, and local reactivity properties according to average local ionization energy surfaces. Based on these results, N24 atom of pyridine ring and oxygen atom O1 were identified with nucleophilic nature. In order to collect the information necessary for the proposition of degradation compounds we also calculated bond dissociation energies (BDE) for hydrogen abstraction and single acyclic bonds as well. According to BDE, the oxidation is likely to occur in the piperidine and cycloheptane rings. MD simulations were used in order to understand the interactions with water through radial distribution functions (RDF). Based on RDFs the most important interactions with solvent are determined for carbon atom C5, chlorine atom Cl15, and oxygen atom O1. Collected results based on DFT calculations and MD simulations provided information important for suggestion of possible degradation compounds. Covalent and noncovalent interactions between LOR and (•)OH have also been investigated.
More Related Videos
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:13
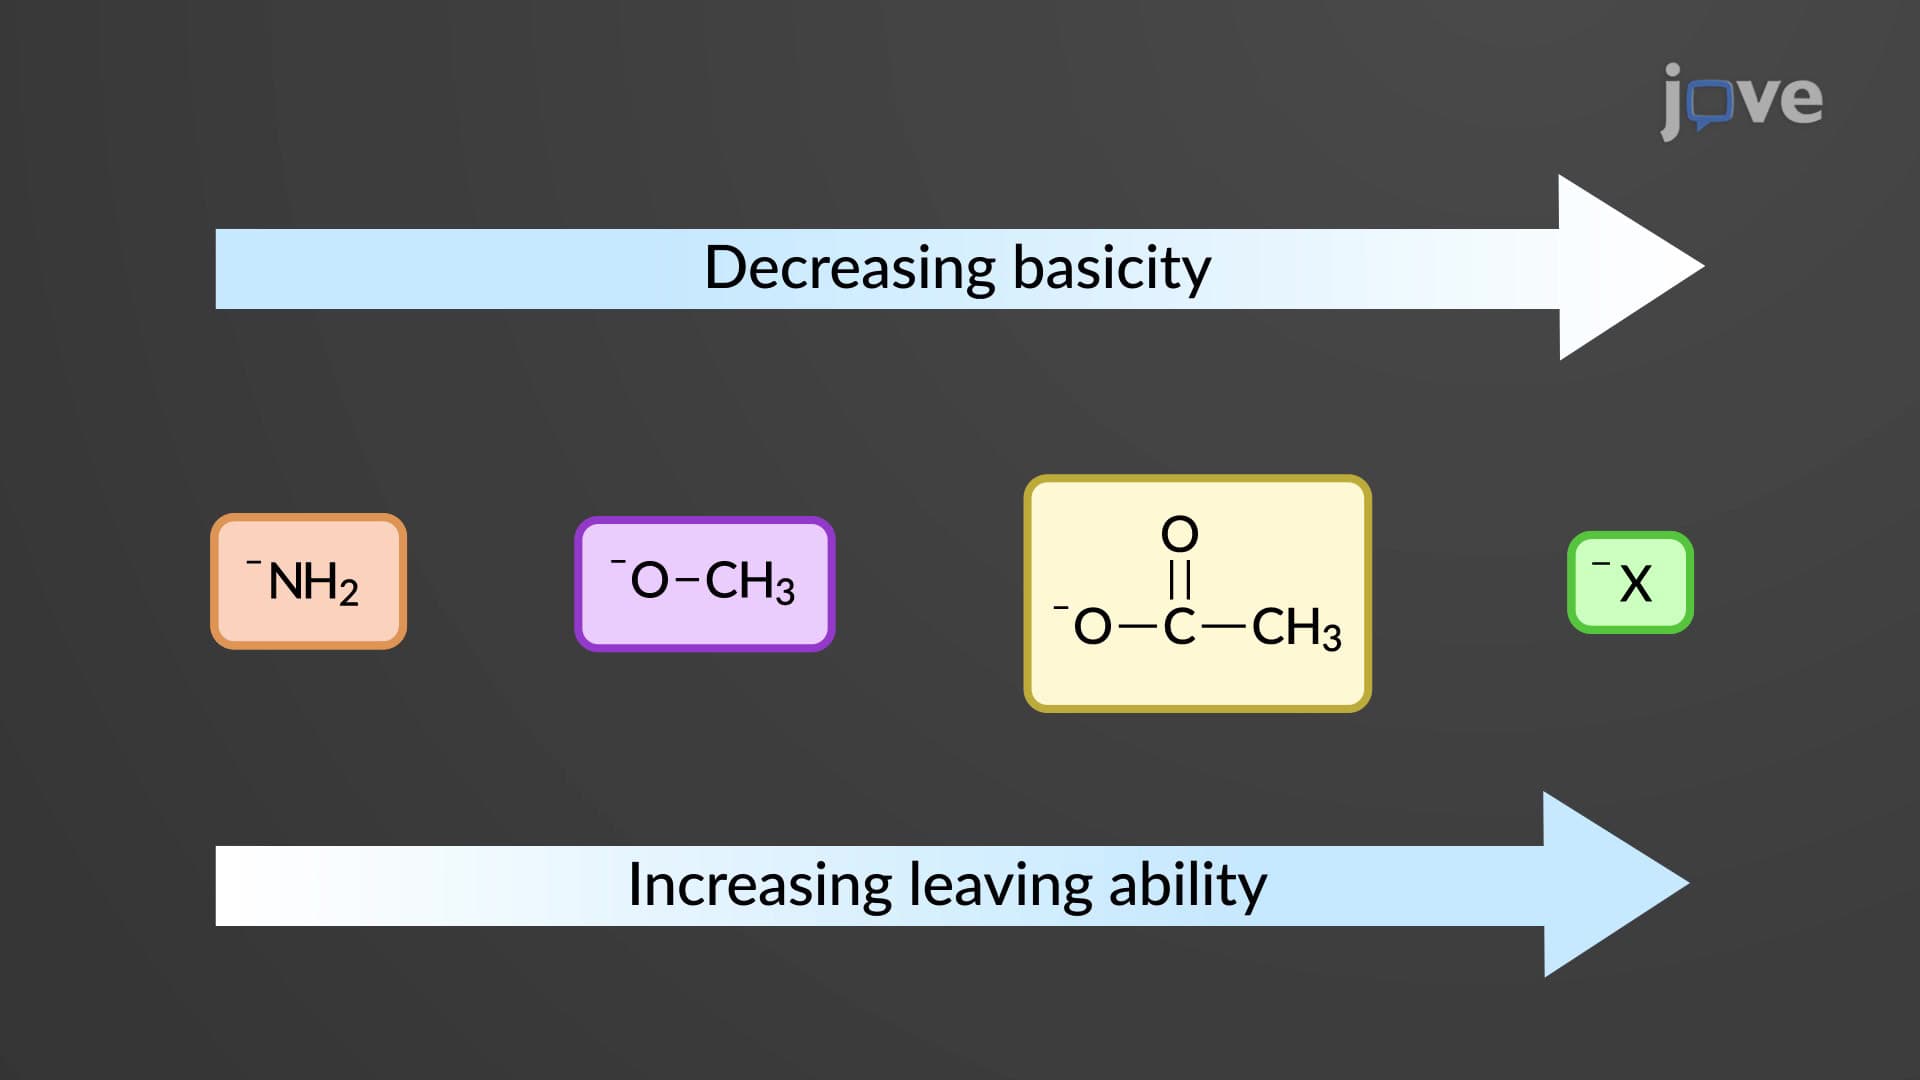
Relative Reactivity of Carboxylic Acid Derivatives
A key factor in assessing the reactivity of the acid derivatives is the basicity of the substituent or the leaving group. The lower the basicity of the leaving group, the higher the reactivity of the derivative. The basicity of the leaving group follows this order:
Halide ions < Acyloxy ions < Alkoxy ions < Amine ions
 01:19
Phase I Reactions: Oxidation of Aliphatic and Aromatic Carbon-Containing Systems
Oxidation reactions are fundamental in aromatic carbon-containing systems. An example is the hydroxylation of phenobarbital, a process that transforms it into...