Related Experiment Video
Updated: Aug 3, 2026

An Intestine/Liver Microphysiological System for Drug Pharmacokinetic and Toxicological Assessment
Published on: December 3, 2020
Azithromycin assay in drug formulations: Validation of a HPTLC method with a quadratic polynomial calibration model
A Bouklouze1, M Kharbach2, Y Cherrah1
1Pharmaceutical and toxicological analysis research team, laboratory of pharmacology and toxicology, faculty of medicine and pharmacy, avenue Mohammed Belarbi Elalaoui, BP, 6203 Rabat, Morocco.

Abstract:
Many different assaying high performance thin layer chromatography (HPTLC) methods have been developed and validated in order to be used in routine analysis in different analytical fields. Validation often starts by the evaluation of the linearity of the calibration curve. Frequently, if the correlation coefficient is close to one, the linear calibration curve model is considered to be proper to predict the unknown concentration in the sample. But is this simple model effective to assess the behavior of the response of an HPTLC method as a function of concentration. To answer this question, a method for the determination of azithromycin by HPTLC has been developed and validated following both the classical approach and that based on the accuracy profile. Silica gel plates with fluorescence indicator F254 and chloroform - ethanol - 25% ammonia 6:14:0.2 (v/v/v) as mobile phase were used. Analysis was carried out in reflectance mode at 483nm. The RF of azithromycin was 0.53. The validation based on the classical approach, shows that the behavior is not linear, even though r2=0.999 because the lack of fit test is significant (P<0.05). Validation based on the accuracy profile approach considering both the straight line and the quadratic regression model, show that the former results is a β-expectation tolerance interval outside the acceptance limits, while with the latter, this interval is within the limits of ±5% acceptability for a range which extends from 0.2 to 1.0μg/zone. With the quadratic model, the method showed to be precise and accurate.
More Related Videos
 08:54
Validated LC-MS/MS Panel for Quantifying 11 Drug-Resistant TB Medications in Small Hair Samples
Published on: May 19, 2020
 07:29
HPLC Coupled with Chemical Fingerprinting for Multi-Pattern Recognition for Identifying the Authenticity of Clematidis Armandii Caulis
Published on: November 11, 2022
Related Concept Videos
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods
 01:13
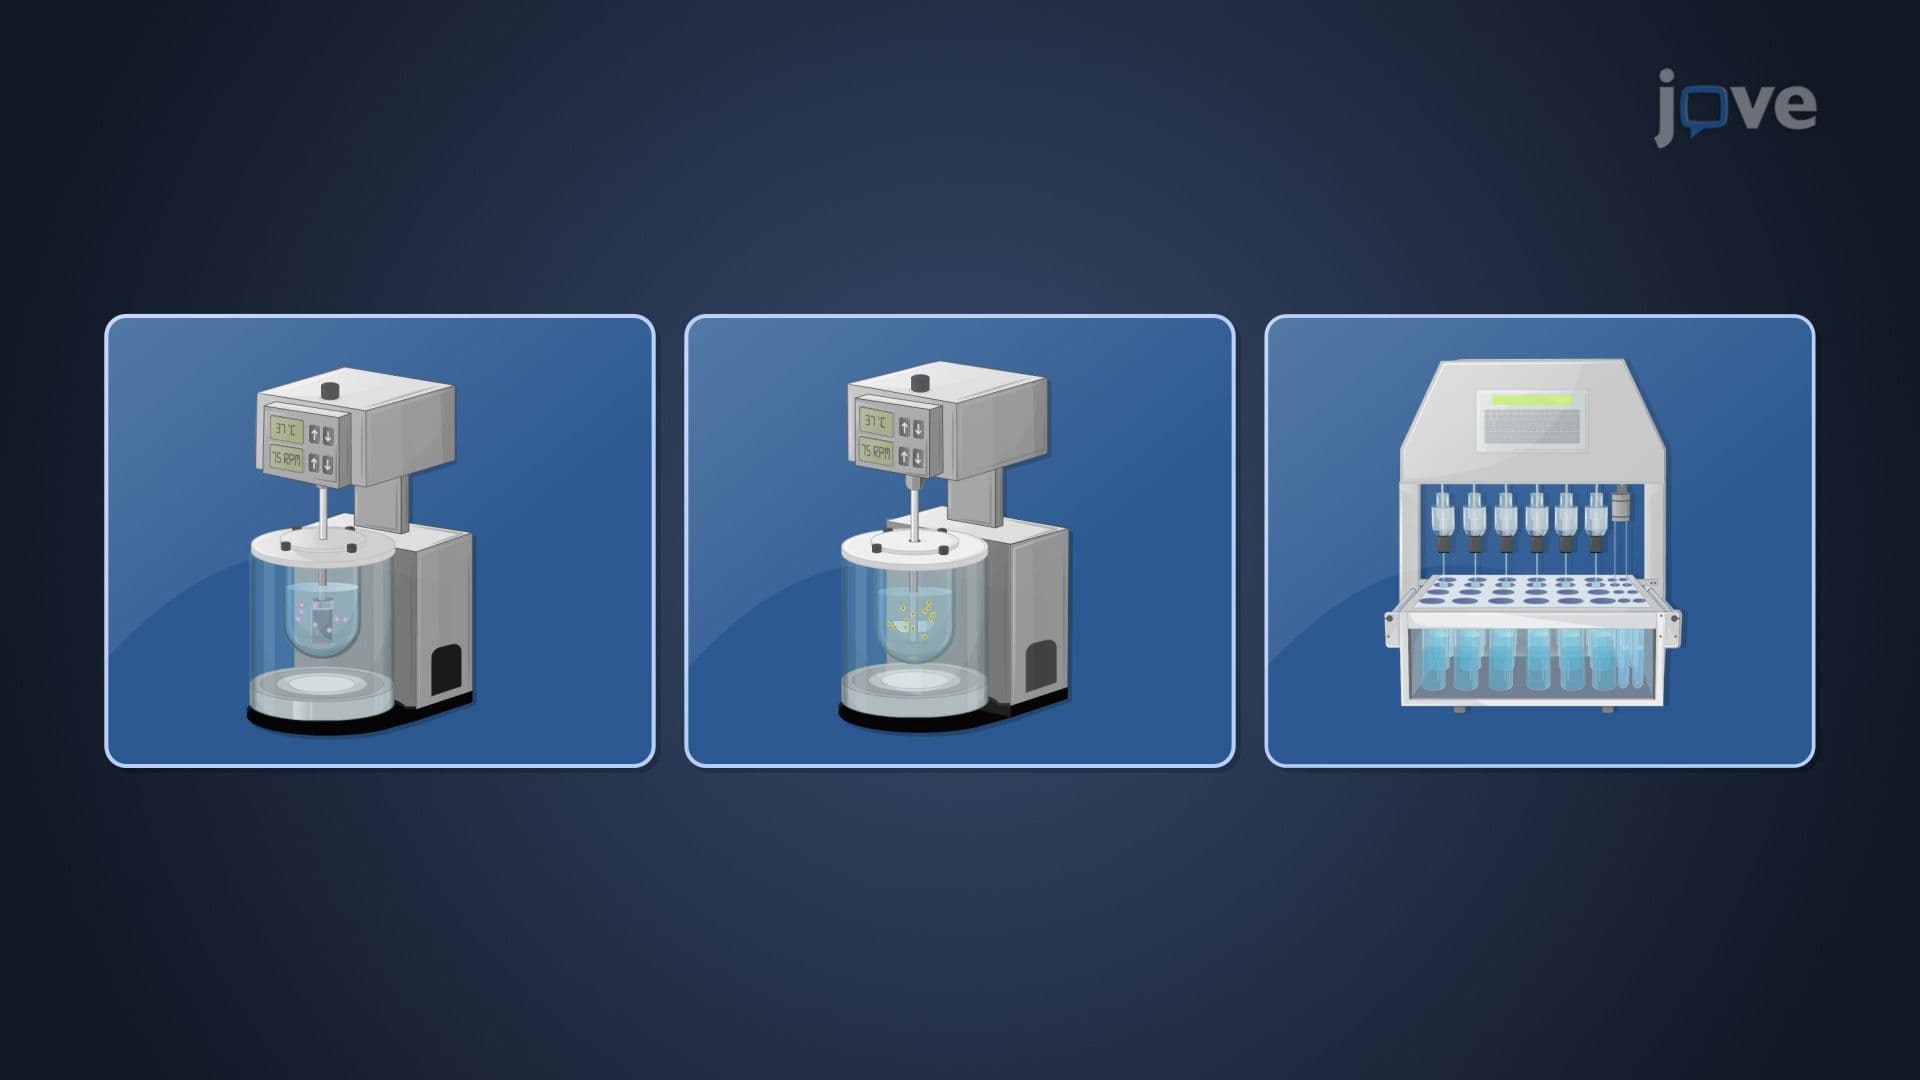
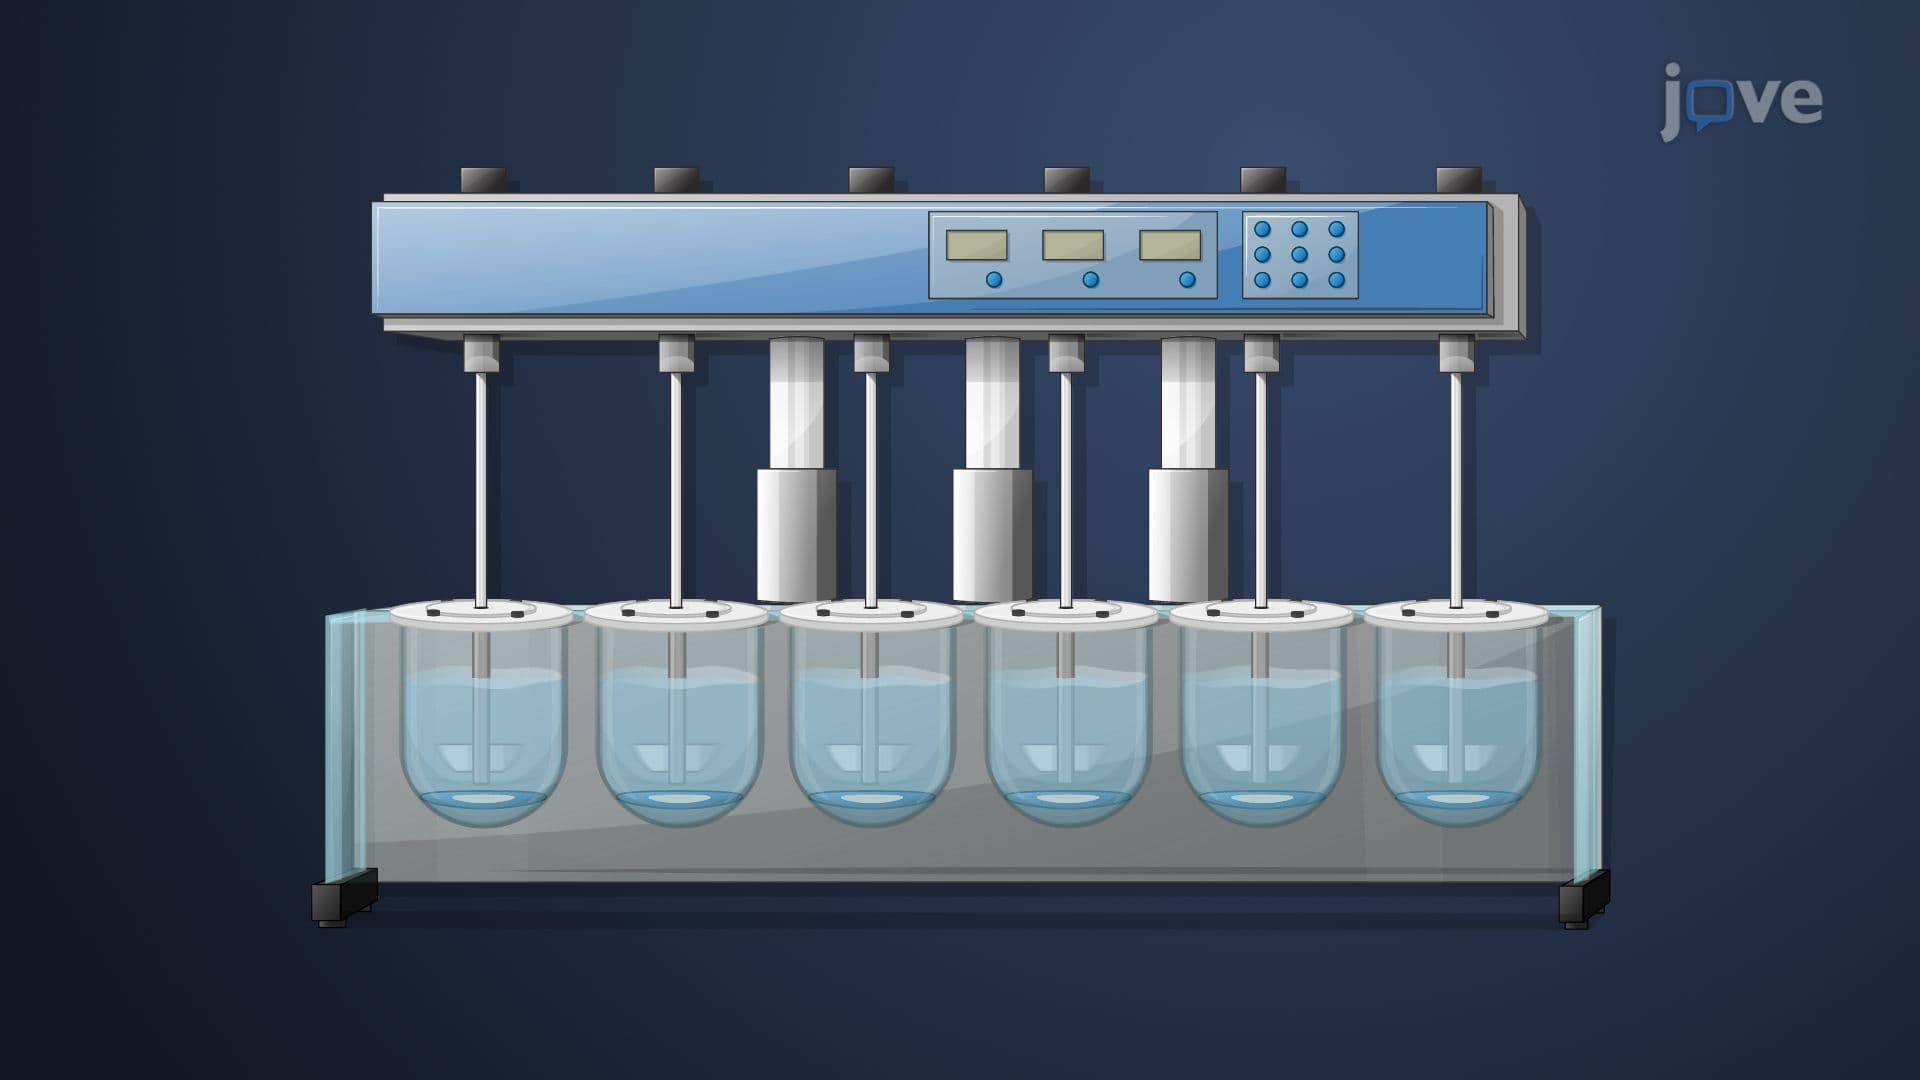
In Vitro Drug Dissolution: Compendial Testing Models I
 01:09
In Vitro Drug Dissolution: Compendial Testing Models II
 01:14
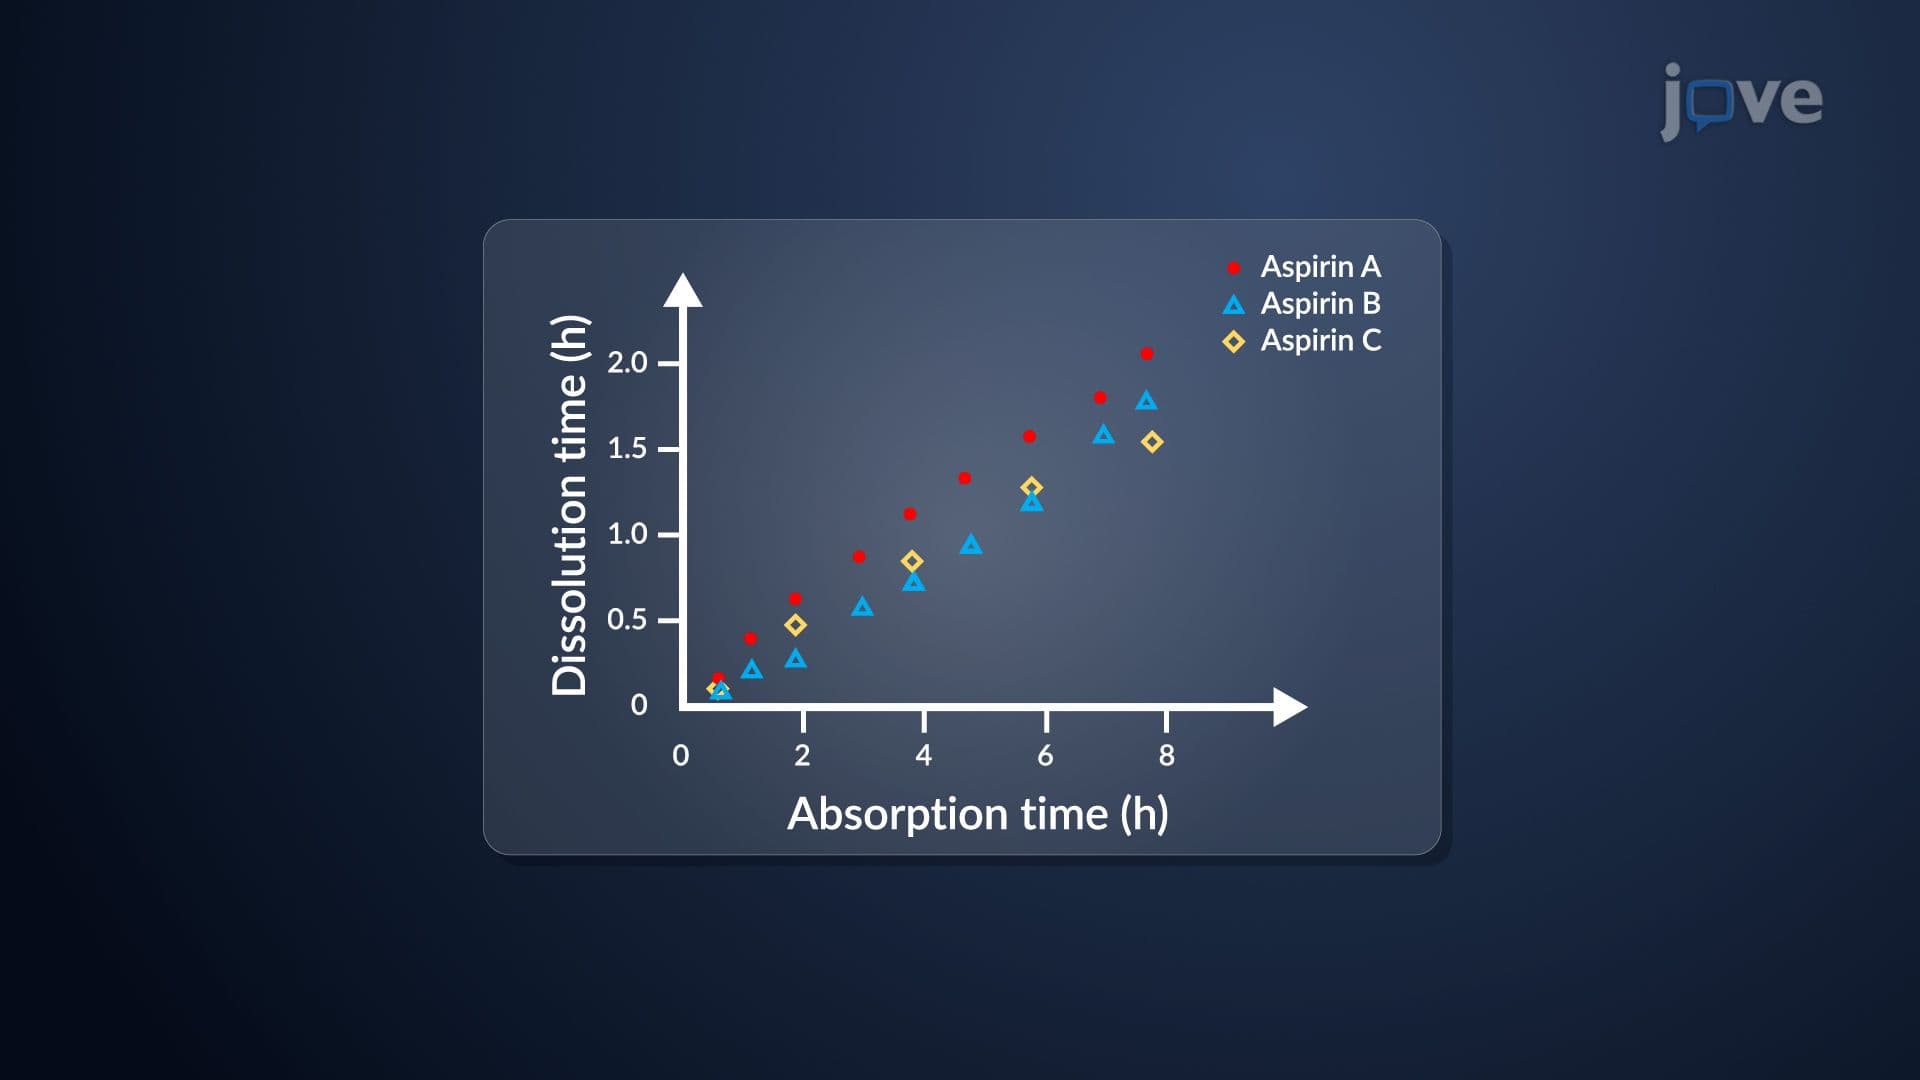
Drug Dissolution: Requirements and Profile Comparison
 01:20
Drug Product Performance: In Vitro–In Vivo Correlation
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model