Related Experiment Video
Updated: Mar 10, 2026

Development of Heterogeneous Enantioselective Catalysts using Chiral Metal-Organic Frameworks MOFs
Published on: January 17, 2020
Catalytic Kinetic Resolution of Saturated N-Heterocycles by Enantioselective Amidation with Chiral Hydroxamic Acids
Imants Kreituss1, Jeffrey W Bode1
1Laboratorium für Organische Chemie, Department of Chemistry and Applied Biosciences, ETH-Zürich , 8093 Zürich, Switzerland.

Abstract:
The preparation of enantioenriched chiral compounds by kinetic resolution dates back to the laboratories of Louis Pasteur in the middle of the 19th century. Unlike asymmetric synthesis, this process can always deliver enantiopure material (ee > 99%) if the reactions are allowed to proceed to sufficient conversion and the selectivity of the process is not unity (s > 1). One of the most appealing and practical variants is acylative kinetic resolution, which affords easily separable reaction products, and several highly efficient enzymatic and small molecule catalysts are available. Unfortunately, this method is applicable to limited substrate classes such as alcohols and primary benzylamines. This Account focuses on our work in catalytic acylative kinetic resolution of saturated N-heterocycles, a class of molecules that has been notoriously difficult to access via asymmetric synthesis. We document the development of hydroxamic acids as suitable catalysts for enantioselective acylation of amines through relay catalysis. Alongside catalyst optimization and reaction development, we present mechanistic studies and theoretical calculation accounting for the origins of selectivity and revealing the concerted nature of many amide-bond forming reactions. Immobilization of the hydroxamic acid to form a polymer supported reagent allows simplification of the experimental setup, improvement in product purification, and extension of the substrate scope. The kinetic resolutions are operationally straight forward: reactions proceed at room temperature and open to air conditions, without generation of difficult-to-remove side products. This was utilized to achieve decagram scale resolution of antimalarial drug mefloquine to prepare more than 50 g of (+)-erythro-meflqouine (er > 99:1) from the racemate. The immobilized quasienantiomeric acyl hydroxamic acid reagents were also exploited for a rare practical implementation of parallel kinetic resolution that affords both enantiomers of the amine products in high enantiopurity. The success of this process relied on identification of two cleavable acyl groups alongside implementation of flow-chemistry techniques to ensure reusability of the resolving agents. The work discussed in this Account has laid foundations for new catalyst design as well as development of desymmetrization and dynamic kinetic resolution processes. In the meantime, as all the requisite reagents are commercially available, we hope that hydroxamic acid promoted acylative kinetic resolution will become a method of choice for preparation of saturated N-heterocycles in enantiopure form.
More Related Videos
 07:06
A Microwave-Assisted Direct Heteroarylation of Ketones Using Transition Metal Catalysis
Published on: February 16, 2020
 06:31
Highly Stereoselective Synthesis of 1,6-Ketoesters Mediated by Ionic Liquids: A Three-component Reaction Enabling Rapid Access to a New Class of Low Molecular Weight Gelators
Published on: November 27, 2015
Related Concept Videos
 02:17
Reduction of Alkenes: Asymmetric Catalytic Hydrogenation
The metal catalyst used can be either heterogeneous or homogeneous. When hydrogenation of an alkene generates a chiral center, a pair of enantiomeric products is expected to form. However, an enantiomeric excess of one of the products can be facilitated using an enantioselective reaction or an...
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 02:30
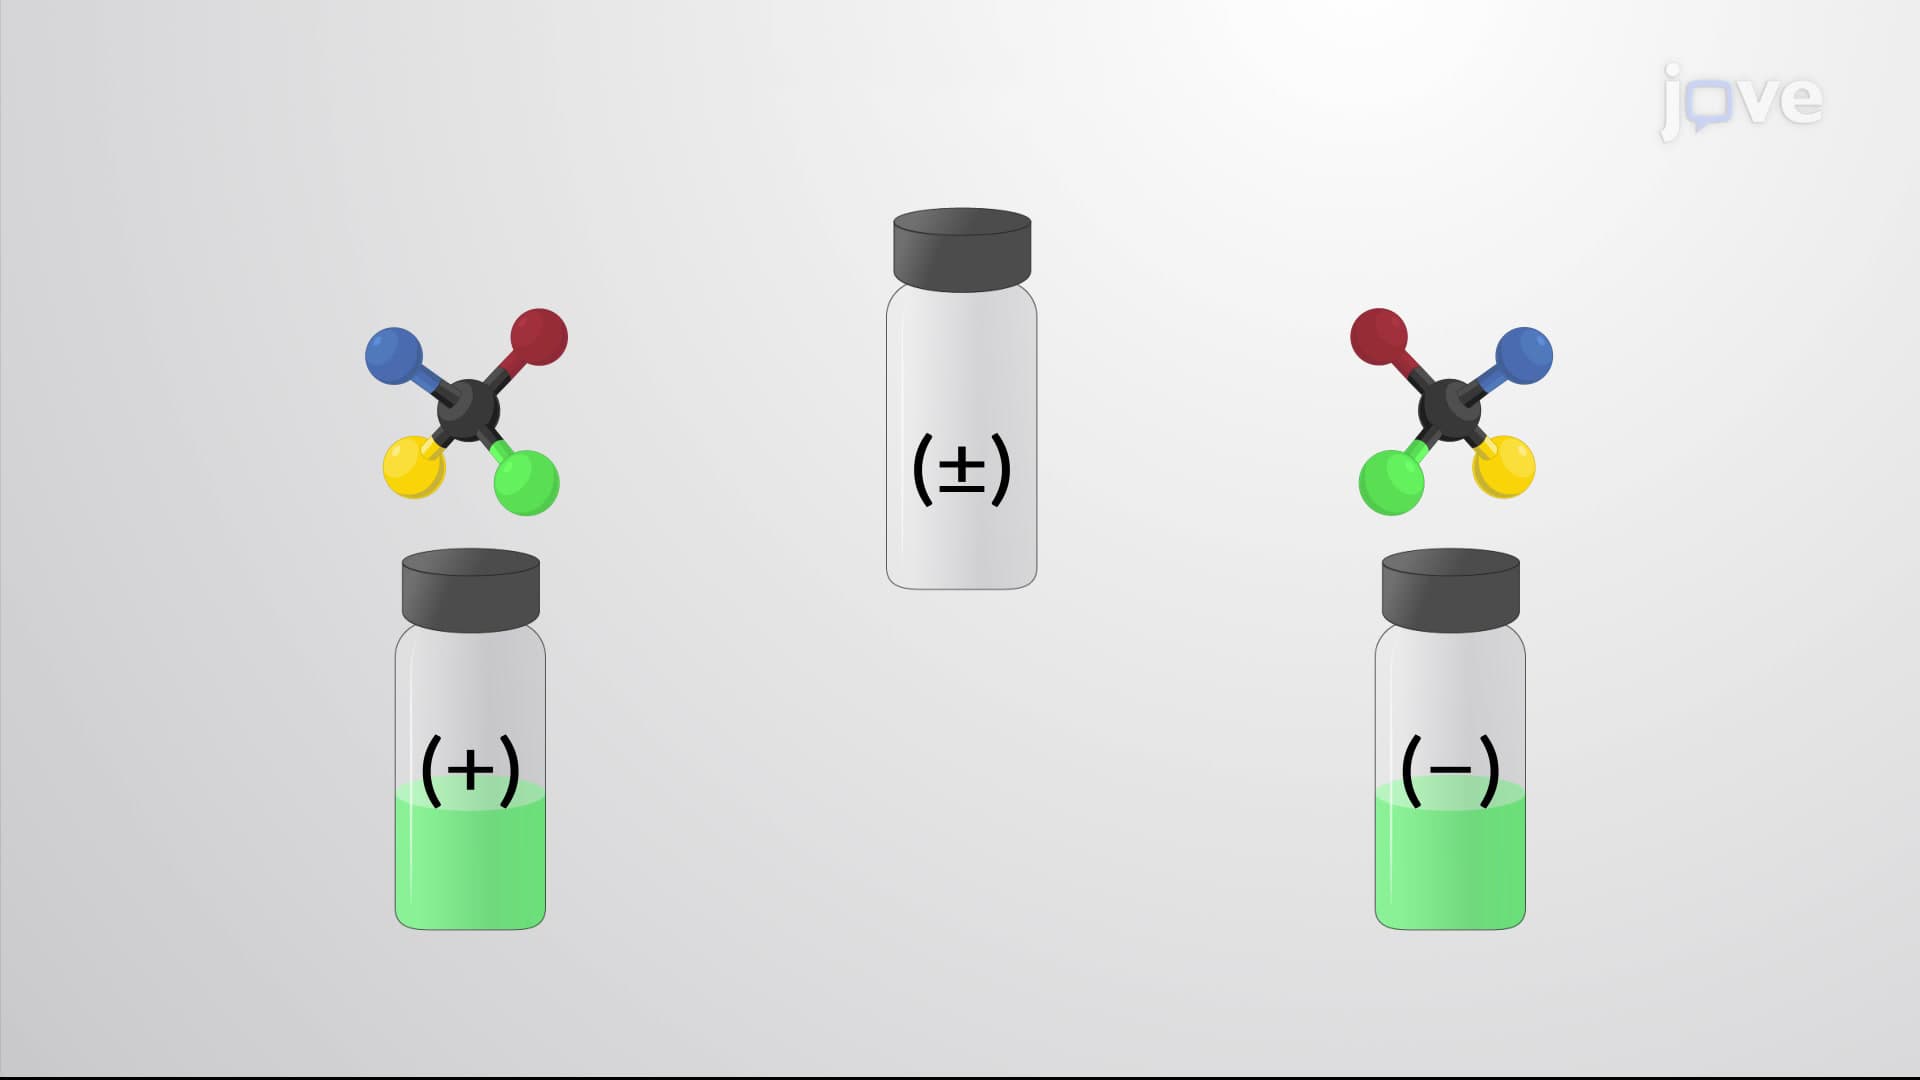
Racemic Mixtures and the Resolution of Enantiomers
 01:12
Stereochemical Effects of Enolization
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 02:05
Prochirality