Independent test assessment using the extreme value distribution theory
Related Experiment Video
Updated: Mar 10, 2026
Measuring the Subjective Value of Risky and Ambiguous Options using Experimental Economics and Functional MRI Methods
Published on: September 19, 2012
Marcio Almeida1, Lucy Blondell1, Juan M Peralta1
1South Texas Diabetes and Obesity Institute, University of Texas at Brownsville, 2700 East Jackson Street, Brownsville, TX 78520 USA.

Abstract:
The new generation of whole genome sequencing platforms offers great possibilities and challenges for dissecting the genetic basis of complex traits. With a very high number of sequence variants, a naïve multiple hypothesis threshold correction hinders the identification of reliable associations by the overreduction of statistical power. In this report, we examine 2 alternative approaches to improve the statistical power of a whole genome association study to detect reliable genetic associations. The approaches were tested using the Genetic Analysis Workshop 19 (GAW19) whole genome sequencing data. The first tested method estimates the real number of effective independent tests actually being performed in whole genome association project by the use of an extreme value distribution and a set of phenotype simulations. Given the familiar nature of the GAW19 data and the finite number of pedigree founders in the sample, the number of correlations between genotypes is greater than in a set of unrelated samples. Using our procedure, we estimate that the effective number represents only 15 % of the total number of independent tests performed. However, even using this corrected significance threshold, no genome-wide significant association could be detected for systolic and diastolic blood pressure traits. The second approach implements a biological relevance-driven hypothesis tested by exploiting prior computational predictions on the effect of nonsynonymous genetic variants detected in a whole genome sequencing association study. This guided testing approach was able to identify 2 promising single-nucleotide polymorphisms (SNPs), 1 for each trait, targeting biologically relevant genes that could help shed light on the genesis of the human hypertension. The first gene, PFH14, associated with systolic blood pressure, interacts directly with genes involved in calcium-channel formation and the second gene, MAP4, encodes a microtubule-associated protein and had already been detected by previous genome-wide association study experiments conducted in an Asian population. Our results highlight the necessity of the development of alternative approached to improve the efficiency on the detection of reasonable candidate associations in whole genome sequencing studies.

Related Concept Videos
 01:21
Introduction to Test of Independence
The test statistic for a test of independence is similar to that of a goodness-of-fit test:
 01:29
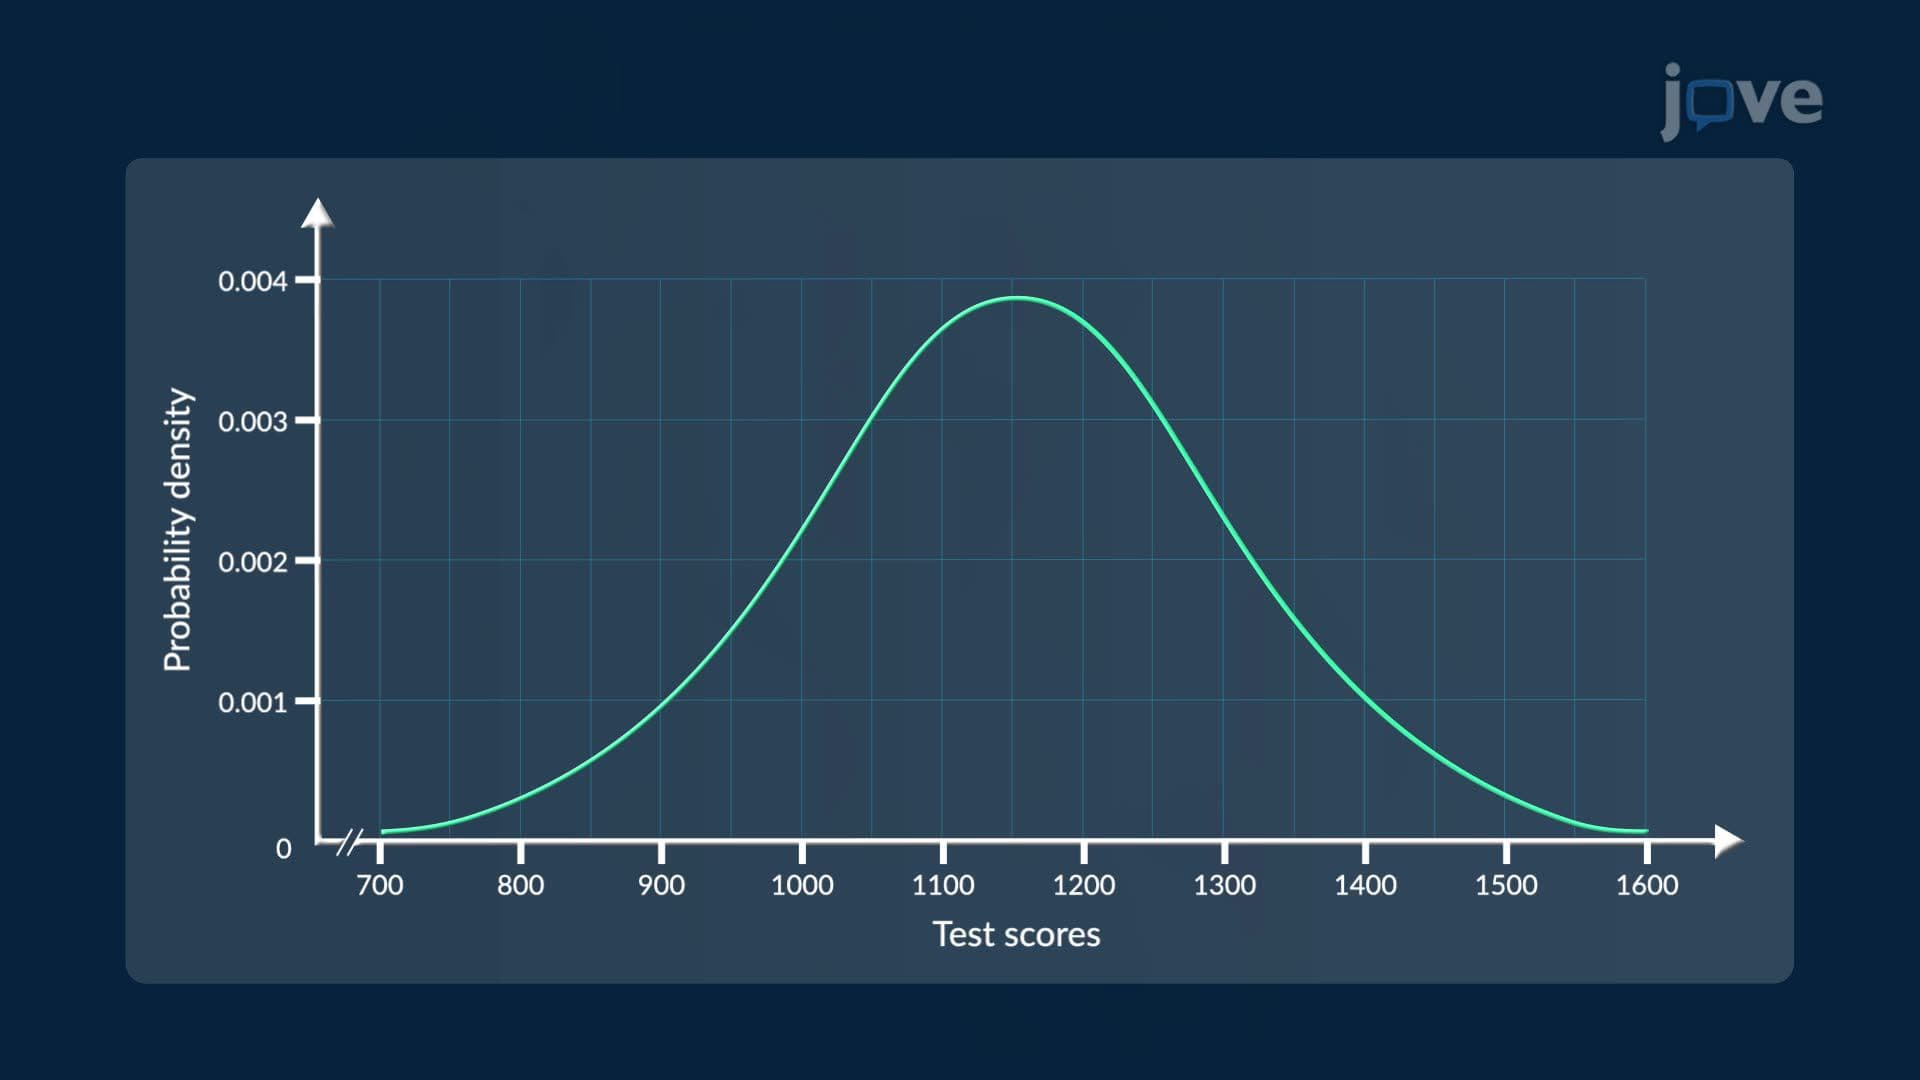
Introduction to Normal Distributions
 01:19
F Distribution
 01:32
Probability Distributions
A discrete probability distribution is a probability distribution of discrete random variables. It can be categorized into binomial probability distribution and Poisson...
 01:16
The Anderson-Darling Test
 01:19
Testing a Claim about Standard Deviation
The hypothesis testing for the claim of population standard deviation (or variance) requires the data and samples to be random and unbiased. The population distribution also must be normal. There is no specific requirement on the sample size as the estimation is based on the chi-square distribution.
As a first step, the hypothesis (null and alternative) concerning the claim about...