Exogenous H2S regulates endoplasmic reticulum-mitochondria cross-talk to inhibit apoptotic pathways in STZ-induced type I diabetes
Related Experiment Video
Updated: Mar 9, 2026
High-resolution Respirometry to Measure Mitochondrial Function of Intact Beta Cells in the Presence of Natural Compounds
Published on: January 23, 2018
Fan Yang1, Xiangjing Yu1, Ting Li1
1Department of Pathophysiology, Harbin Medical University, Harbin, China; and.

Abstract:
The upregulation of reactive oxygen species (ROS) is a primary cause of cardiomyocyte apoptosis in diabetes cardiomyopathy (DCM). Mitofusin-2 (Mfn-2) is a key protein that bridges the mitochondria and endoplasmic reticulum (ER). Hydrogen sulfide (H2S)-mediated cardioprotection is related to antioxidant effects. The present study demonstrated that H2S inhibited the interaction between the ER and mitochondrial apoptotic pathway. This study investigated cardiac function, ultrastructural changes in the ER and mitochondria, apoptotic rate using TUNEL, and the expression of ER stress-associated proteins and mitochondrial apoptotic proteins in cardiac tissues in STZ-induced type I diabetic rats treated with or without NaHS (donor of H2S). Mitochondria of cardiac tissues were isolated, and MPTP opening and cytochrome c (cyt C) and Mfn-2 expression were also detected. Our data showed that hyperglycemia decreased the cardiac function by ultrasound cardiogram, and the administration of exogenous H2S ameliorated these changes. We demonstrated that the expression of ER stress sensors and apoptotic rates were elevated in cardiac tissue of DCM and cultured H9C2 cells, but the expression of these proteins was reduced following exogenous H2S treatment. The expression of mitochondrial apoptotic proteins, cyt C, and mPTP opening was decreased following treatment with exogenous H2S. In our experiment, the expression and immunofluorescence of Mfn-2 were both decreased after transfection with Mfn-2-siRNA. Hyperglycemia stimulated ER interactions and mitochondrial apoptotic pathways, which were inhibited by exogenous H2S treatment through the regulation of Mfn-2 expression.
Insights
Hydrogen sulfide (H2S) protects the heart in diabetic cardiomyopathy by reducing oxidative stress and inhibiting ER-mitochondrial apoptosis. H2S treatment, using NaHS, improved cardiac function and normalized key protein expressions in diabetic rats.
Area of Science:
- Cardiovascular Research
- Molecular Biology
- Endocrinology
Background:
- Diabetes cardiomyopathy (DCM) involves cardiomyocyte apoptosis driven by reactive oxygen species (ROS).
- Mitofusin-2 (Mfn-2) links mitochondria and endoplasmic reticulum (ER), crucial for apoptosis.
- Hydrogen sulfide (H2S) exhibits cardioprotective effects attributed to its antioxidant properties.
Purpose of the Study:
- To investigate the protective role of exogenous H2S against cardiac dysfunction in STZ-induced diabetic rats.
- To elucidate the mechanism of H2S in regulating ER-mitochondrial interactions and apoptosis in DCM.
Main Methods:
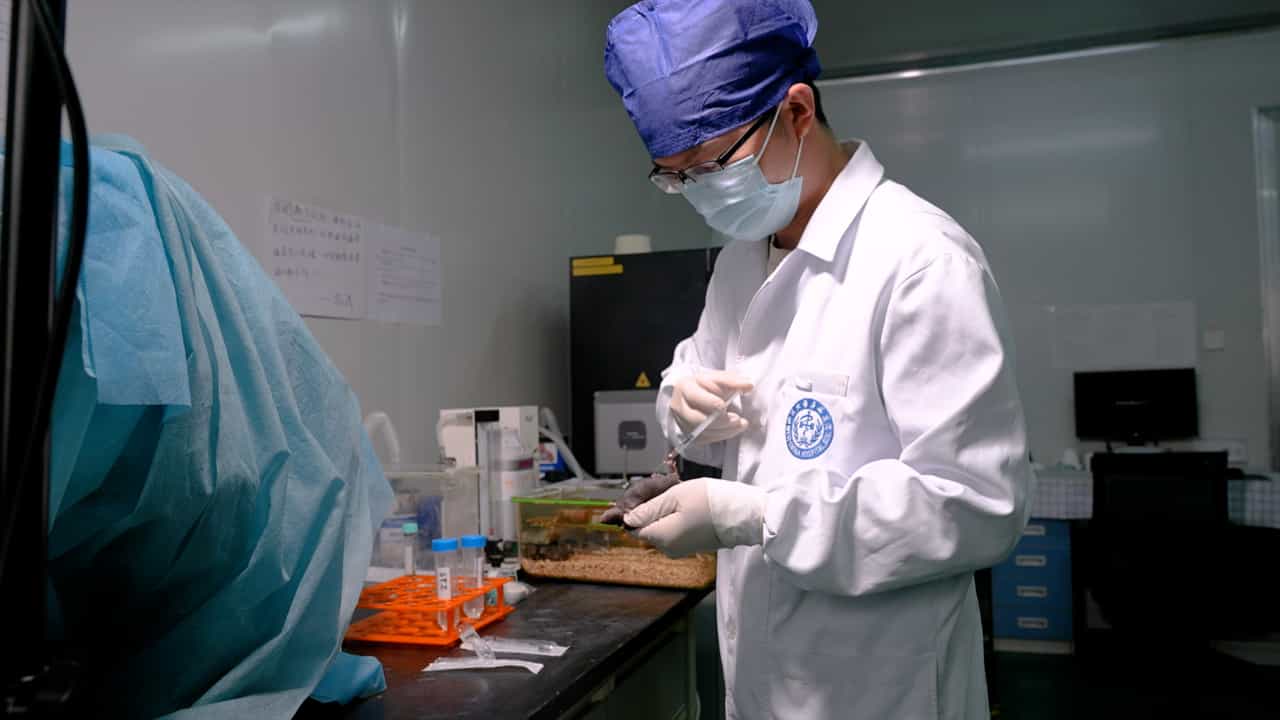
- Induction of type I diabetes in rats using streptozotocin (STZ).
- Administration of sodium hydrosulfide (NaHS) as an H2S donor.
- Assessment of cardiac function, ultrastructural changes, apoptosis (TUNEL), ER stress markers, mitochondrial proteins (cytochrome c, Mfn-2), and mitochondrial permeability transition pore (MPTP) opening.
Main Results:
- Hyperglycemia impaired cardiac function and increased cardiomyocyte apoptosis, ER stress, and Mfn-2 downregulation.
- Exogenous H2S administration ameliorated cardiac dysfunction and reduced apoptosis and ER stress markers.
- H2S treatment decreased mitochondrial apoptotic protein release and MPTP opening, while restoring Mfn-2 expression.
Conclusions:
- Exogenous H2S protects against DCM by inhibiting the ER-mitochondrial apoptotic pathway.
- H2S exerts its cardioprotective effects through the regulation of Mfn-2 expression and mitigation of oxidative stress.