Related Experiment Video
Updated: Mar 9, 2026

Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
Molecular Simulations Identify Binding Poses and Approximate Affinities of Stapled α-Helical Peptides to MDM2 and
Joseph A Morrone1, Alberto Perez1, Qiaolin Deng2
1Laufer Center for Physical and Quantitative Biology, Stony Brook University , Stony Brook, New York 11794, United States.

Abstract:
Traditionally, computing the binding affinities of proteins to even relatively small and rigid ligands by free-energy methods has been challenging due to large computational costs and significant errors. Here, we apply a new molecular simulation acceleration method called MELD (Modeling by Employing Limited Data) to study the binding of stapled α-helical peptides to the MDM2 and MDMX proteins. We employ free-energy-based molecular dynamics simulations (MELD-MD) to identify binding poses and calculate binding affinities. Even though stapled peptides are larger and more complex than most protein ligands, the MELD-MD simulations can identify relevant binding poses and compute relative binding affinities. MELD-MD appears to be a promising method for computing the binding properties of peptide ligands with proteins.
More Related Videos
 09:17
Structure-Based Simulation and Sampling of Transcription Factor Protein Movements along DNA from Atomic-Scale Stepping to Coarse-Grained Diffusion
Published on: March 1, 2022
 05:56
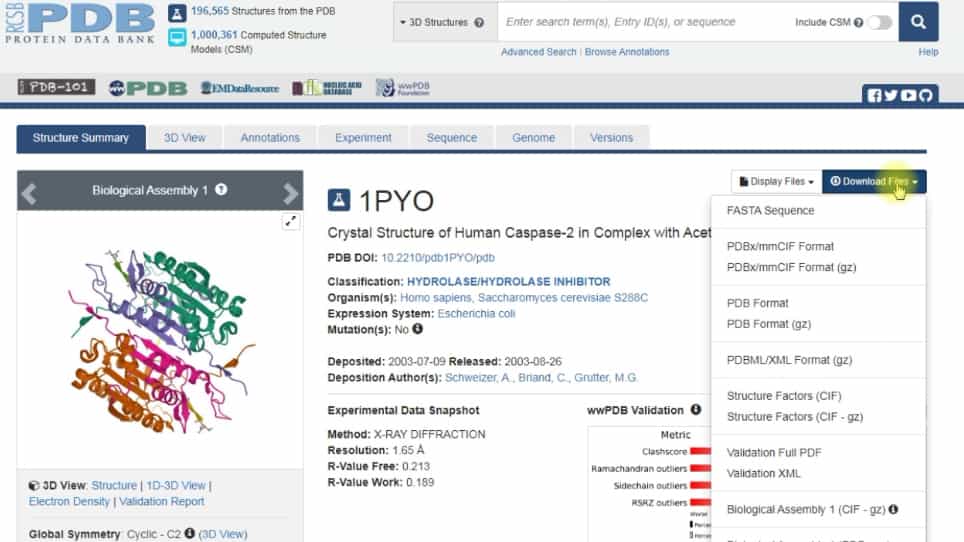
Exploring Caspase Mutations and Post-Translational Modification by Molecular Modeling Approaches
Published on: October 13, 2022