Related Experiment Video
Updated: Mar 8, 2026
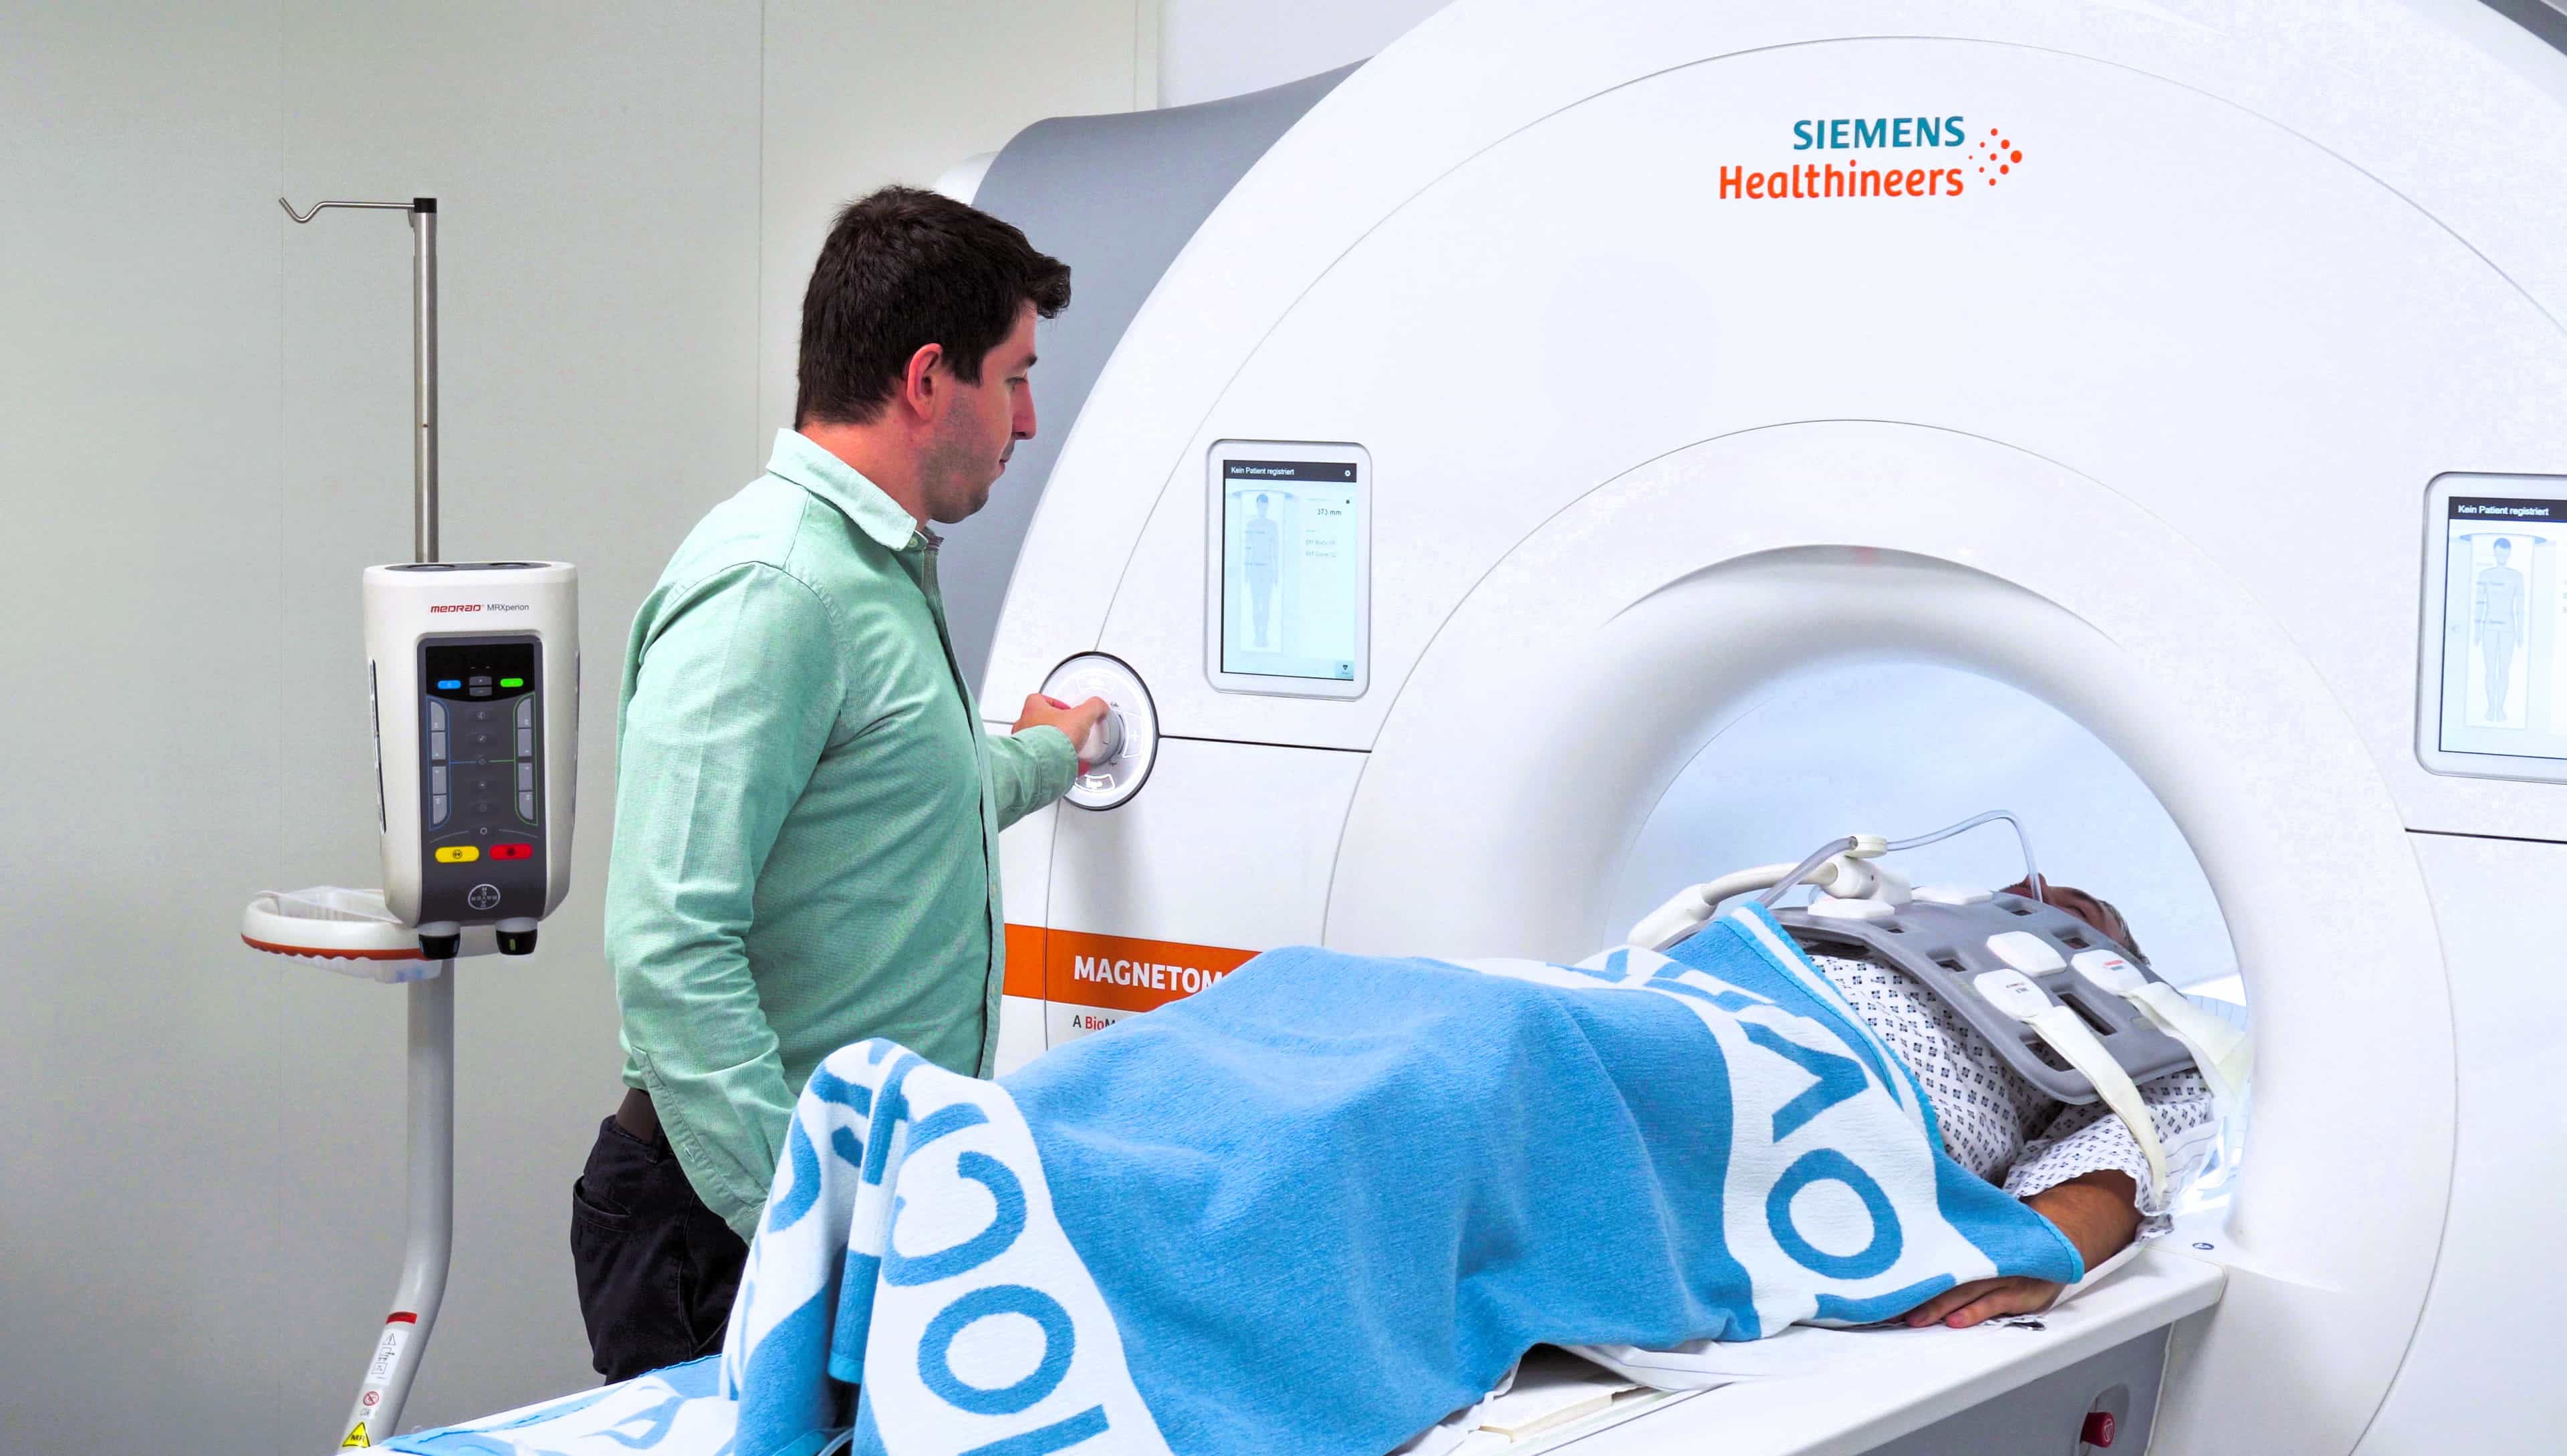
Three-Dimensional Phase Resolved Functional Lung Magnetic Resonance Imaging
Published on: June 21, 2024
Quantitative computed tomography analysis of the airways in patients with cystic fibrosis using automated software:
Marcel Koenigkam Santos1, Danilo Lemos Cruvinel2, Marcelo Bezerra de Menezes3
1PhD, MD, Radiologist, Collaborating Professor at the Center for Imaging Sciences and Medical Physics of the Hospital das Clínicas da Faculdade de Medicina de Ribeirão Preto da Universidade de São Paulo (HCFMRP-USP), Ribeirão Preto, SP, Brazil.

Objective:
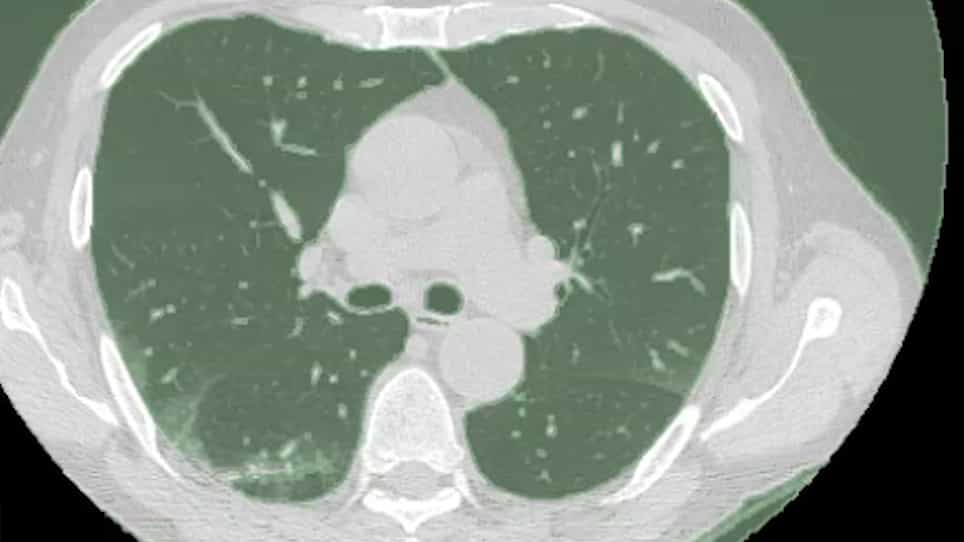
To perform a quantitative analysis of the airways using automated software, in computed tomography images of patients with cystic fibrosis, correlating the results with spirometric findings.
Materials And Methods:
Thirty-four patients with cystic fibrosis were studied-20 males and 14 females; mean age 18 ± 9 years-divided into two groups according to the spirometry findings: group I (n = 21), without severe airflow obstruction (forced expiratory volume in first second [FEV1] > 50% predicted), and group II (n = 13), with severe obstruction (FEV1 ≤ 50% predicted). The following tracheobronchial tree parameters were obtained automatically: bronchial diameter, area, thickness, and wall attenuation.
Results:
On average, 52 bronchi per patient were studied. The number of bronchi analyzed was higher in group II. The correlation with spirometry findings, especially between the relative wall thickness of third to eighth bronchial generation and predicted FEV1, was better in group I.
Conclusion:
Quantitative analysis of the airways by computed tomography can be useful for assessing disease severity in cystic fibrosis patients. In patients with severe airflow obstruction, the number of bronchi studied by the method is higher, indicating more bronchiectasis. In patients without severe obstruction, the relative bronchial wall thickness showed a good correlation with the predicted FEV1.
Related Concept Videos
 01:27
Chronic Obstructive Pulmonary Disease-IV: Assessement and Diagnostic Studies
Medical History
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:25
Pulmonary Function Tests
Pulmonary Function Tests are crucial diagnostic tools for assessing respiratory function, particularly in patients with chronic respiratory disorders. They comprehensively evaluate lung volumes, ventilatory function, breathing mechanics, diffusion, and gas exchange. These tests help diagnose pulmonary diseases and play a significant role in monitoring disease progression, evaluating disability, and assessing response to therapy.
PFTs involve using a spirometer, a...