Related Experiment Video
Updated: Mar 3, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Multi-state nonadiabatic deactivation mechanism of coumarin revealed by ab initio on-the-fly trajectory surface
Yanzhen Gan1, Ling Yue, Xugeng Guo
1State Key Laboratory of Physical Chemistry of Solid Surfaces and Fujian Province Key Lab of Theoretical and Computational Chemistry, College of Chemistry and Chemical Engineering, Xiamen University, Xiamen 361005, P. R. China. zxcao@xmu.edu.cn.

Abstract:
An on-the-fly trajectory surface hopping dynamic simulation has been performed for revealing the multi-state nonadiabatic deactivation mechanism of coumarin. The mechanism involves three adiabatic excited states, S3(ππ*Lb), S2(nπ*, ππ*La) and S1(ππ*La, nπ*), and the ground state S0 at the four state-averaged complete active space self-consistent field, SA4-CASSCF(12,10)/6-31G* level of theory. Upon photoexcitation to the third excited state S3(ππ*Lb) in the Franck-Condon region, 80% sampling trajectories decay to the dark S2(nπ*) state within an average of 5 fs via the conical intersection S3(ππ*Lb)/S2(nπ*), while 20% decay to the S2(ππ*La) state within an average of 11 fs via the conical intersection S3(ππ*Lb)/S2(ππ*La). Then, sampling trajectories via S2(nπ*)/S1(ππ*La) continue with ultrafast decay processes to give a final distribution of quantum yields as follows: 42% stay on the dark S1(nπ*) state, 43.3% go back to the ground S0 state, 12% undergo a ring-opening reaction to the Z-form S0(Z) state, and 2.7% go to the E-form S0(E) state. The lifetimes of the excited states are estimated as follows: the S3 state is about 12 fs on average, the S2 state is about 80 fs, and the S1 state has a fast component of about 160 fs and a slow component of 15 ps. The simulated ultrafast radiationless deactivation pathways of photoexcited coumarin immediately interpret the experimentally observed weak fluorescence emission.
More Related Videos

Related Concept Videos
 01:25
Deactivation Processes: Jablonski Diagram
 01:12
Cycloaddition Reactions: MO Requirements for Photochemical Activation
 02:13
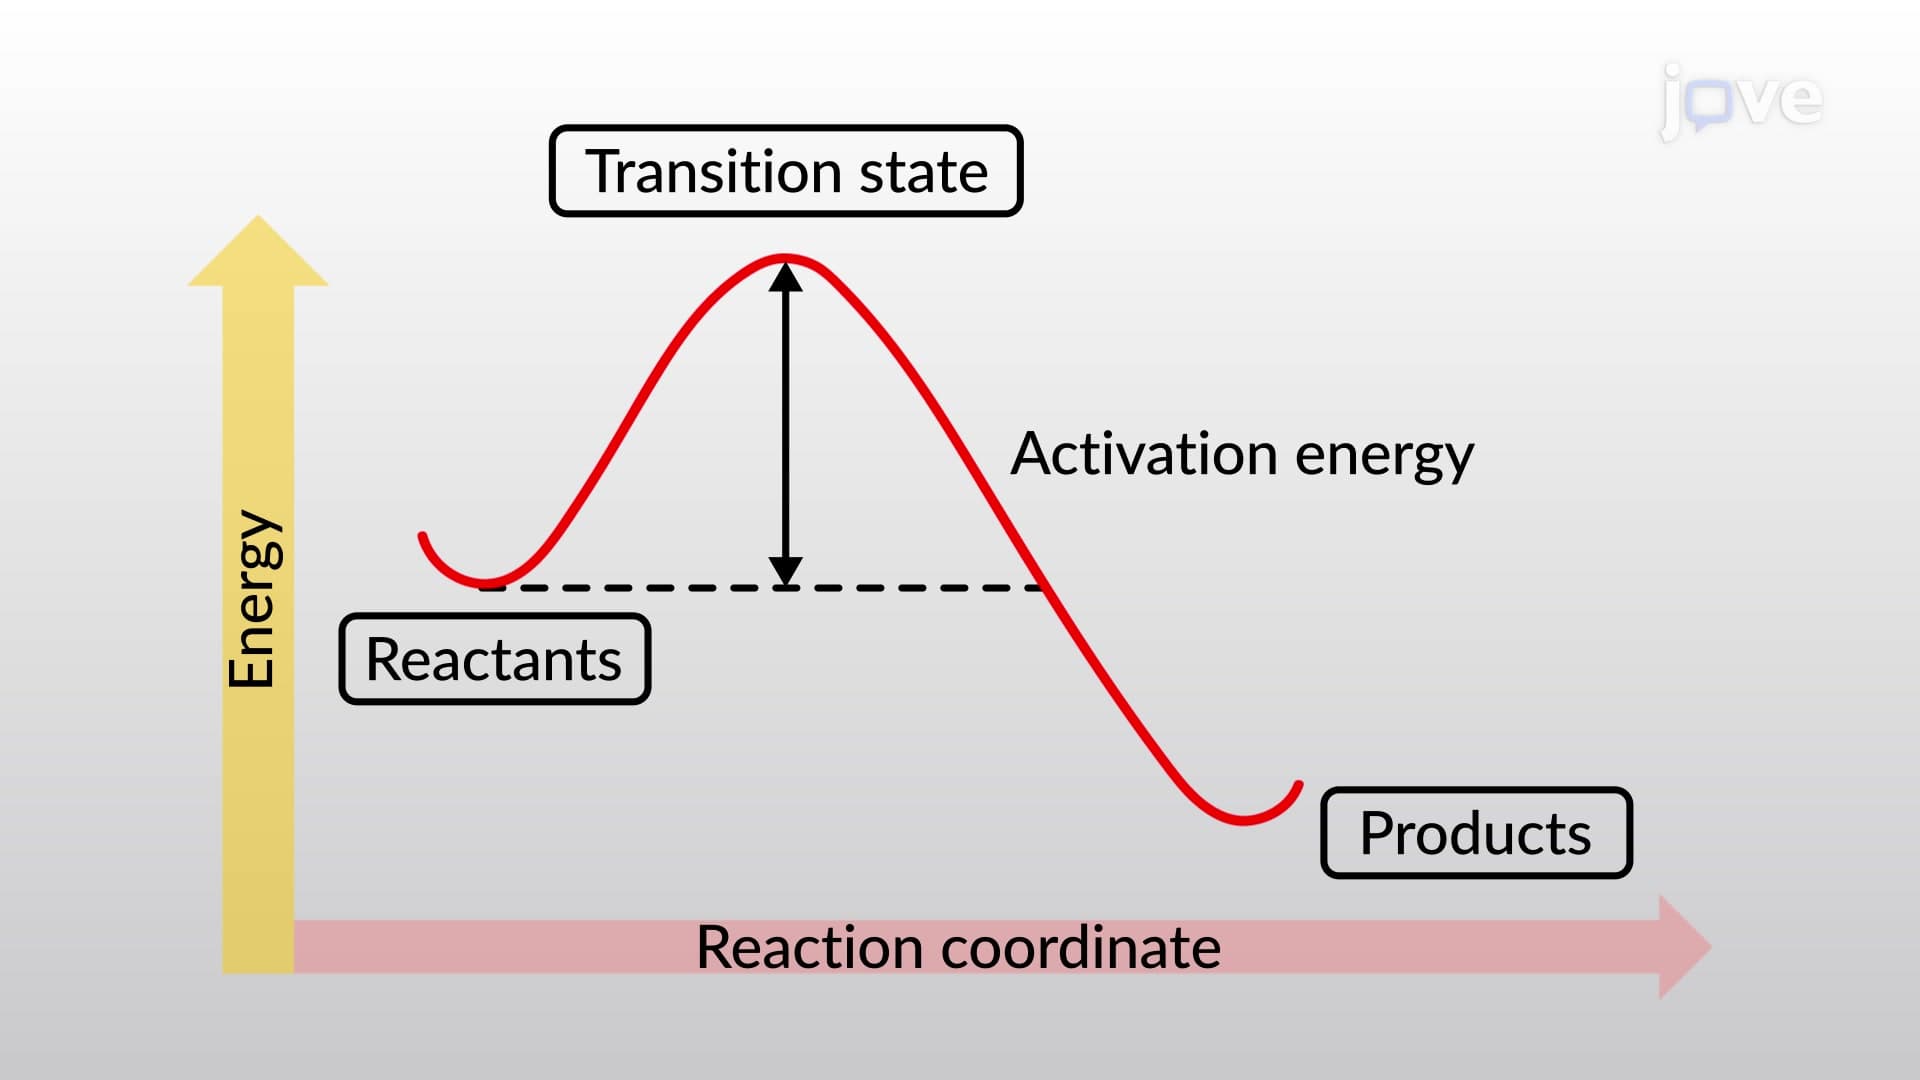
Bond Dissociation Energy and Activation Energy
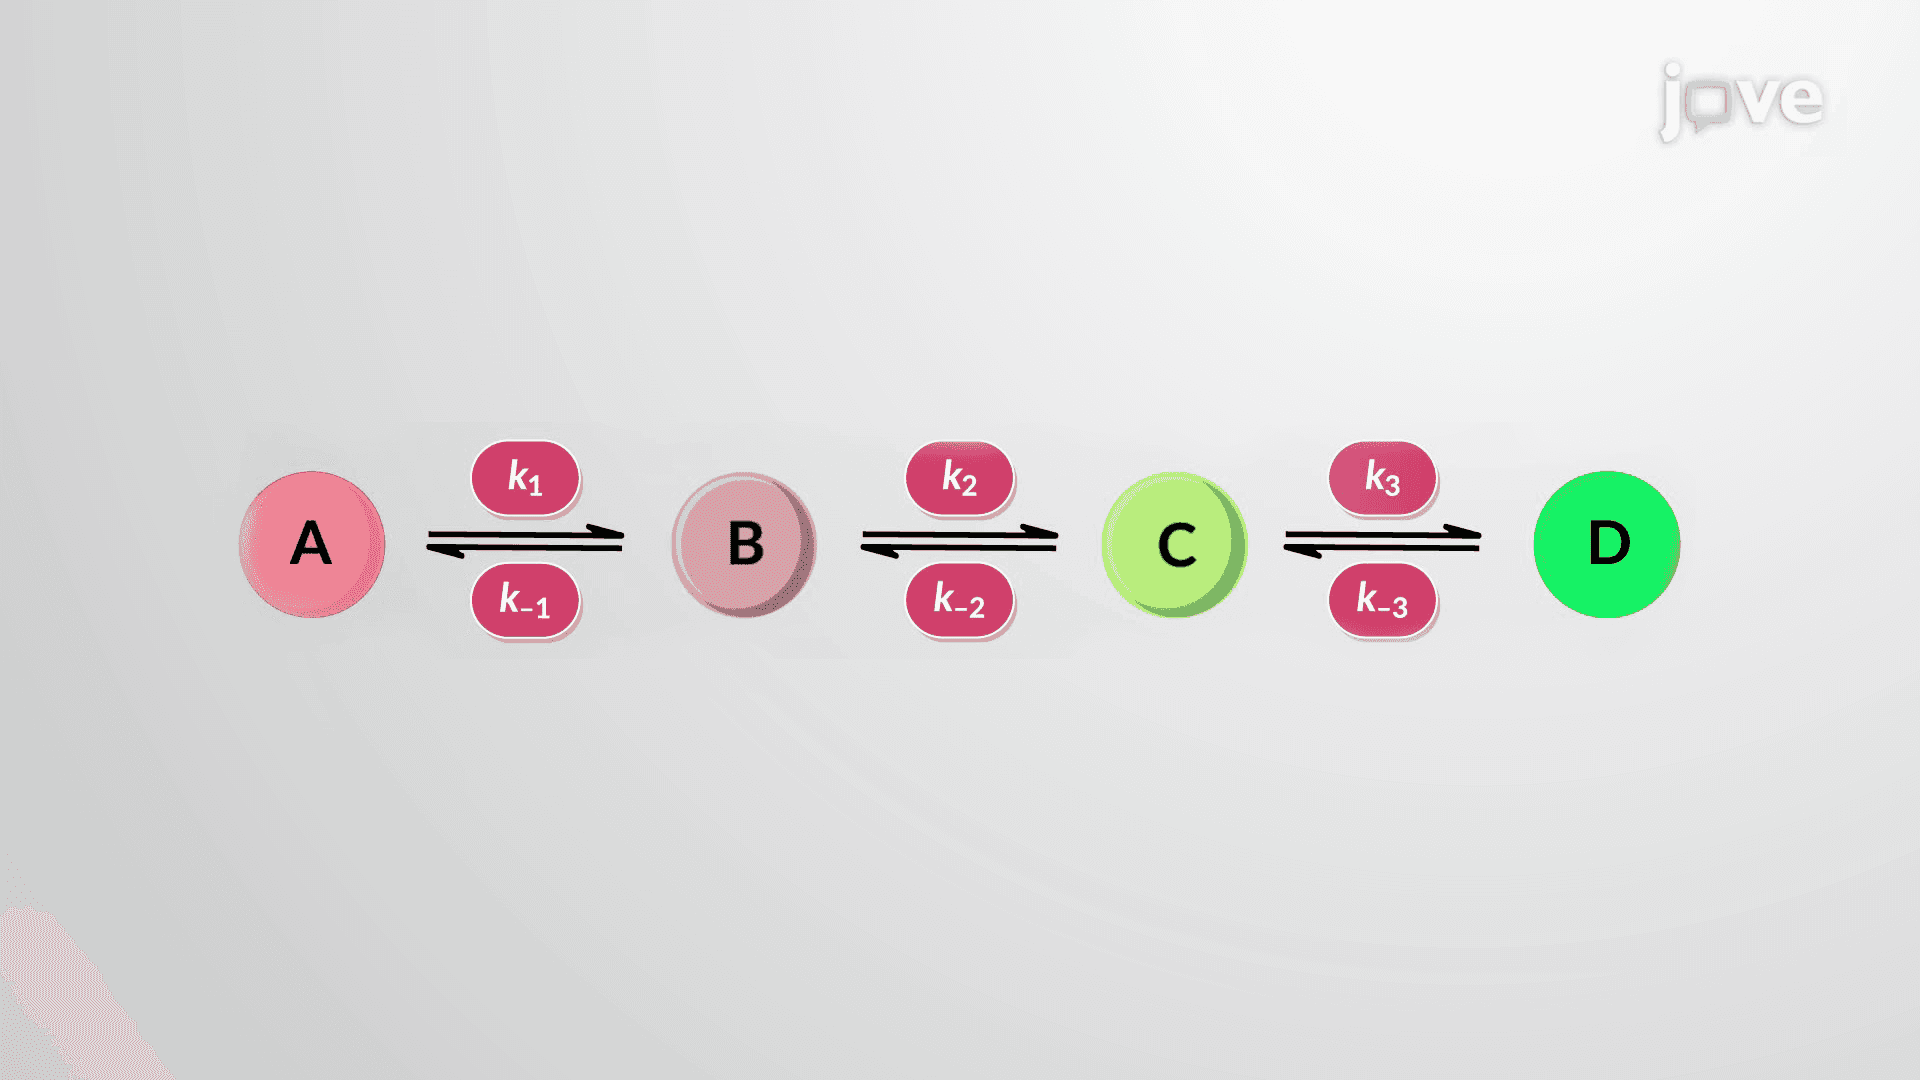 01:29
Reaction Mechanisms: Rate-limiting Step Approximation
 01:16
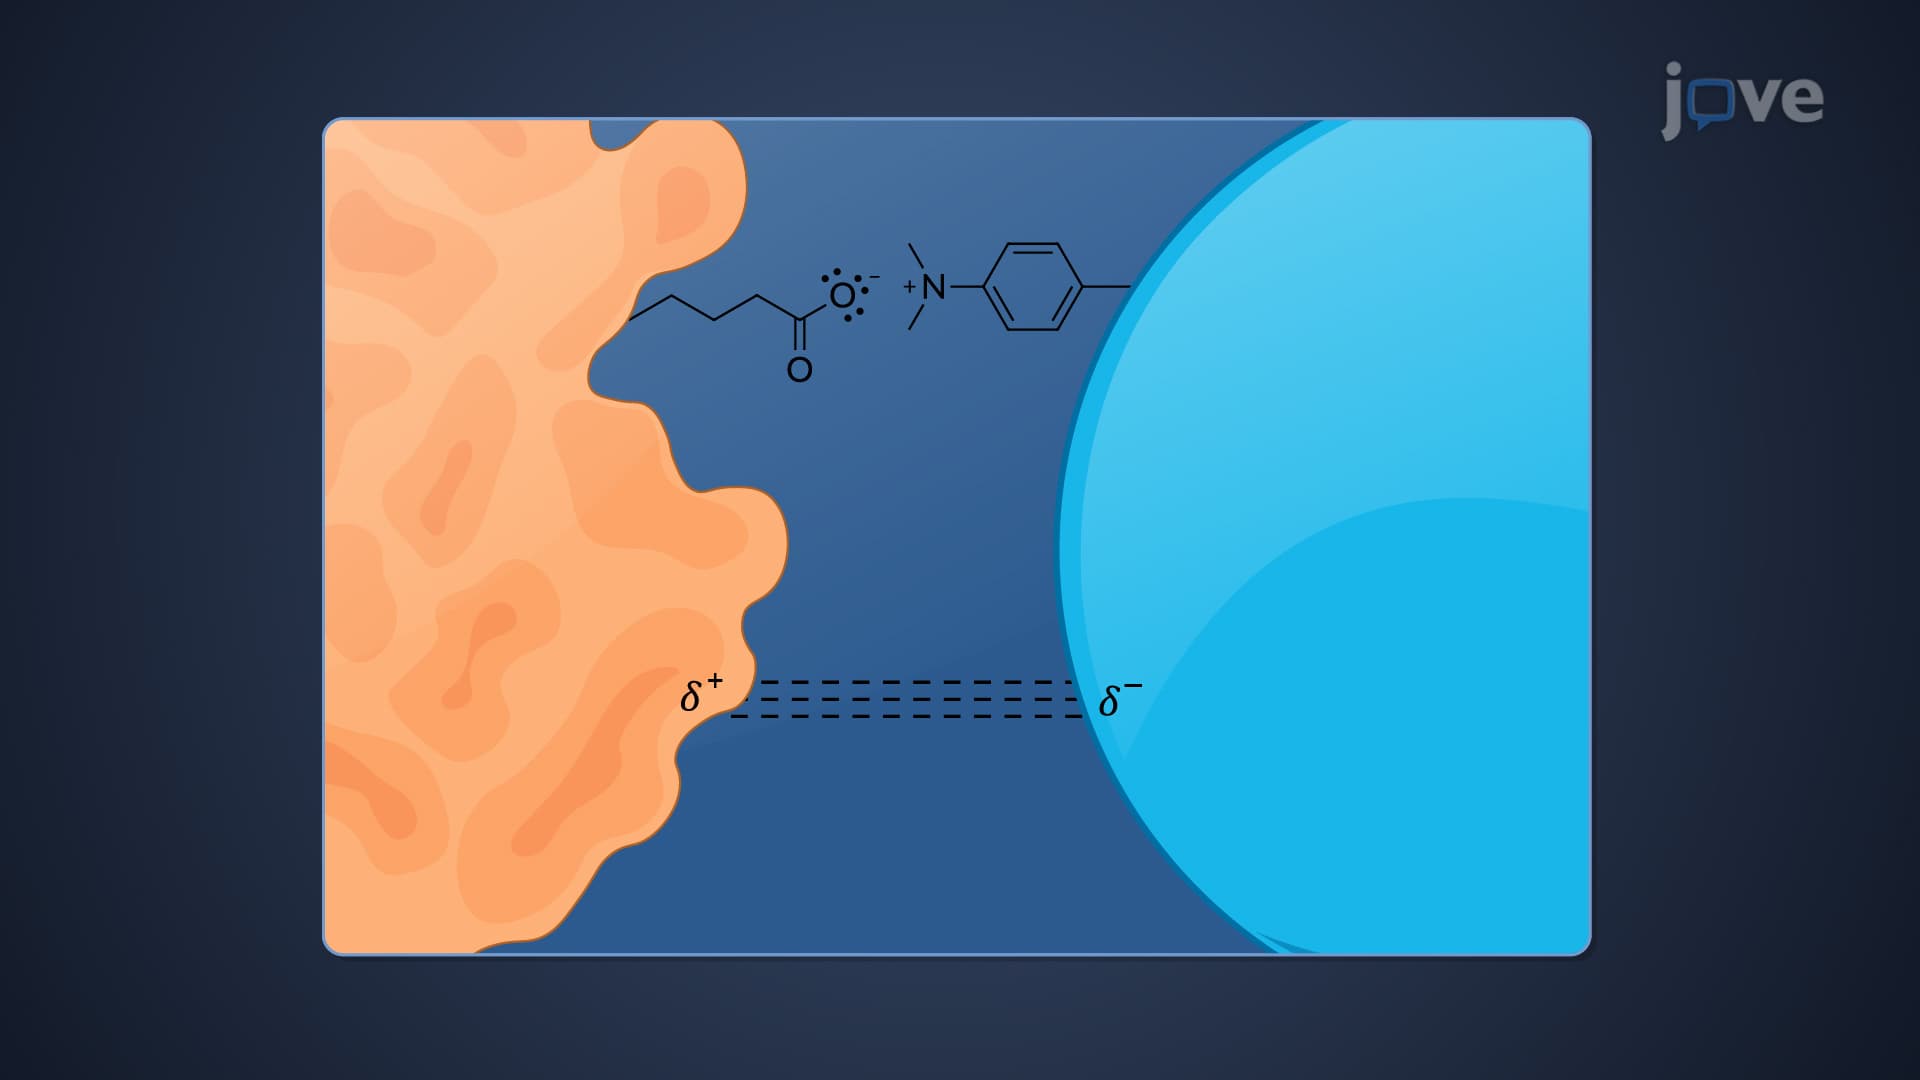
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:58
Cooperative Allosteric Transitions