Related Experiment Video
Updated: Mar 3, 2026

Evaluation of LC3-II Release via Extracellular Vesicles in Relation to the Accumulation of Intracellular LC3-positive Vesicles
Published on: October 18, 2024
Polyglutamine tracts regulate beclin 1-dependent autophagy
Avraham Ashkenazi1, Carla F Bento1, Thomas Ricketts1
1Department of Medical Genetics, Cambridge Institute for Medical Research (CIMR), University of Cambridge, Cambridge, UK.

Abstract:
Nine neurodegenerative diseases are caused by expanded polyglutamine (polyQ) tracts in different proteins, such as huntingtin in Huntington's disease and ataxin 3 in spinocerebellar ataxia type 3 (SCA3). Age at onset of disease decreases with increasing polyglutamine length in these proteins and the normal length also varies. PolyQ expansions drive pathogenesis in these diseases, as isolated polyQ tracts are toxic, and an N-terminal huntingtin fragment comprising exon 1, which occurs in vivo as a result of alternative splicing, causes toxicity. Although such mutant proteins are prone to aggregation, toxicity is also associated with soluble forms of the proteins. The function of the polyQ tracts in many normal cytoplasmic proteins is unclear. One such protein is the deubiquitinating enzyme ataxin 3 (refs 7, 8), which is widely expressed in the brain. Here we show that the polyQ domain enables wild-type ataxin 3 to interact with beclin 1, a key initiator of autophagy. This interaction allows the deubiquitinase activity of ataxin 3 to protect beclin 1 from proteasome-mediated degradation and thereby enables autophagy. Starvation-induced autophagy, which is regulated by beclin 1, was particularly inhibited in ataxin-3-depleted human cell lines and mouse primary neurons, and in vivo in mice. This activity of ataxin 3 and its polyQ-mediated interaction with beclin 1 was competed for by other soluble proteins with polyQ tracts in a length-dependent fashion. This competition resulted in impairment of starvation-induced autophagy in cells expressing mutant huntingtin exon 1, and this impairment was recapitulated in the brains of a mouse model of Huntington's disease and in cells from patients. A similar phenomenon was also seen with other polyQ disease proteins, including mutant ataxin 3 itself. Our data thus describe a specific function for a wild-type polyQ tract that is abrogated by a competing longer polyQ mutation in a disease protein, and identify a deleterious function of such mutations distinct from their propensity to aggregate.
More Related Videos
Related Concept Videos
 01:36
Delivery Pathways to the Lysosome
Endocytosis
In endocytosis, the cell membrane takes up macromolecules and particles from the surrounding medium. Clathrin-mediated...
 01:27
Autophagy
An autophagic pathway consists of a series of signaling events activated in response to diverse stress and physiological conditions such as food deprivation,...
 01:18
Autophagic Cell Death
Autophagy and Apoptosis
Autophagy can activate apoptosis. In normal conditions, the autophagy activating protein Beclin-1 and...
 02:58
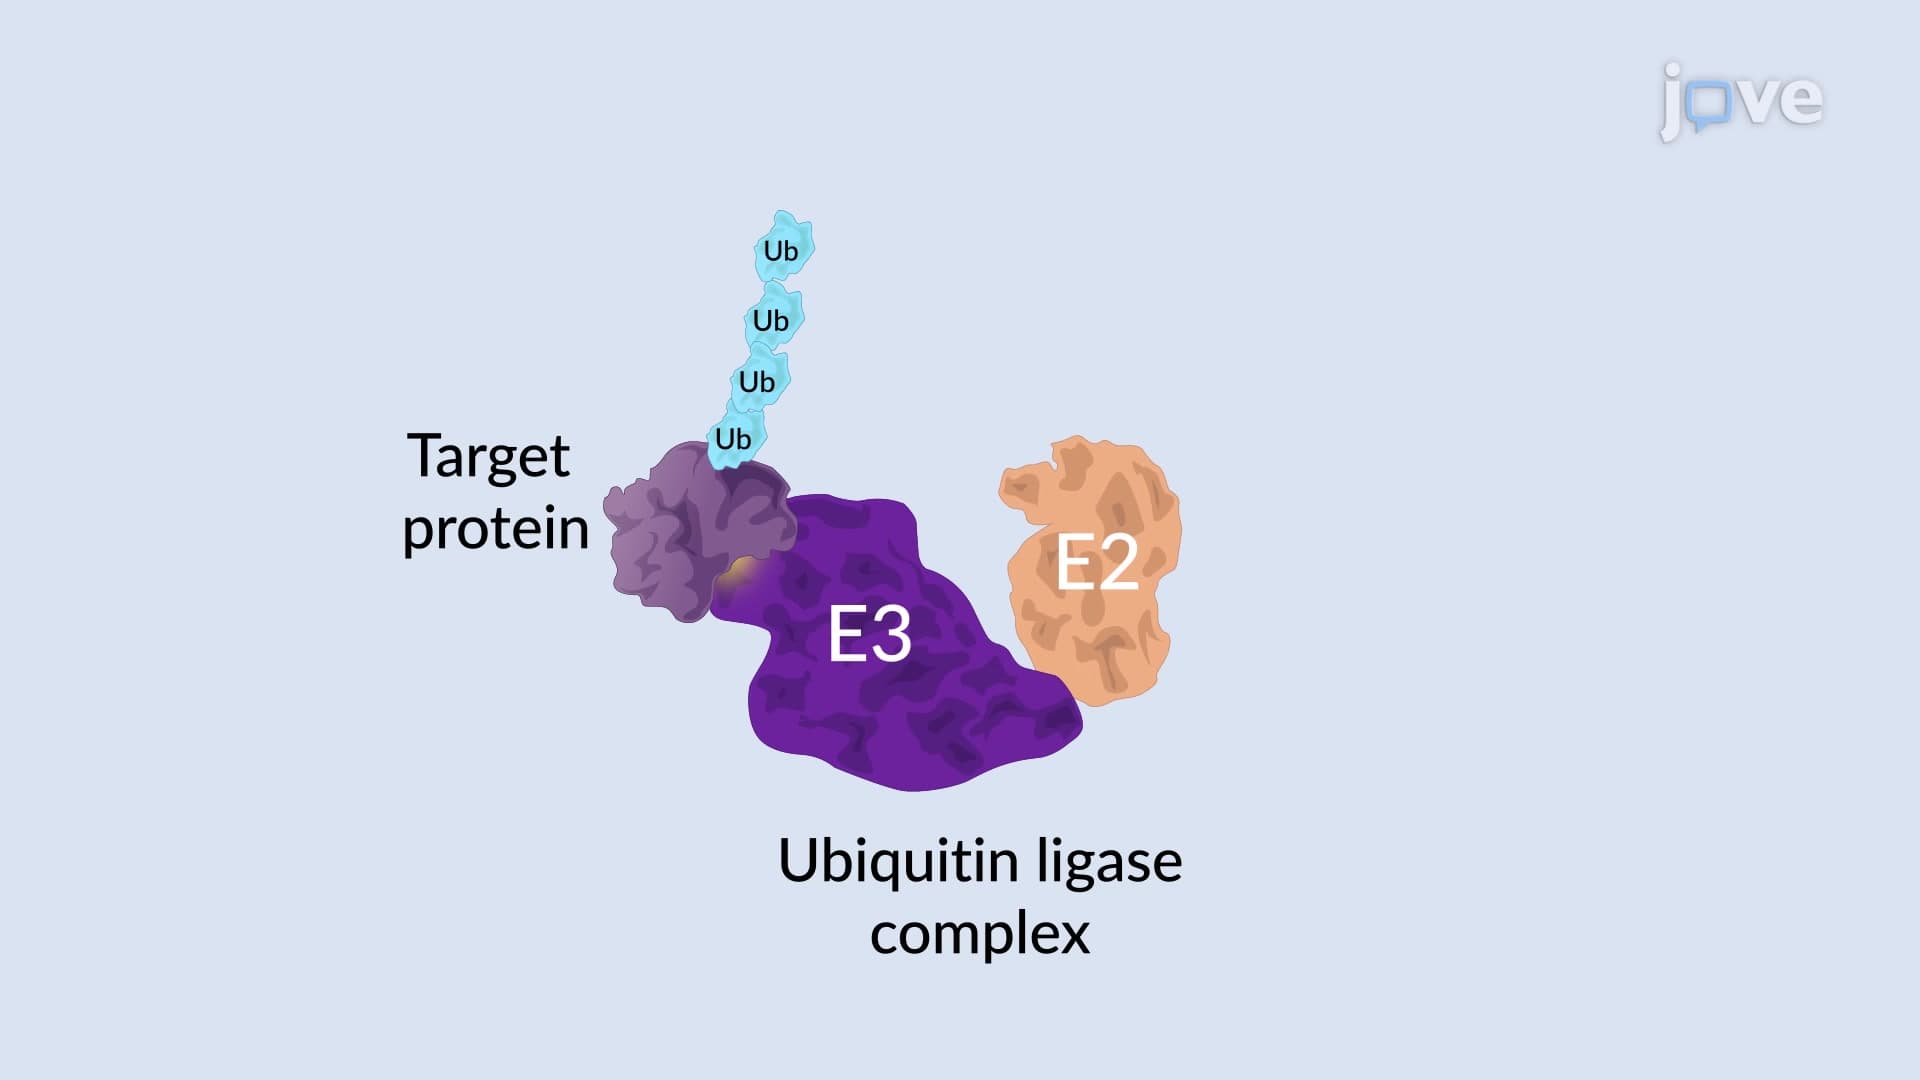
Regulated Protein Degradation
Protein degradation plays two important roles in the cells. It helps to protect cells from misfolded or damaged proteins before they lead to a...
02:58Regulated Protein Degradation
 01:13
The Proteasome
In this pathway, the target proteins are first tagged with small proteins called ubiquitin. This involves participation of a series of enzymes including— E1 (ubiquitin-activating enzyme), E2 (ubiquitin-conjugating enzyme), and E3...