Related Experiment Video
Updated: Feb 27, 2026

Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
Using force-based adaptive resolution simulations to calculate solvation free energies of amino acid sidechain
Raffaele Fiorentini1, Kurt Kremer1, Raffaello Potestio1
1Max Planck Institute for Polymer Research, Ackermannweg 10, 55128 Mainz, Germany.

Abstract:
The calculation of free energy differences is a crucial step in the characterization and understanding of the physical properties of biological molecules. In the development of efficient methods to compute these quantities, a promising strategy is that of employing a dual-resolution representation of the solvent, specifically using an accurate model in the proximity of a molecule of interest and a simplified description elsewhere. One such concurrent multi-resolution simulation method is the Adaptive Resolution Scheme (AdResS), in which particles smoothly change their resolution on-the-fly as they move between different subregions. Before using this approach in the context of free energy calculations, however, it is necessary to make sure that the dual-resolution treatment of the solvent does not cause undesired effects on the computed quantities. Here, we show how AdResS can be used to calculate solvation free energies of small polar solutes using Thermodynamic Integration (TI). We discuss how the potential-energy-based TI approach combines with the force-based AdResS methodology, in which no global Hamiltonian is defined. The AdResS free energy values agree with those calculated from fully atomistic simulations to within a fraction of kBT. This is true even for small atomistic regions whose size is on the order of the correlation length, or when the properties of the coarse-grained region are extremely different from those of the atomistic region. These accurate free energy calculations are possible because AdResS allows the sampling of solvation shell configurations which are equivalent to those of fully atomistic simulations. The results of the present work thus demonstrate the viability of the use of adaptive resolution simulation methods to perform free energy calculations and pave the way for large-scale applications where a substantial computational gain can be attained.
More Related Videos
 05:57
Author Spotlight: In Silico Creation and Impact of Carbonylated Amino Acids on Protein Structure and Function
Published on: April 26, 2024
 05:08
Solubility of Hydrophobic Compounds in Aqueous Solution Using Combinations of Self-assembling Peptide and Amino Acid
Published on: September 20, 2017
Related Concept Videos
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 02:12
Solvating Effects
 02:05
Entropy and Solvation
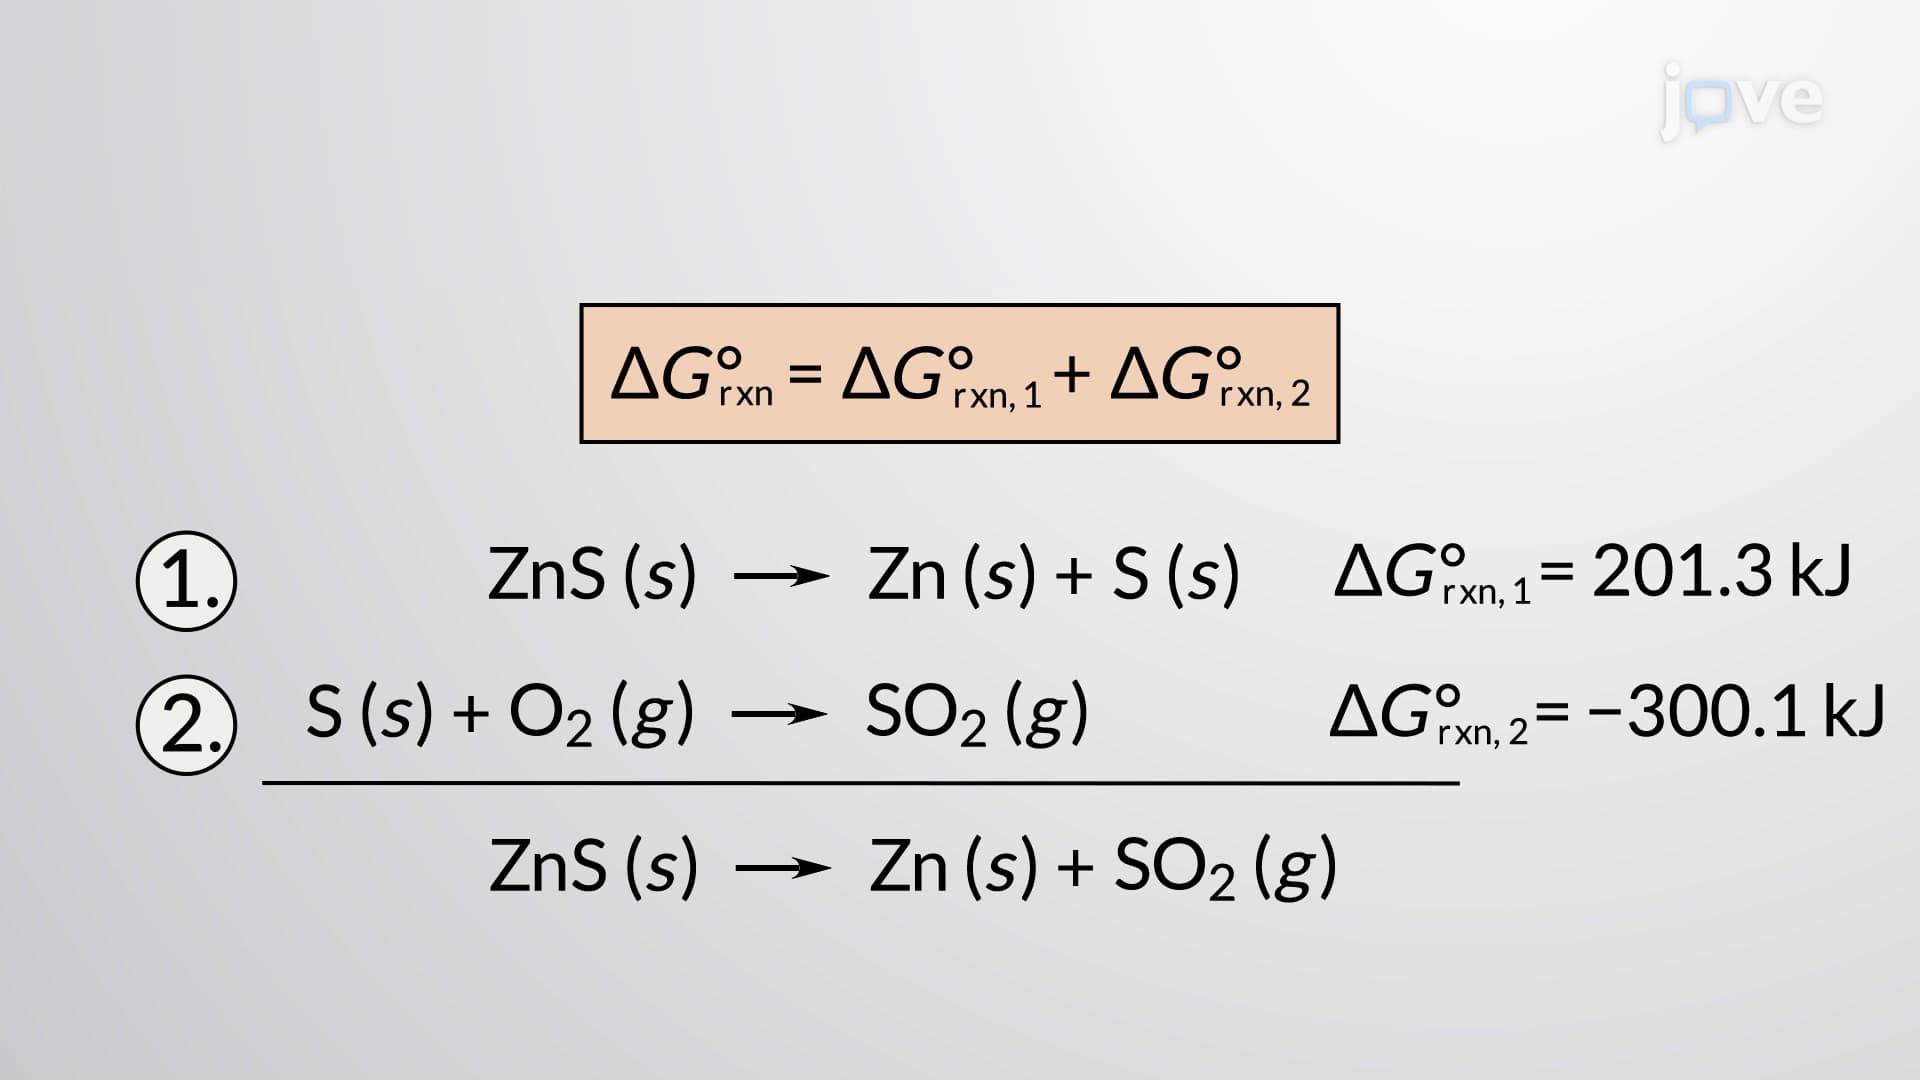 02:49
Calculating Standard Free Energy Changes
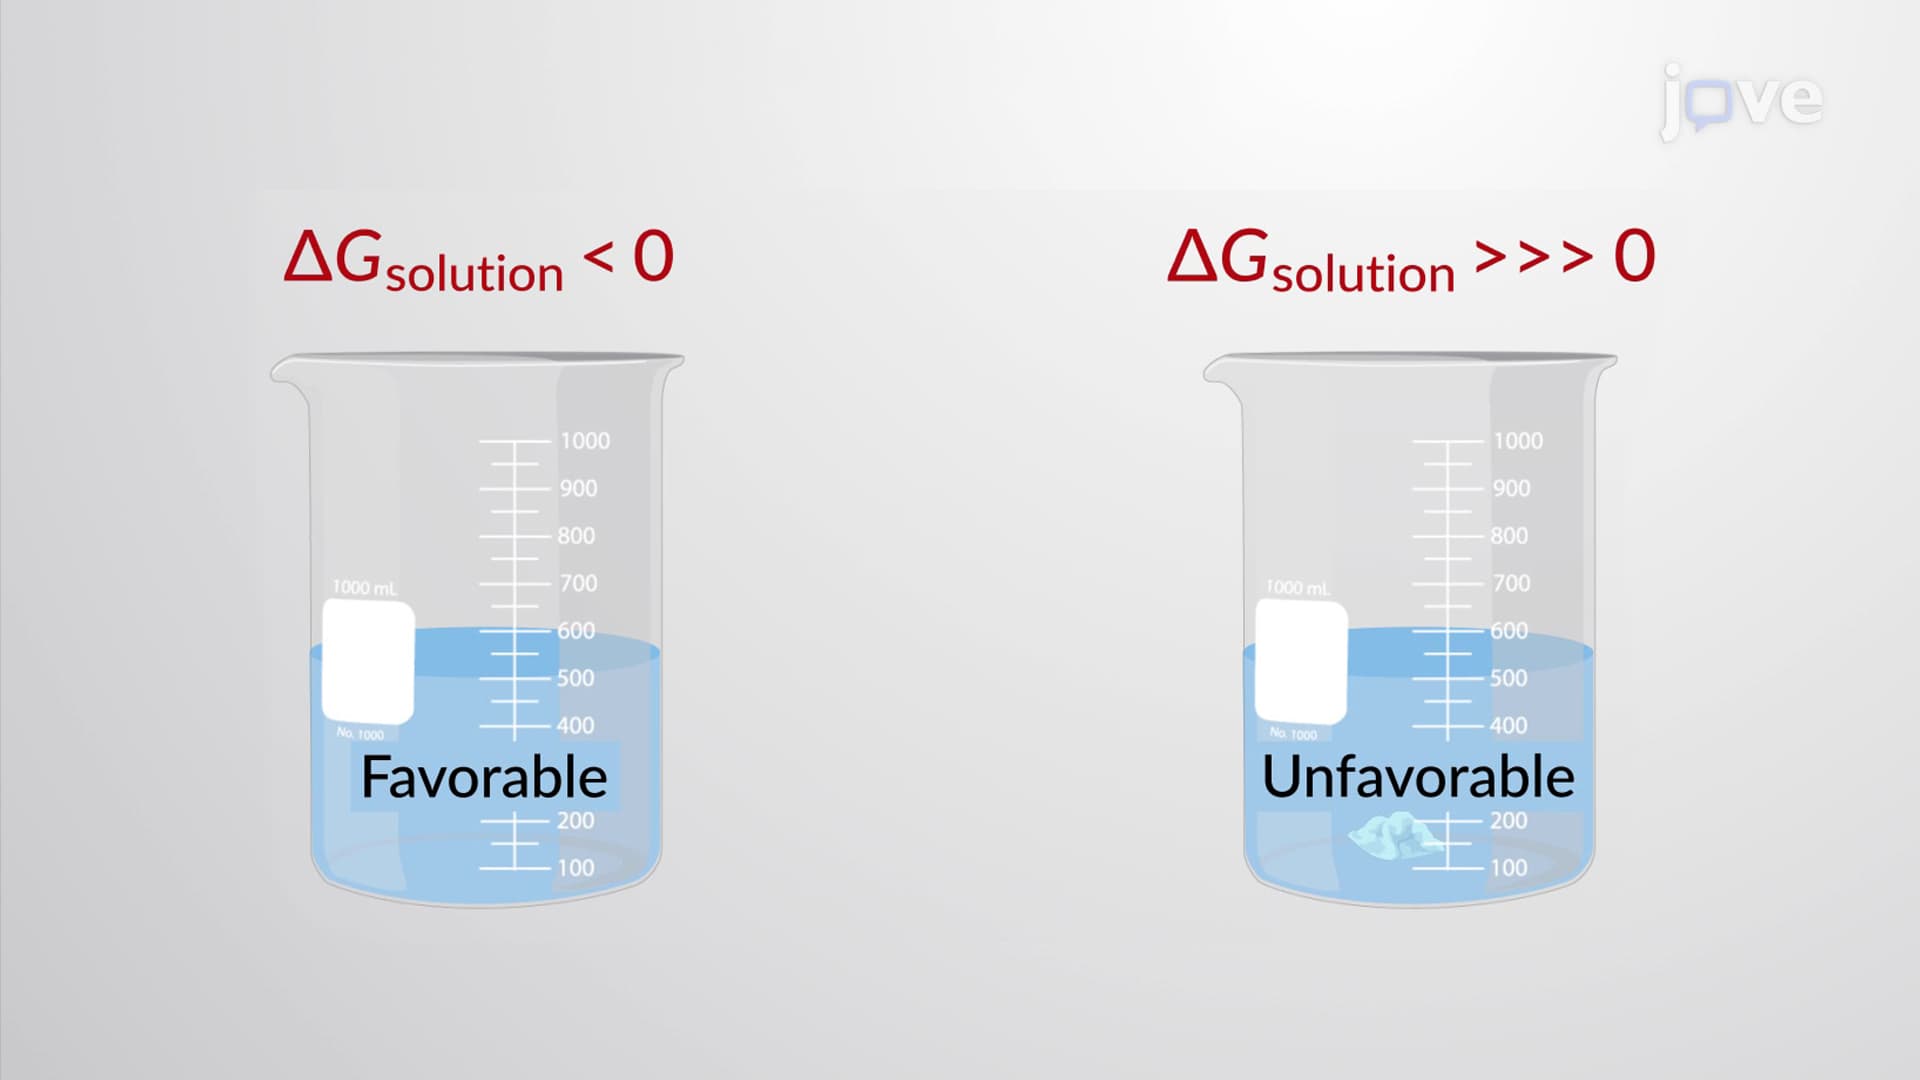 02:21
Chemical and Solubility Equilibria