Evaluating amber force fields using computed NMR chemical shifts
Related Experiment Video
Updated: Feb 27, 2026
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
1Computational and Systems Biology, University of Pittsburgh, Pittsburgh, Pennsylvania, 15260.

Abstract:
NMR chemical shifts can be computed from molecular dynamics (MD) simulations using a template matching approach and a library of conformers containing chemical shifts generated from ab initio quantum calculations. This approach has potential utility for evaluating the force fields that underlie these simulations. Imperfections in force fields generate flawed atomic coordinates. Chemical shifts obtained from flawed coordinates have errors that can be traced back to these imperfections. We use this approach to evaluate a series of AMBER force fields that have been refined over the course of two decades (ff94, ff96, ff99SB, ff14SB, ff14ipq, and ff15ipq). For each force field a series of MD simulations are carried out for eight model proteins. The calculated chemical shifts for the 1 H, 15 N, and 13 Ca atoms are compared with experimental values. Initial evaluations are based on root mean squared (RMS) errors at the protein level. These results are further refined based on secondary structure and the types of atoms involved in nonbonded interactions. The best chemical shift for identifying force field differences is the shift associated with peptide protons. Examination of the model proteins on a residue by residue basis reveals that force field performance is highly dependent on residue position. Examination of the time course of nonbonded interactions at these sites provides explanations for chemical shift differences at the atomic coordinate level. Results show that the newer ff14ipq and ff15ipq force fields developed with the implicitly polarized charge method perform better than the older force fields.
Related Concept Videos
 01:15
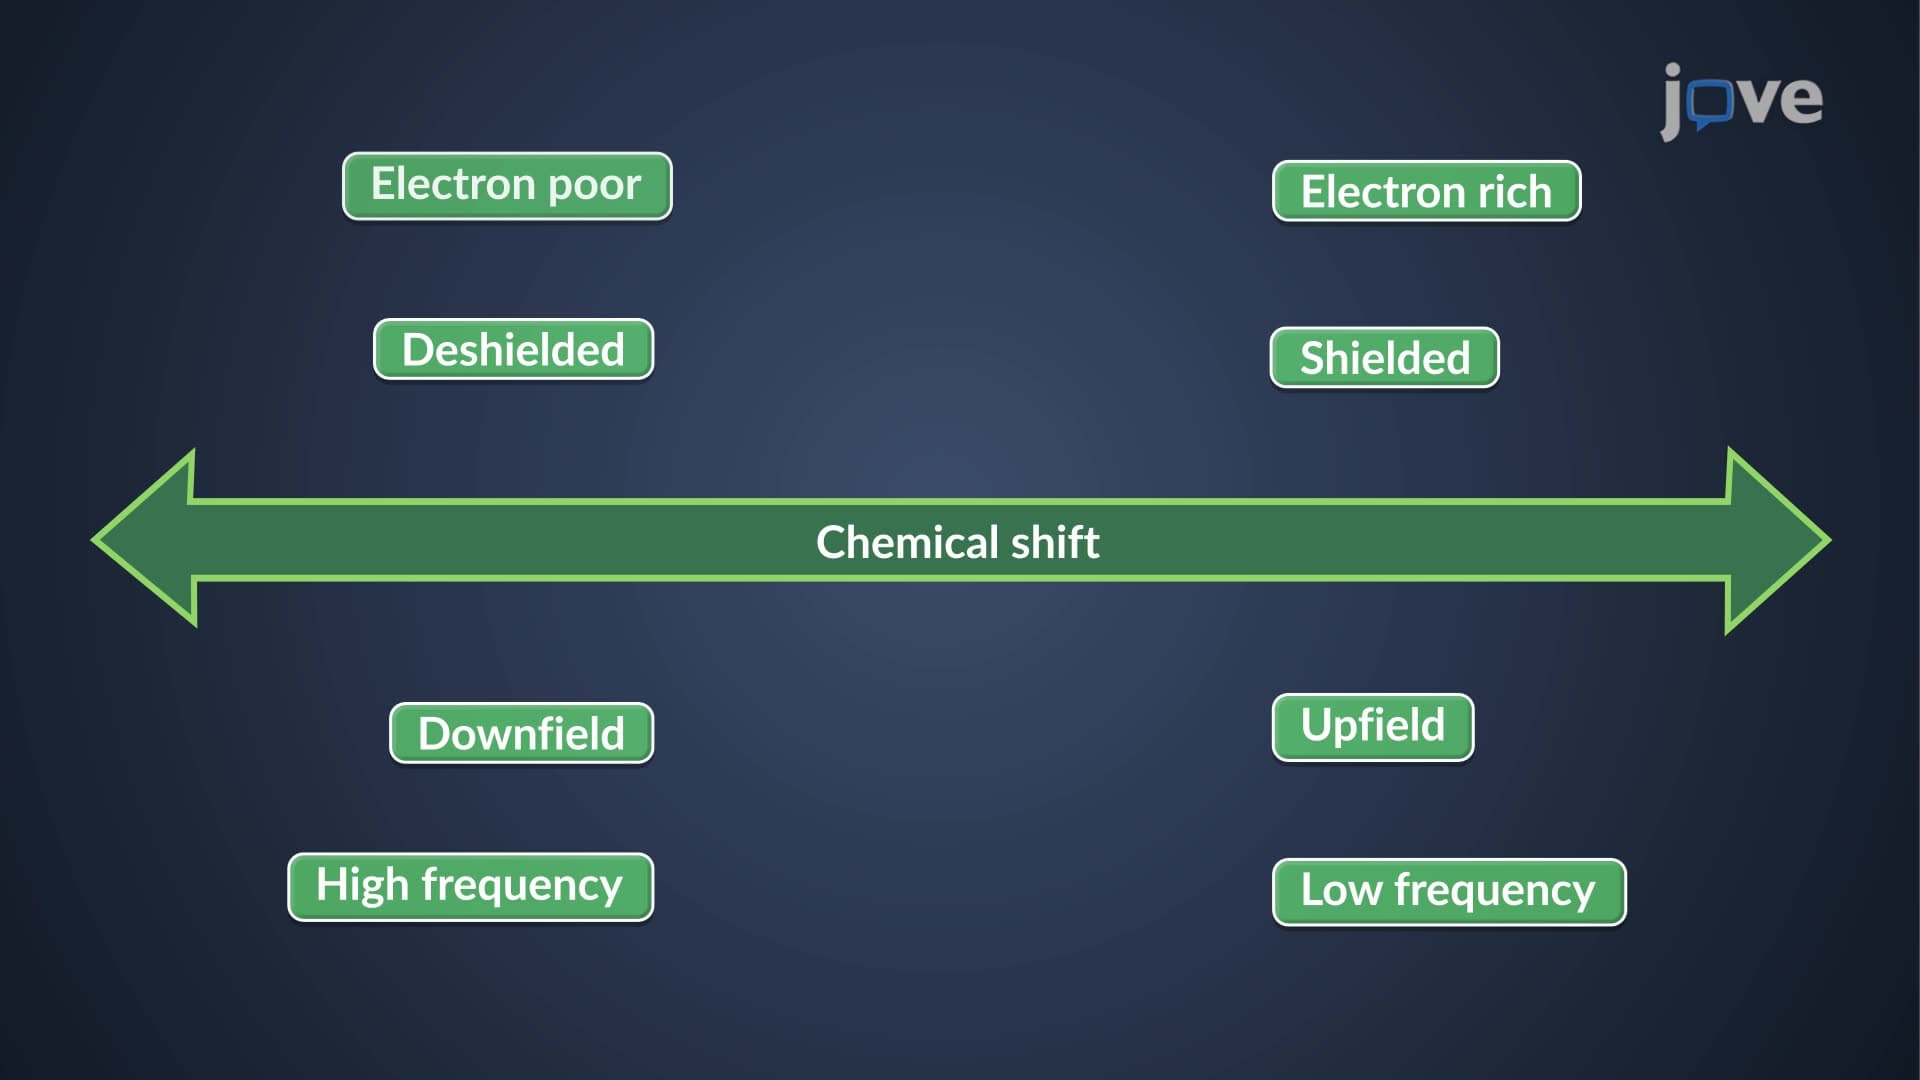
NMR Spectroscopy: Chemical Shift Overview
For instance, the proton...
 01:19
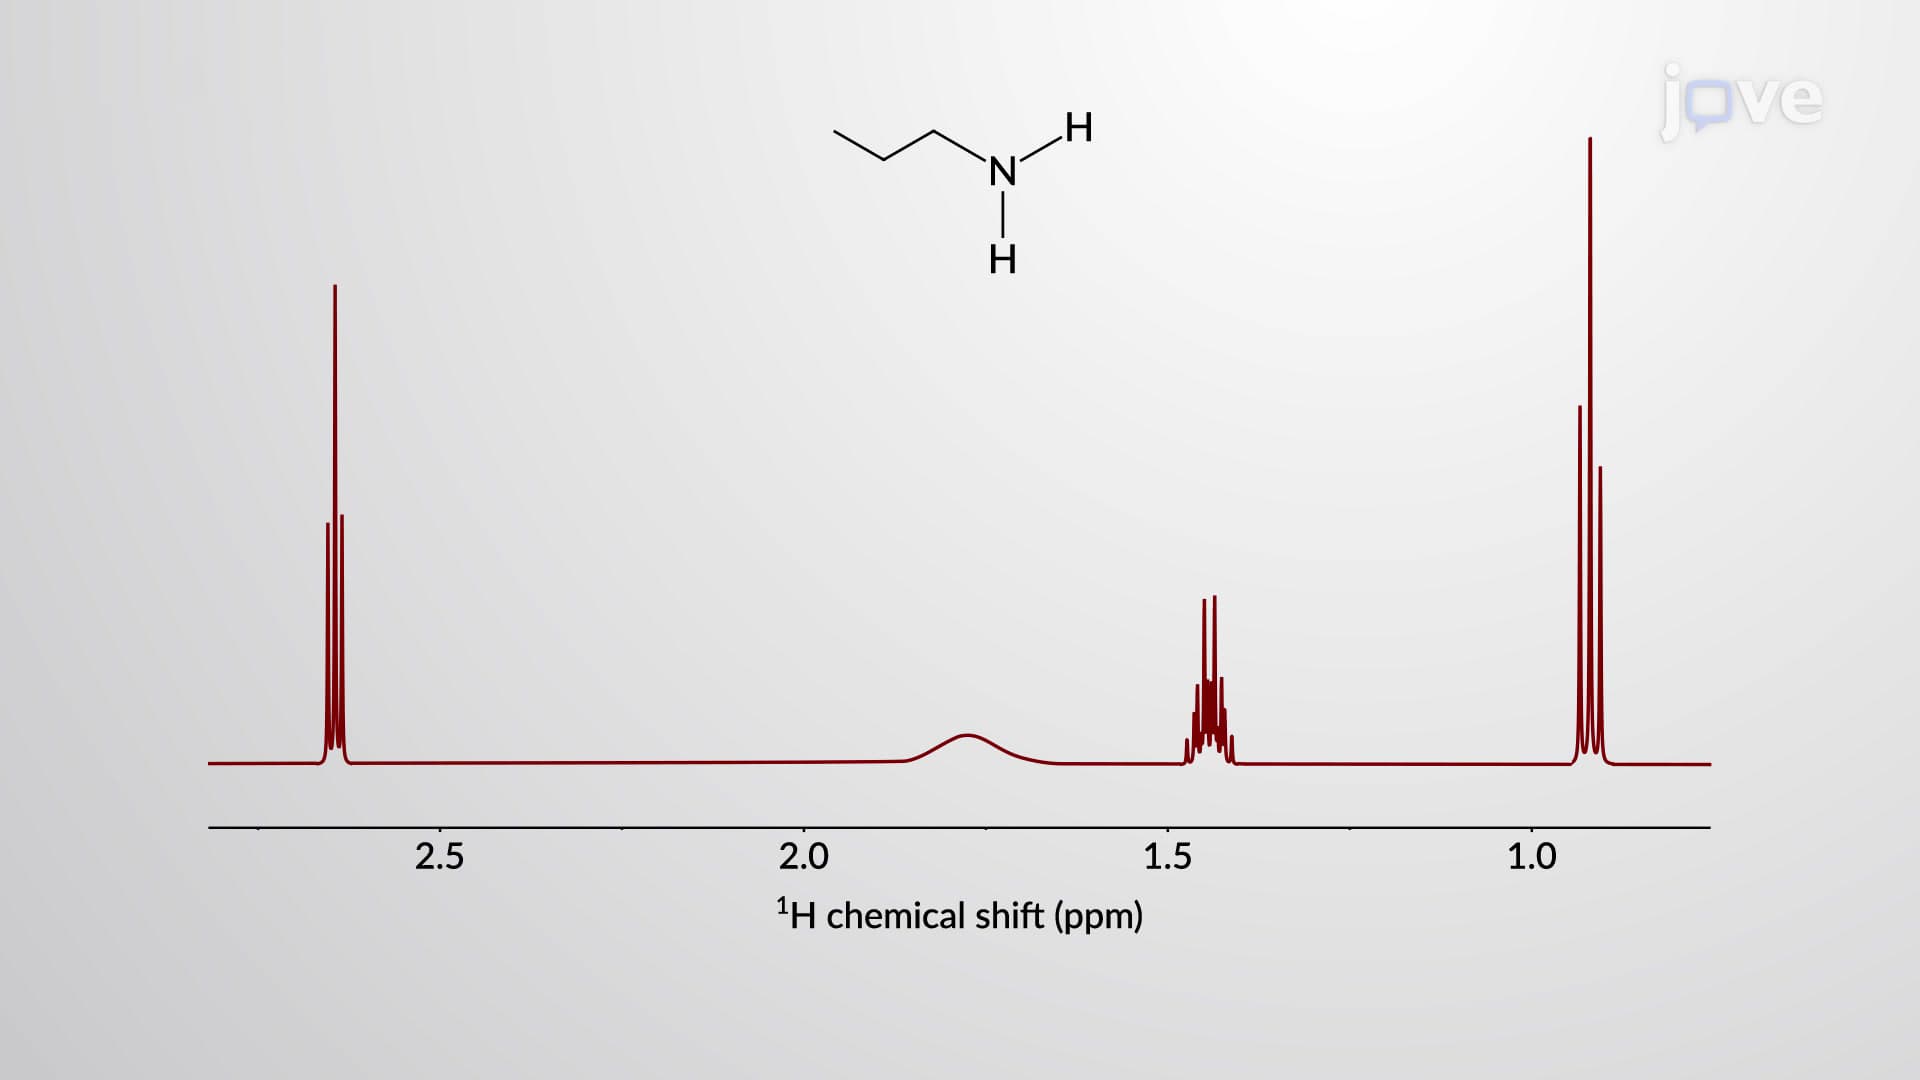
NMR Spectroscopy Of Amines
 01:02
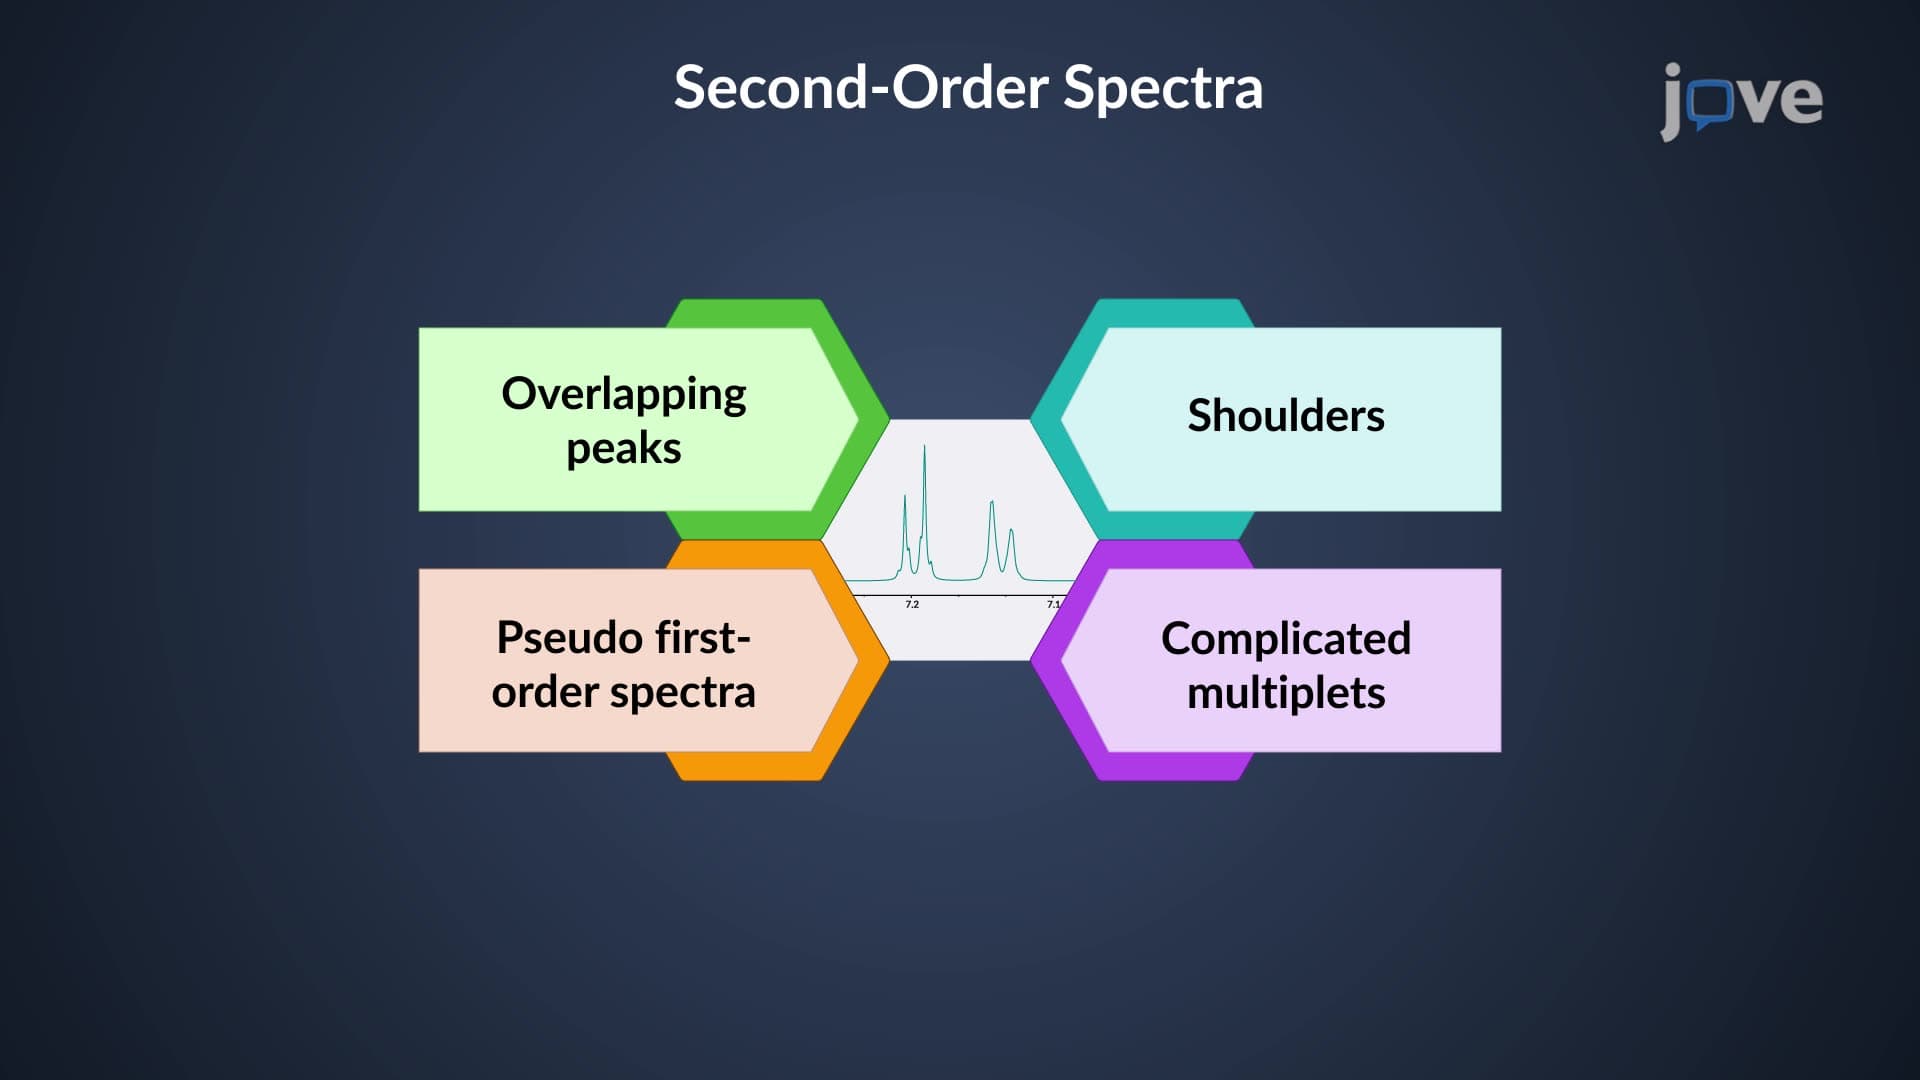
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:07
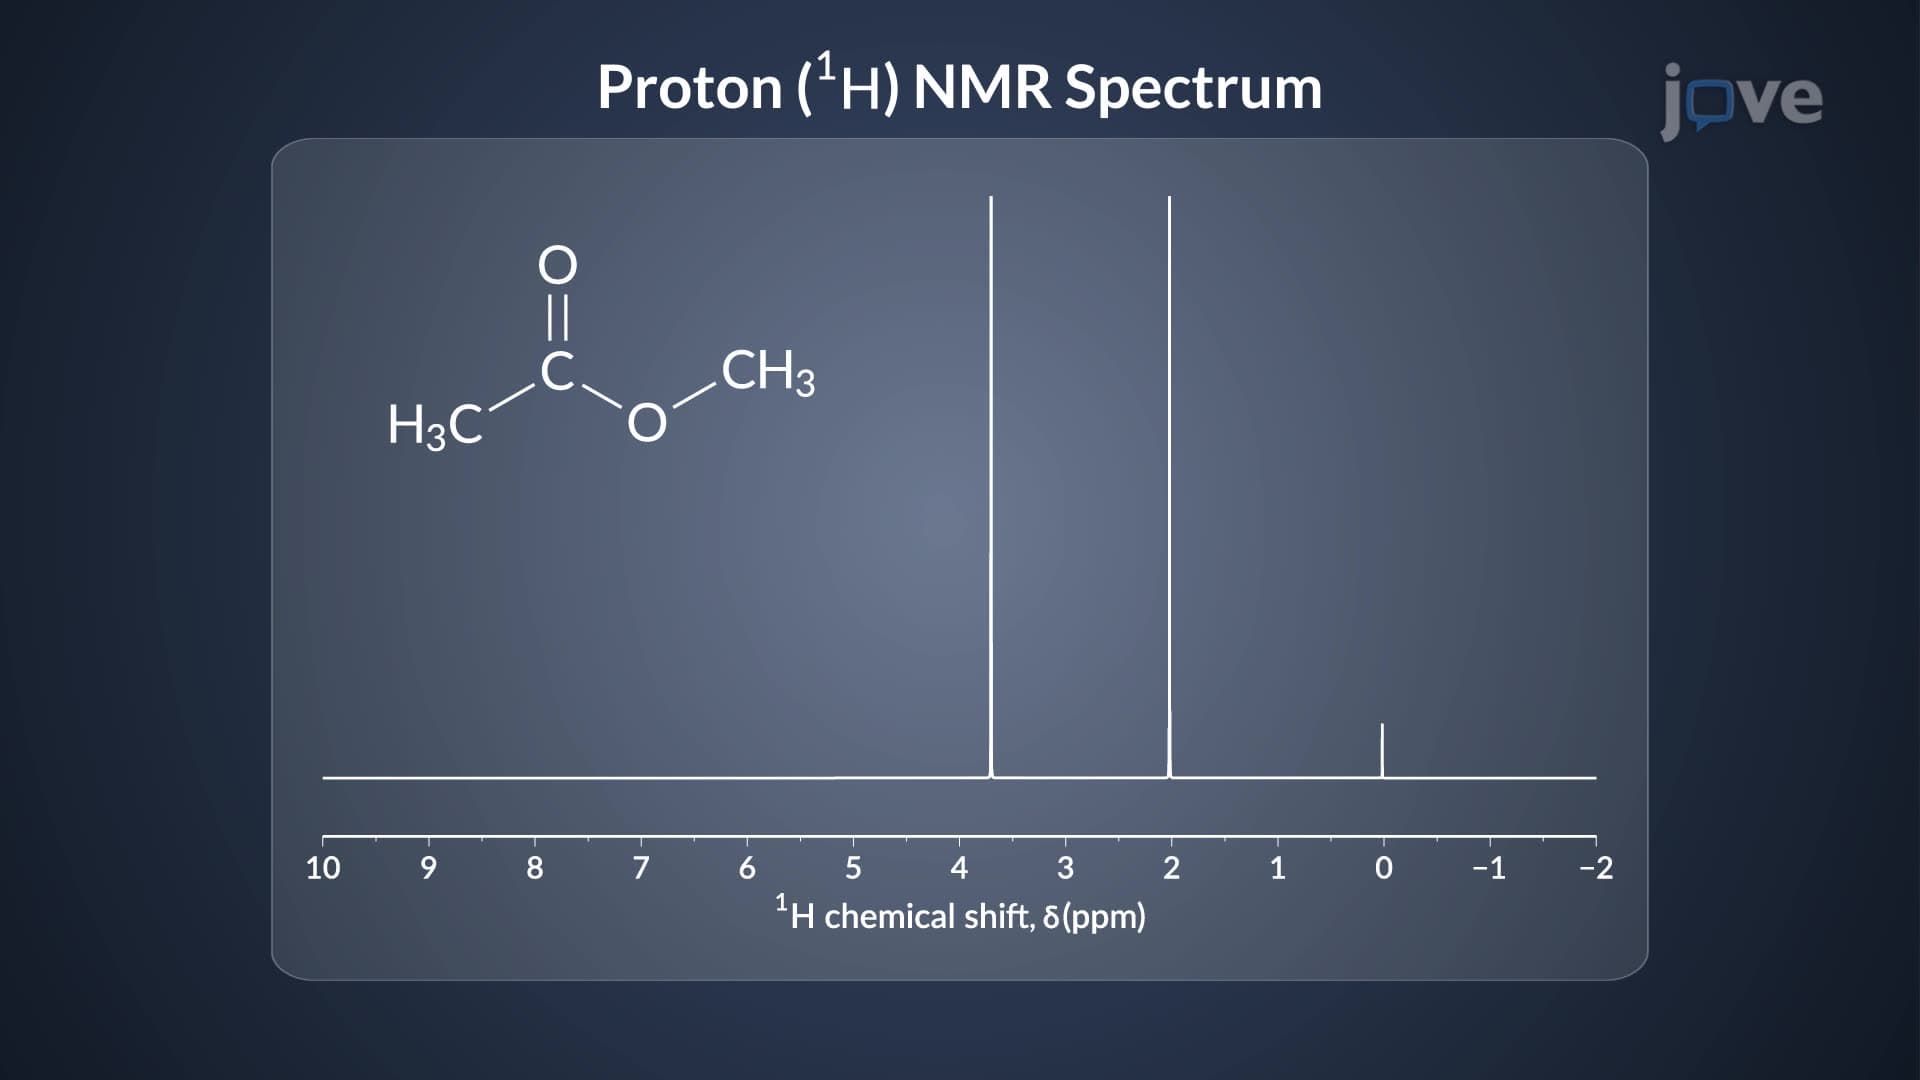
Proton (¹H) NMR: Chemical Shift
Absorption signals of all the protium nuclei...
 01:10
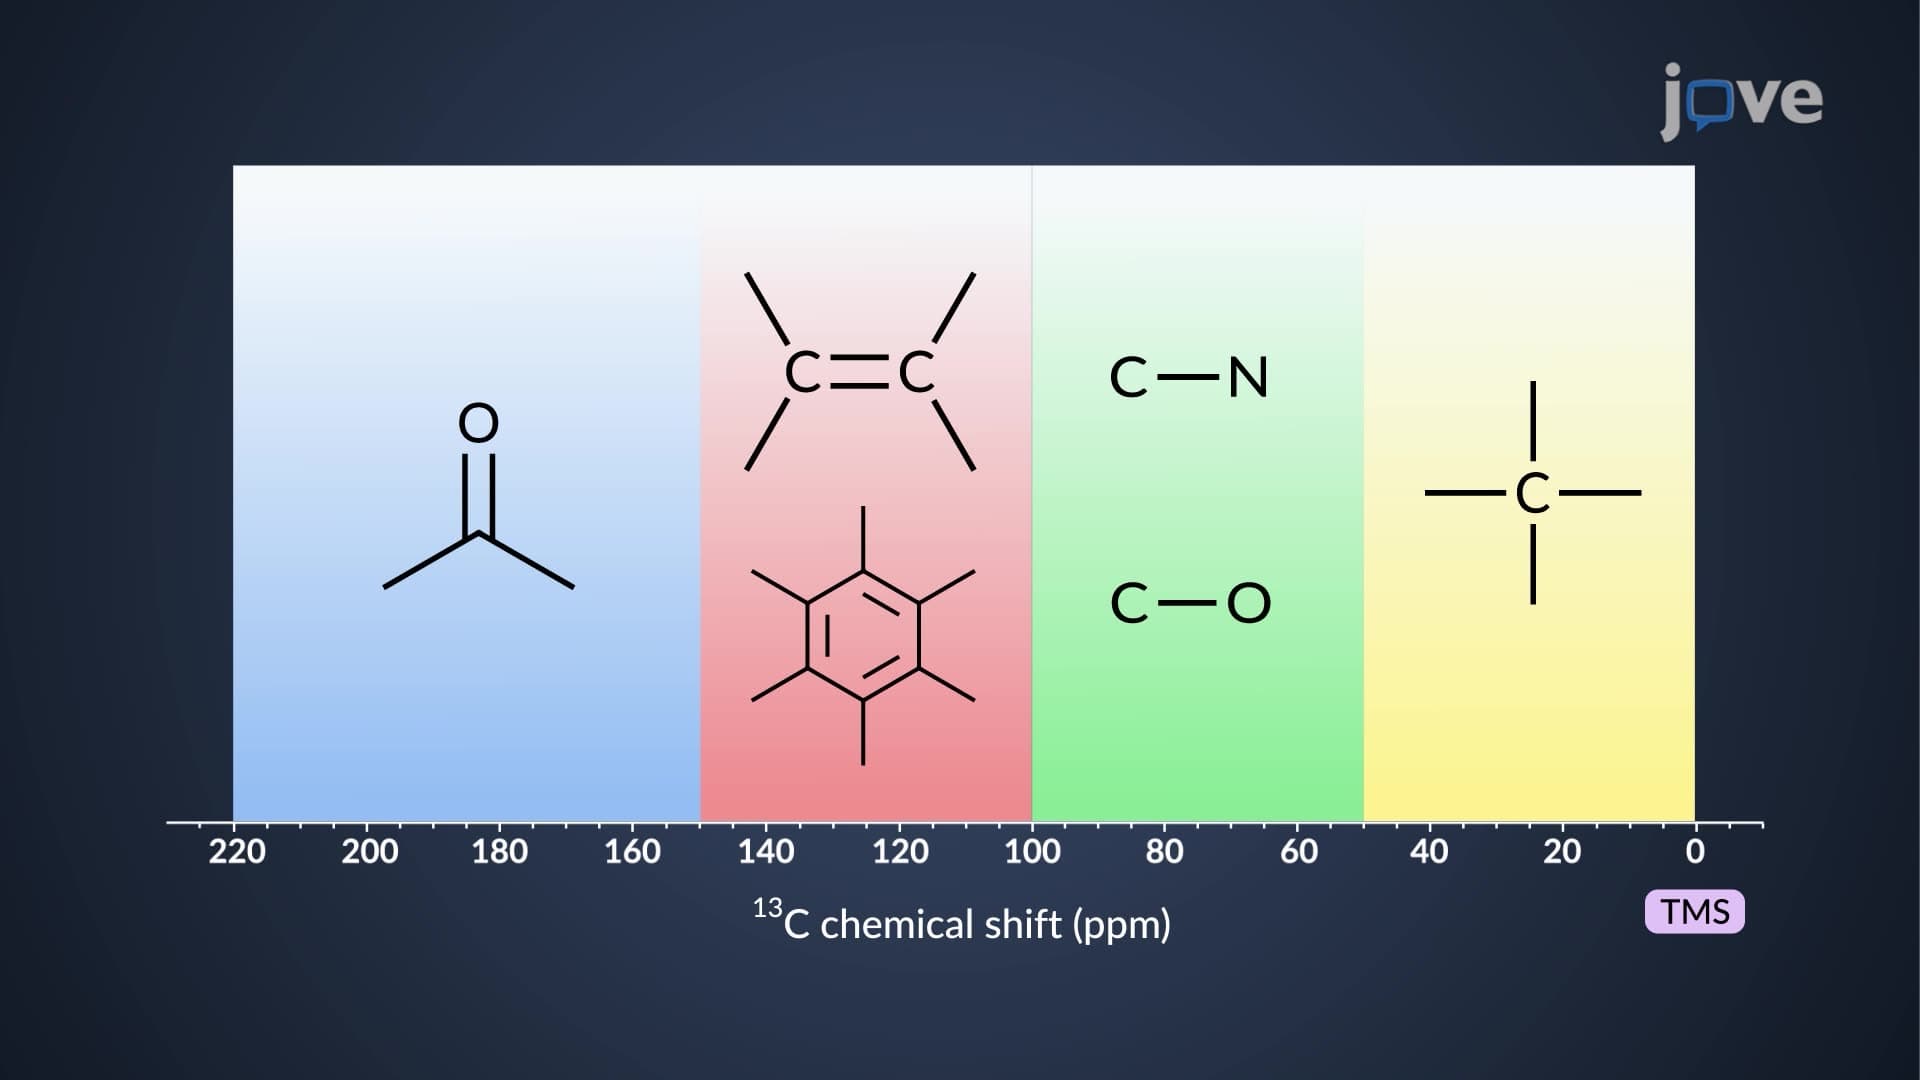
Carbon-13 (¹³C) NMR: Overview
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...